Related Experiment Video
Updated: Jun 30, 2025


Technical Aspect of the Automated Synthesis and Real-Time Kinetic Evaluation of [11C]SNAP-7941
Published on: April 28, 2019
Pharmacology and pharmacokinetics of tazemetostat
Marco Orleni1,2,3, Jan H Beumer4,5,6
1Cancer Therapeutics Program, UPMC Hillman Cancer Center, Room G27E, Hillman Research Pavilion, 5117 Centre Avenue, Pittsburgh, PA, 15213-1863, USA.

Abstract:
Tazemetostat, a novel oral selective inhibitor of enhancer of zeste homolog 2 (EZH2), was approved by the Food and Drug Administration (FDA) in 2020 for use in patients with advanced epithelioid sarcoma or relapsed/refractory (R/R) EZH2-mutated follicular lymphoma. These indications were approved by the FDA trough accelerated approval based on objective response rate and duration of response that resulted from phase 2 clinical trials. Tazemetostat competes with S-adenosylmethionine (SAM) cofactor to inhibit EZH2, reducing the levels of trimethylated lysine 27 of histone 3 (H3K27me3), considered as pharmacodynamic marker. Tazemetostat is orally bioavailable, characterized by rapid absorption and dose-proportional exposure, which is not influenced by coadministration with food or gastric acid reducing agents. It highly distributes in tissues, but with limited access to central nervous system. Tazemetostat is metabolized by CYP3A in the liver to 3 major inactive metabolites (M1, M3, and M5), has a short half-life and is mainly excreted in feces. Drug-drug interactions were shown with moderate CYP3A inhibitors as fluconazole, leading the FDA to recommend a 50% dose reduction, while studies investigating coadministration of tazemetostat with strong inhibitors/inducers are ongoing. No dosage modifications are recommended based on renal or hepatic dysfunctions. Overall, tazemetostat is the first-in-class EZH2 inhibitor approved by the FDA for cancer treatment. Current clinical studies are evaluating combination therapies in patients with several malignancies.
Insights
Tazemetostat is a new FDA-approved EZH2 inhibitor for specific cancers. It works by blocking EZH2, a key protein, and shows promising results in clinical trials for advanced epithelioid sarcoma and follicular lymphoma.
Area of Science:
- Oncology
- Pharmacology
- Epigenetics
Background:
- Tazemetostat is a novel oral selective inhibitor of enhancer of zeste homolog 2 (EZH2).
- EZH2 inhibition is a therapeutic strategy for certain cancers, including epithelioid sarcoma and EZH2-mutated follicular lymphoma.
- FDA approval in 2020 was based on accelerated approval pathways utilizing objective response rate and duration of response from Phase 2 trials.
Purpose of the Study:
- To review the pharmacology, pharmacokinetics, and clinical applications of tazemetostat.
- To highlight its mechanism of action as a first-in-class EZH2 inhibitor.
- To discuss its approved indications and ongoing research in combination therapies.
Main Methods:
- Review of preclinical and clinical data on tazemetostat.
- Analysis of its mechanism of action, competing with S-adenosylmethionine (SAM) to inhibit EZH2.
- Evaluation of pharmacokinetic properties including bioavailability, metabolism, excretion, and drug interactions.
Main Results:
- Tazemetostat effectively reduces H3K27me3, a pharmacodynamic marker of EZH2 inhibition.
- It is orally bioavailable with rapid absorption and dose-proportional exposure, unaffected by food or gastric acid reducers.
- Metabolized by CYP3A, with limited CNS penetration; no dose adjustments needed for renal or hepatic dysfunction.
Conclusions:
- Tazemetostat is the first approved EZH2 inhibitor for cancer treatment.
- Its pharmacokinetic profile supports oral administration, with specific considerations for drug interactions (e.g., fluconazole).
- Ongoing studies are exploring tazemetostat in combination therapies for various malignancies.
Related Concept Videos
 01:20
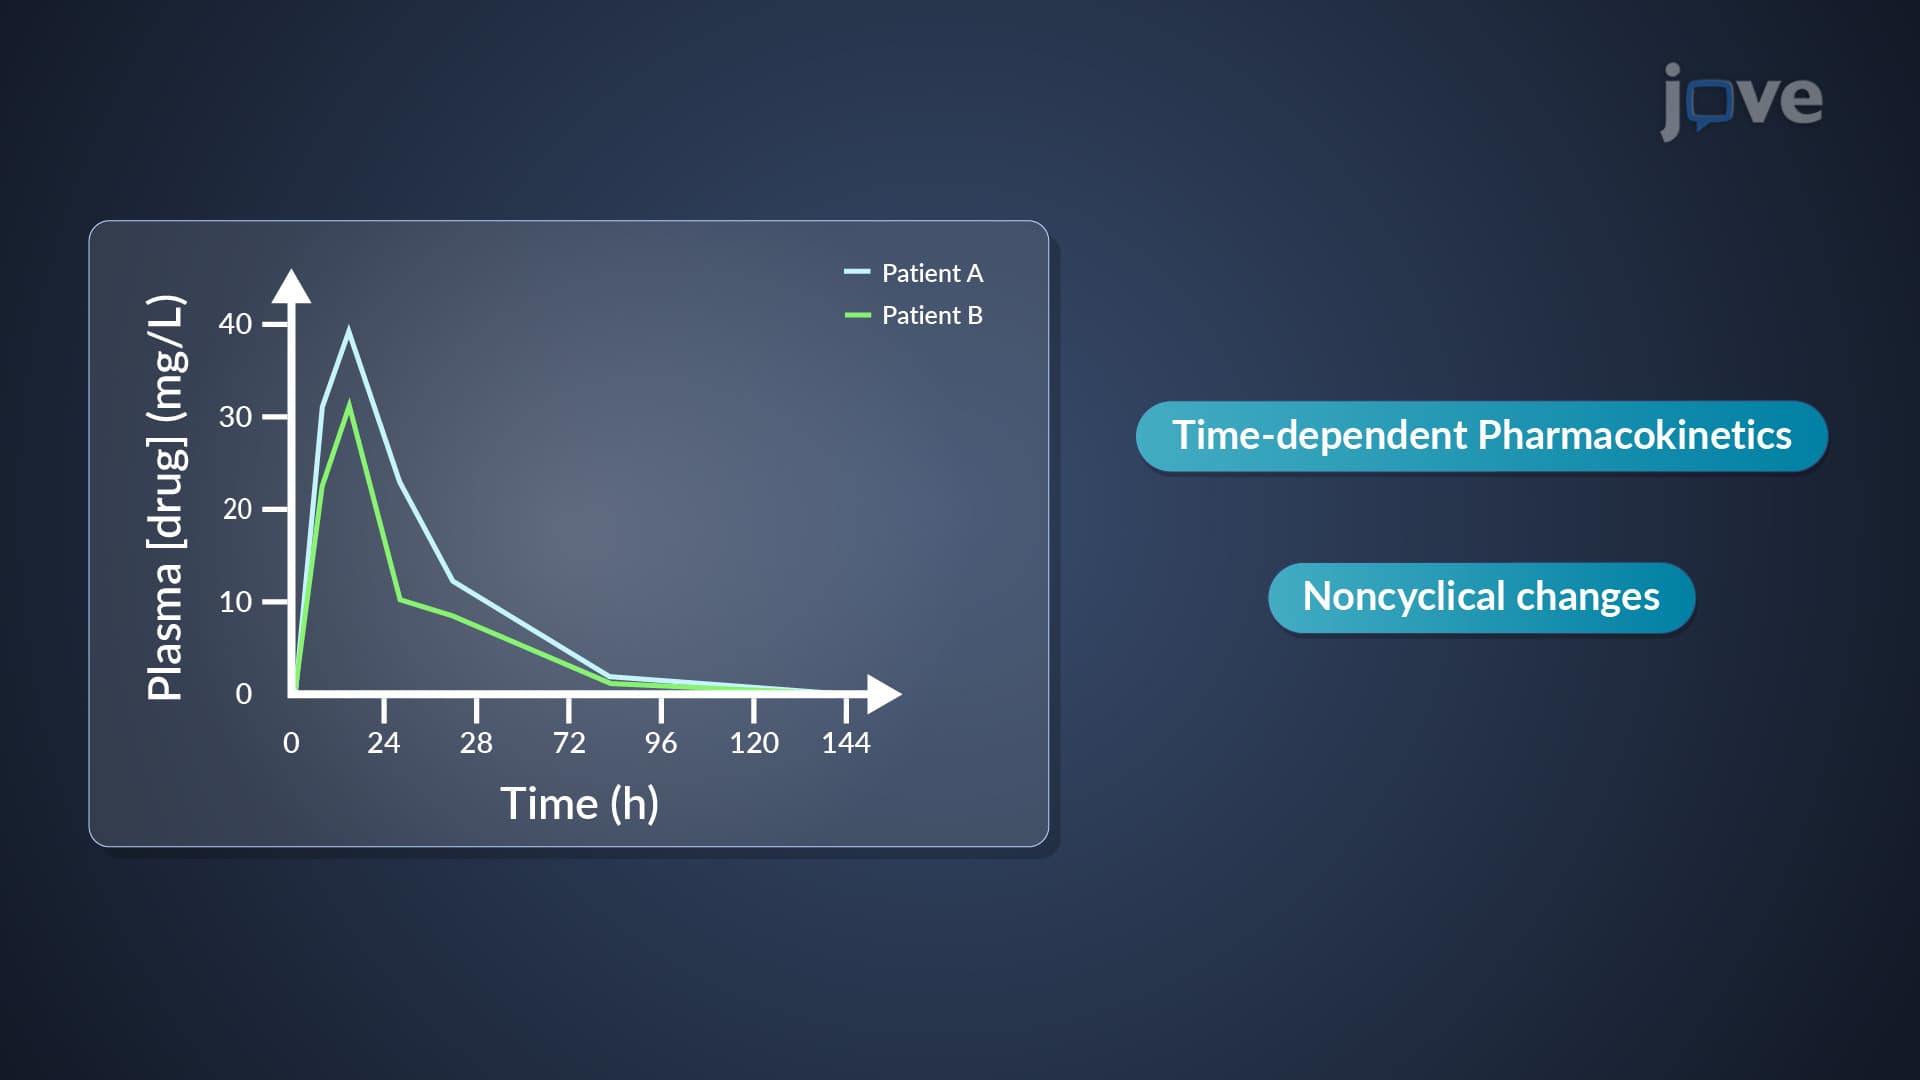
Chronopharmacokinetics: Time-Dependent Pharmacokinetics
Time-dependent pharmacokinetics refers to non-cyclical changes in drug rate processes over a period of time. It can lead to nonlinear pharmacokinetics, where the relationship between drug concentration and time is not proportional. Non-cyclical...
 01:24
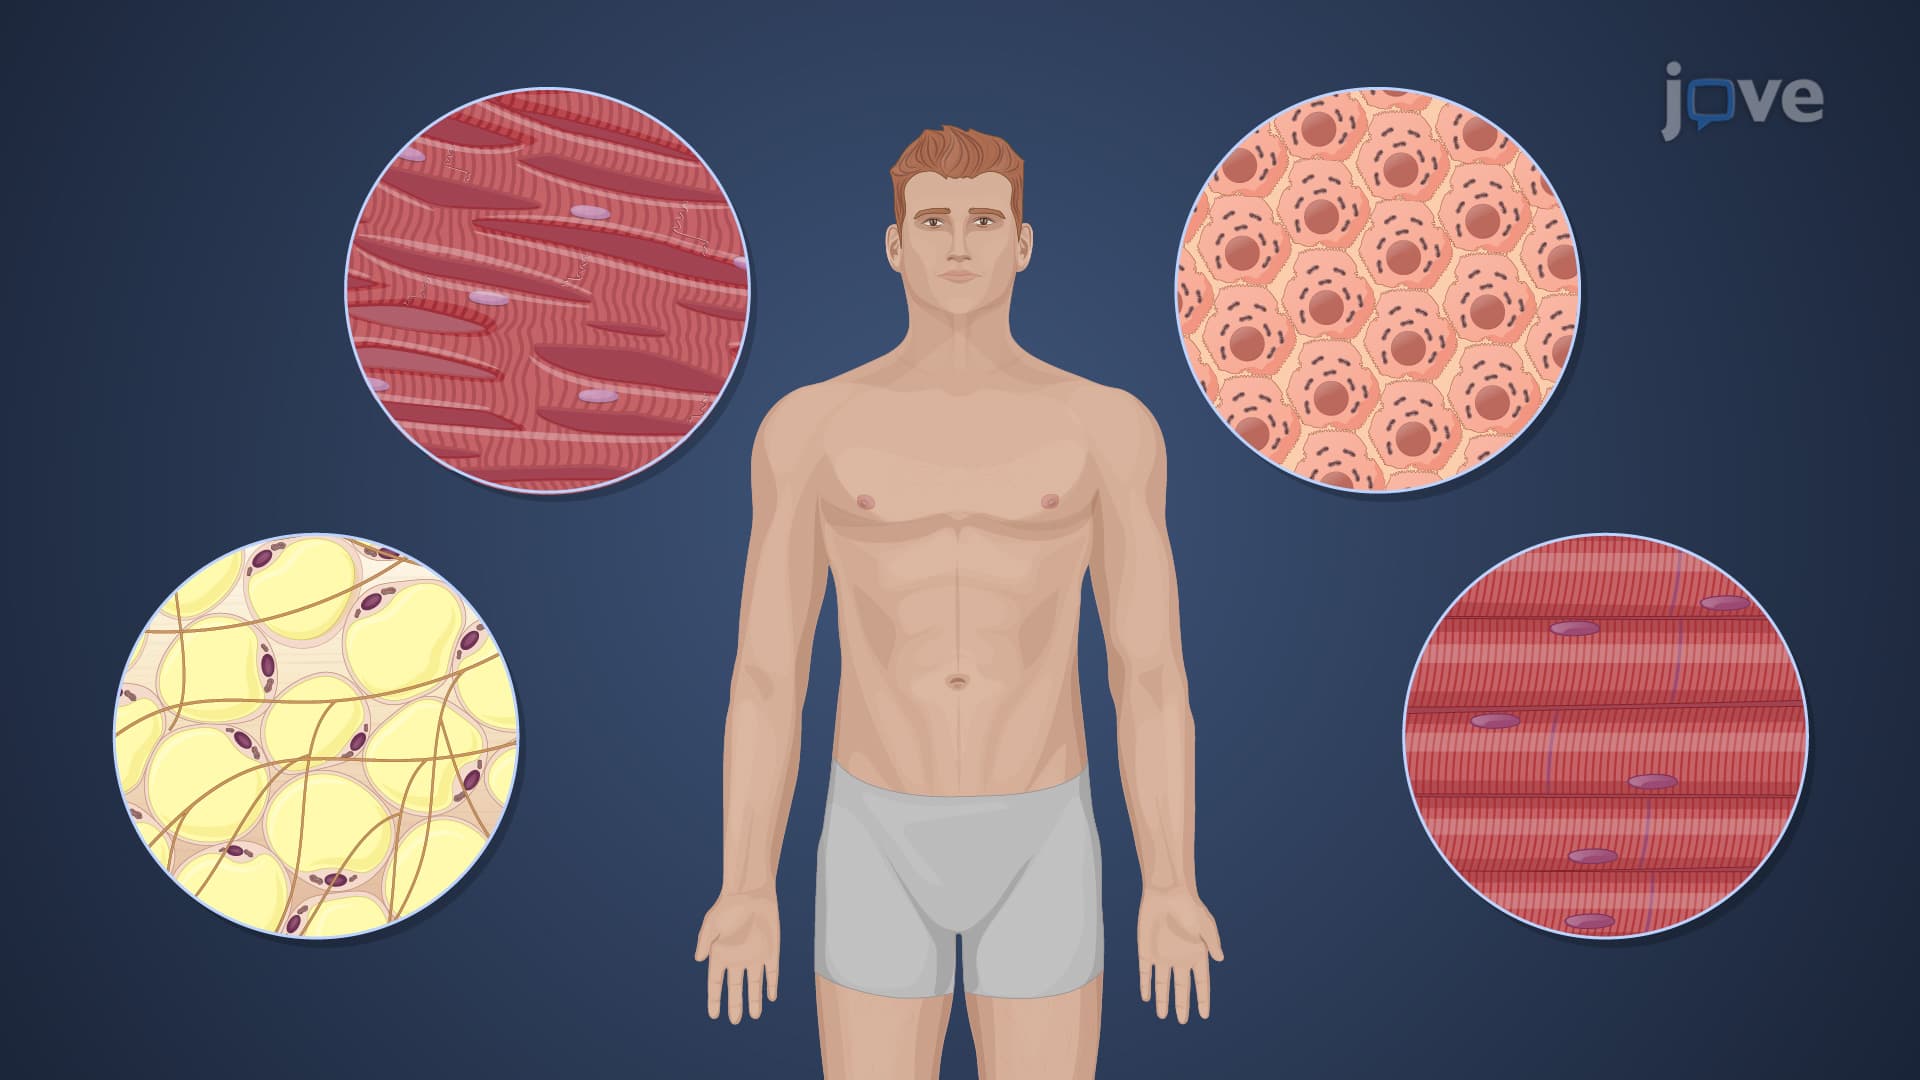
Tissue-Drug Binding: Localization of Drugs and its Significance
Drugs can bind to different tissue components, enhancing their distribution and localization. The factors influencing drug localization in tissues include the drug's lipophilicity, structural characteristics, tissue perfusion rate, and pH differences. These factors determine...
 01:10
Pharmacokinetics: Overview
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:16
Drug Biotransformation: Overview
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...