Related Experiment Video
Updated: Jun 29, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Noise-independent route toward the genesis of a COMPACT ansatz for molecular energetics: A dynamic approach
Dipanjali Halder1, Dibyendu Mondal1, Rahul Maitra1,2
1Department of Chemistry, Indian Institute of Technology Bombay, Powai, Mumbai 400076, India.

Abstract:
Recent advances in quantum information and quantum science have inspired the development of various compact, dynamically structured ansätze that are expected to be realizable in Noisy Intermediate-Scale Quantum (NISQ) devices. However, such ansätze construction strategies hitherto developed involve considerable measurements, and thus, they deviate significantly in the NISQ platform from their ideal structures. Therefore, it is imperative that the usage of quantum resources be minimized while retaining the expressivity and dynamical structure of the ansatz that can adapt itself depending on the degree of correlation. We propose a novel ansatz construction strategy based on the ab initio many-body perturbation theory that requires no pre-circuit measurement and, thus, remains structurally unaffected by any hardware noise. The accuracy and quantum complexity associated with the ansatz are solely dictated by a pre-defined perturbative order, as desired, and, hence, are tunable. Furthermore, the underlying perturbative structure of the ansatz construction pipeline enables us to decompose any high-rank excitation that appears in higher perturbative orders into the product of various low-rank operators, and it thus keeps the execution gate-depth to its minimum. With a number of challenging applications on strongly correlated systems, we demonstrate that our ansatz performs significantly better, both in terms of accuracy, parameter count, and circuit depth, in comparison to the allied unitary coupled cluster based ansätze.
More Related Videos
 05:00
Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 03:25
Free Energy Changes for Nonstandard States
where R is the gas constant (8.314 J/K·mol), T is the absolute temperature in kelvin, and Q is the reaction quotient. This equation may be used to predict the spontaneity of a process under any given set of conditions.
Reaction Quotient...
 02:34
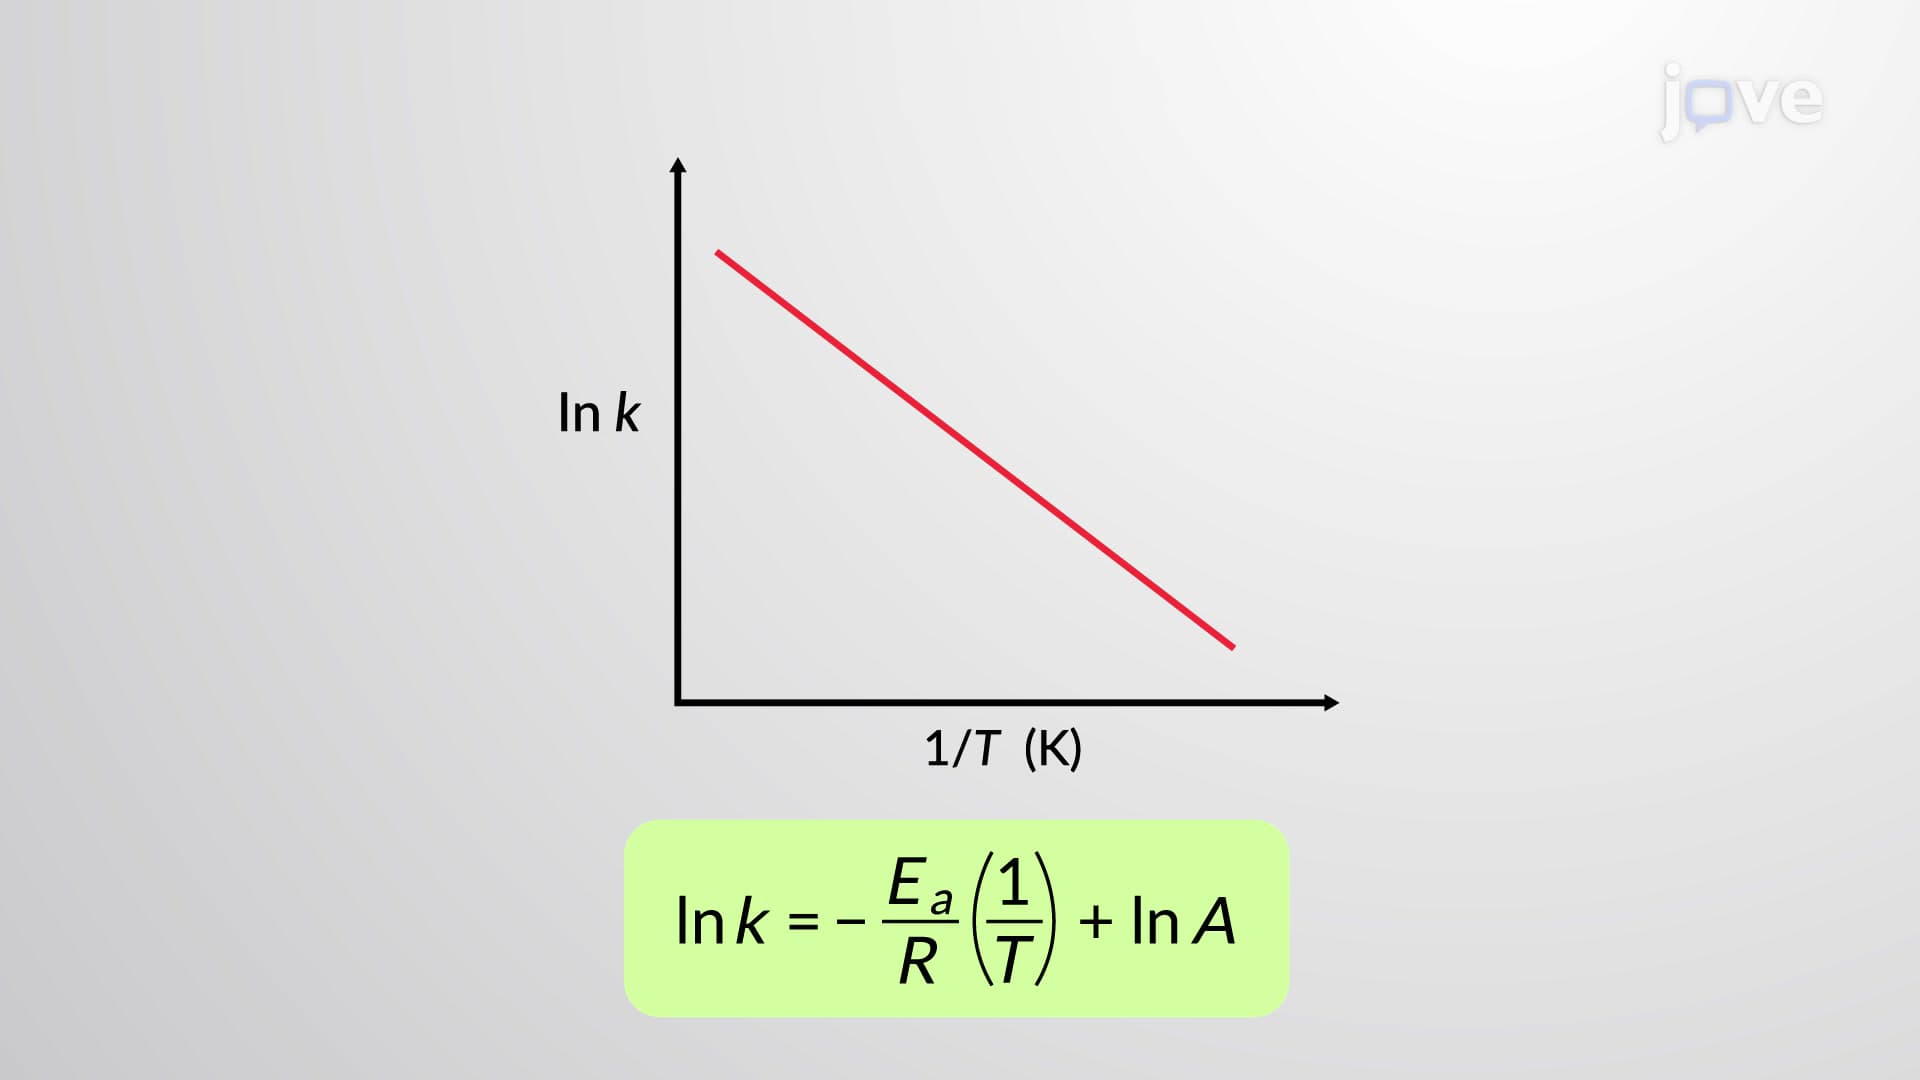
Arrhenius Plots
The Arrhenius equation can be used...
 02:18
The Bohr Model
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
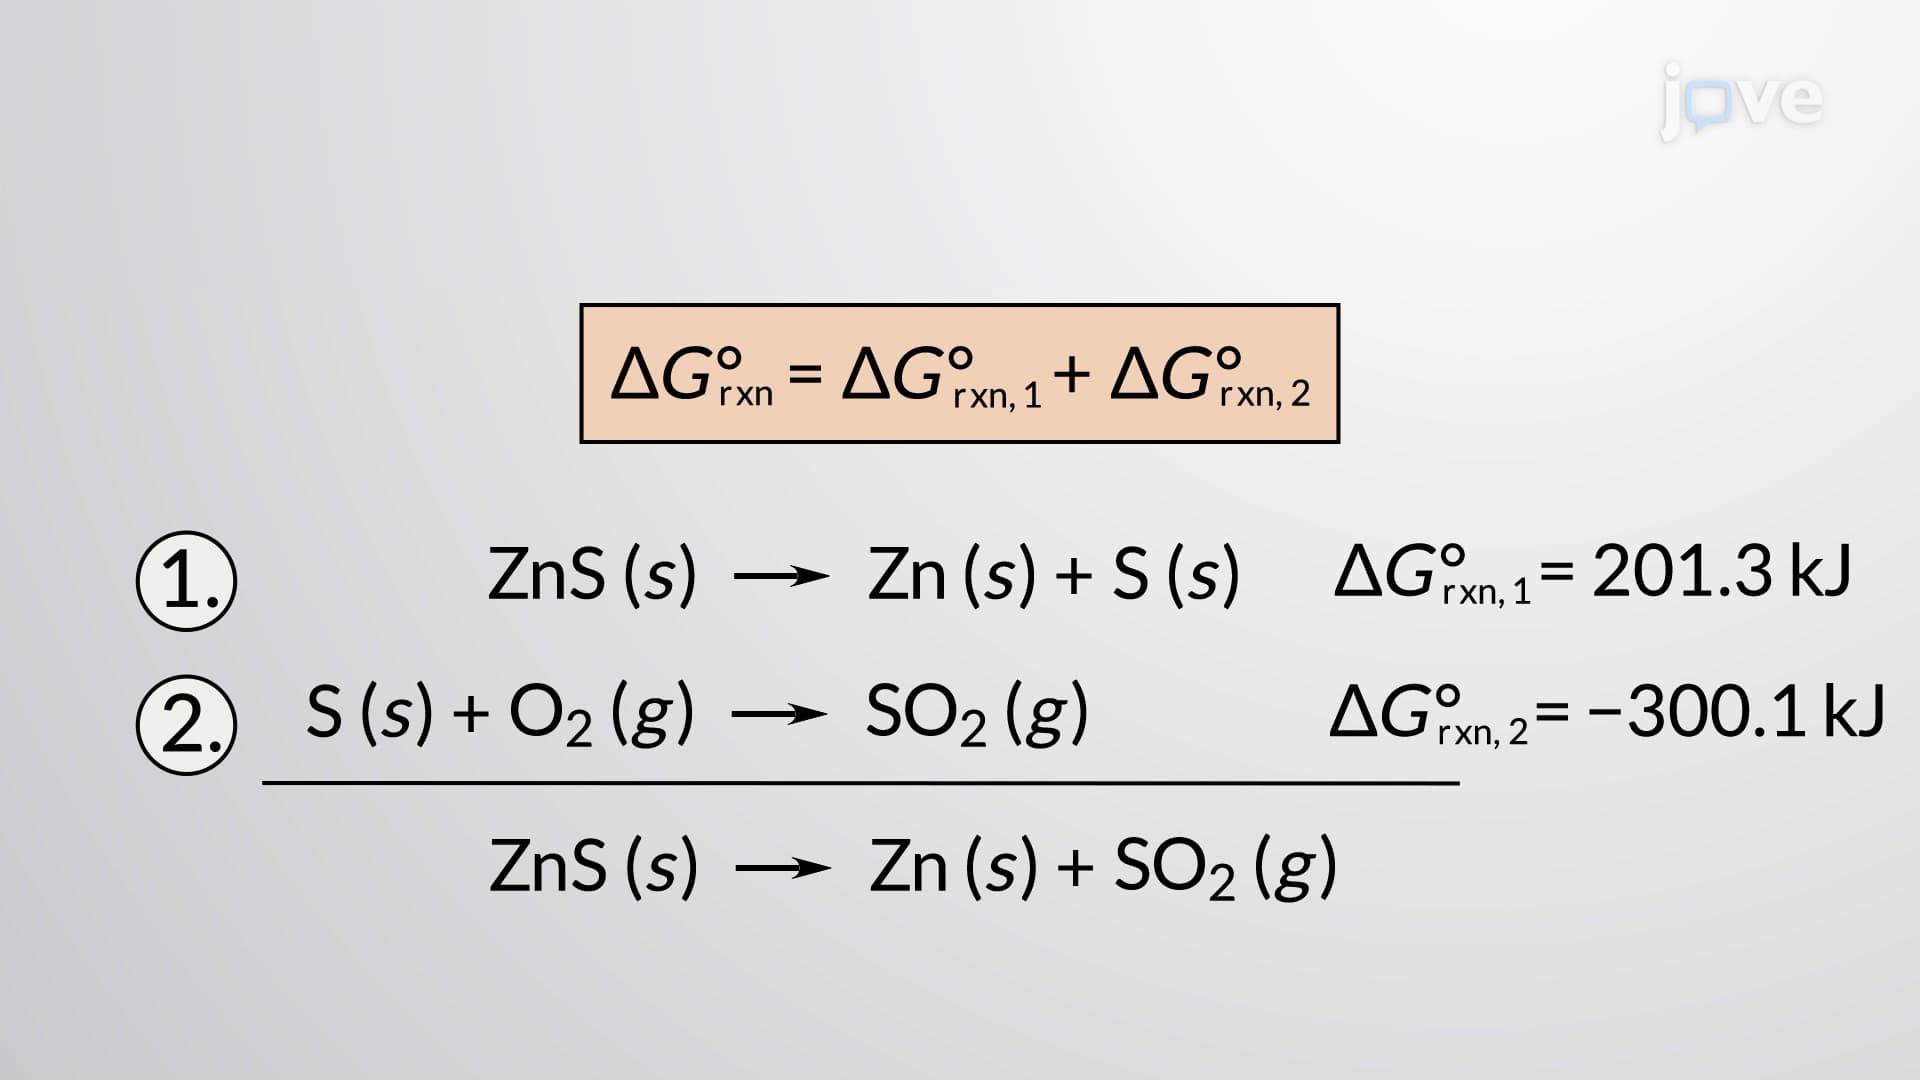 02:49
Calculating Standard Free Energy Changes
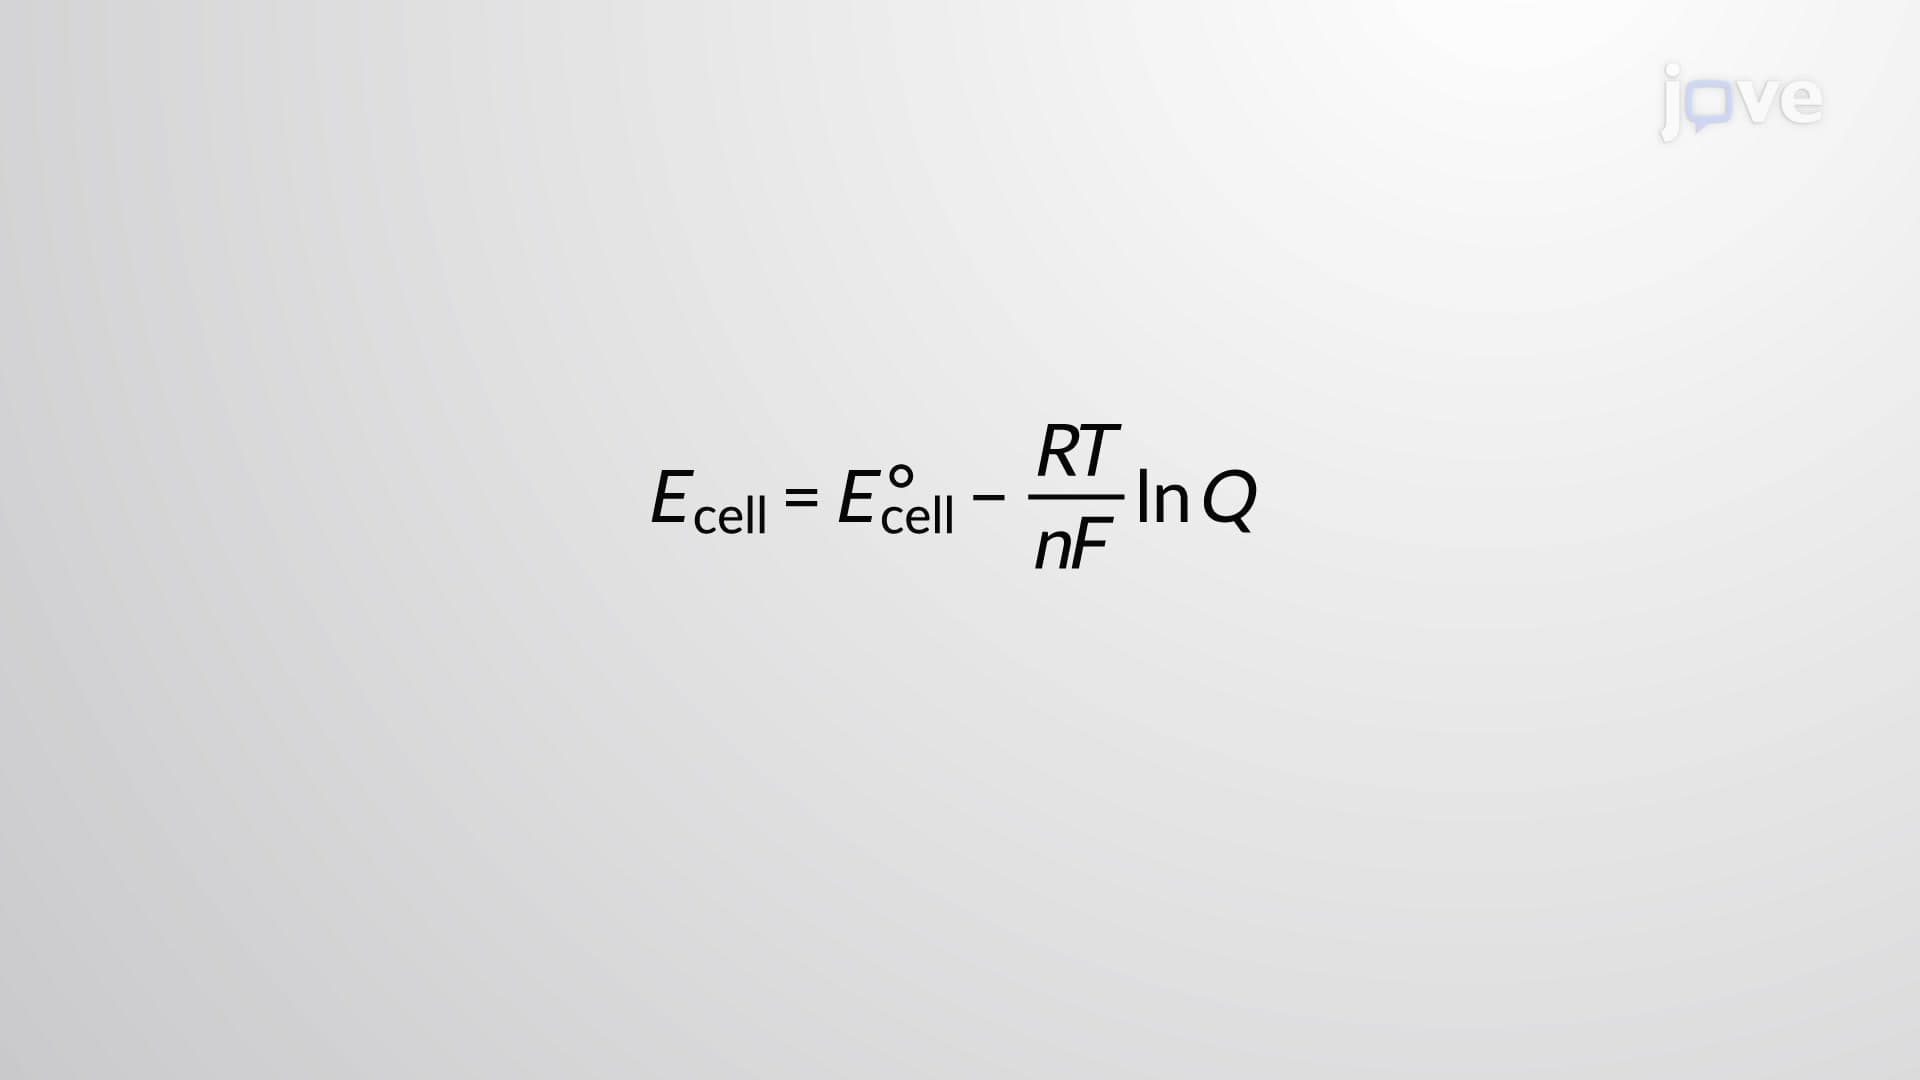 02:59
The Nernst Equation
The interconnection between standard cell potentials and various thermodynamic parameters such as the standard free energy change ΔG° and equilibrium constant K has been previously explored. For example, a redox reaction involving zinc(II) and tin(II) ions at 1 M concentration with Eºcell = +0.291 V and ΔG° = −56.2 kJ is spontaneous.