Related Experiment Video
Updated: Jun 28, 2025

Screening and Identification of Small Peptides Targeting Fibroblast Growth Factor Receptor2 using a Phage Display Peptide Library
Published on: September 30, 2019
Development of Highly Potent and Selective Covalent FGFR4 Inhibitors Based on SNAr Electrophiles
Moritz Schwarz1, Maksym Kurkunov1,2, Florian Wittlinger1
1Department of Pharmaceutical/Medicinal Chemistry, Institute of Pharmaceutical Sciences, Eberhard Karls University Tübingen, 72076 Tübingen, Germany.

Abstract:
Fibroblast growth factor receptor 4 (FGFR4) is thought to be a driver in several cancer types, most notably in hepatocellular carcinoma. One way to achieve high potency and isoform selectivity for FGFR4 is covalently targeting a rare cysteine (C552) in the hinge region of its kinase domain that is not present in other FGFR family members (FGFR1-3). Typically, this cysteine is addressed via classical acrylamide electrophiles. We demonstrate that noncanonical covalent "warheads" based on nucleophilic aromatic substitution (SNAr) chemistry can be employed in a rational manner to generate highly potent and (isoform-)selective FGFR4 inhibitors with a low intrinsic reactivity. Key compounds showed low to subnanomolar potency, efficient covalent inactivation kinetics, and excellent selectivity against the other FGFRs, the kinases with an equivalent cysteine, and a representative subset of the kinome. Moreover, these compounds achieved nanomolar potencies in cellular assays and demonstrated good microsomal stability, highlighting the potential of SNAr-based approaches in covalent inhibitor design.
Insights
Novel covalent inhibitors targeting Fibroblast Growth Factor Receptor 4 (FGFR4) were developed using SNAr chemistry. These potent and selective FGFR4 inhibitors show promise for cancer therapy, particularly hepatocellular carcinoma.
Area of Science:
- Oncology
- Medicinal Chemistry
- Biochemistry
Background:
- Fibroblast Growth Factor Receptor 4 (FGFR4) is implicated as a driver in various cancers, including hepatocellular carcinoma.
- Targeting the unique cysteine residue (C552) in FGFR4 offers a strategy for potent and isoform-selective inhibition.
- Traditional covalent inhibitors often utilize acrylamide electrophiles.
Purpose of the Study:
- To explore noncanonical covalent warheads for FGFR4 inhibition.
- To develop highly potent and isoform-selective FGFR4 inhibitors with reduced intrinsic reactivity.
- To assess the potential of nucleophilic aromatic substitution (SNAr) chemistry in covalent inhibitor design.
Main Methods:
- Design and synthesis of novel covalent inhibitors utilizing SNAr chemistry.
- Biochemical assays to determine potency, selectivity, and inactivation kinetics.
- Cellular assays to evaluate compound efficacy and microsomal stability.
Main Results:
- Compounds achieved low to subnanomolar potency against FGFR4.
- Demonstrated efficient covalent inactivation kinetics and excellent selectivity over FGFR1-3 and other kinases.
- Exhibited nanomolar potency in cellular assays and good microsomal stability.
Conclusions:
- SNAr-based covalent warheads represent a viable strategy for designing potent and selective FGFR4 inhibitors.
- These novel inhibitors show significant potential for therapeutic applications in FGFR4-driven cancers.
- This approach offers an alternative to classical electrophilic warheads in covalent drug discovery.
More Related Videos
 06:17
Covalent Fragment Screening Using the Quantitative Irreversible Tethering Assay
Published on: February 28, 2025
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
Related Concept Videos
 01:25
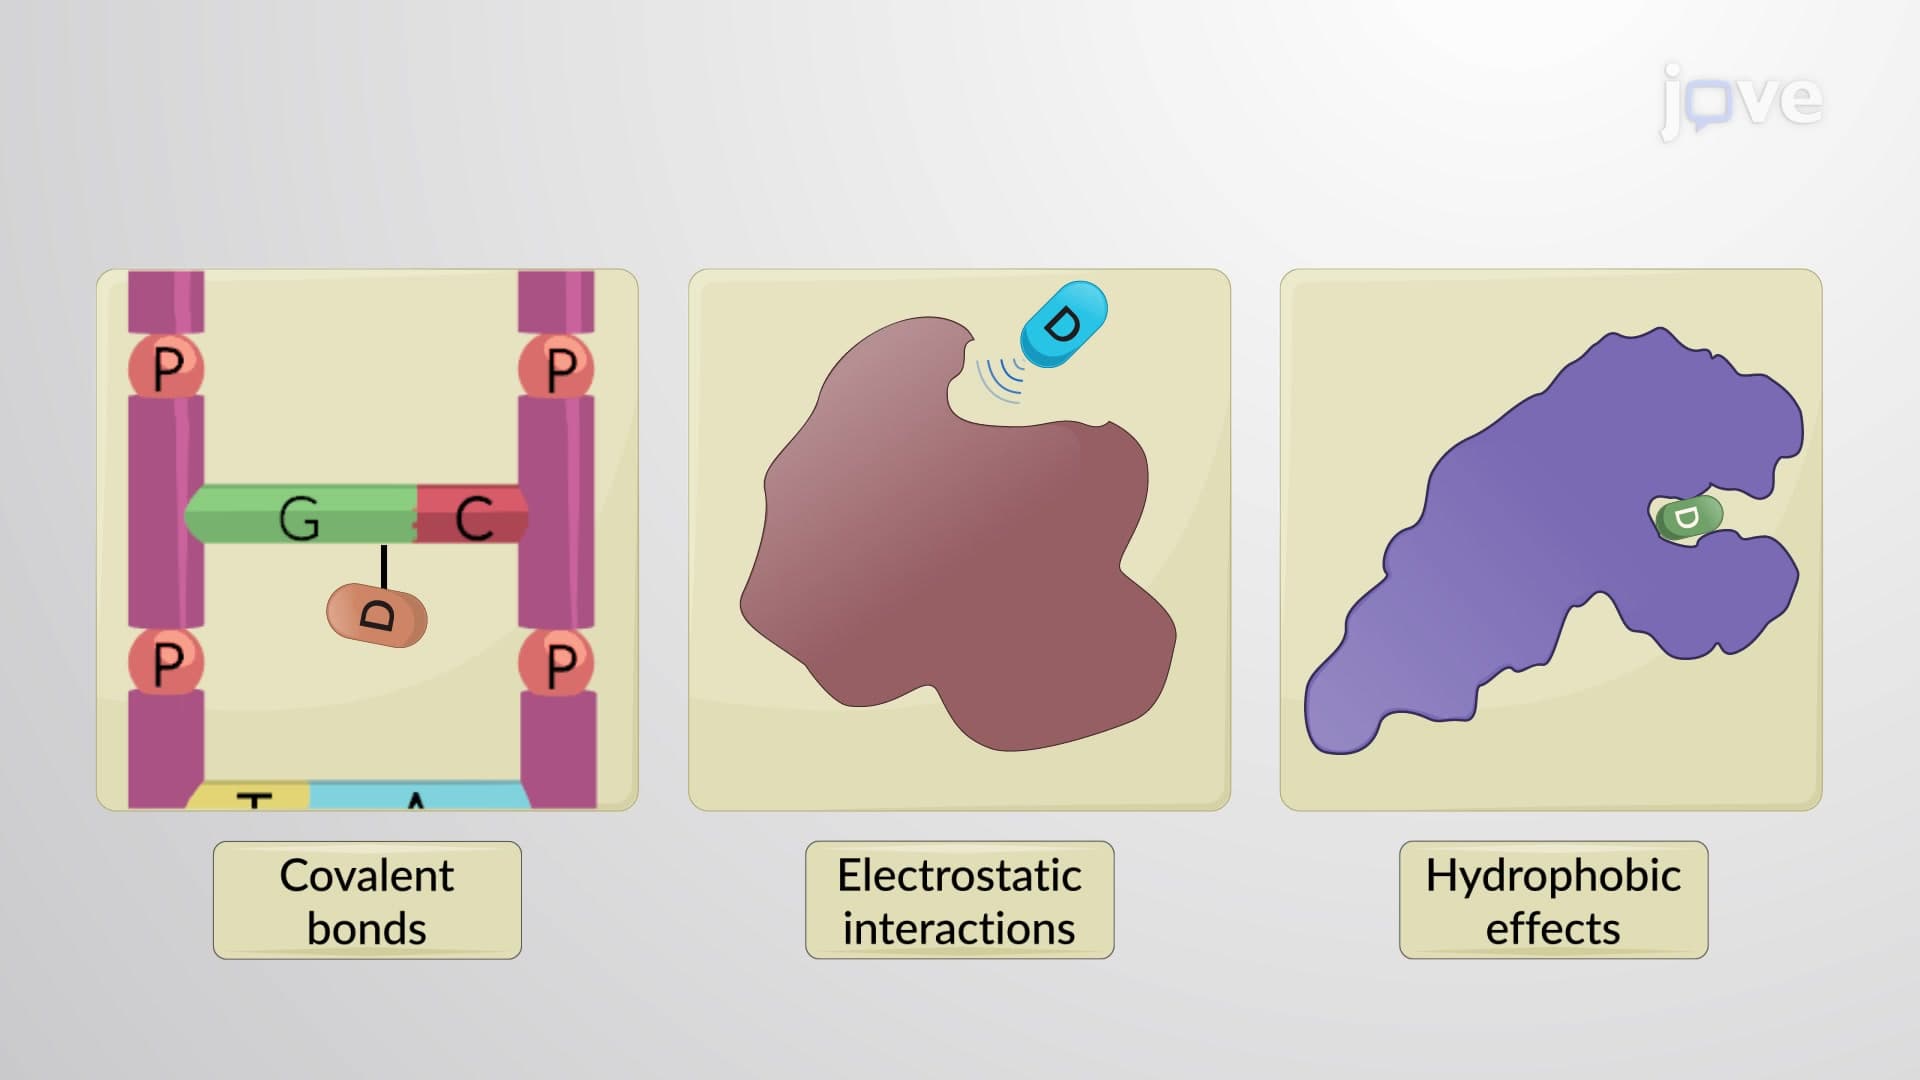
Drug-Receptor Bonds
In...
 01:30
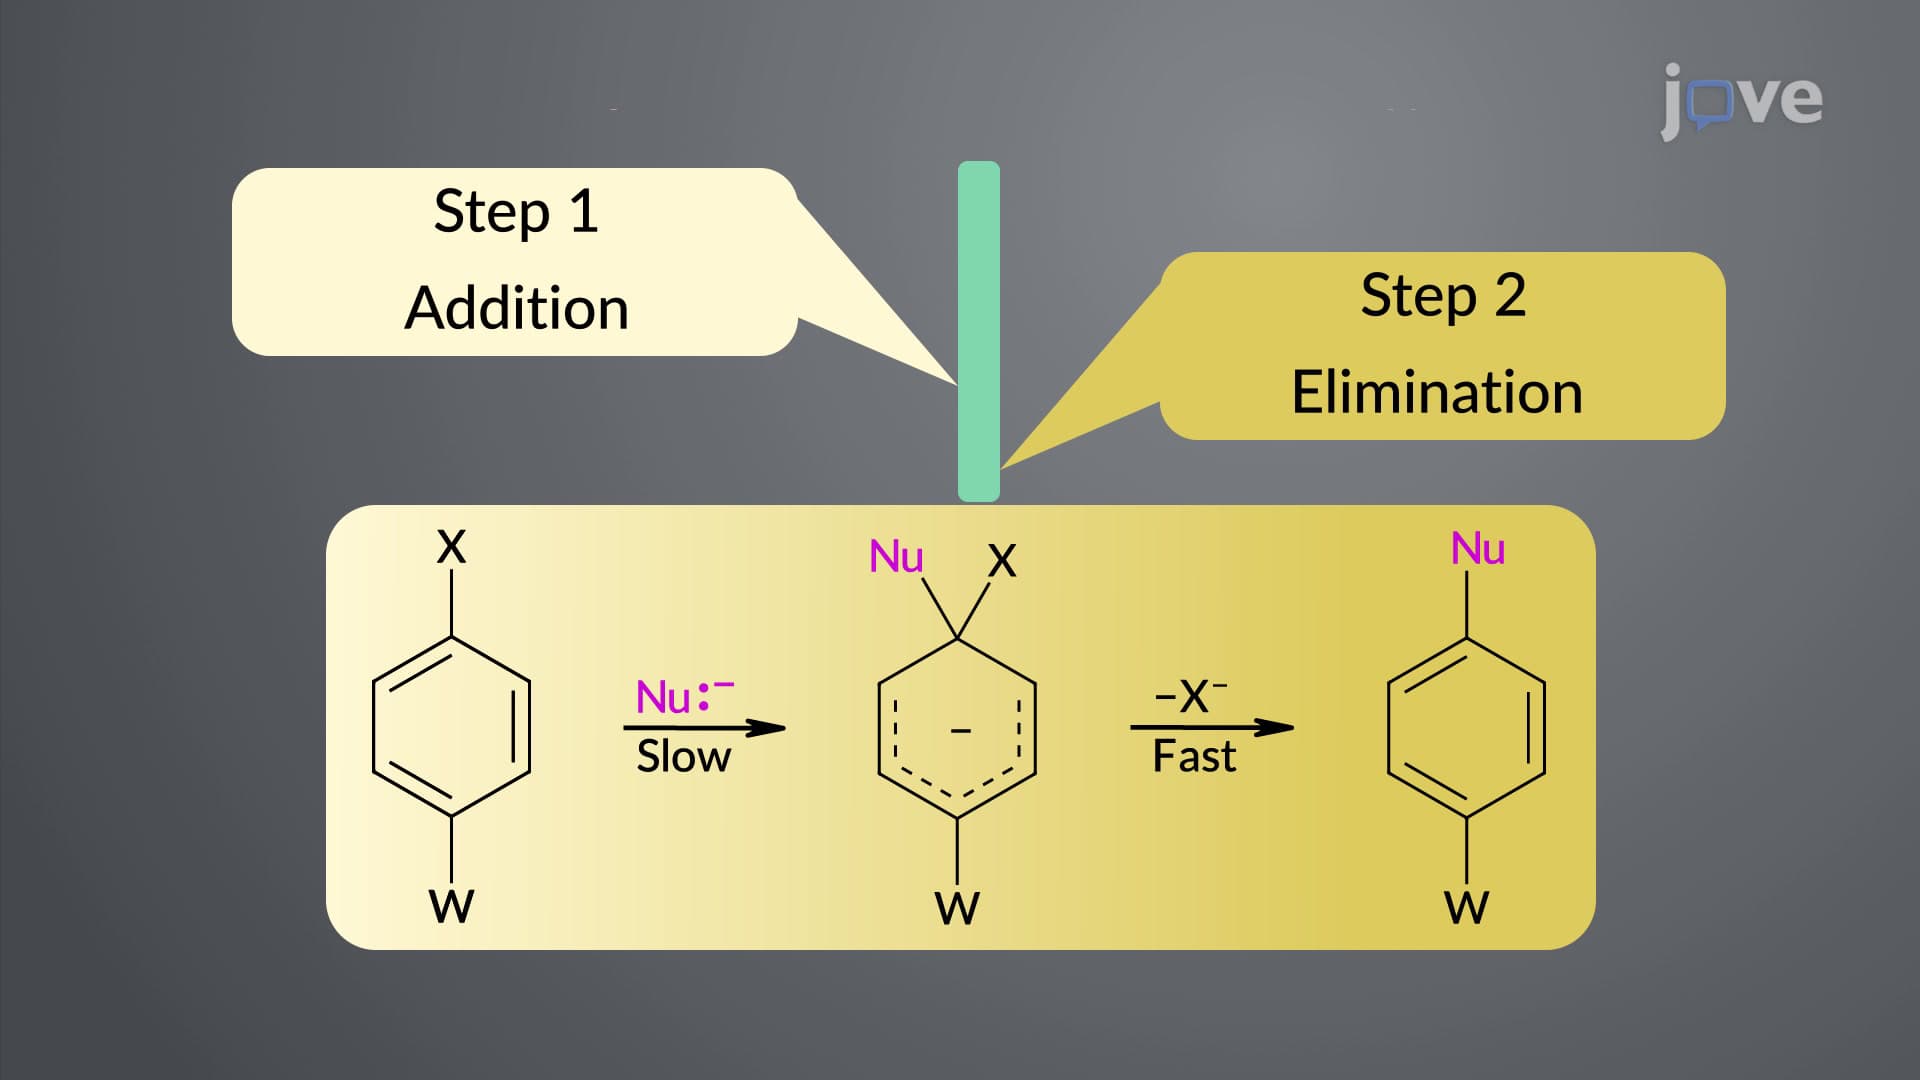
Nucleophilic Aromatic Substitution: Addition–Elimination (SNAr)
The reaction begins with an attack of the nucleophile on the carbon that holds the leaving group. This results in the delocalization of the π electrons over the ring carbons. The resonance interaction between...