Related Experiment Video
Updated: May 9, 2026
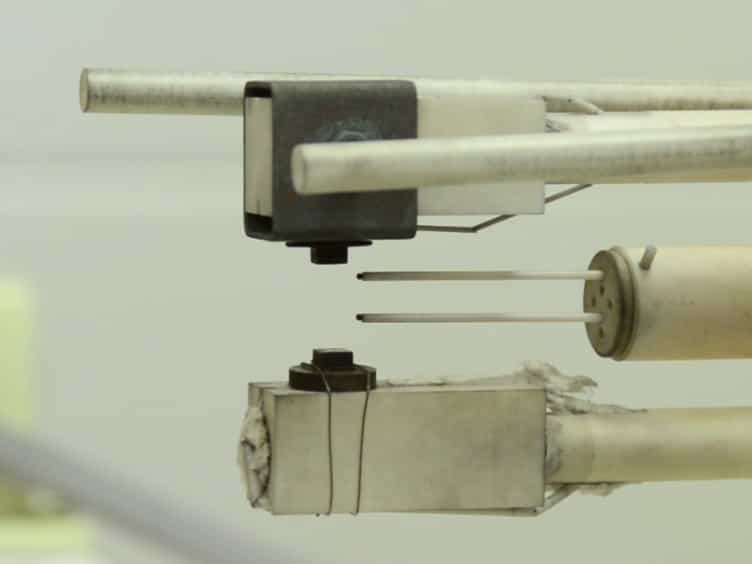

Synthesis of Non-uniformly Pr-doped SrTiO3 Ceramics and Their Thermoelectric Properties
Published on: August 15, 2015
Photogenerated carrier dynamics of Mn2+ doped CsPbBr3 assembled with TiO2 systems: Effect of Mn doping content
Luchao Du1, Jie An1, Tetsuro Katayama2
1Institute of Atomic and Molecular Physics, Jilin University, 2699 Qianjin Street, Changchun 130012, China.

Abstract:
In recent years, all-inorganic perovskite materials have become an ideal choice for new thin film solar cells due to their excellent photophysical properties and have become a research hotspot. Studying the ultrafast dynamics of photo-generated carriers is of great significance for further improving the performance of such devices. In this work, we focus on the transient dynamic process of CsPbBr3/TiO2 composite systems with different Mn2+ doping contents using femtosecond transient absorption spectroscopy technology. We used singular value decomposition and global fitting to analyze the transient absorption spectra and obtained three components, which are classified as hot carrier cooling, charge transfer, and charge recombination processes, respectively. We found that the doping concentration of Mn2+ has an impact on all three processes. We think that the following two factors are responsible: one is the density of defect states and the other is the bandgap width of perovskite. As the concentration of doped Mn2+ increases, the charge transfer time constant shows a trend of initially increasing, followed by a subsequent decrease, reaching a turning point. This indicates that an appropriate amount of Mn2+ doping can effectively improve the photoelectric performance of solar cell systems. We proposed a possible charge transfer mechanism model and further elucidated the microscopic mechanism of the effect of Mn2+ doping on the interface charge transfer process of the CsPbBr3/TiO2 solar cell system.
Related Concept Videos
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...