Related Experiment Video
Updated: Jun 28, 2025

Demonstration of the Sequence Alignment to Predict Across Species Susceptibility Tool for Rapid Assessment of Protein Conservation
Published on: February 10, 2023
Predicting FFAR4 agonists using structure-based machine learning approach based on molecular fingerprints
Zaid Anis Sherwani1, Syeda Sumayya Tariq1, Mamona Mushtaq1
1Dr. Panjwani Center for Molecular Medicine and Drug Research, International Center for Chemical and Biological Sciences, University of Karachi, Karachi, 75270, Pakistan.

Abstract:
Free Fatty Acid Receptor 4 (FFAR4), a G-protein-coupled receptor, is responsible for triggering intracellular signaling pathways that regulate various physiological processes. FFAR4 agonists are associated with enhancing insulin release and mitigating the atherogenic, obesogenic, pro-carcinogenic, and pro-diabetogenic effects, normally associated with the free fatty acids bound to FFAR4. In this research, molecular structure-based machine-learning techniques were employed to evaluate compounds as potential agonists for FFAR4. Molecular structures were encoded into bit arrays, serving as molecular fingerprints, which were subsequently analyzed using the Bayesian network algorithm to identify patterns for screening the data. The shortlisted hits obtained via machine learning protocols were further validated by Molecular Docking and via ADME and Toxicity predictions. The shortlisted compounds were then subjected to MD Simulations of the membrane-bound FFAR4-ligand complexes for 100 ns each. Molecular analyses, encompassing binding interactions, RMSD, RMSF, RoG, PCA, and FEL, were conducted to scrutinize the protein-ligand complexes at the inter-atomic level. The analyses revealed significant interactions of the shortlisted compounds with the crucial residues of FFAR4 previously documented. FFAR4 as part of the complexes demonstrated consistent RMSDs, ranging from 3.57 to 3.64, with minimal residue fluctuations 5.27 to 6.03 nm, suggesting stable complexes. The gyration values fluctuated between 22.8 to 23.5 nm, indicating structural compactness and orderliness across the studied systems. Additionally, distinct conformational motions were observed in each complex, with energy contours shifting to broader energy basins throughout the simulation, suggesting thermodynamically stable protein-ligand complexes. The two compounds CHEMBL2012662 and CHEMBL64616 are presented as potential FFAR4 agonists, based on these insights and in-depth analyses. Collectively, these findings advance our comprehension of FFAR4's functions and mechanisms, highlighting these compounds as potential FFAR4 agonists worthy of further exploration as innovative treatments for metabolic and immune-related conditions.
Insights
Machine learning identified two compounds, CHEMBL2012662 and CHEMBL64616, as potential agonists for Free Fatty Acid Receptor 4 (FFAR4). These compounds show promise for treating metabolic and immune-related conditions.
Area of Science:
- Pharmacology
- Computational Chemistry
- Biophysics
Background:
- Free Fatty Acid Receptor 4 (FFAR4) is a G-protein-coupled receptor involved in regulating physiological processes.
- FFAR4 agonists may enhance insulin release and reduce metabolic disease risks.
Purpose of the Study:
- To identify novel FFAR4 agonists using molecular structure-based machine learning.
- To validate potential agonists through molecular docking, ADME/Toxicity predictions, and molecular dynamics simulations.
Main Methods:
- Machine learning (Bayesian network) applied to molecular fingerprints for initial screening.
- Molecular docking and ADME/Toxicity predictions for hit validation.
- 100 ns Molecular Dynamics (MD) simulations of FFAR4-ligand complexes.
Main Results:
- Machine learning identified promising candidate compounds.
- MD simulations revealed stable FFAR4-ligand complexes with significant interactions at crucial residues.
- Analyses indicated compact structures (RMSD 3.57-3.64 nm, fluctuations 5.27-6.03 nm) and thermodynamic stability.
Conclusions:
- Two compounds, CHEMBL2012662 and CHEMBL64616, are identified as potential FFAR4 agonists.
- These compounds warrant further investigation for therapeutic applications in metabolic and immune disorders.


Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:25
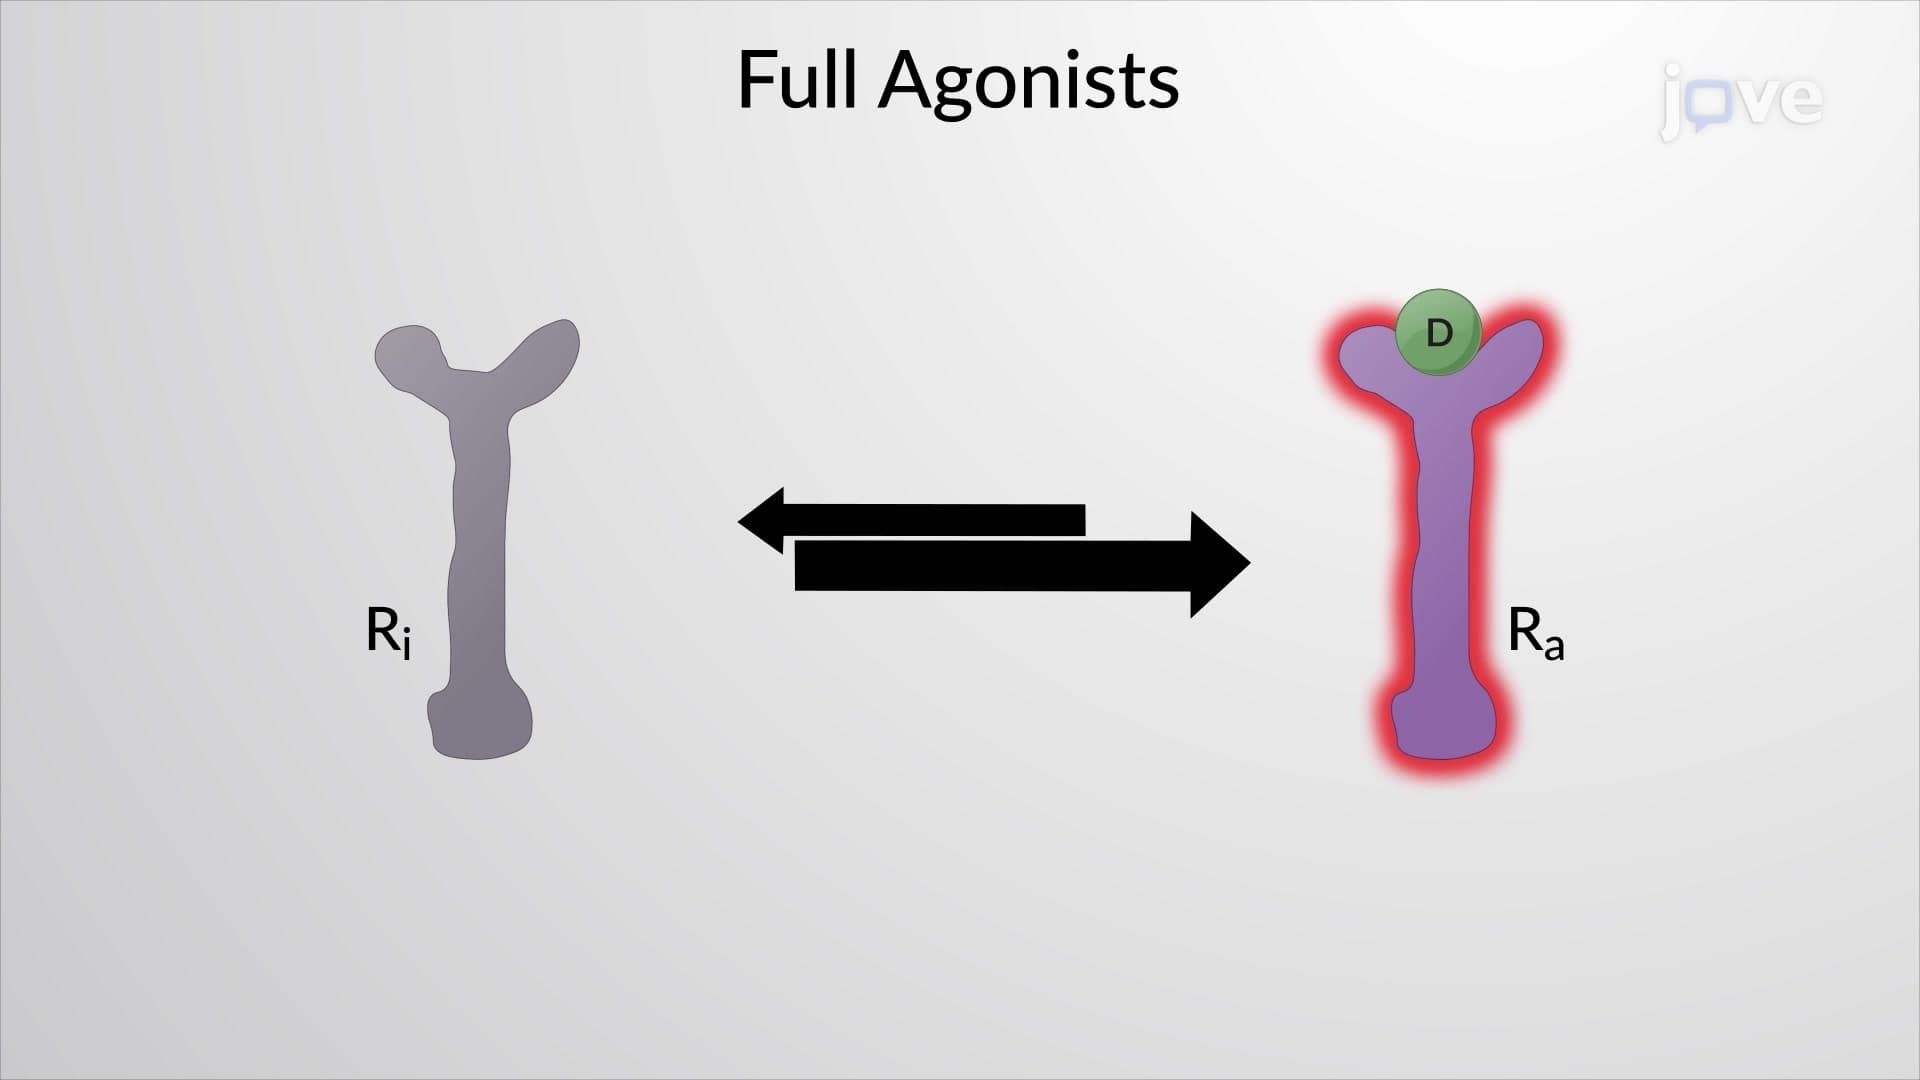
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...