Related Experiment Video
Updated: Jun 27, 2025

Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Quantification of the thermodynamic effects of the low-spin - high-spin interaction in molecular crystals of a
Juliusz A Wolny1, Konstantin Gröpl1, Jonathan Kiehl2
1Department of Physics, University of Kaiserslautern-Landau, Erwin-Schrödinger-Str. 46, 67663 Kaiserslautern, Germany. wolny@rptu.de.

Abstract:
A method is proposed to estimate the energetic and entropic effects of spins of neighbouring molecules on the spin transition of a mononuclear spin crossover (SCO) complex in a molecular crystal. Density functional theory (DFT) methods have been used to model the SCO material [FeII(Lnpdtz)2(NCS)2] (Lnpdtz = 2-naphthyl-5-pyridyl-1,2,4-thiadiazole) exhibiting numerous π-π interactions using a 2D arrangement of 15 molecules. The modelling considers only the effects in the crystallographical ac plane with a particularly pronounced stacking but paves the way for future work with 3D arrangements which are computational much more costly. It involves the optimisation and normal mode calculation of the molecules in a rigid matrix of both low-spin (LS) and high-spin (HS) neighbours. This procedure has been used to calculate the previously defined cooperativity parameter Hcoop (S. Rackwitz, W. Klopper, V. Schünemann and J. A. Wolny, Phys. Chem. Chem. Phys., 2013, 15, 15450). For [FeII(Lnpdtz)2(NCS)] we obtain Hcoop = 11 kJ mol-1, a value which is comparable to those found for 3D polynuclear spin crossover materials. A normal mode analysis of the optimised centrally located molecule indicates that the vibrational entropy of the spin transition is somewhat higher (5 J K-1 mol-1) for the LS to HS transition in the LS matrix than in the HS one. The calculations show that the interactions with the neighbours influence the low-frequency modes with wave numbers <65-70 cm-1. These cause the main difference in the vibrational entropy of the spin transition for the vicinity of high- and low-spin molecules. Furthermore, a deformation of the coordination sphere of the central molecule is observed when the spins of the surrounding centres are switched. This deformation is accompanied by a change in the equatorial Fe-N bond lengths.
More Related Videos
Related Concept Videos
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:22
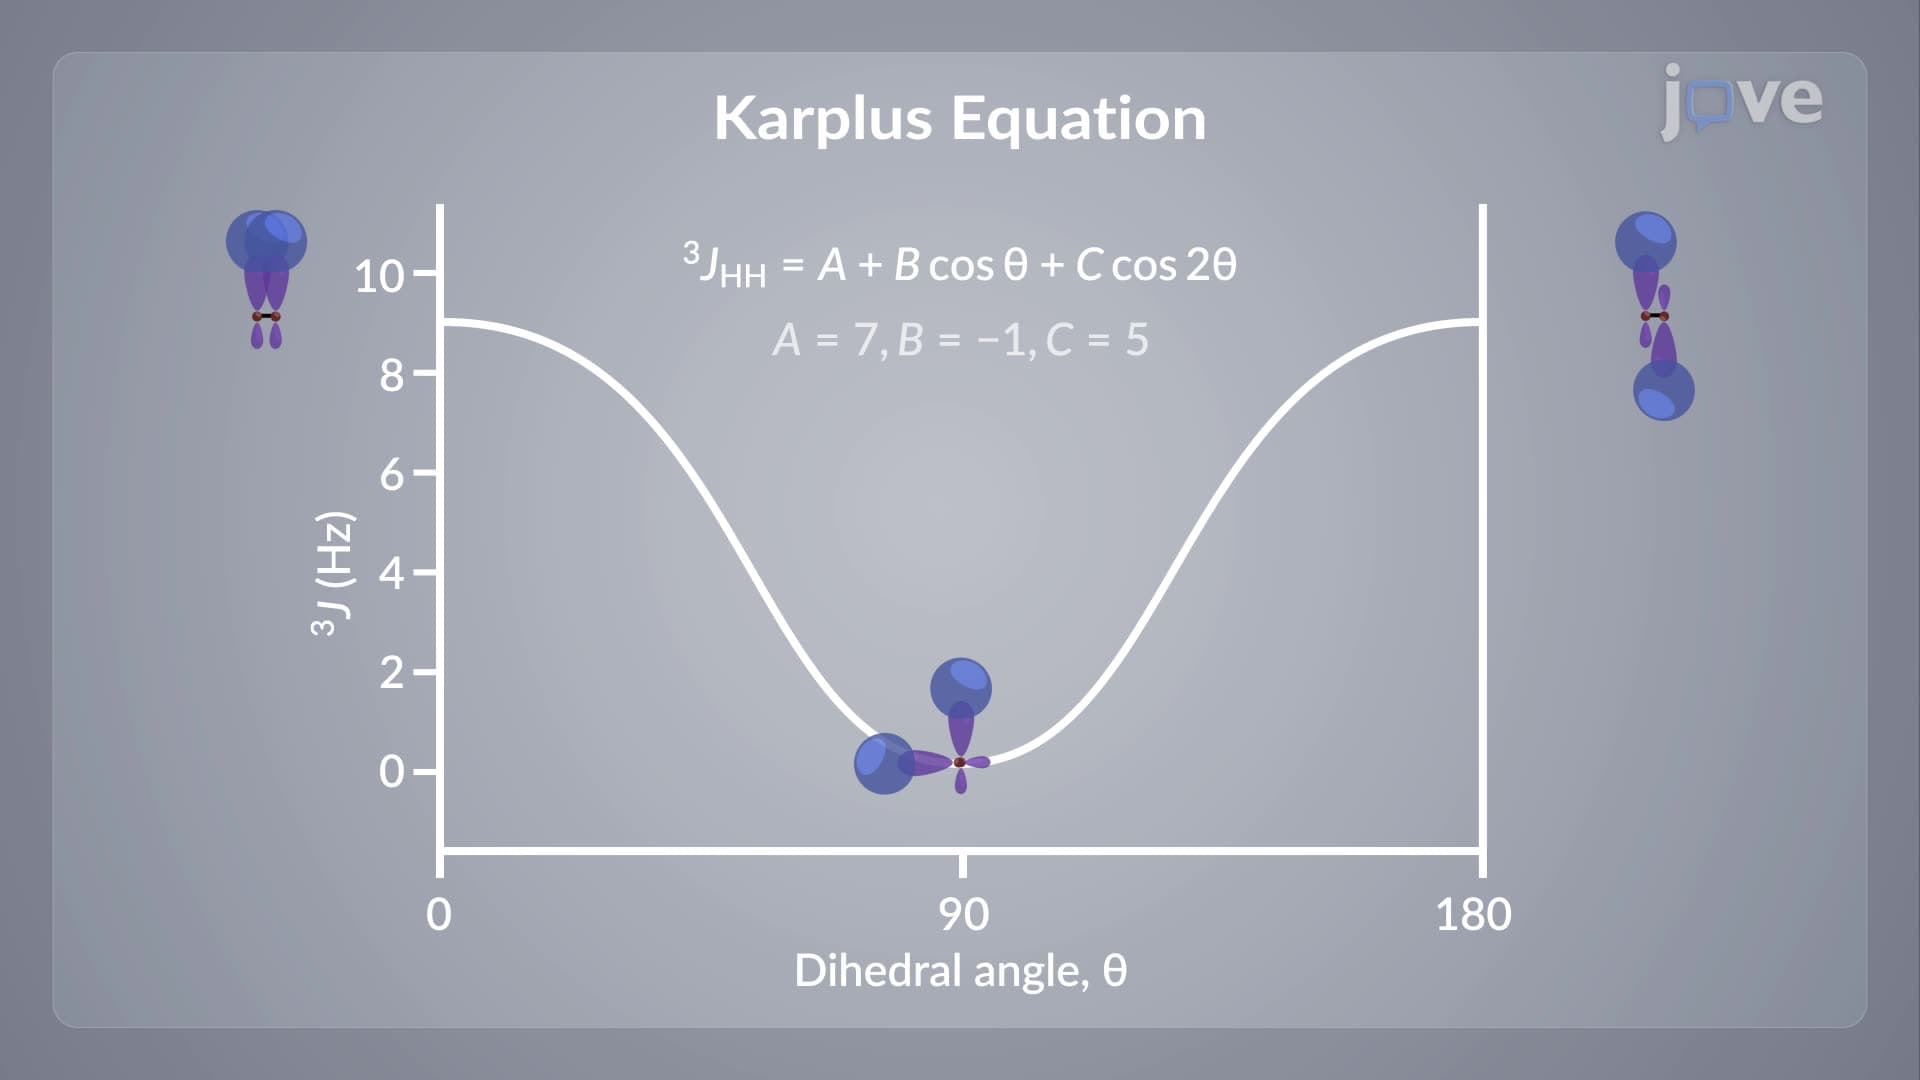
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:08
NMR Spectroscopy: Spin–Spin Coupling
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...