MzDOCK: A free ready-to-use GUI-based pipeline for molecular docking simulations
Related Experiment Video
Updated: Jul 6, 2026
Author Spotlight: Network Pharmacology and Molecular Docking to Decipher the Action of Jiawei Shengjiang San Against Diabetic Kidney Disease
Published on: May 10, 2024
Muzammil Kabier1, Nicola Gambacorta2, Daniela Trisciuzzi3
1Department of Pharmaceutical Chemistry, Amrita School of Pharmacy, Amrita Vishwa Vidyapeetham, AIMS Health Sciences Campus, Kochi, India.

Abstract:
Molecular docking is by far the most preferred approach in structure-based drug design for its effectiveness to predict the scoring and posing of a given bioactive small molecule into the binding site of its pharmacological target. Herein, we present MzDOCK, a new GUI-based pipeline for Windows operating system, designed with the intent of making molecular docking easier to use and higher reproducible even for inexperienced people. By harmonic integration of python and batch scripts, which employs various open source packages such as Smina (docking engine), OpenBabel (file conversion) and PLIP (analysis), MzDOCK includes many practical options such as: binding site configuration based on co-crystallized ligands; generation of enantiomers from SMILES input; application of different force fields (MMFF94, MMFF94s, UFF, GAFF, Ghemical) for energy minimization; retention of selectable ions and cofactors; sidechain flexibility of selectable binding site residues; multiple input file format (SMILES, PDB, SDF, Mol2, Mol); generation of reports and of pictures for interactive visualization. Users can download for free MzDOCK at the following link: https://github.com/Muzatheking12/MzDOCK.
More Related Videos
 05:00
Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
 10:29
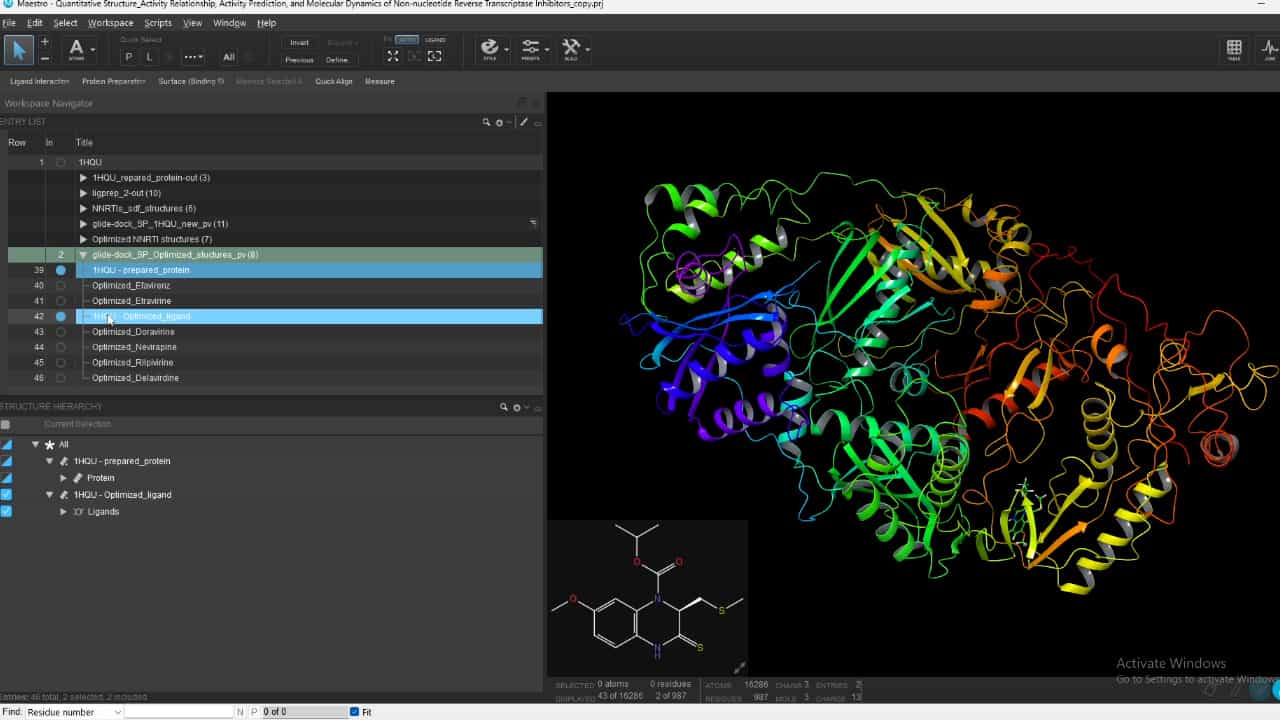
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
