Mix-Key: graph mixup with key structures for molecular property prediction
Related Experiment Video
Updated: Jun 27, 2025
Curation of Computational Chemical Libraries Demonstrated with Alpha-Amino Acids
Published on: April 13, 2022
Tianyi Jiang1,2, Zeyu Wang1,2, Wenchao Yu3
1Institute of Cyberspace Security, College of Information Engineering, Zhejiang University of Technology, 310023, Hangzhou, China.

Abstract:
Molecular property prediction faces the challenge of limited labeled data as it necessitates a series of specialized experiments to annotate target molecules. Data augmentation techniques can effectively address the issue of data scarcity. In recent years, Mixup has achieved significant success in traditional domains such as image processing. However, its application in molecular property prediction is relatively limited due to the irregular, non-Euclidean nature of graphs and the fact that minor variations in molecular structures can lead to alterations in their properties. To address these challenges, we propose a novel data augmentation method called Mix-Key tailored for molecular property prediction. Mix-Key aims to capture crucial features of molecular graphs, focusing separately on the molecular scaffolds and functional groups. By generating isomers that are relatively invariant to the scaffolds or functional groups, we effectively preserve the core information of molecules. Additionally, to capture interactive information between the scaffolds and functional groups while ensuring correlation between the original and augmented graphs, we introduce molecular fingerprint similarity and node similarity. Through these steps, Mix-Key determines the mixup ratio between the original graph and two isomers, thus generating more informative augmented molecular graphs. We extensively validate our approach on molecular datasets of different scales with several Graph Neural Network architectures. The results demonstrate that Mix-Key consistently outperforms other data augmentation methods in enhancing molecular property prediction on several datasets.
More Related Videos

Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:00
Molecular Models
 01:13
¹H NMR: Complex Splitting
Splitting diagrams or splitting tree diagrams are routinely used to depict such complex couplings. While drawing splitting diagrams, the splitting with the larger coupling constant is usually applied...
 02:36
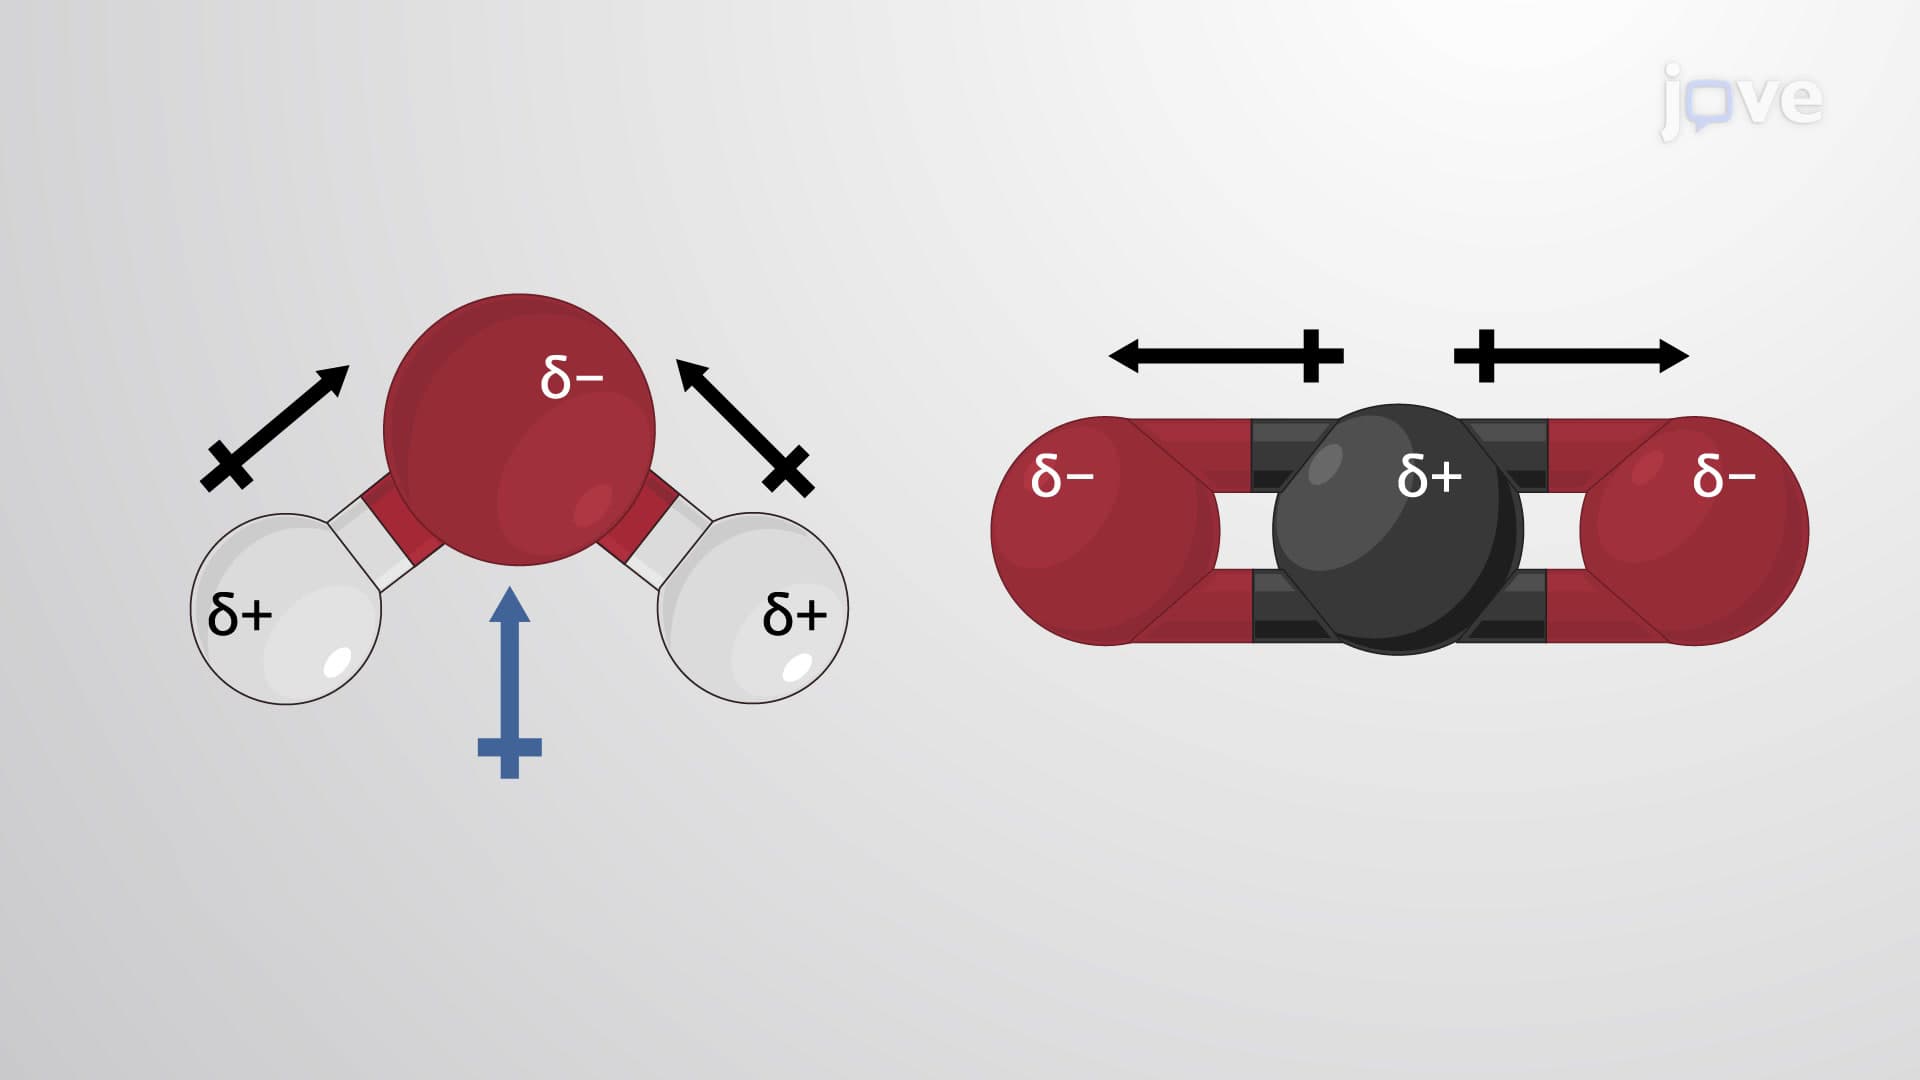
Molecular Geometry and Dipole Moments
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:03
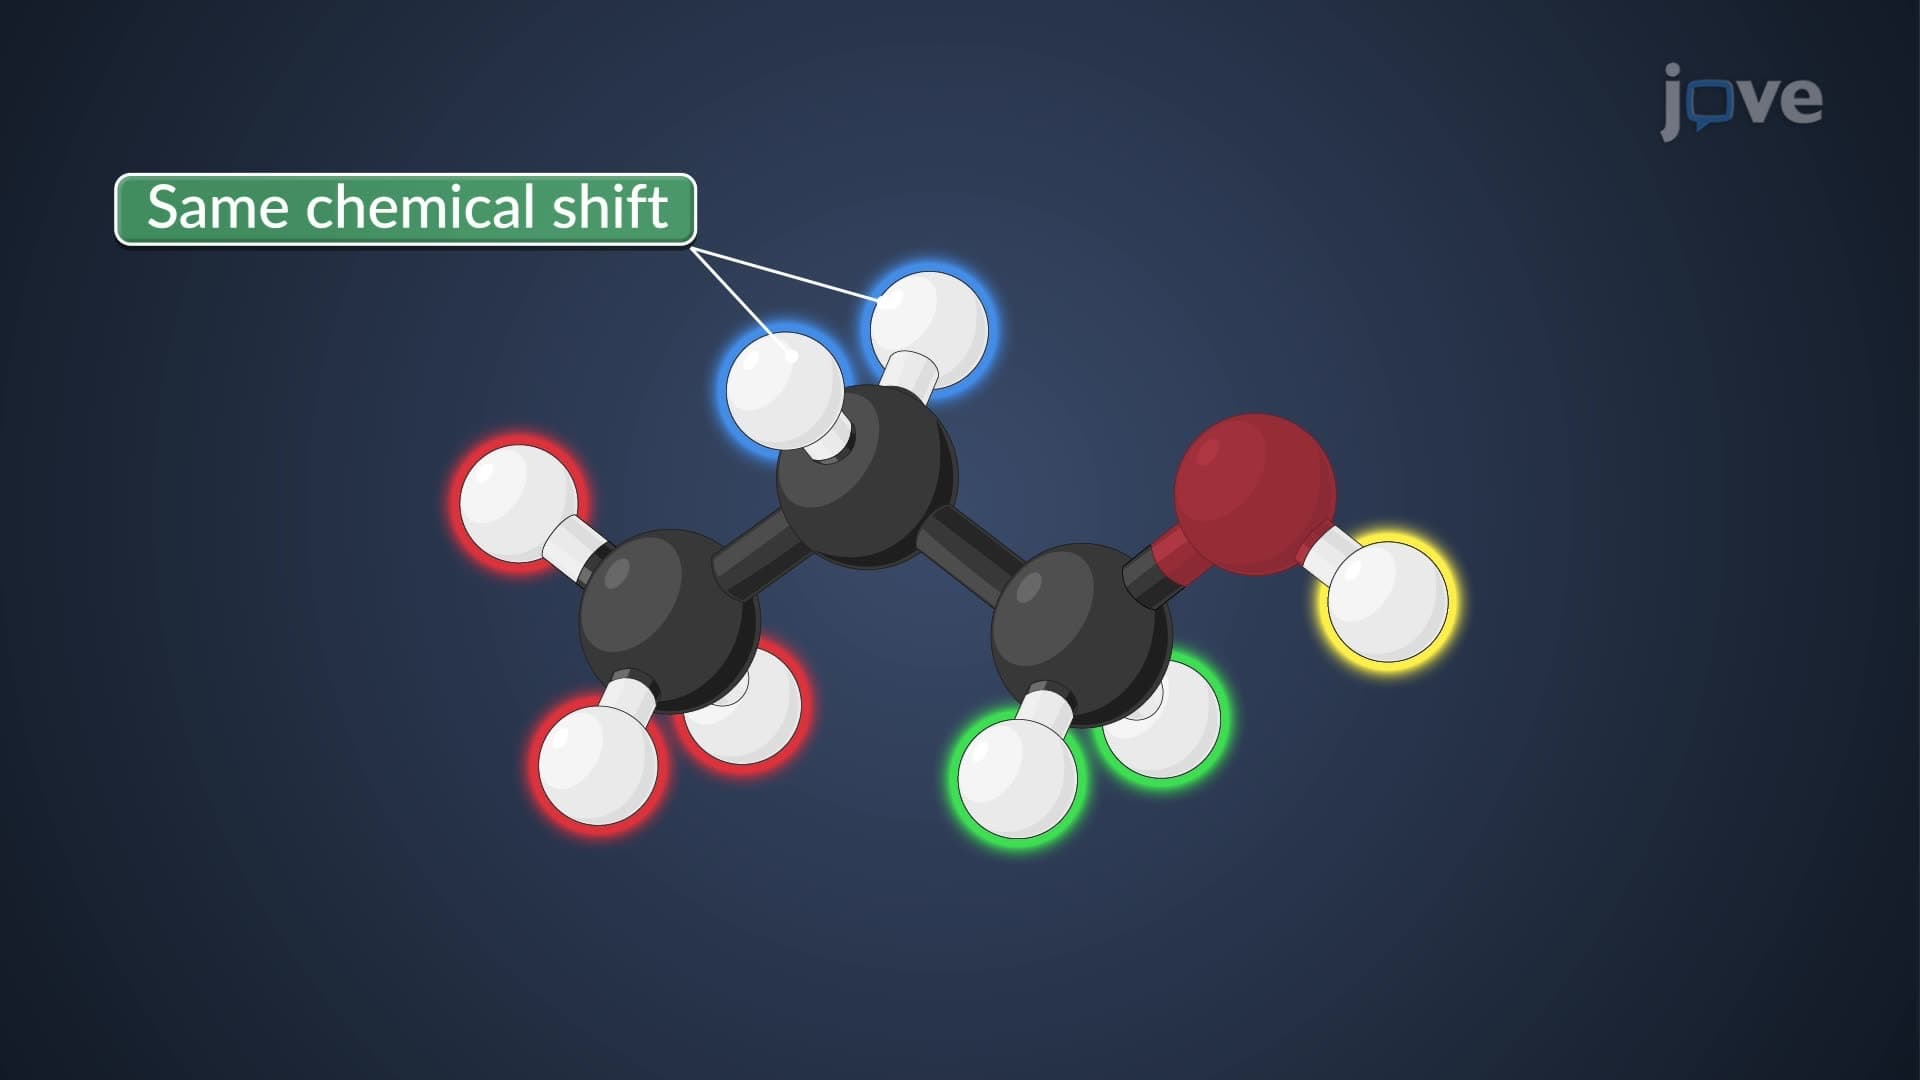
¹H NMR Chemical Shift Equivalence: Homotopic and Heterotopic Protons