Related Experiment Video
Updated: Jun 26, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Predicting fundamental gaps accurately from density functional theory with non-empirical local range separation
Moritz Brütting1, Hilke Bahmann2, Stephan Kümmel1
1Theoretical Physics IV, University of Bayreuth, 95440 Bayreuth, Germany.

Abstract:
We present an exchange-correlation approximation in which the Coulomb interaction is split into long- and short-range components and the range separation is determined by a non-empirical density functional. The functional respects important constraints, such as the homogeneous and slowly varying density limits, leads to the correct long-range potential, and eliminates one-electron self-interaction. Our approach is designed for spectroscopic purposes and closely approximates the piecewise linearity of the energy as a function of the particle number. The functional's accuracy for predicting the fundamental gap in generalized Kohn-Sham theory is demonstrated for a large number of systems, including organic semiconductors with a notoriously difficult electronic structure.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:52
VSEPR Theory and the Basic Shapes
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:40
MO Theory and Covalent Bonding
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:02
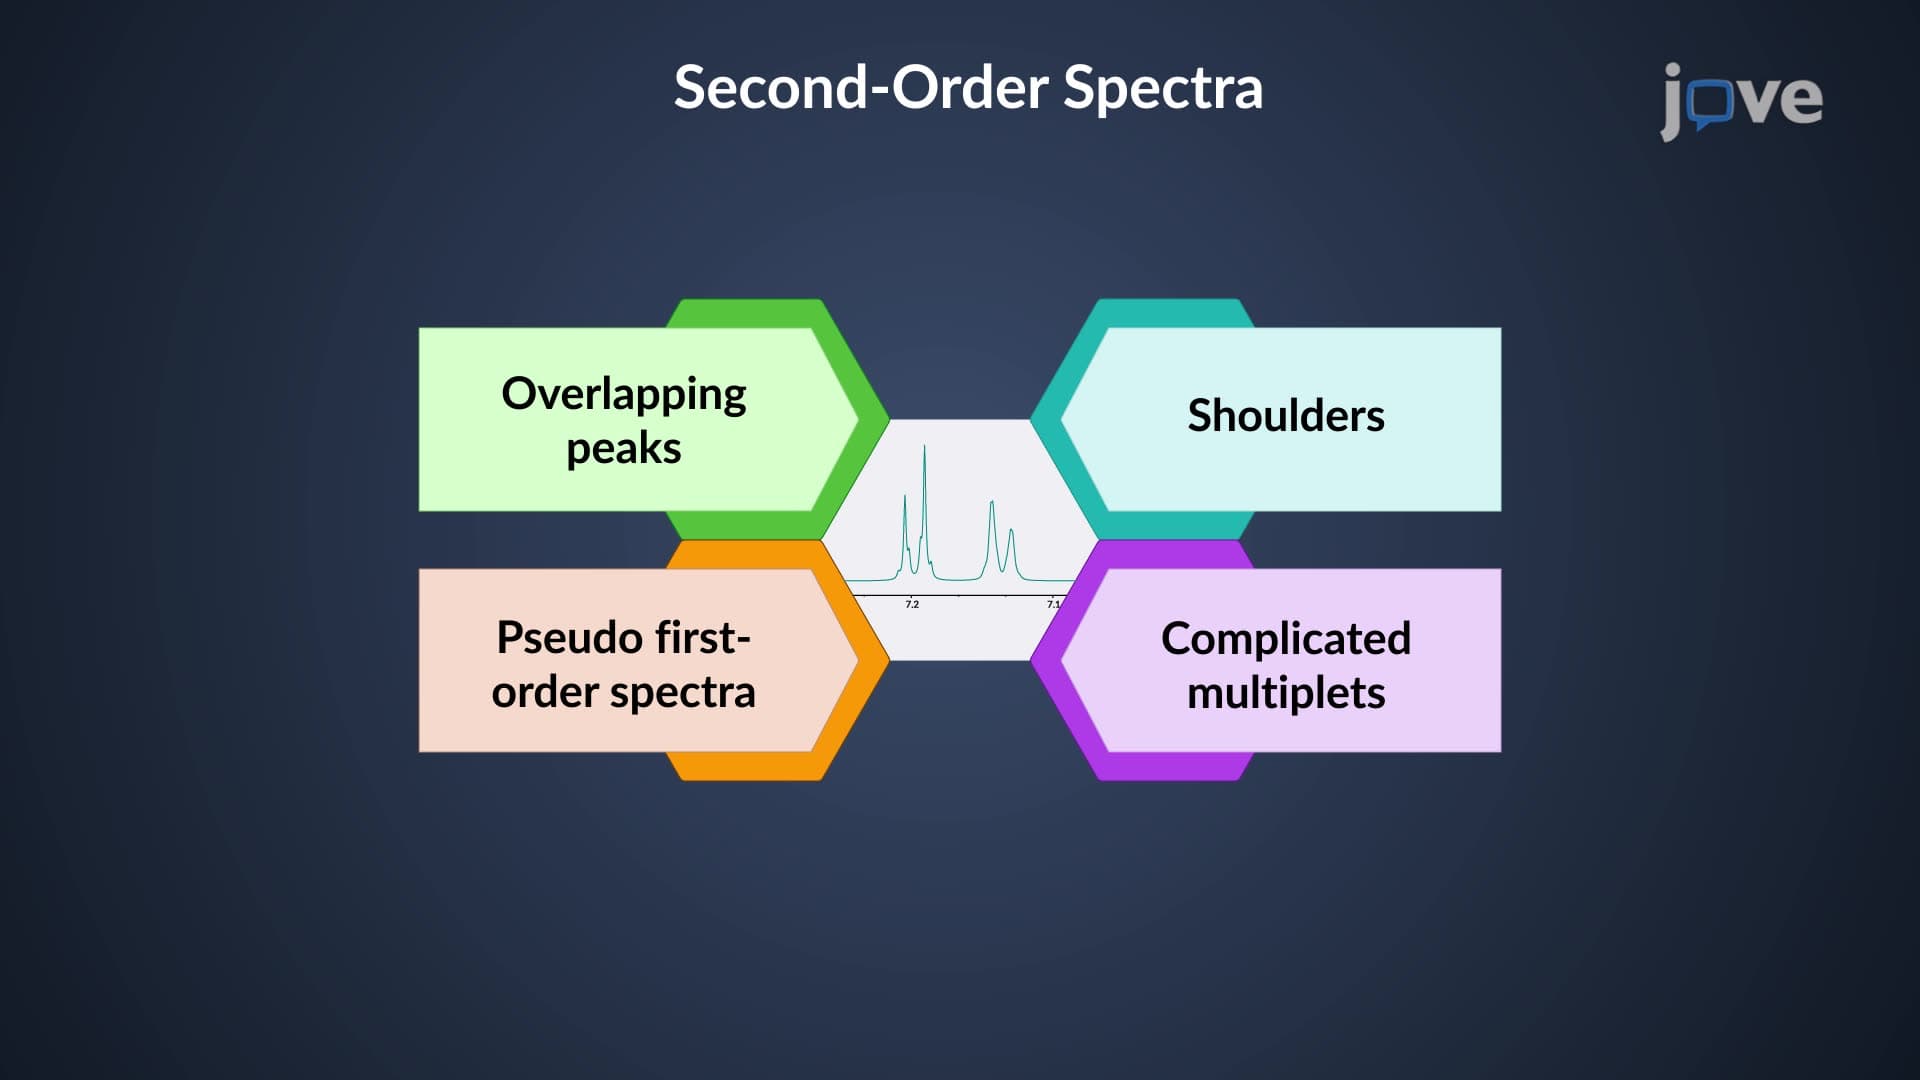
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...