Related Experiment Video
Updated: Jun 26, 2025


Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Leveraging an all-fixed transfer framework to predict the interpretable formation energy of MXenes with hybrid
Zihao Song1, Xiaobin Niu1, Haiyuan Chen1
1School of Materials and Energy, University of Electronic Science and Technology of China, Chengdu 611731, China. hychen@uestc.edu.cn.

Abstract:
MXenes have attracted substantial attention for their various applications in energy storage, sensors, and catalysts. Experimental exploration of MXenes with hybrid terminal surfaces offers a unique means of property tailoring that is crucial for expanding the performance space of MXenes, wherein the formation energy of an MXene with mixed surface terminals plays a key role in determining its relative stability and practical applications. However, the challenge of identifying energetically stable MXenes with multifunctional surfaces persists, primarily due to the absence of precise surface modification during experiments and the vast structural space for DFT calculations. Herein, we use an all-fixed transfer (AFT) framework combined with first-principles calculations to predict the formation energies of MXenes terminated by binary elements from groups VIA and VIIA. The trained model exhibits a high average R2 of 0.99, maintaining transferability and accuracy in predicting larger supercells from smaller-sized MXenes and datasets despite the structural imbalance between the training and prediction sets. The underlying interpretation of the high accuracy is revealed through the capture of main attributes and comparison of node features. Additionally, it is important to mention that the factors influencing the average formation energy include the types of element pairs, the ratio of terminal groups, and the distribution of terminations on two surfaces, with the first one being dominant. Finally, we successfully streamline the diverse structural cardinality of a large hybrid terminated MXene space of over 700 million, thereby facilitating the rapid screening of the top 5 stable MXene classes with binary terminal elements (FO, FCl, FBr, FS, and FSe). Besides, in the scenarios of lithium storage, the TL-predicted MXene can enhance its relative stability by increasing the fluorine ratio where the capacity can be optimized by different surface group combinations. All results indicate that the AFT framework has the advantages of screening functional MXenes with a huge structure space from smaller and imbalanced data sets.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:13
Energy Diagrams, Transition States, and Intermediates
 02:34
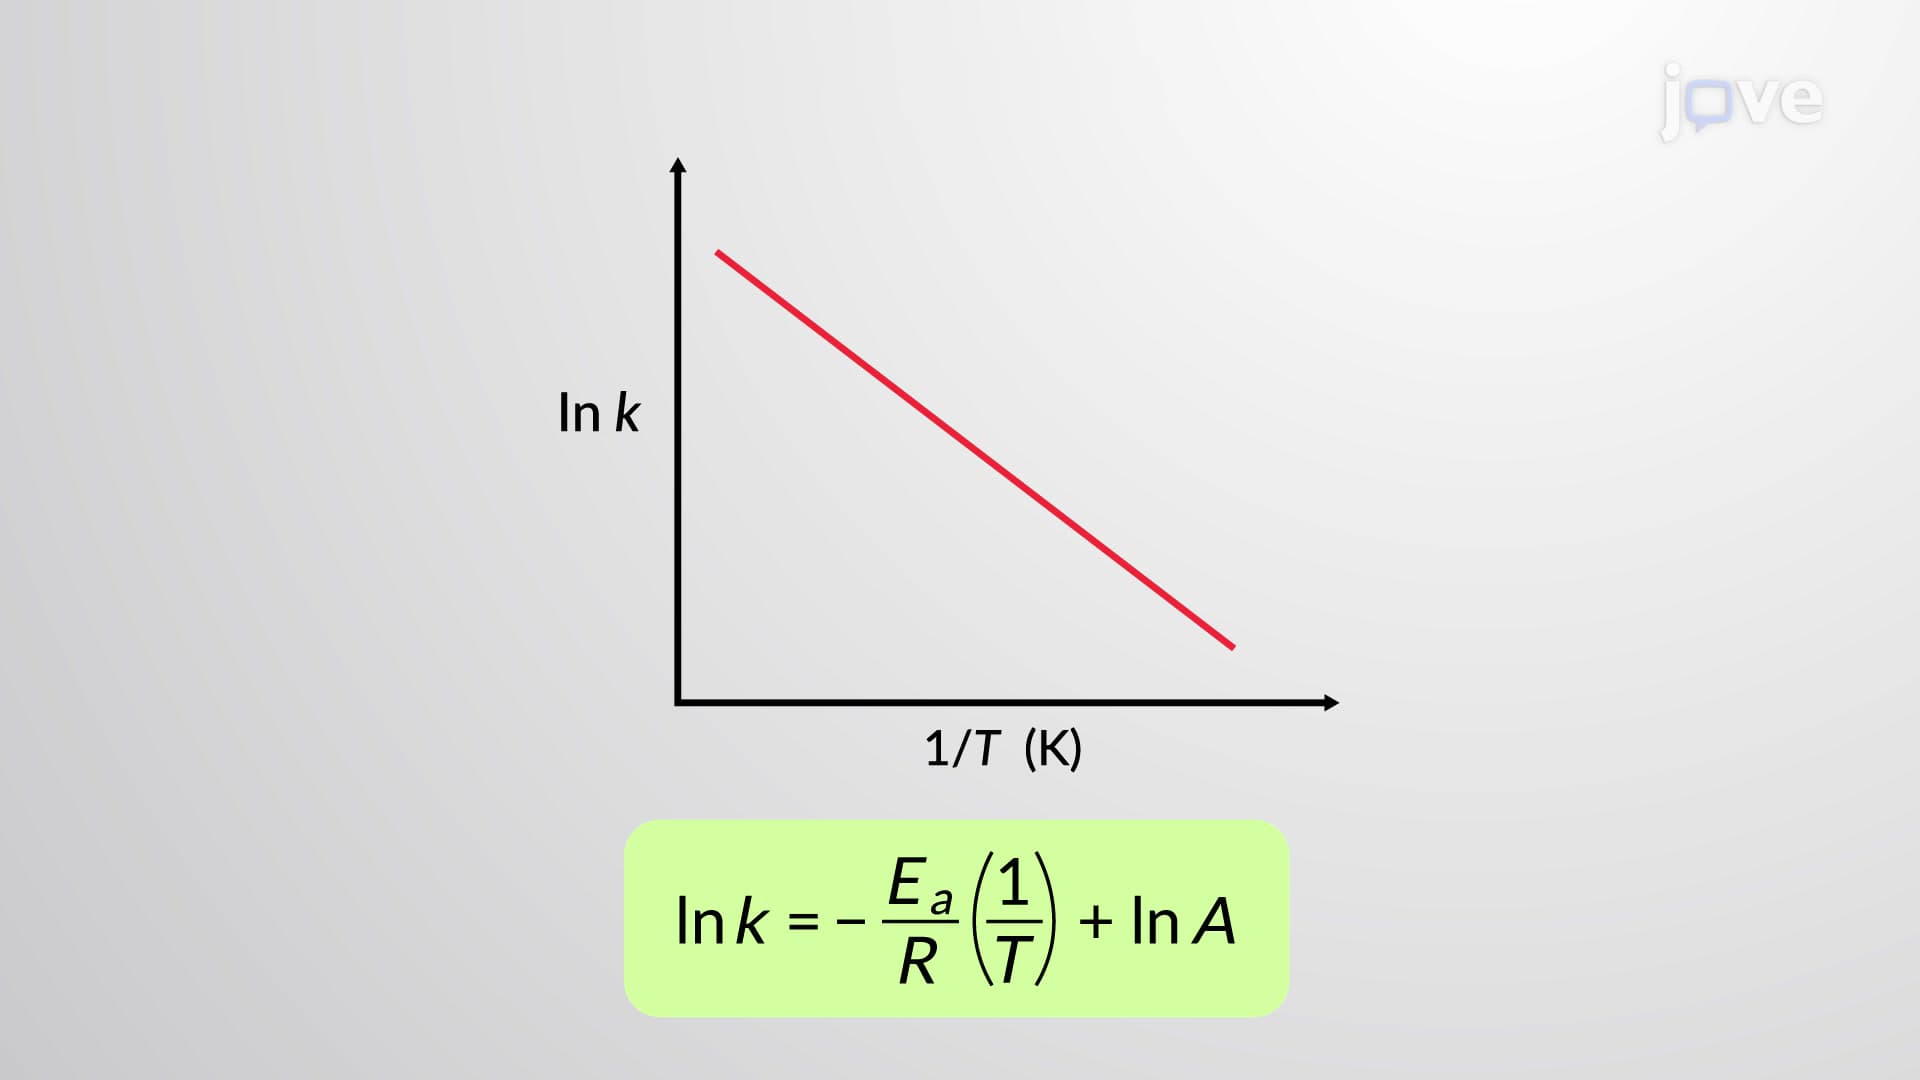
Arrhenius Plots
The Arrhenius equation can be used...
 03:51
Molecular Orbital Theory II
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...