Related Experiment Video
Updated: Jun 26, 2025
Facile Synthesis of Colloidal Lead Halide Perovskite Nanoplatelets via Ligand-Assisted Reprecipitation
Published on: October 1, 2019
Disentangling the effects of structure and lone-pair electrons in the lattice dynamics of halide perovskites
Sebastián Caicedo-Dávila1, Adi Cohen2, Silvia G Motti3,4
1Physics Department, TUM School of Natural Sciences, Technical University of Munich, Garching, Germany.

Abstract:
Halide perovskites show great optoelectronic performance, but their favorable properties are paired with unusually strong anharmonicity. It was proposed that this combination derives from the ns2 electron configuration of octahedral cations and associated pseudo-Jahn-Teller effect. We show that such cations are not a prerequisite for the strong anharmonicity and low-energy lattice dynamics encountered in these materials. We combine X-ray diffraction, infrared and Raman spectroscopies, and molecular dynamics to contrast the lattice dynamics of CsSrBr3 with those of CsPbBr3, two compounds that are structurally similar but with the former lacking ns2 cations with the propensity to form electron lone pairs. We exploit low-frequency diffusive Raman scattering, nominally symmetry-forbidden in the cubic phase, as a fingerprint of anharmonicity and reveal that low-frequency tilting occurs irrespective of octahedral cation electron configuration. This highlights the role of structure in perovskite lattice dynamics, providing design rules for the emerging class of soft perovskite semiconductors.
Related Concept Videos
 02:42
Ionic Crystal Structures
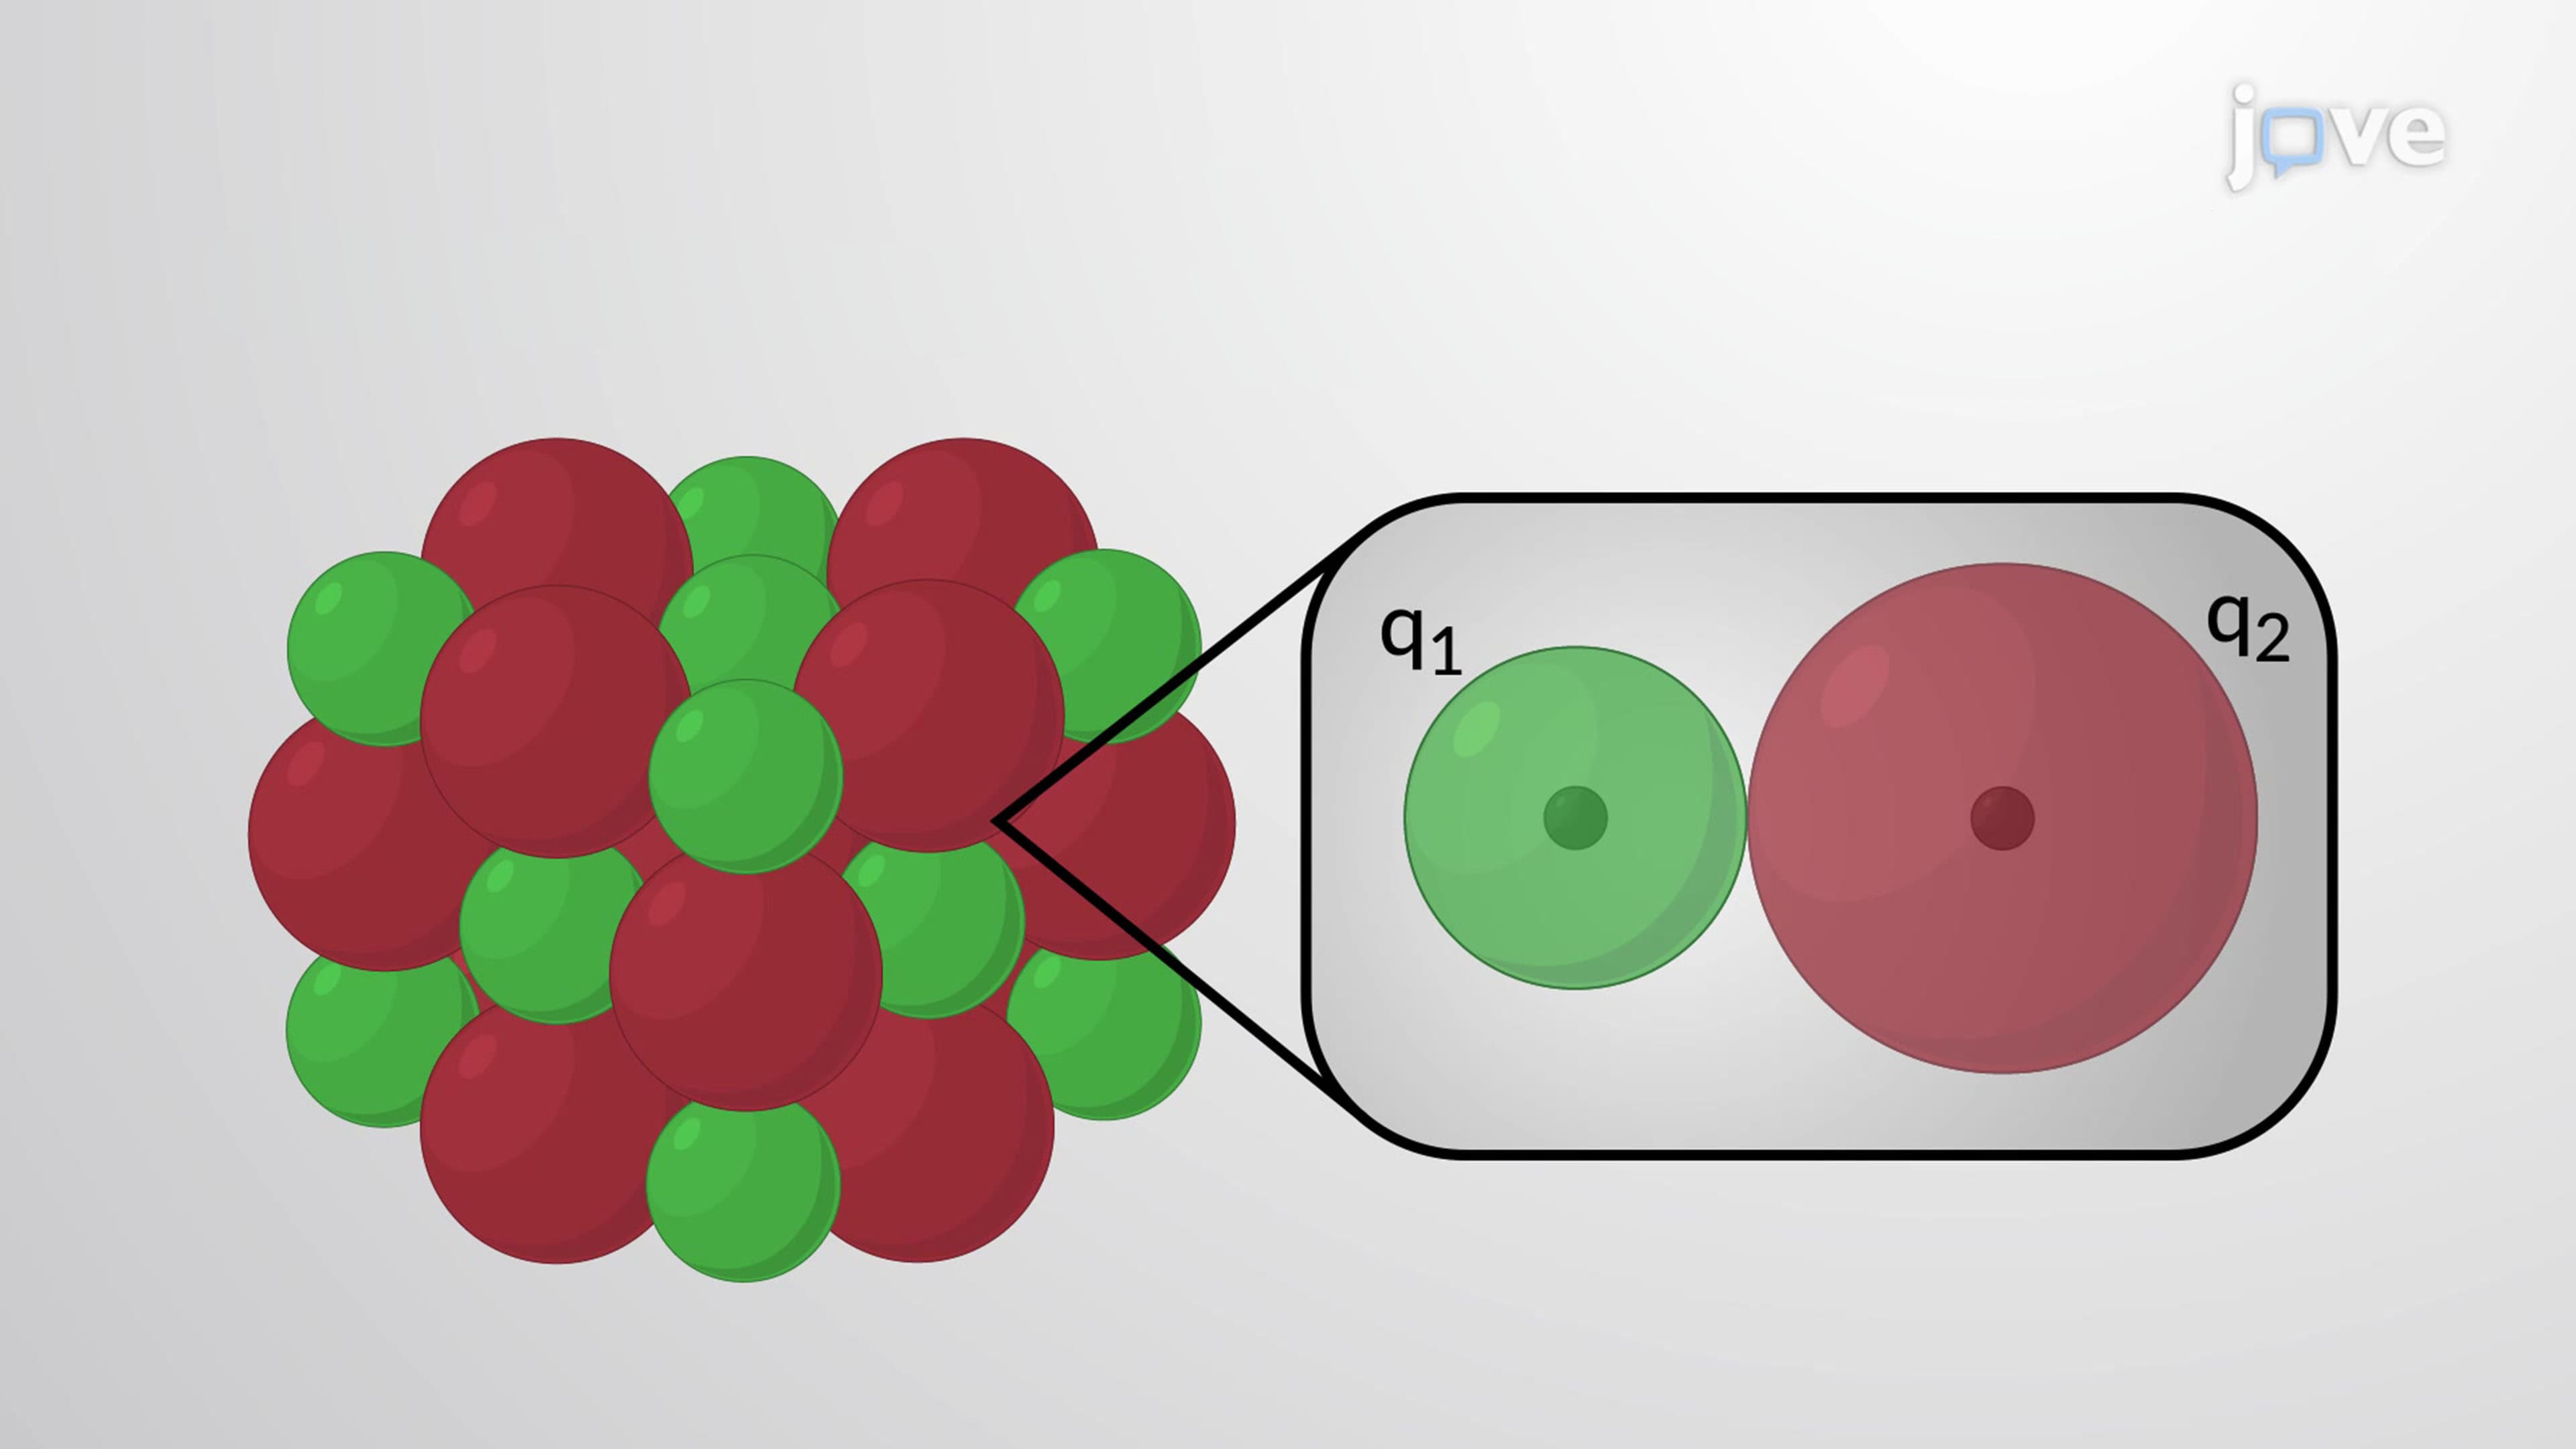
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 04:01
VSEPR Theory and the Effect of Lone Pairs
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:27
Predicting Molecular Geometry
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...