Related Experiment Video
Updated: Jun 25, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Going Beyond the GW Approximation Using the Time-Dependent Hartree-Fock Vertex
Simone Vacondio1,2, Daniele Varsano2, Alice Ruini1,2
1Dipartimento di Scienze Fisiche, Informatiche e Matematiche, Università degli Studi di Modena e Reggio Emilia, Via G. Campi 213/a, 41125 Modena, Italy.

Abstract:
The time-dependent Hartree-Fock (TDHF) vertex of many-body perturbation theory (MBPT) makes it possible to extend TDHF theory to charged excitations. Here we assess its performance by applying it to spherical atoms in their neutral electronic configuration. On a theoretical level, we recast the TDHF vertex as a reducible vertex, highlighting the emergence of a self-energy expansion purely in orders of the bare Coulomb interaction; then, on a numerical level, we present results for polarizabilities, ionization energies (IEs), and photoemission satellites. We confirm the superiority of THDF over simpler methods such as the random phase approximation for the prediction of atomic polarizabilities. We then find that the TDHF vertex reliably provides better IEs than GW and low-order self-energies do in the light-atom, few-electron regime; its performance degrades in heavier, many-electron atoms instead, where an expansion in orders of an unscreened Coulomb interaction becomes less justified. New relevant features are introduced in the satellite spectrum by the TDHF vertex, but the experimental spectra are not fully reproduced due to a missing account of nonlinear effects connected to hole relaxation. We also explore various truncations of the self-energy given by the TDHF vertex, but do not find them to be more convenient than low-order approximations such as GW and second Born (2B), suggesting that vertex corrections should be carried out consistently both in the self-energy and in the polarizability.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:12
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:10
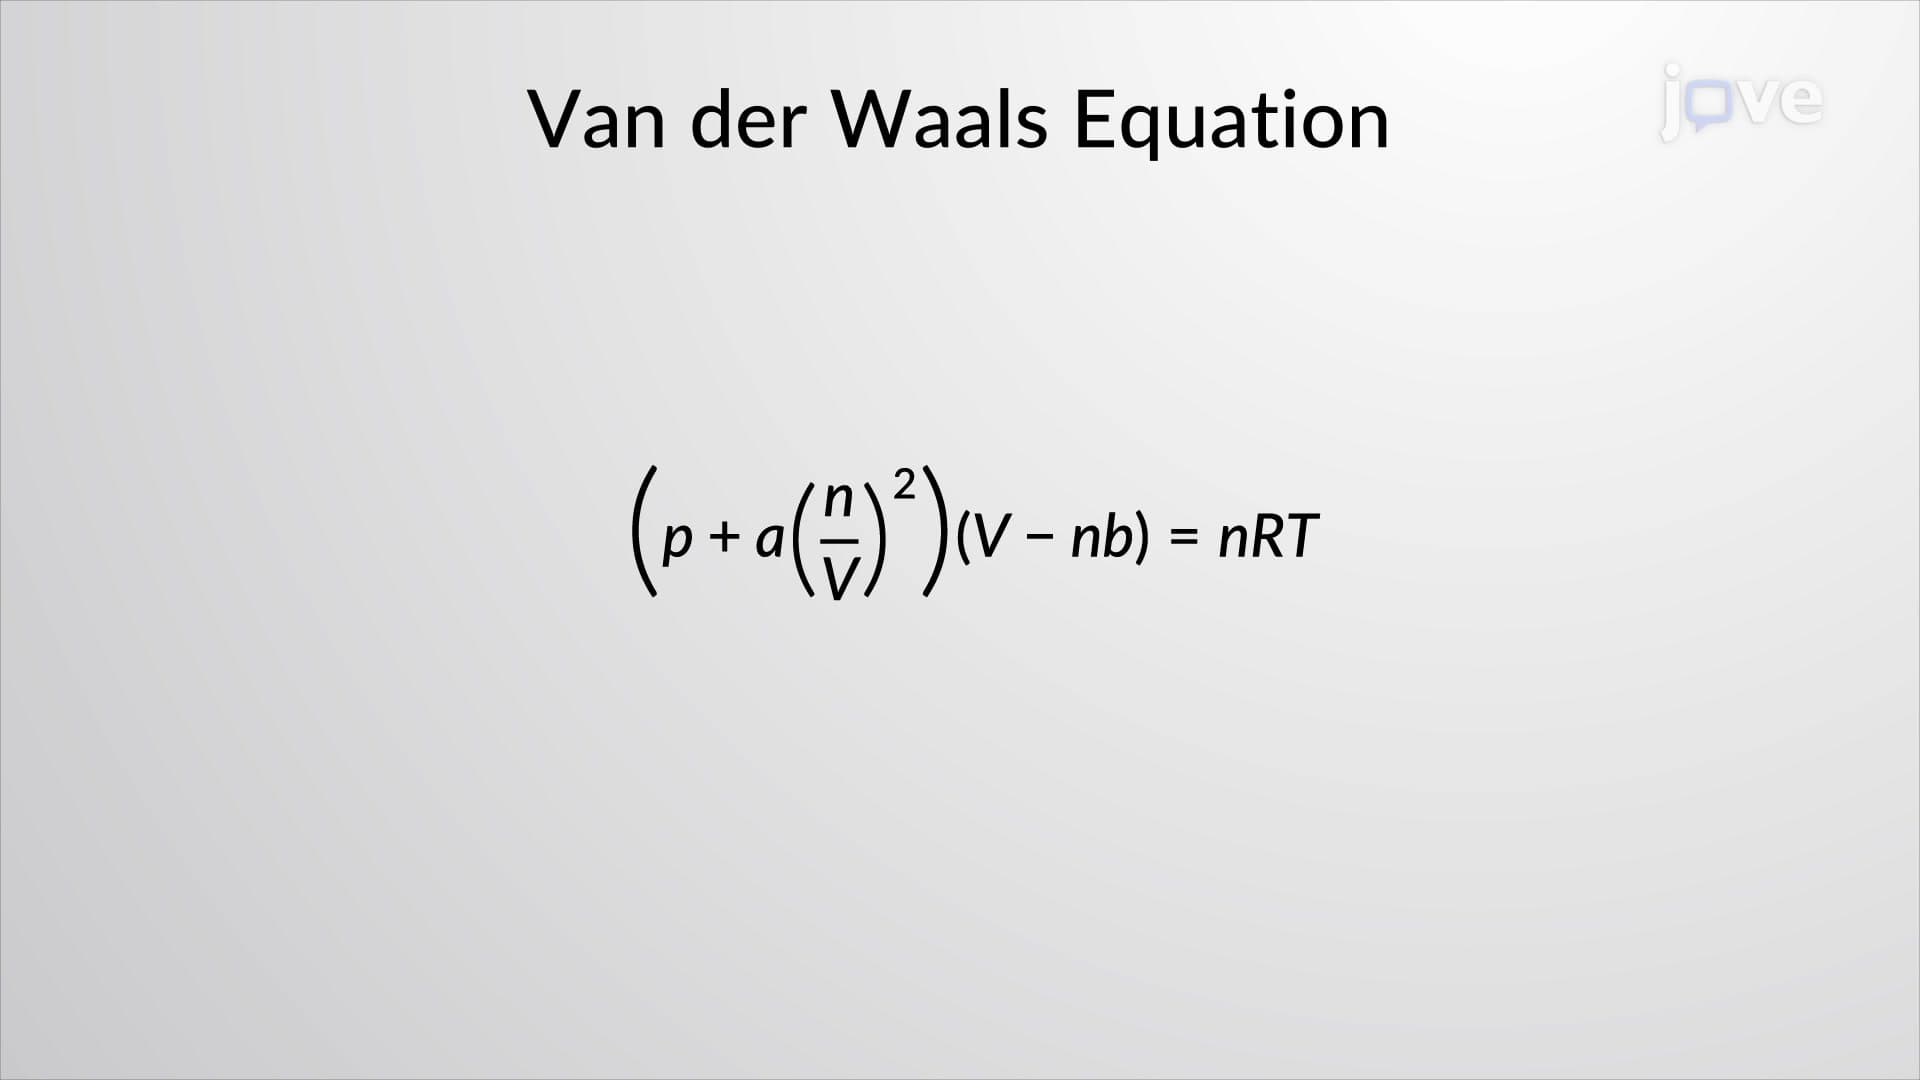
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation