Related Experiment Video
Updated: Jun 25, 2025

Multicolor Flow Cytometry-based Quantification of Mitochondria and Lysosomes in T Cells
Published on: January 9, 2019
Upregulation of cholesterol synthesis by lysosomal defects requires a functional mitochondrial respiratory chain
Francesco Agostini1, Leonardo Pereyra2, Justin Dale1
1Department of Cellular and Molecular Physiology, Penn State College of Medicine, Hershey, Pennsylvania, USA.

Abstract:
Mitochondria and lysosomes are two organelles that carry out both signaling and metabolic roles in cells. Recent evidence has shown that mitochondria and lysosomes are dependent on one another, as primary defects in one cause secondary defects in the other. Although there are functional impairments in both cases, the signaling consequences of primary mitochondrial dysfunction and lysosomal defects are dissimilar. Here, we used RNA sequencing to obtain transcriptomes from cells with primary mitochondrial or lysosomal defects to identify the global cellular consequences associated with mitochondrial or lysosomal dysfunction. We used these data to determine the pathways affected by defects in both organelles, which revealed a prominent role for the cholesterol synthesis pathway. We observed a transcriptional upregulation of this pathway in cellular and murine models of lysosomal defects, while it is transcriptionally downregulated in cellular and murine models of mitochondrial defects. We identified a role for the posttranscriptional regulation of transcription factor SREBF1, a master regulator of cholesterol and lipid biosynthesis, in models of mitochondrial respiratory chain deficiency. Furthermore, we found that retention of Ca2+ in lysosomes of cells with mitochondrial respiratory chain defects contributes to the differential regulation of the cholesterol synthesis pathway in the mitochondrial and lysosomal defects tested. Finally, we verified in vivo, using a model of mitochondria-associated disease in Caenorhabditis elegans that normalization of lysosomal Ca2+ levels results in partial rescue of the developmental delay induced by the respiratory chain deficiency.
More Related Videos
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:39
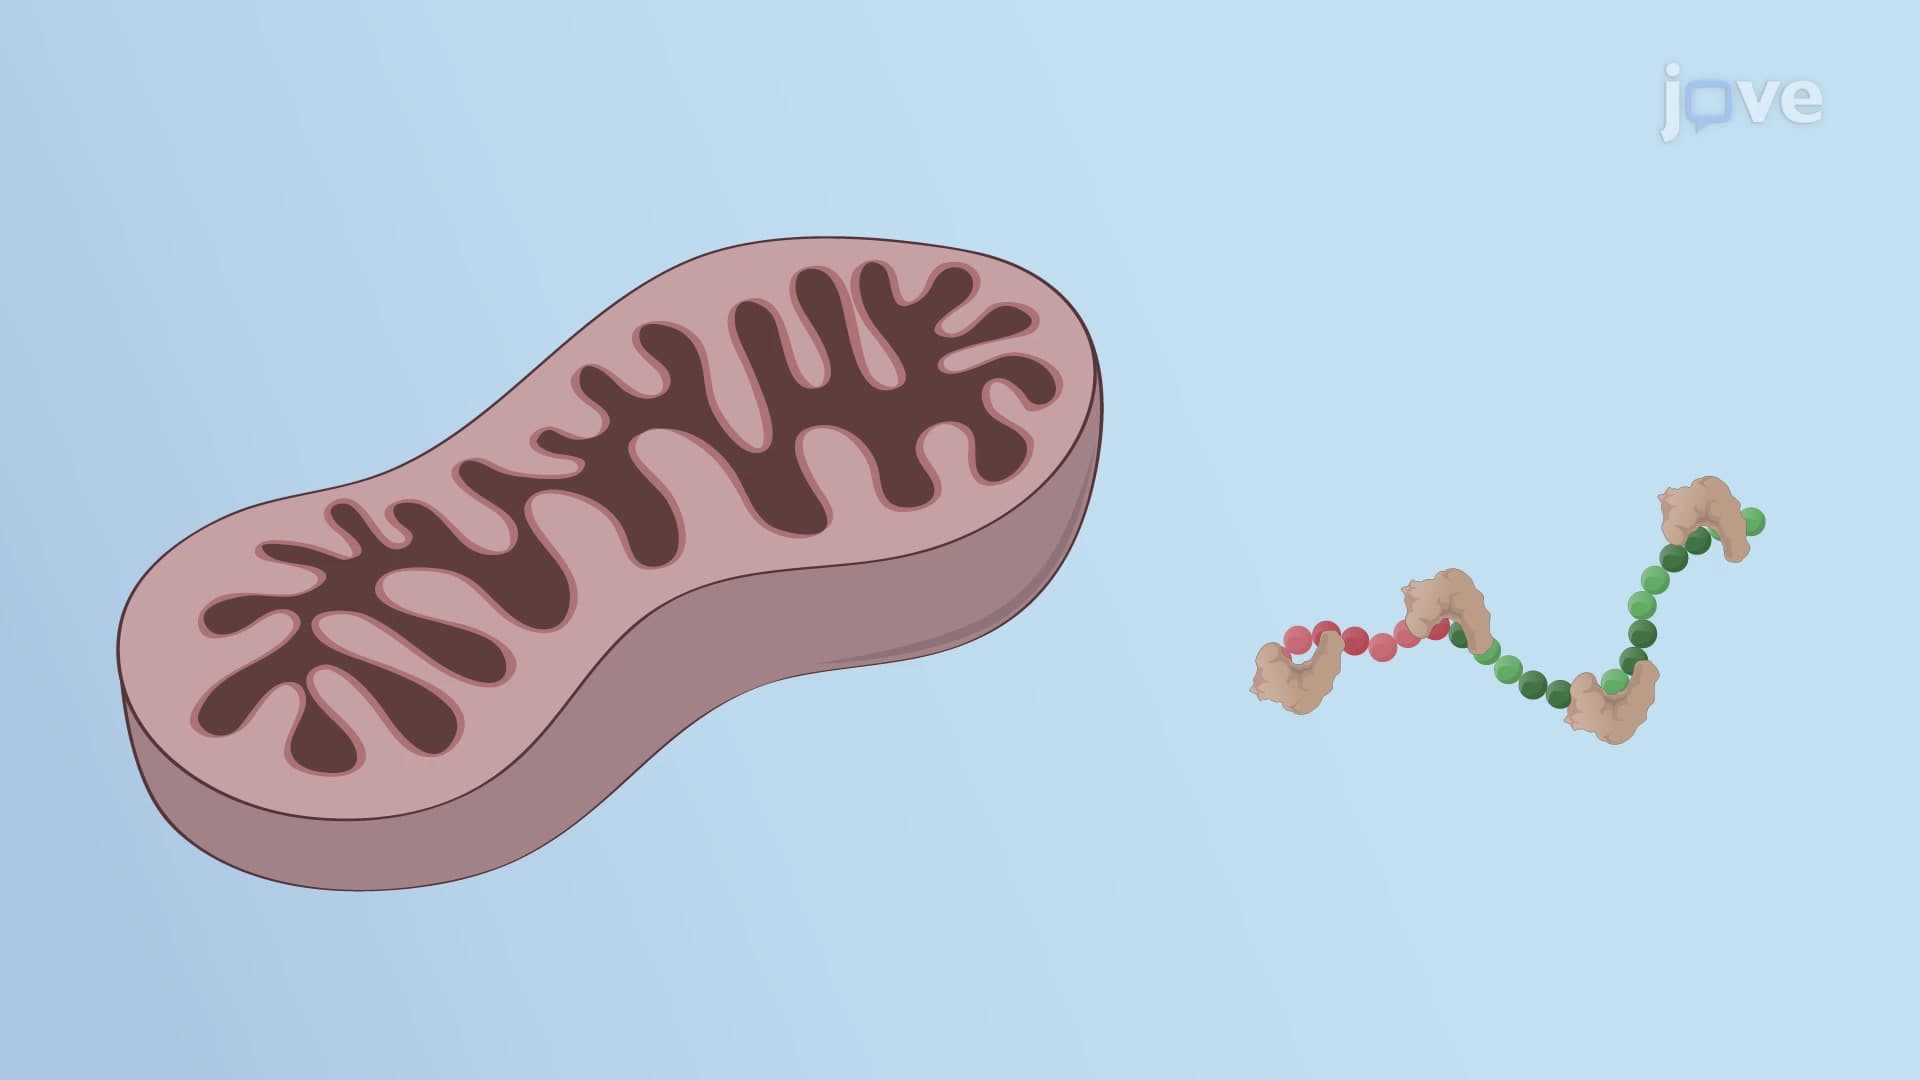
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:41
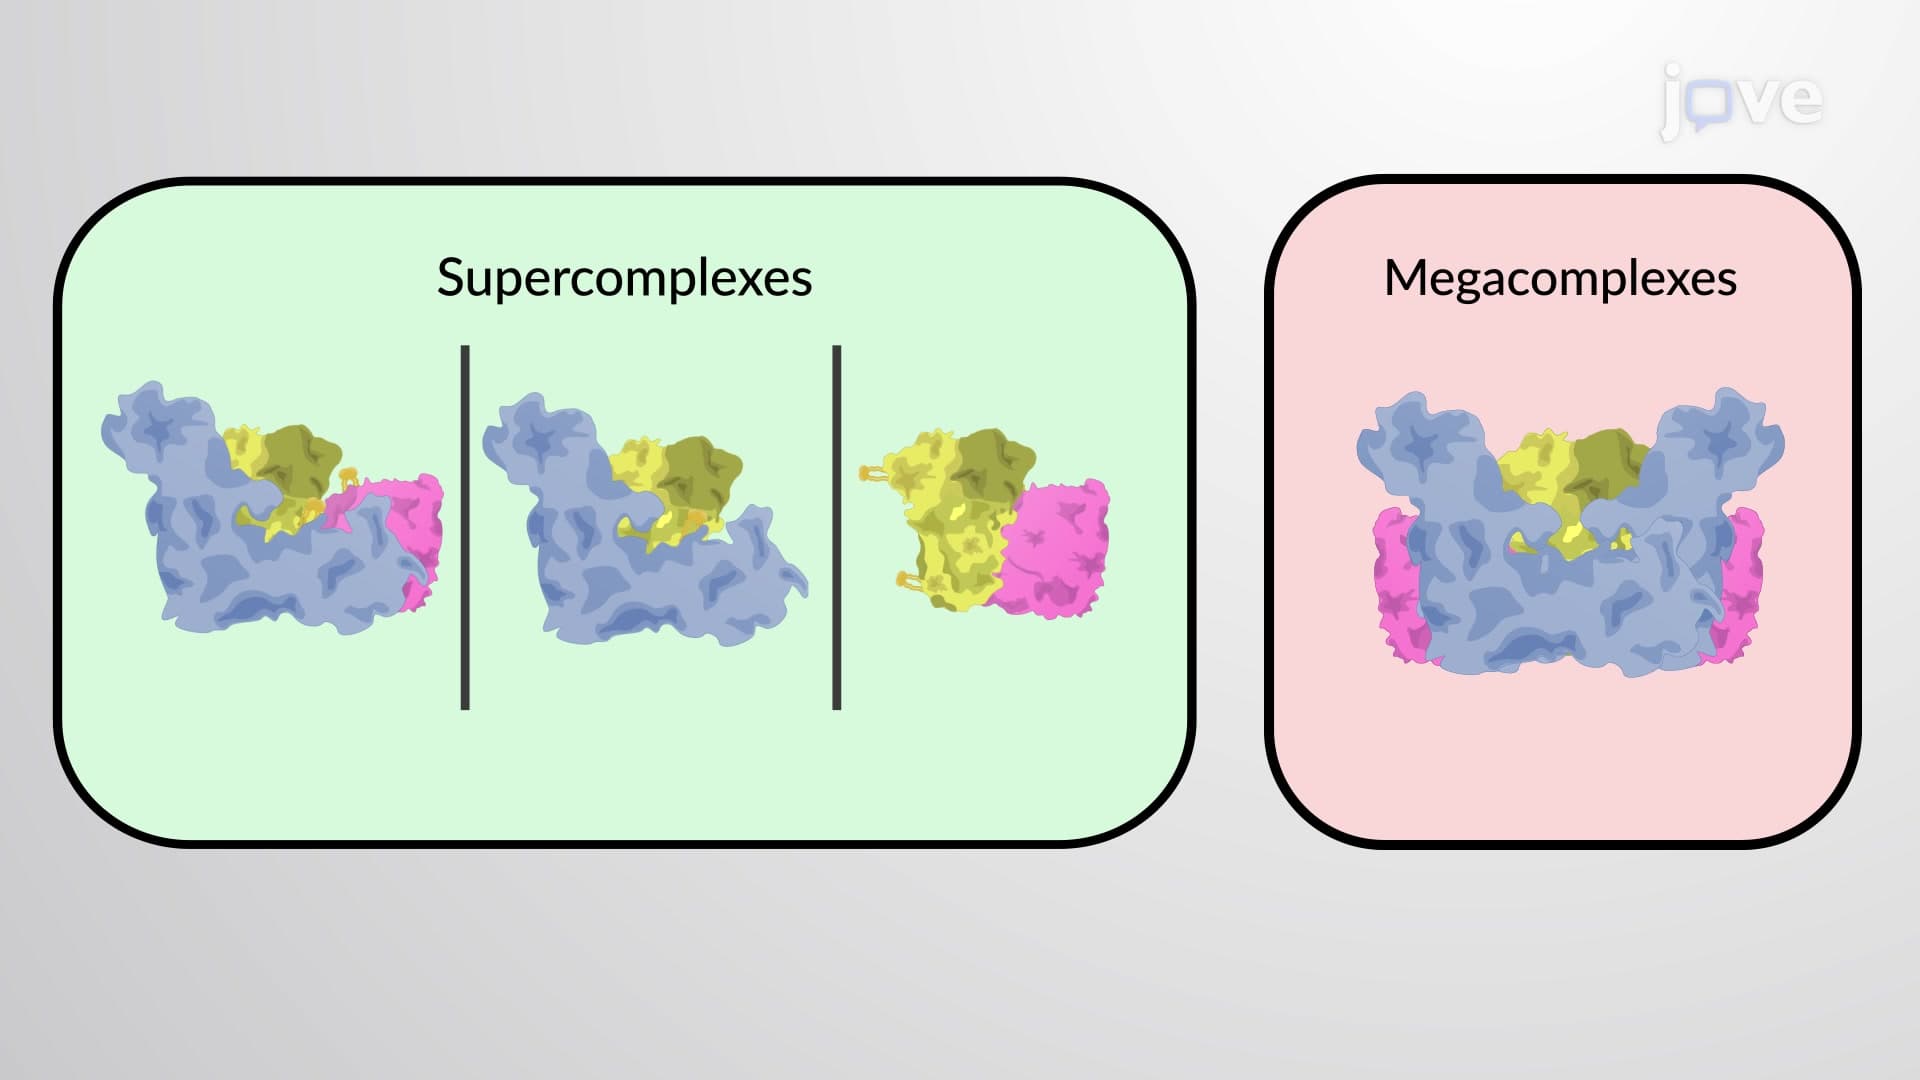
The Supercomplexes in the Crista Membrane
 01:30
The Electron Transport Chain
Inhibitors of the electron transport chain
Rotenone, a widely used pesticide, prevents electron transfer from Fe-S cluster to ubiquinone or Q...
 01:45
Mitochondrial Membranes