Related Experiment Video
Updated: Jun 24, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Amber free energy tools: Interoperable software for free energy simulations using generalized quantum
Yujun Tao1, Timothy J Giese1, Şölen Ekesan1
1Laboratory for Biomolecular Simulation Research, Institute for Quantitative Biomedicine and Department of Chemistry and Chemical Biology, Rutgers University, Piscataway, New Jersey 08854, USA.

Abstract:
We report the development and testing of new integrated cyberinfrastructure for performing free energy simulations with generalized hybrid quantum mechanical/molecular mechanical (QM/MM) and machine learning potentials (MLPs) in Amber. The Sander molecular dynamics program has been extended to leverage fast, density-functional tight-binding models implemented in the DFTB+ and xTB packages, and an interface to the DeePMD-kit software enables the use of MLPs. The software is integrated through application program interfaces that circumvent the need to perform "system calls" and enable the incorporation of long-range Ewald electrostatics into the external software's self-consistent field procedure. The infrastructure provides access to QM/MM models that may serve as the foundation for QM/MM-ΔMLP potentials, which supplement the semiempirical QM/MM model with a MLP correction trained to reproduce ab initio QM/MM energies and forces. Efficient optimization of minimum free energy pathways is enabled through a new surface-accelerated finite-temperature string method implemented in the FE-ToolKit package. Furthermore, we interfaced Sander with the i-PI software by implementing the socket communication protocol used in the i-PI client-server model. The new interface with i-PI allows for the treatment of nuclear quantum effects with semiempirical QM/MM-ΔMLP models. The modular interoperable software is demonstrated on proton transfer reactions in guanine-thymine mispairs in a B-form deoxyribonucleic acid helix. The current work represents a considerable advance in the development of modular software for performing free energy simulations of chemical reactions that are important in a wide range of applications.
More Related Videos

Related Concept Videos
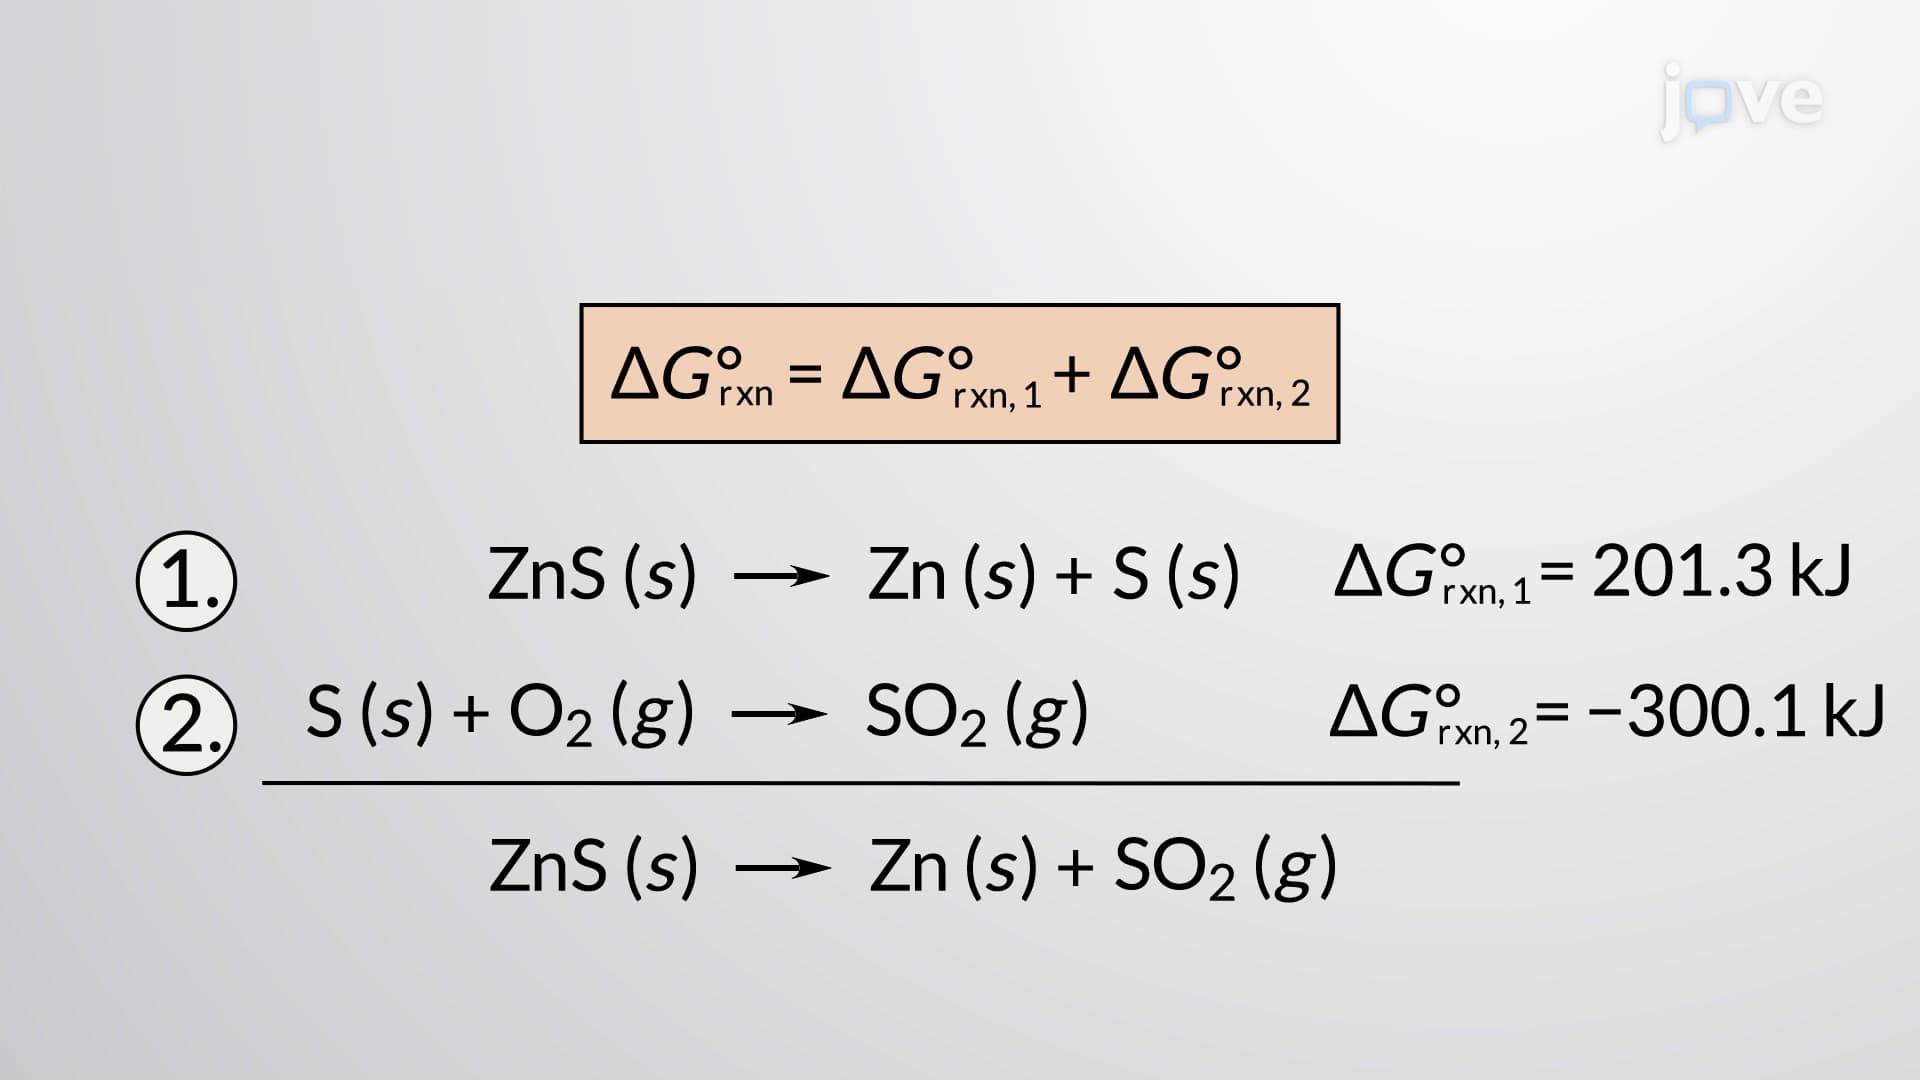 02:49
Calculating Standard Free Energy Changes
 01:26
Thermodynamic Potentials
 03:25
Free Energy Changes for Nonstandard States
where R is the gas constant (8.314 J/K·mol), T is the absolute temperature in kelvin, and Q is the reaction quotient. This equation may be used to predict the spontaneity of a process under any given set of conditions.
Reaction Quotient...
 01:21
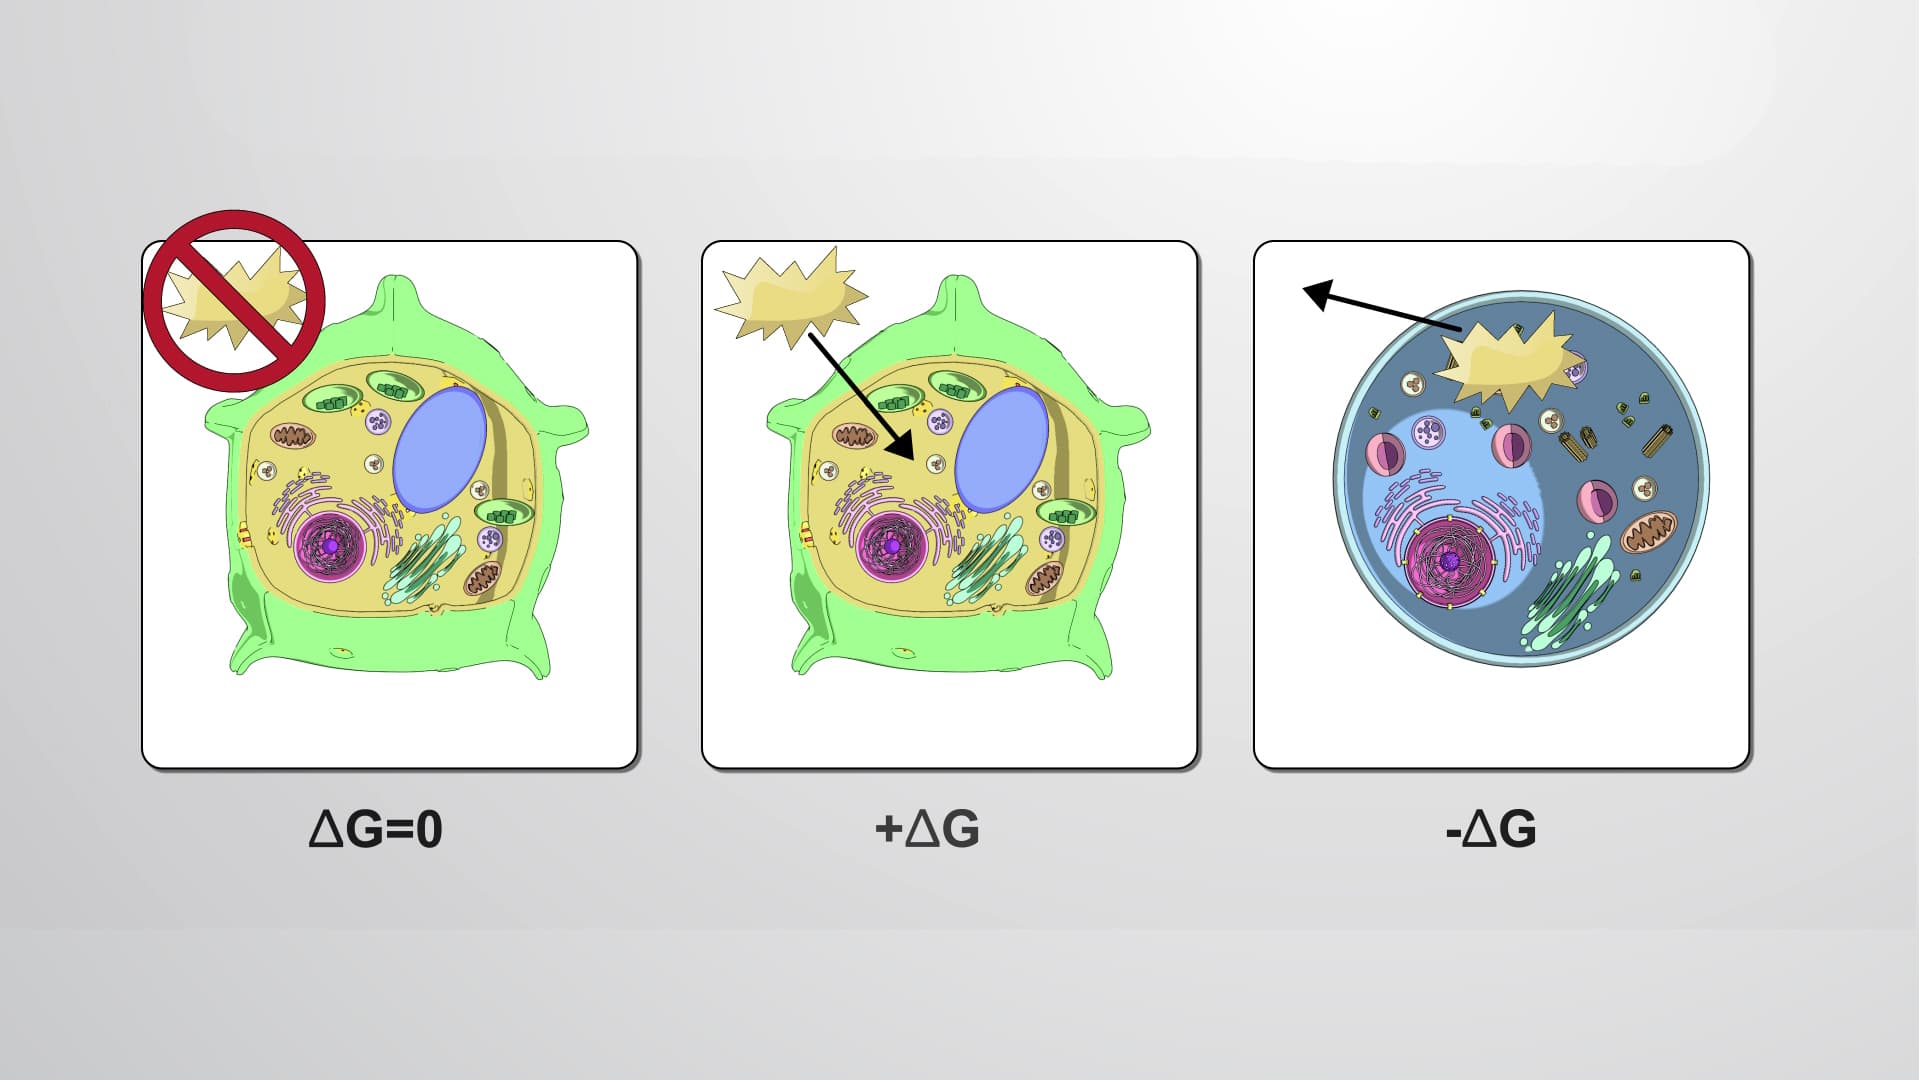
Free Energy
 01:05
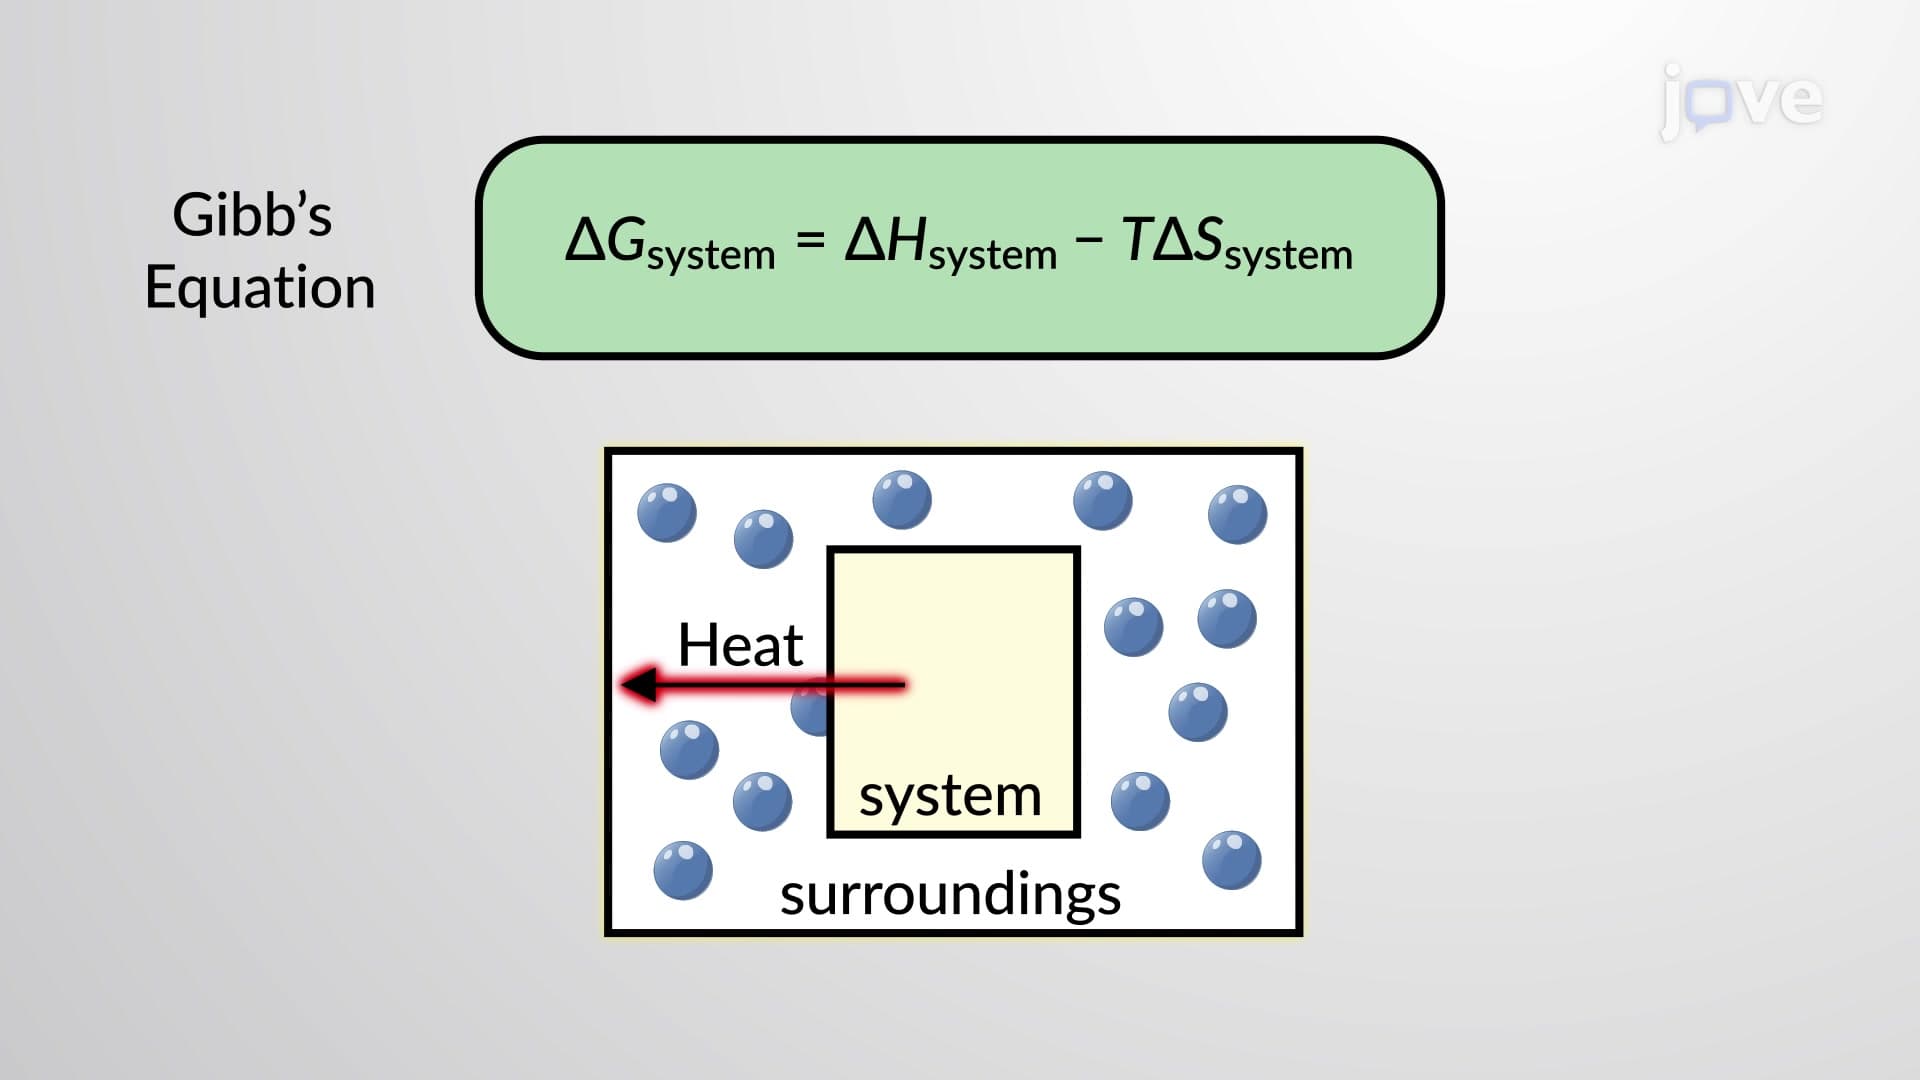
An Introduction to Free Energy
 01:21
Molecular Kinetic Energy