Related Experiment Video
Updated: Jun 23, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A many-body energy decomposition analysis (MB-EDA) scheme based on a target state optimization self-consistent field
Zhen Tang1,2, Hong Zhu1,2, Zhijun Pan2
1Peking University Shenzhen Graduate School, Shenzhen, Guangdong 518055, People's Republic of China. jiali@jiaoligao.org.

Abstract:
In this paper, we combine an energy decomposition analysis (EDA) scheme with many-body expansion (MBE) to develop a MB-EDA method to study the cooperative and anti-cooperative effects in molecular cluster systems. Based on the target state optimization self-consistent field (TSO-SCF) method, the intermolecular interaction energy can be decomposed into five chemically meaningful terms, i.e., electrostatic, exchange, polarization, charge transfer and dispersion interaction energies. MB-EDA can decompose each of these terms in MBE. This MB-EDA has been applied to 3 different cluster systems: water clusters, ionic liquid clusters, and acetonitrile-methane clusters. This reveals that electrostatic, exchange, and dispersion interactions are highly pairwise additive in all systems. In water and ionic liquid clusters, the many-body effects are significant in both polarization and charge transfer interactions, but are cooperative and anti-cooperative, respectively. For acetonitrile-methane clusters, which do not involve hydrogen bonds or charge-charge Coulombic interactions, the many-body effects are quite small. The chemical origins of different many-body effects are deeply analyzed. The MB-EDA method has been implemented in Qbics (https://qbics.info) and can be a useful tool for understanding the many-body behavior in molecular aggregates at the quantum chemical level of theory.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 03:25
Free Energy Changes for Nonstandard States
where R is the gas constant (8.314 J/K·mol), T is the absolute temperature in kelvin, and Q is the reaction quotient. This equation may be used to predict the spontaneity of a process under any given set of conditions.
Reaction Quotient...
 01:16
Potential-Energy Criterion for Equilibrium
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:12
Fermi Level Dynamics
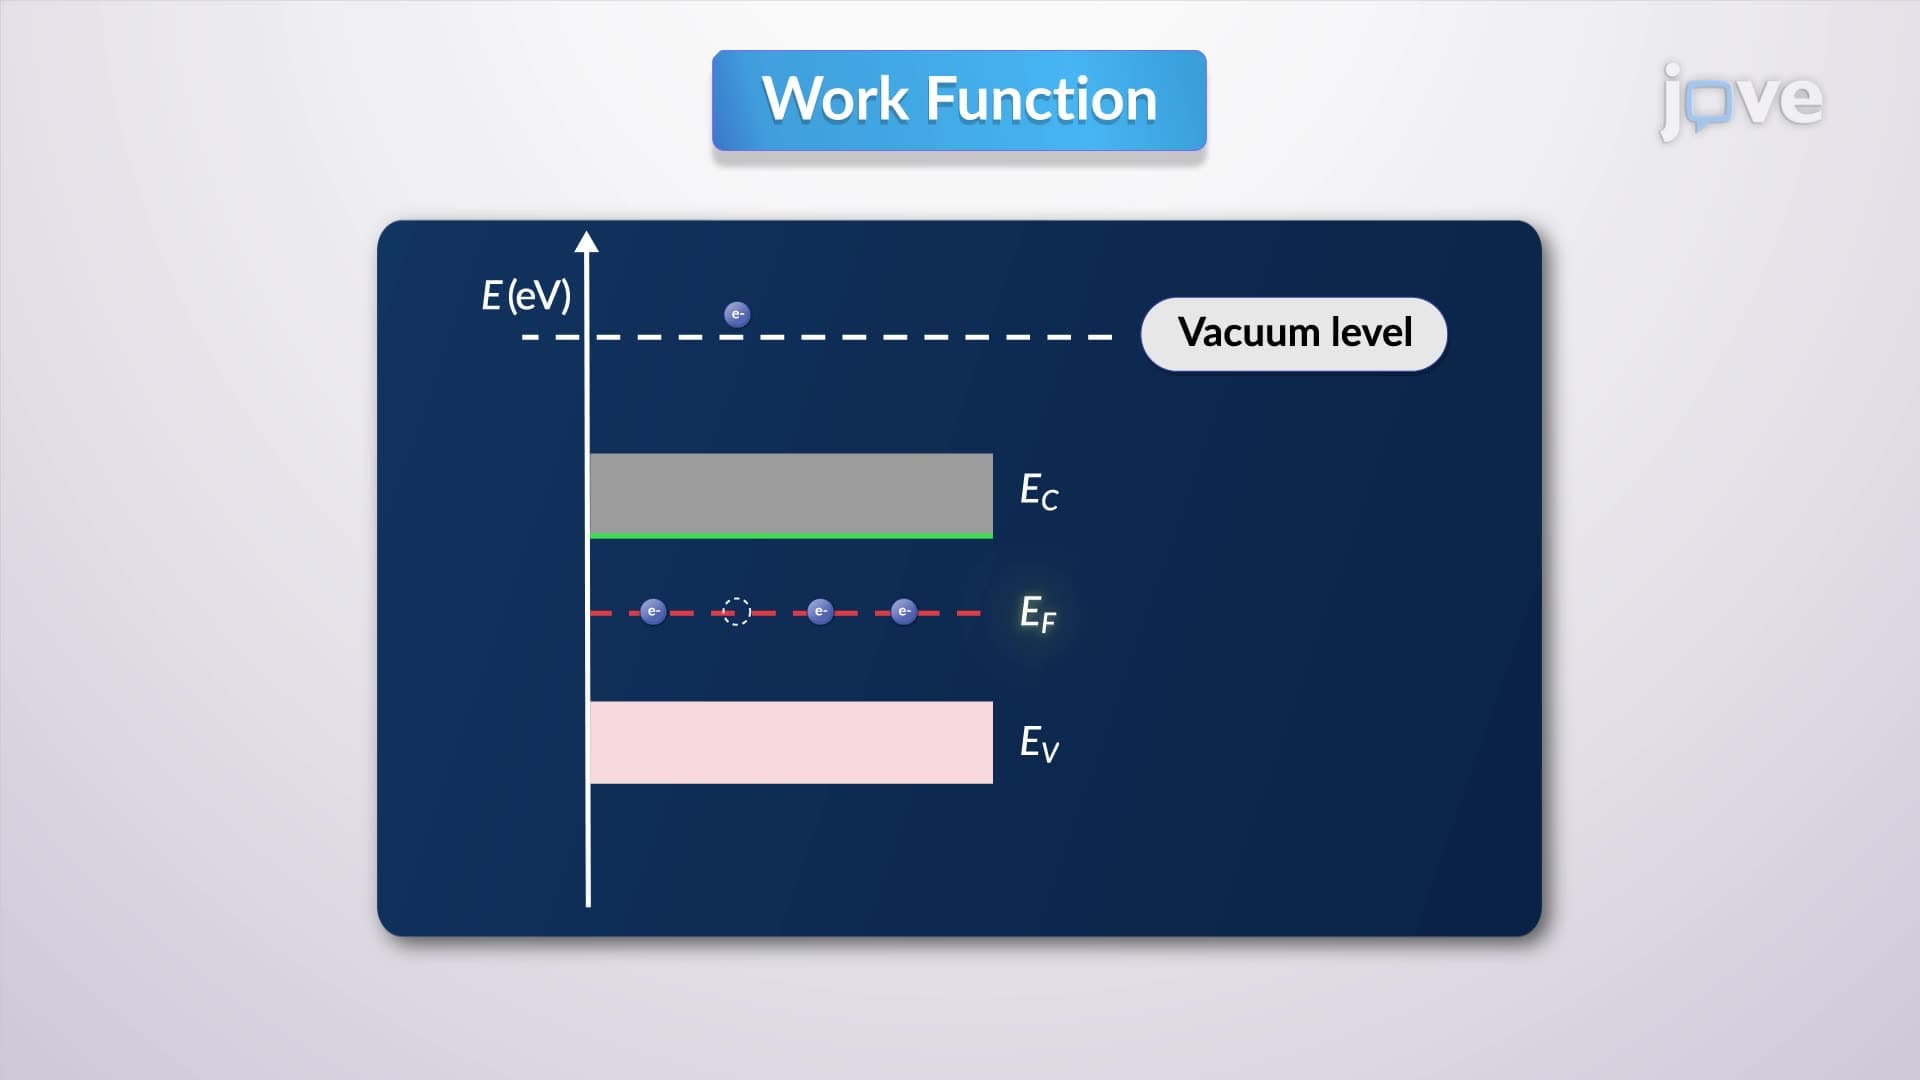
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 01:13
Force and Potential Energy in One Dimension