Related Experiment Video
Updated: Jun 23, 2025


Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Predicting spin states of iron porphyrins with DFT methods including crystal packing effects and thermodynamic
1Faculty of Chemistry, Jagiellonian University, Kraków, Poland. mradon@chemia.uj.edu.pl.

Abstract:
Accurate computational treatment of spin states for transition metal complexes, exemplified by iron porphyrins, lies at the heart of quantum bioinorganic chemistry, but at the same time represents a great challenge for approximate density functional theory (DFT) methods, which are predominantly used. Here, the accuracy of DFT methods for spin-state splittings in iron porphyrin is assessed by probing the ability to correctly predict the ground states for six FeIII or FeII complexes experimentally characterized in solid state. For each case, molecular and periodic DFT calculations are employed to quantify the effect of porphyrin side substituents and the crystal packing effect (CPE) on the spin-state splitting. It is proposed to partition the total CPE into additive components, the direct and structural one, the importance of which is shown to significantly vary from case to case. By knowing the substituent effect, the CPE, and the Gibbs free energy thermodynamic correction from calculations, one can employ the experimental ground-state information in order to derive a quantitative constraint on the electronic energy difference for a simplified (porphin) model of the experimentally characterized metalloporphyrin. The constraints derived in such a way-in the form of single or double inequalities-are used to assess the accuracy of dispersion-corrected DFT methods for 6 spin-state splittings of [FeIII(P)(2-MeIm)2]+, [FeIII(P)(2-MeIm)]+, [FeII(P)(THF)2] and [FeII(P)] models (where P is porphin, 2-MeIm is 2-methylimidazole, THF is tetrahydrofuran). These data constitute the new benchmark set of spin states for crystalline iron porphyrins (SSCIP6). The highest accuracy is obtained in the case of double-hybrid functionals (B2PLYP-D3, DSD-PBEB95-D3), whereas hybrid functionals, especially those with reduced admixture of the exact exchange (B3LYP*-D3, TPSSh-D3), are found to considerably overstabilize the intermediate spin state, leading to incorrect ground-state prediction in FeIII porphyrins. The present approach, which can be generalized to other transition metal complexes, is not only useful in method benchmarking, but also sheds light on the interpretations of experimental data for metalloporphyrins, which are important models to understand the electronic properties of heme proteins.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:27
Predicting Molecular Geometry
 01:22
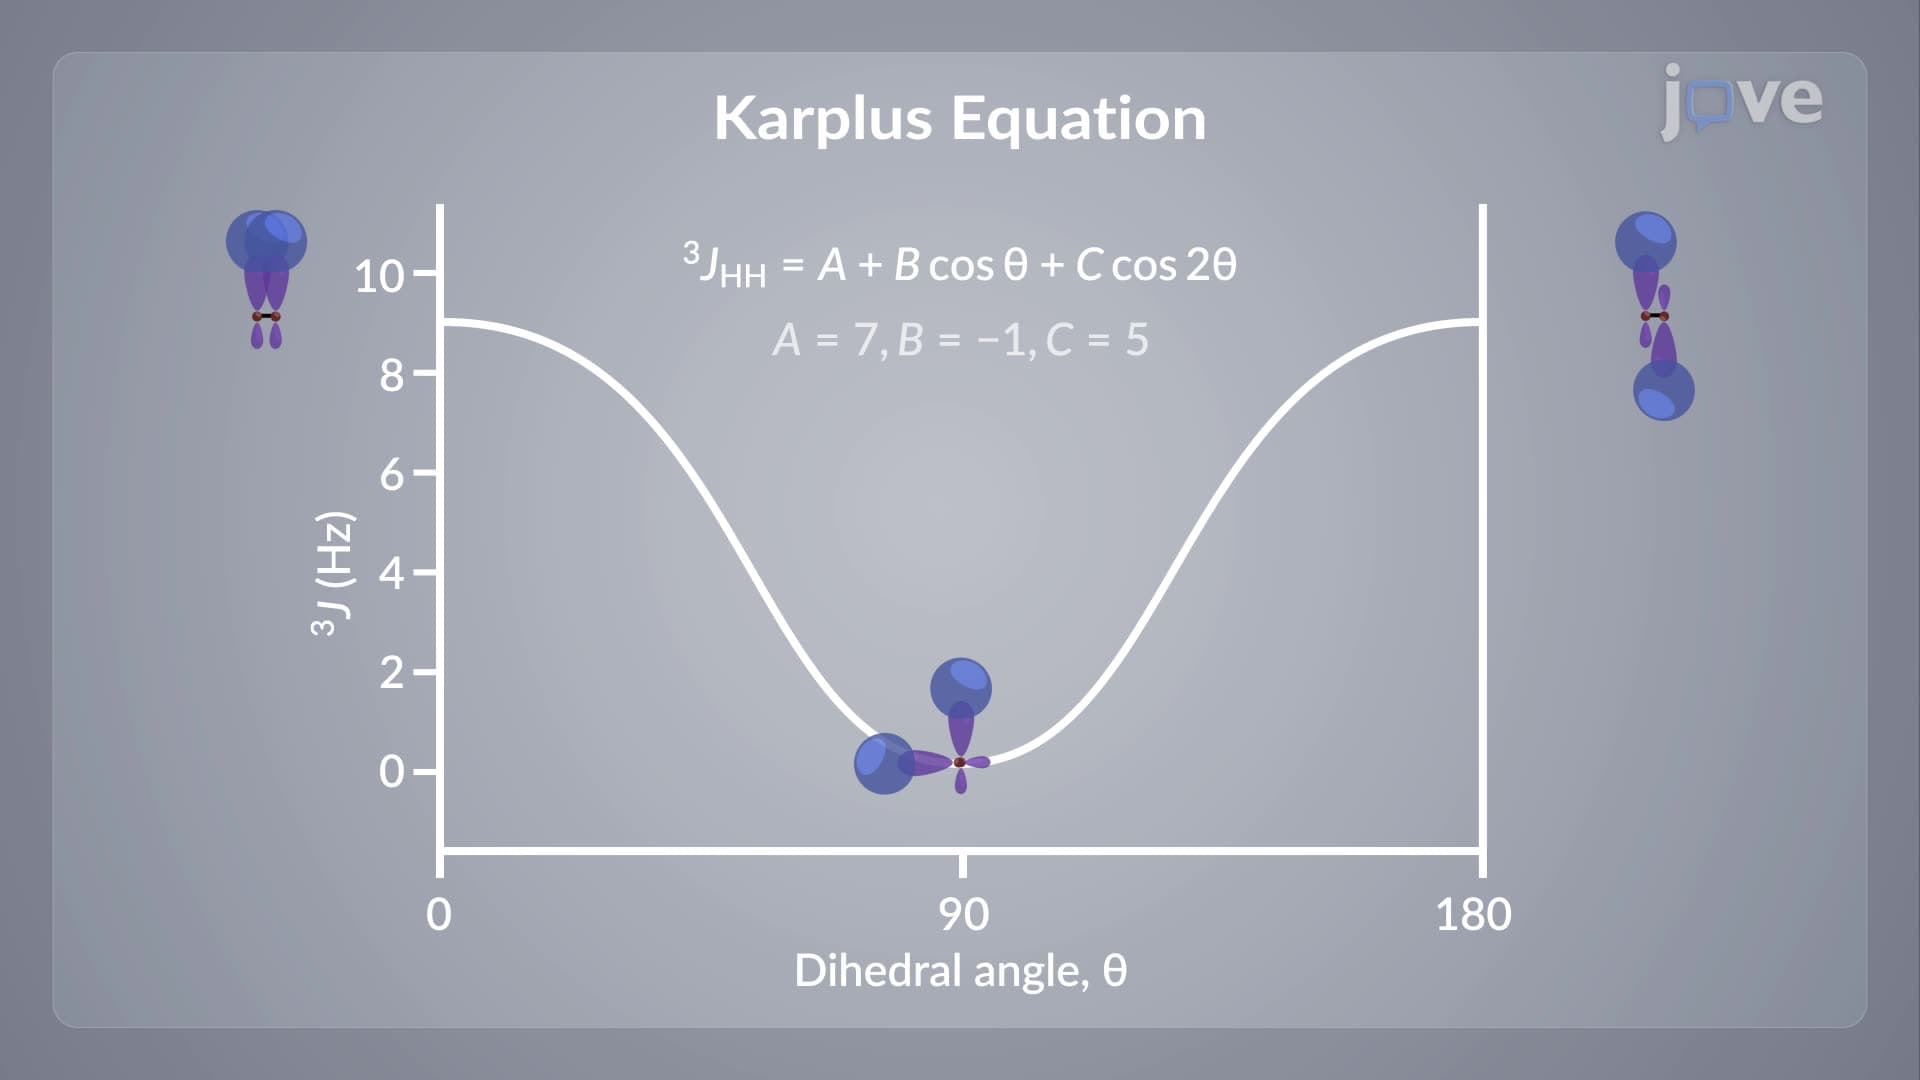
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...