Related Experiment Video
Updated: Jun 19, 2025
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Enhanced molecular first hyperpolarizabilities with Reichardt's type of zwitterions: a computational study on roles
1Department of Chemical Sciences, Auckland Park Kingsway (APK) Campus, University of Johannesburg, PO Box 524, Auckland Park, 2006, Johannesburg, South Africa.

Context:
This work reports structure-property correlations of 27 zwitterions Reichardt's types of zwitterions. Focuses are twofold, to see the (1) impacts of metamerism with Reichardt's vs Brooker's types of zwitterions and (2) impacts of monocyclic aromatic rings as bridges. All the molecules considered here have pyridinium (common acceptor: A) and p-phenylene-dicyanomethanide (common donor: D). Fundamental molecular properties like dipole moments (μ), polarizabilities (α), hyperpolarizabilities (β), and adiabatic absorptions were computed only for the Reichardt types and compared with the literature reported respective Brooker's types of zwitterions. As an impact of metamerism, in general 2-3 times enhanced hyperpolarizabilities (β) were observed for Reichardt's compared to Brooker's types. Exceptions were observed with some triazine bridges and furan bridge, where Brooker's types were found to be more efficient. As impacts of aromatic bridges, in general, 6-sevenfold enhanced β compared to well-known traditional bridges and enhanced β were observed compared to D-A directly connected zwitterion (benzene bridge: sixfold enhanced β). Current findings show that the aromatic bridge control with Reichardt's types of zwitterions is more efficient and thus may be employed as an effective strategy for the designing of functional molecular chromophores for various other fundamental areas.
Methods:
All computations were performed with Gaussian 09. Geometry optimizations and computations of fundamental properties were carried out with HF, B3LYP, CAM-B3LYP, and ωB97xD methodologies, with 6-31G(d,p) and aug-cc-pVDZ basis sets. For adiabatic excitations, computations were carried out using TDDFT and TDHF approaches. For the computations of the response properties (like the nonlinear optical responses), CPHF approach was used.
Related Concept Videos
 01:18
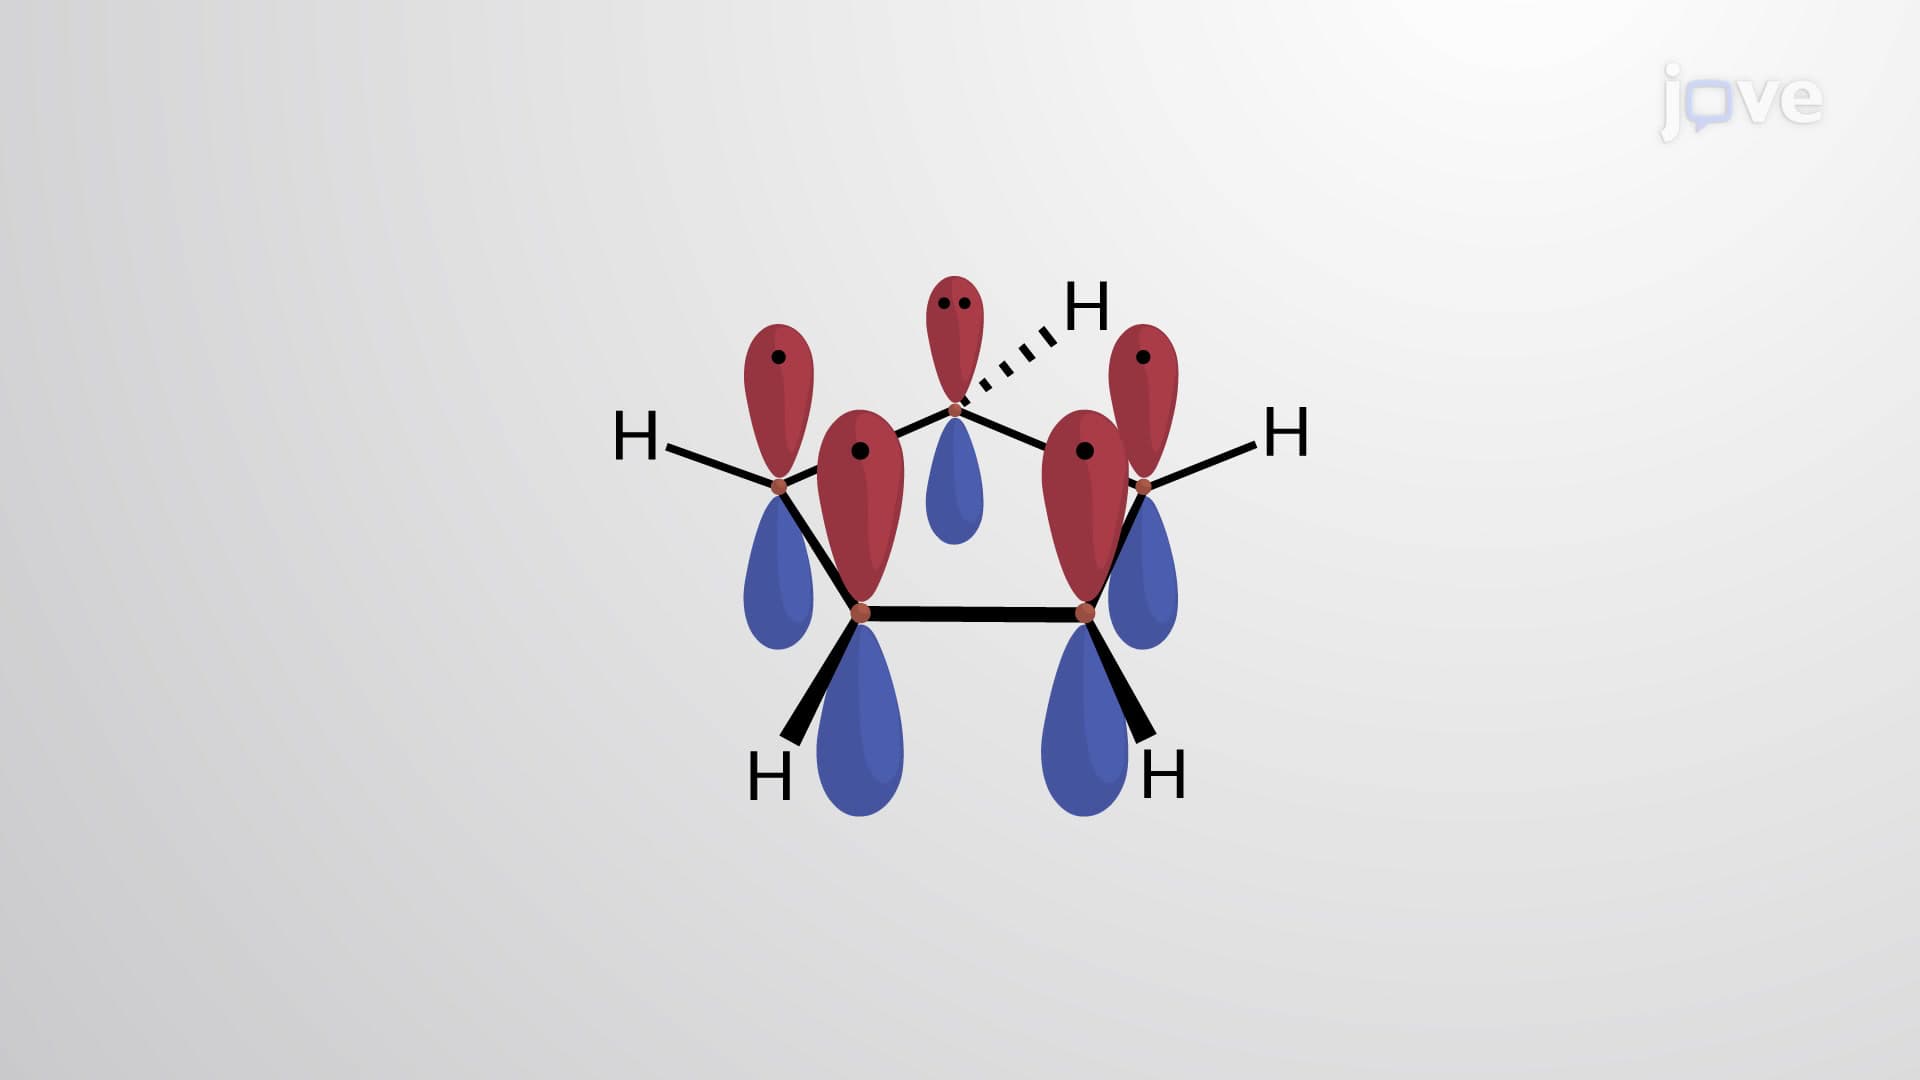
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:20
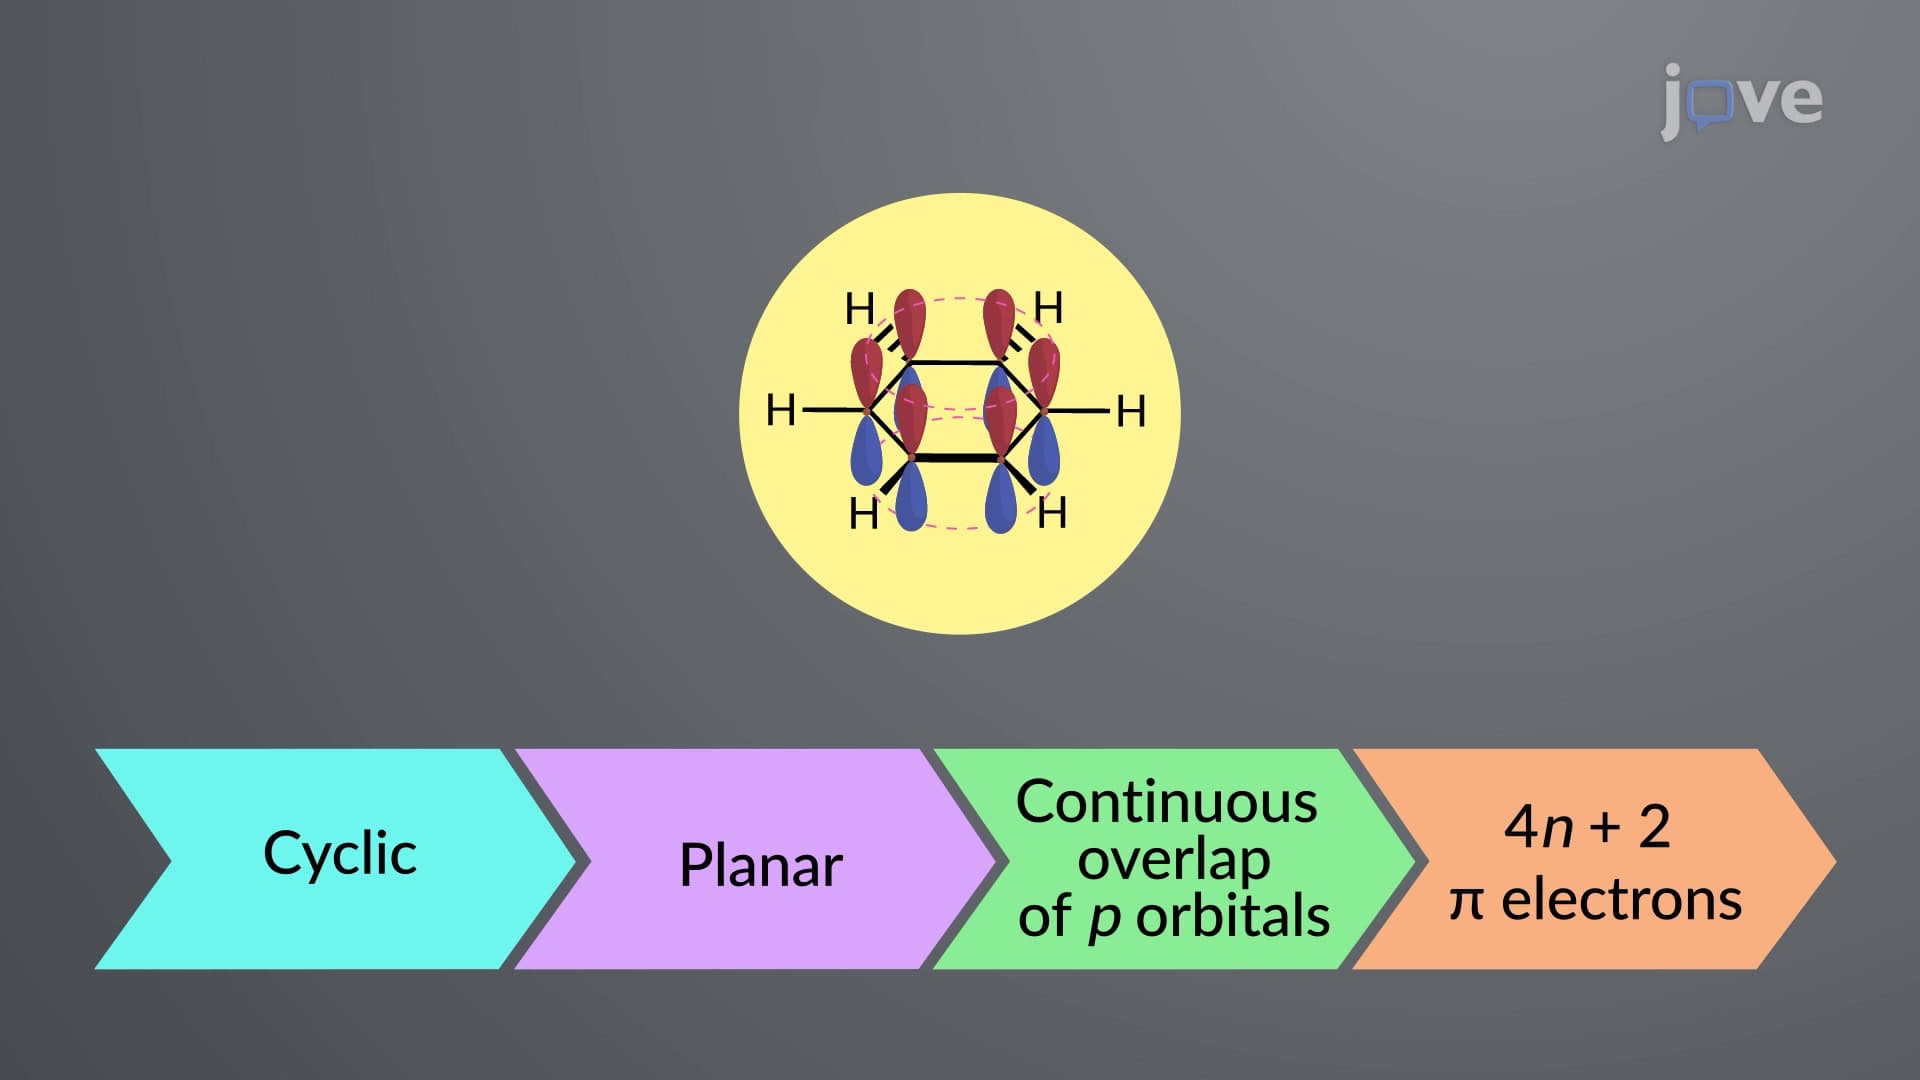
Criteria for Aromaticity and the Hückel 4n + 2 Rule
For the first time, Eric Hückel, a German chemical physicist, derived a set of structural features for a compound to be classified as aromatic. This is now known as...
 01:25
Basicity of Heterocyclic Aromatic Amines
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...