Related Experiment Video
Updated: Jun 18, 2025

Facile Preparation of 2Z,4E-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Visible-Light-Promoted Enantioselective α-Amidation of Aldehydes by Harnessing Organo-Iron Dual Catalysis
Soumyadip Hore1,2, Jiwoo Jeong1,2, Dongwook Kim1,2
1Center for Catalytic Hydrocarbon Functionalizations, Institute for Basic Science (IBS), Daejeon 34141, South Korea.

Abstract:
The strategic integration of organocatalysis with transition-metal catalysis to achieve otherwise unattainable stereoselective transformations may serve as a powerful synthetic tool. Herein, we present a synthetically versatile α-amidation of aldehydes by leveraging dual iron and chiral enamine catalysis in an enantioselective manner (up to >99:1 er). Experimental and computational studies have led us to propose a new mechanistic platform, wherein visible-light-promoted LMCT generates [Fe(II)Cl3-], which effectively activates dioxazolones to form an iron-acylnitrenoid radical that inserts into chiral enamine intermediates.
Related Concept Videos
 01:10
α-Alkylation of Ketones via Enolate Ions
 02:44
Oxidation of Alkenes: Syn Dihydroxylation with Osmium Tetraoxide
 01:14
Aldehydes and Ketones with Amines: Enamine Formation Mechanism
 01:20
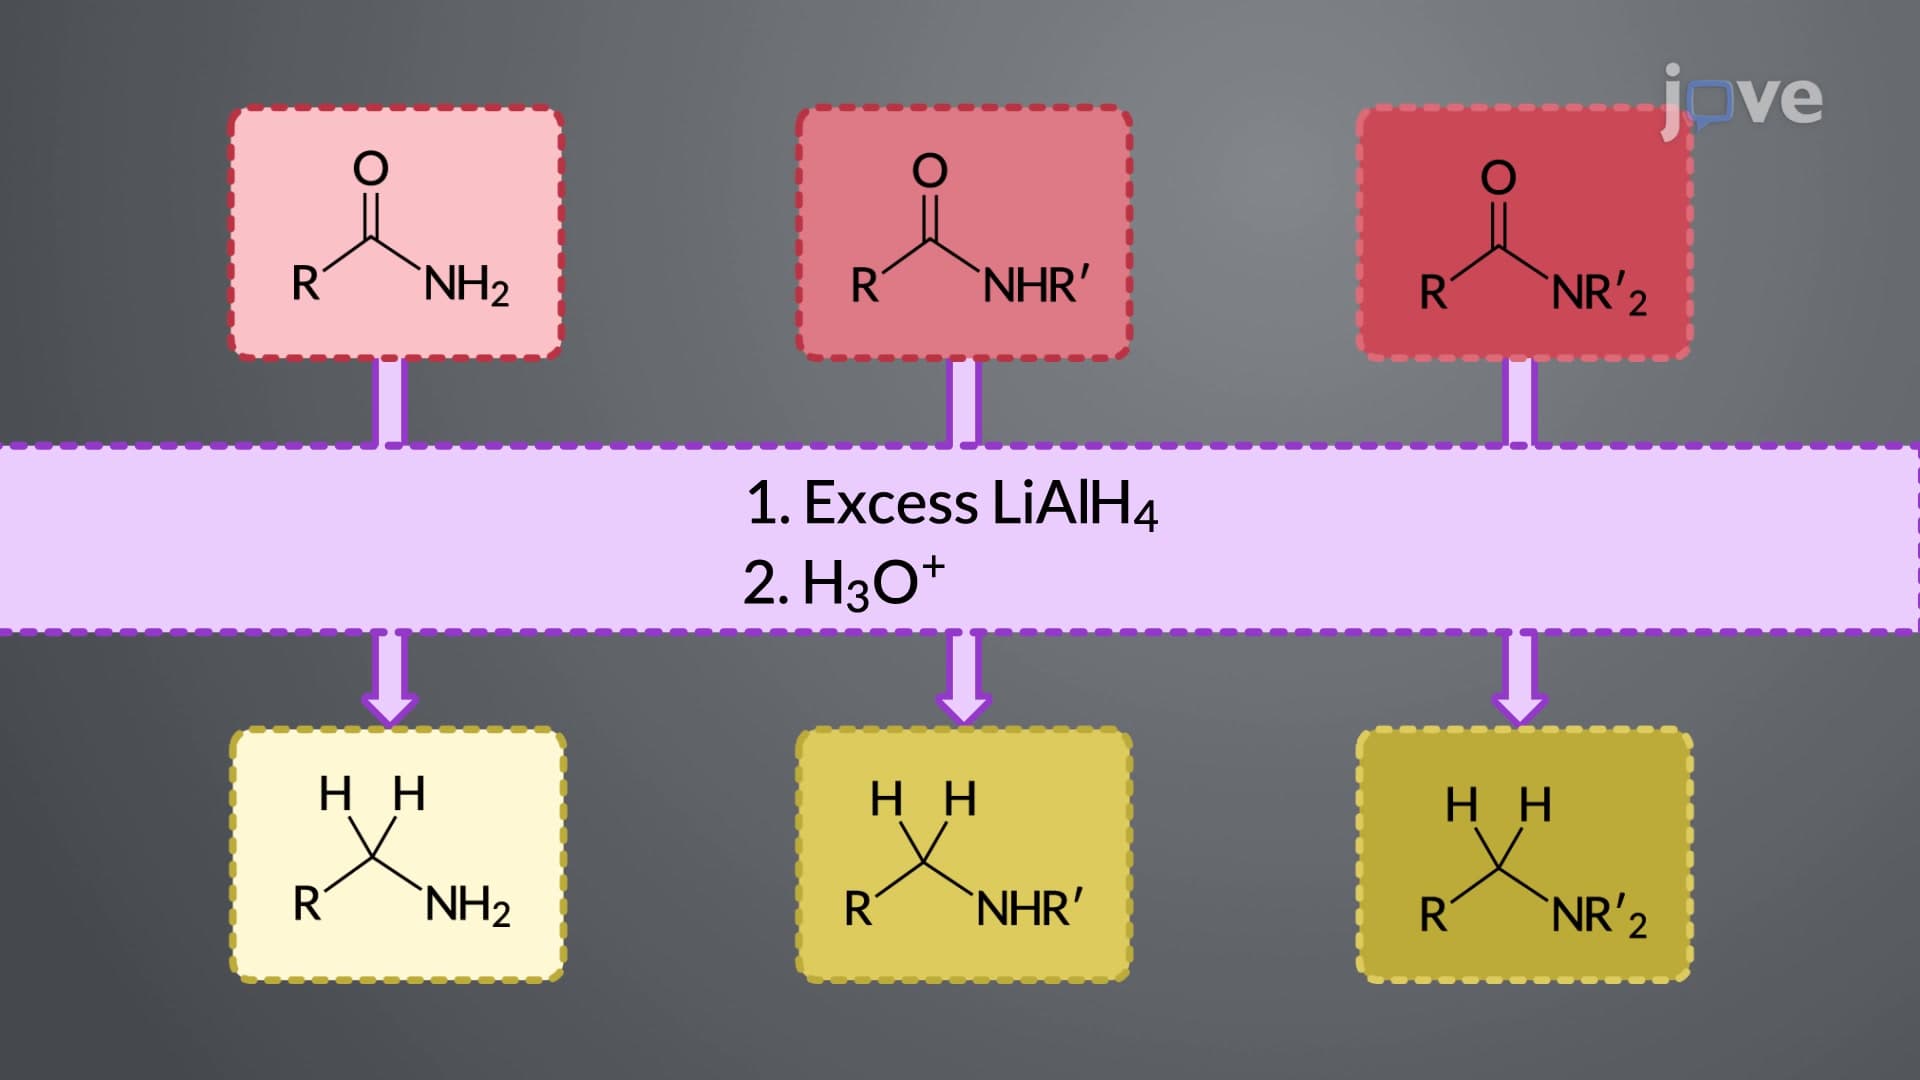
Amides to Amines: LiAlH4 Reduction
Amide reduction requires two equivalents of the reducing agent, acting as a source of hydride ions. As shown in the figure, the reaction is initiated with a nucleophilic attack by the hydride ion at the carbonyl carbon to form a tetrahedral intermediate.
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 01:21
Acid-Catalyzed α-Halogenation of Aldehydes and Ketones
In the first step of the mechanism, the acid protonates the carbonyl oxygen resulting in a resonance-stabilized cation, which subsequently loses an α-hydrogen to form an enol tautomer. The C=C bond in an enol is highly nucleophilic because of the electron-donating nature of the –OH group. Consequently, the double bond attacks an electrophilic halogen to form a...