Related Experiment Video
Updated: Jun 18, 2025

Analyzing Oxygen Consumption Rate in Primary Cultured Mouse Neonatal Cardiomyocytes Using an Extracellular Flux Analyzer
Published on: February 13, 2019
Opa1 processing is dispensable in mouse development but is protective in mitochondrial cardiomyopathy
Sofia Ahola1, Lilli A Pazurek1, Fiona Mayer1
1Max Planck Institute for Biology of Ageing, Cologne, Germany.

Abstract:
Mitochondrial fusion and fission accompany adaptive responses to stress and altered metabolic demands. Inner membrane fusion and cristae morphogenesis depends on optic atrophy 1 (Opa1), which is expressed in different isoforms and is cleaved from a membrane-bound, long to a soluble, short form. Here, we have analyzed the physiological role of Opa1 isoforms and Opa1 processing by generating mouse lines expressing only one cleavable Opa1 isoform or a non-cleavable variant thereof. Our results show that expression of a single cleavable or non-cleavable Opa1 isoform preserves embryonic development and the health of adult mice. Opa1 processing is dispensable under metabolic and thermal stress but prolongs life span and protects against mitochondrial cardiomyopathy in OXPHOS-deficient Cox10-/- mice. Mechanistically, loss of Opa1 processing disturbs the balance between mitochondrial biogenesis and mitophagy, suppressing cardiac hypertrophic growth in Cox10-/- hearts. Our results highlight the critical regulatory role of Opa1 processing, mitochondrial dynamics, and metabolism for cardiac hypertrophy.
Insights
Optic atrophy 1 (Opa1) processing is vital for longevity and preventing heart disease in mice with mitochondrial defects. This mitochondrial dynamics regulation impacts cardiac health and lifespan.
Area of Science:
- Mitochondrial dynamics and cellular metabolism.
- Genetics and physiology of mitochondrial proteins.
- Cardiovascular research and disease mechanisms.
Background:
- Mitochondrial fusion and fission are crucial for cellular adaptation to stress and metabolic changes.
- Optic atrophy 1 (Opa1) is essential for inner mitochondrial membrane fusion and cristae structure.
- Opa1 exists in multiple isoforms and undergoes processing from a long, membrane-bound form to a short, soluble form.
Purpose of the Study:
- To investigate the physiological roles of different Opa1 isoforms and its processing.
- To determine the impact of Opa1 processing on mouse development, stress response, and lifespan.
- To elucidate the role of Opa1 processing in mitochondrial cardiomyopathy in OXPHOS-deficient mice.
Main Methods:
- Generation of mouse lines expressing single cleavable or non-cleavable Opa1 isoforms.
- Analysis of embryonic development and adult mouse health under normal and stress conditions (metabolic, thermal).
- Assessment of mitochondrial cardiomyopathy in Cox10-/- mice with altered Opa1 processing.
Main Results:
- Expression of a single Opa1 isoform (cleavable or non-cleavable) supports normal embryonic development and adult health.
- Opa1 processing is not essential for metabolic or thermal stress adaptation but extends lifespan.
- Loss of Opa1 processing protects against mitochondrial cardiomyopathy in Cox10-/- mice by altering mitochondrial biogenesis and mitophagy balance, suppressing cardiac hypertrophy.
Conclusions:
- Opa1 processing is dispensable for basic survival and stress adaptation but critically regulates lifespan and cardiac health.
- The balance between mitochondrial biogenesis and mitophagy, influenced by Opa1 processing, is key to preventing cardiac hypertrophy in mitochondrial disease.
- Opa1 processing, mitochondrial dynamics, and metabolism are critical regulators of cardiac hypertrophy.
Related Concept Videos
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:19
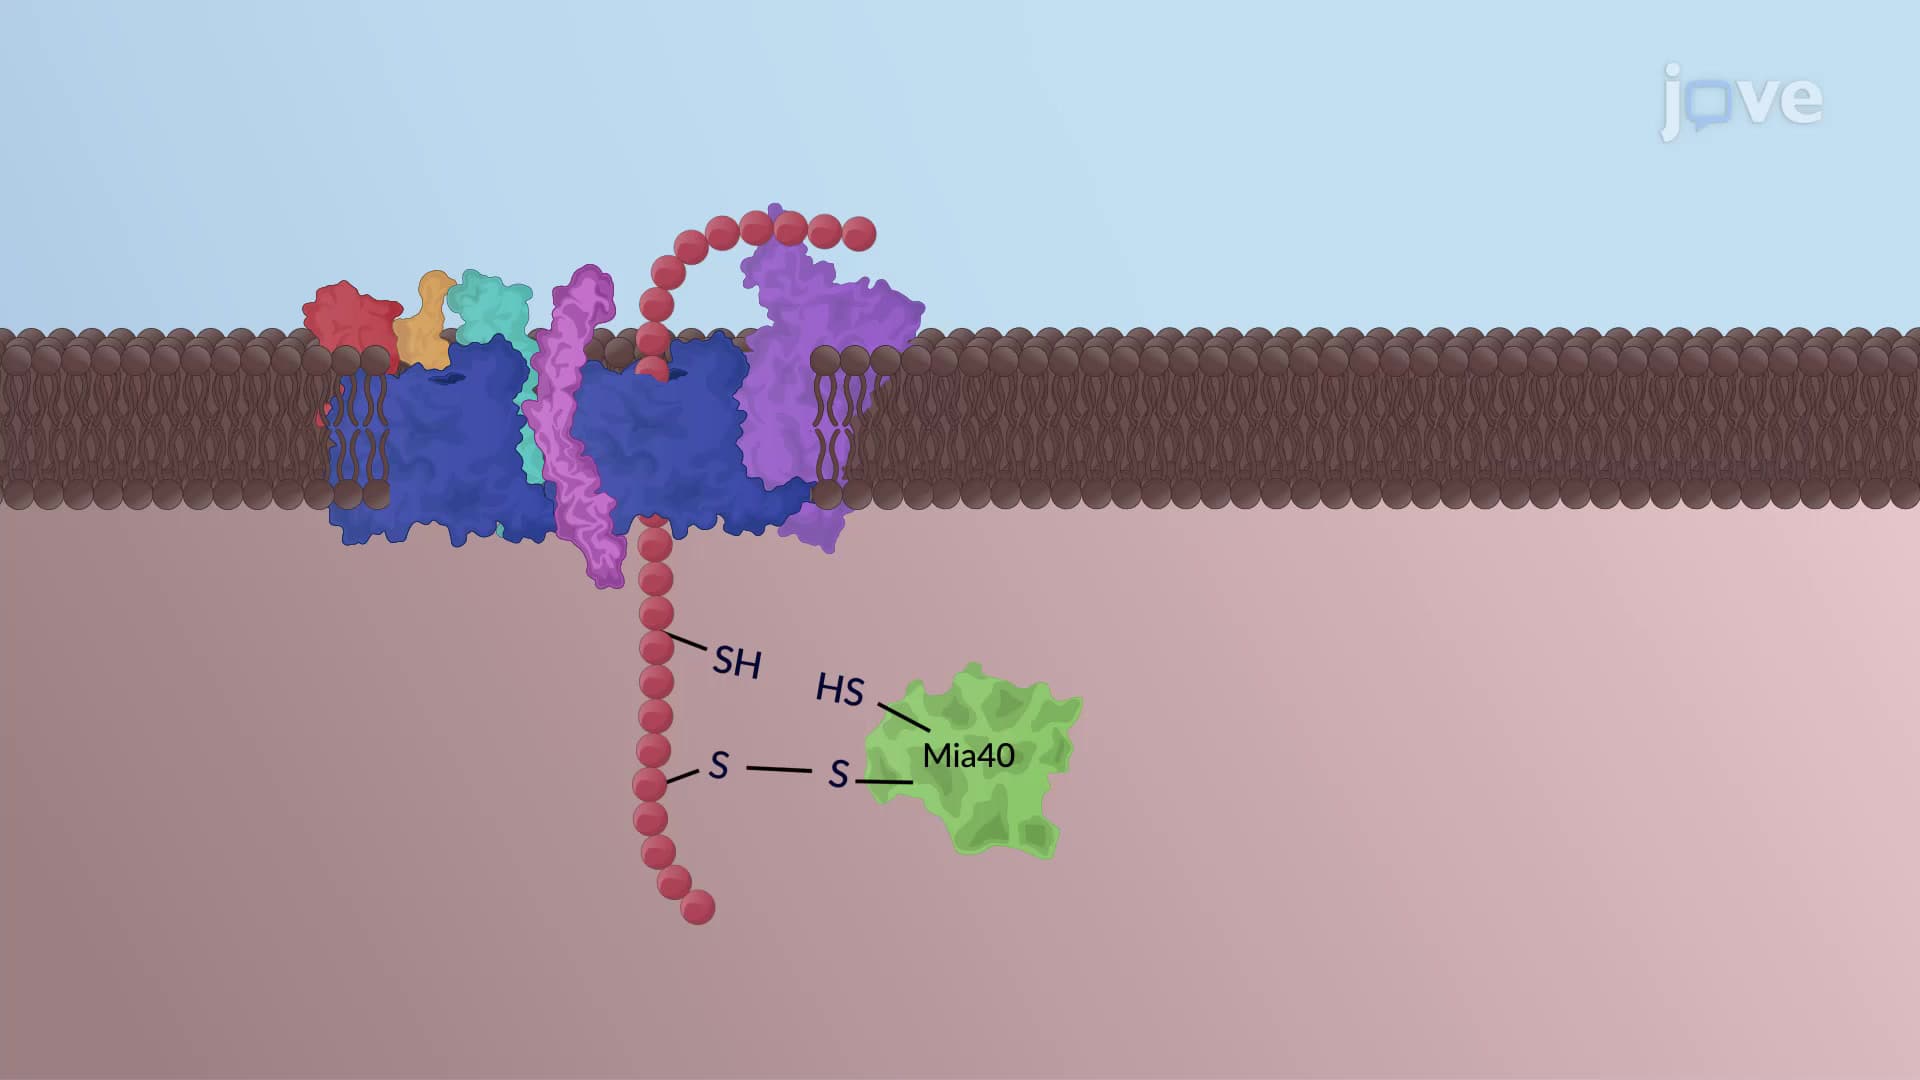
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:48
ATP Synthase: Mechanism
 01:39
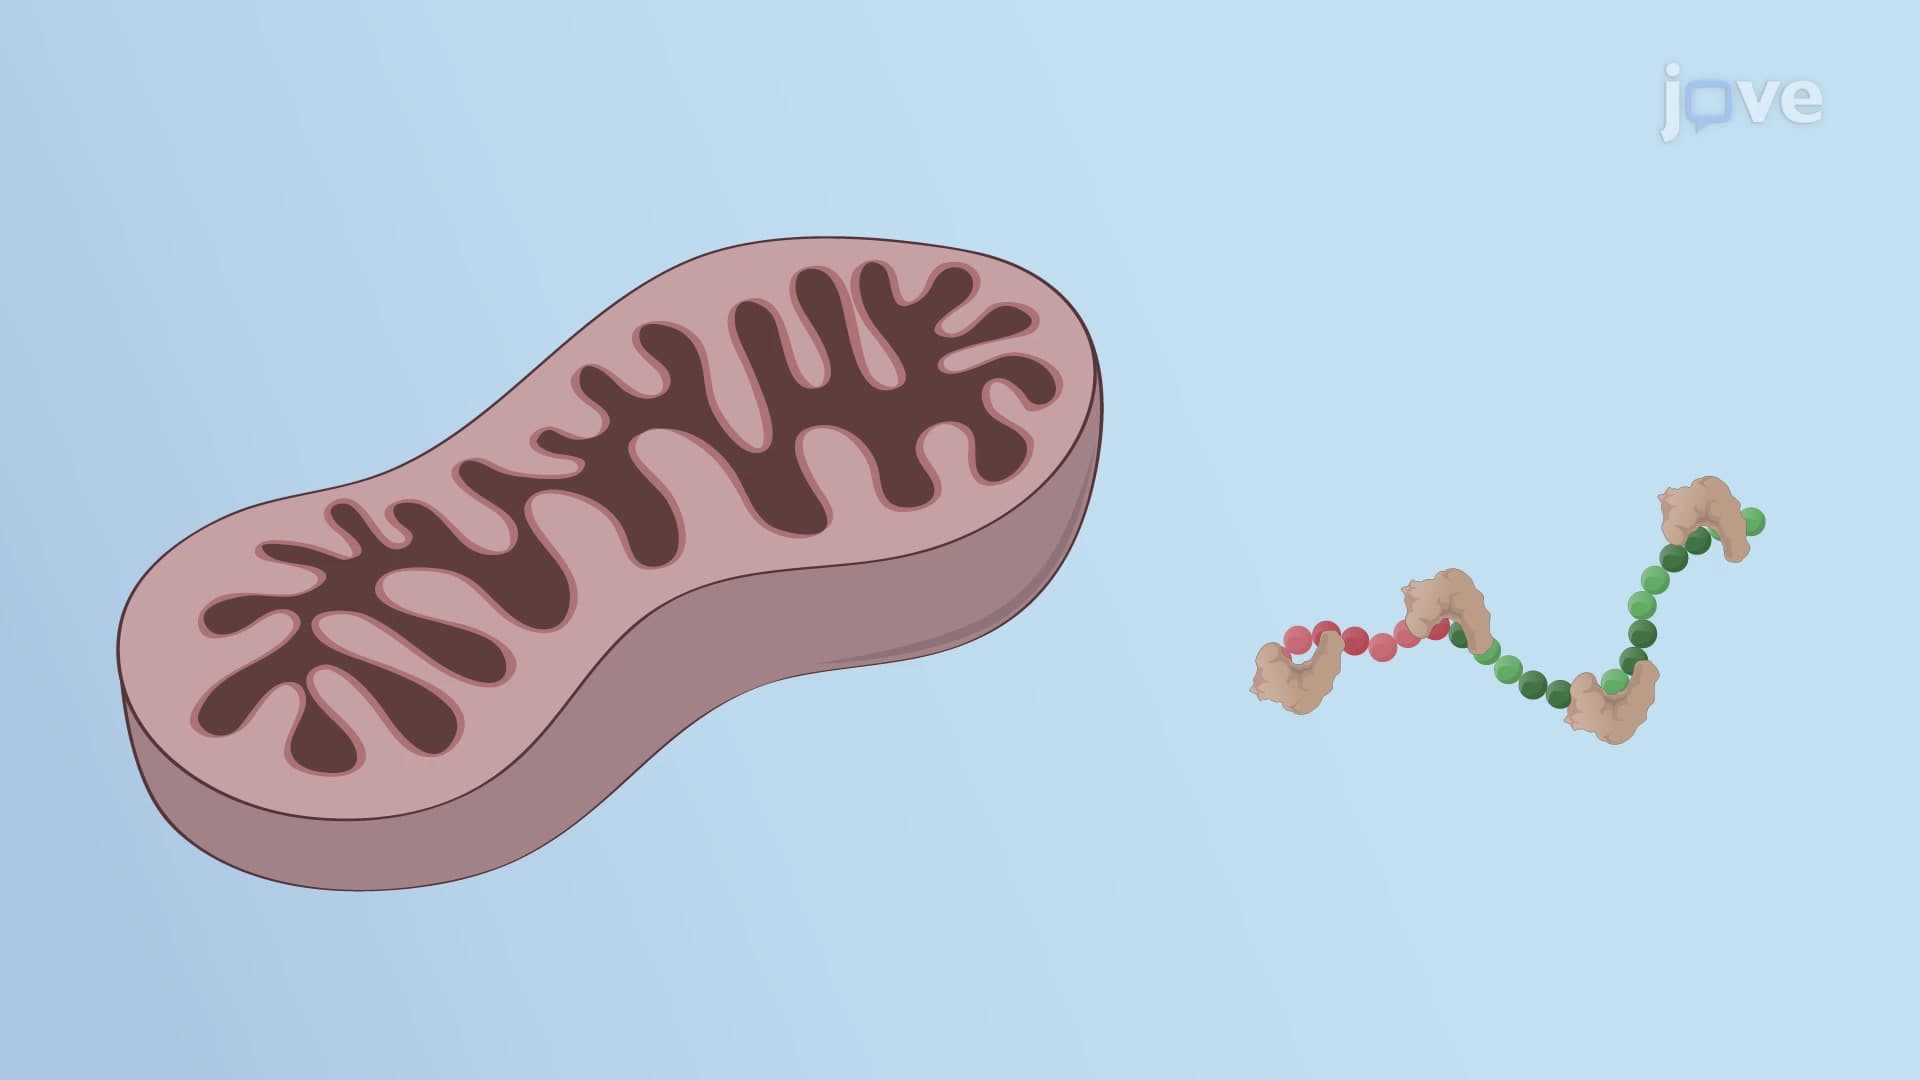
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:12
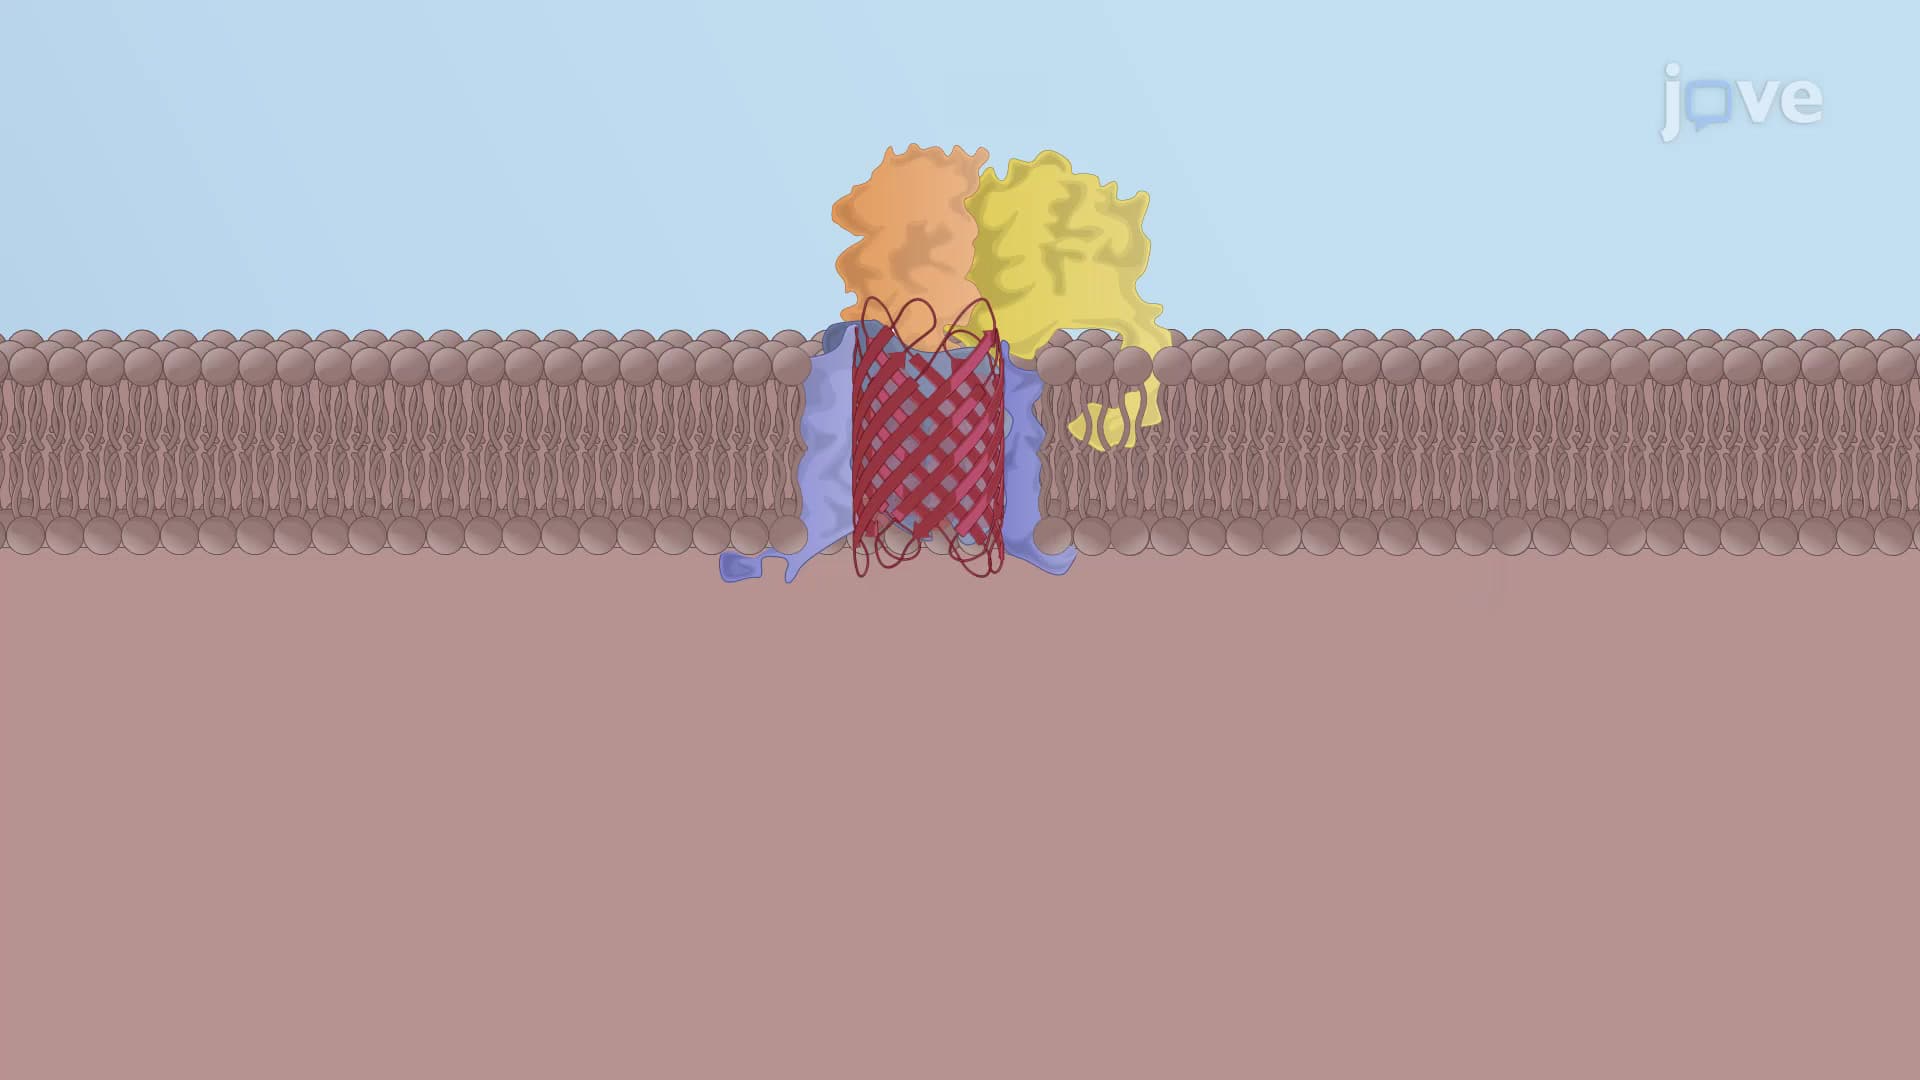
Porin Insertion in the Outer Mitochondrial Membrane
Three models describe the assembly of porins by the SAM complex and their insertion into the outer membrane. Model 1 suggests that porins are assembled outside the SAM channel as the...
 02:59
Animal Mitochondrial Genetics