Related Experiment Video
Updated: Jun 17, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Monte Carlo simulations for free energies of hydration: Past to present
1Department of Chemistry, Yale University, New Haven, Connecticut 06520-8107, USA.

Abstract:
A summary of the development of Monte Carlo statistical mechanics simulations for the computation of free energies of hydration of organic molecules is followed by presentation of results with the latest version of the optimized potentials for liquid simulations-all atom force field and the TIP4P water model. Scaling of the Lennard-Jones interactions between water, oxygen, and carbon atoms by a factor of 1.25 is found to improve the accuracy of free energies of hydration for 50 prototypical organic molecules from a mean unsigned error of 1.0-1.2 to 0.4 kcal/mol.

Related Concept Videos
 02:42
Aqueous Solutions and Heats of Hydration
When ionic compounds dissolve in water, the ions in the solid separate and disperse uniformly throughout the solution because water molecules surround and solvate the ions, reducing the strong electrostatic forces between them. This process...
 02:49
Calculating Standard Free Energy Changes
 02:45
Acid-Catalyzed Hydration of Alkenes
 01:20
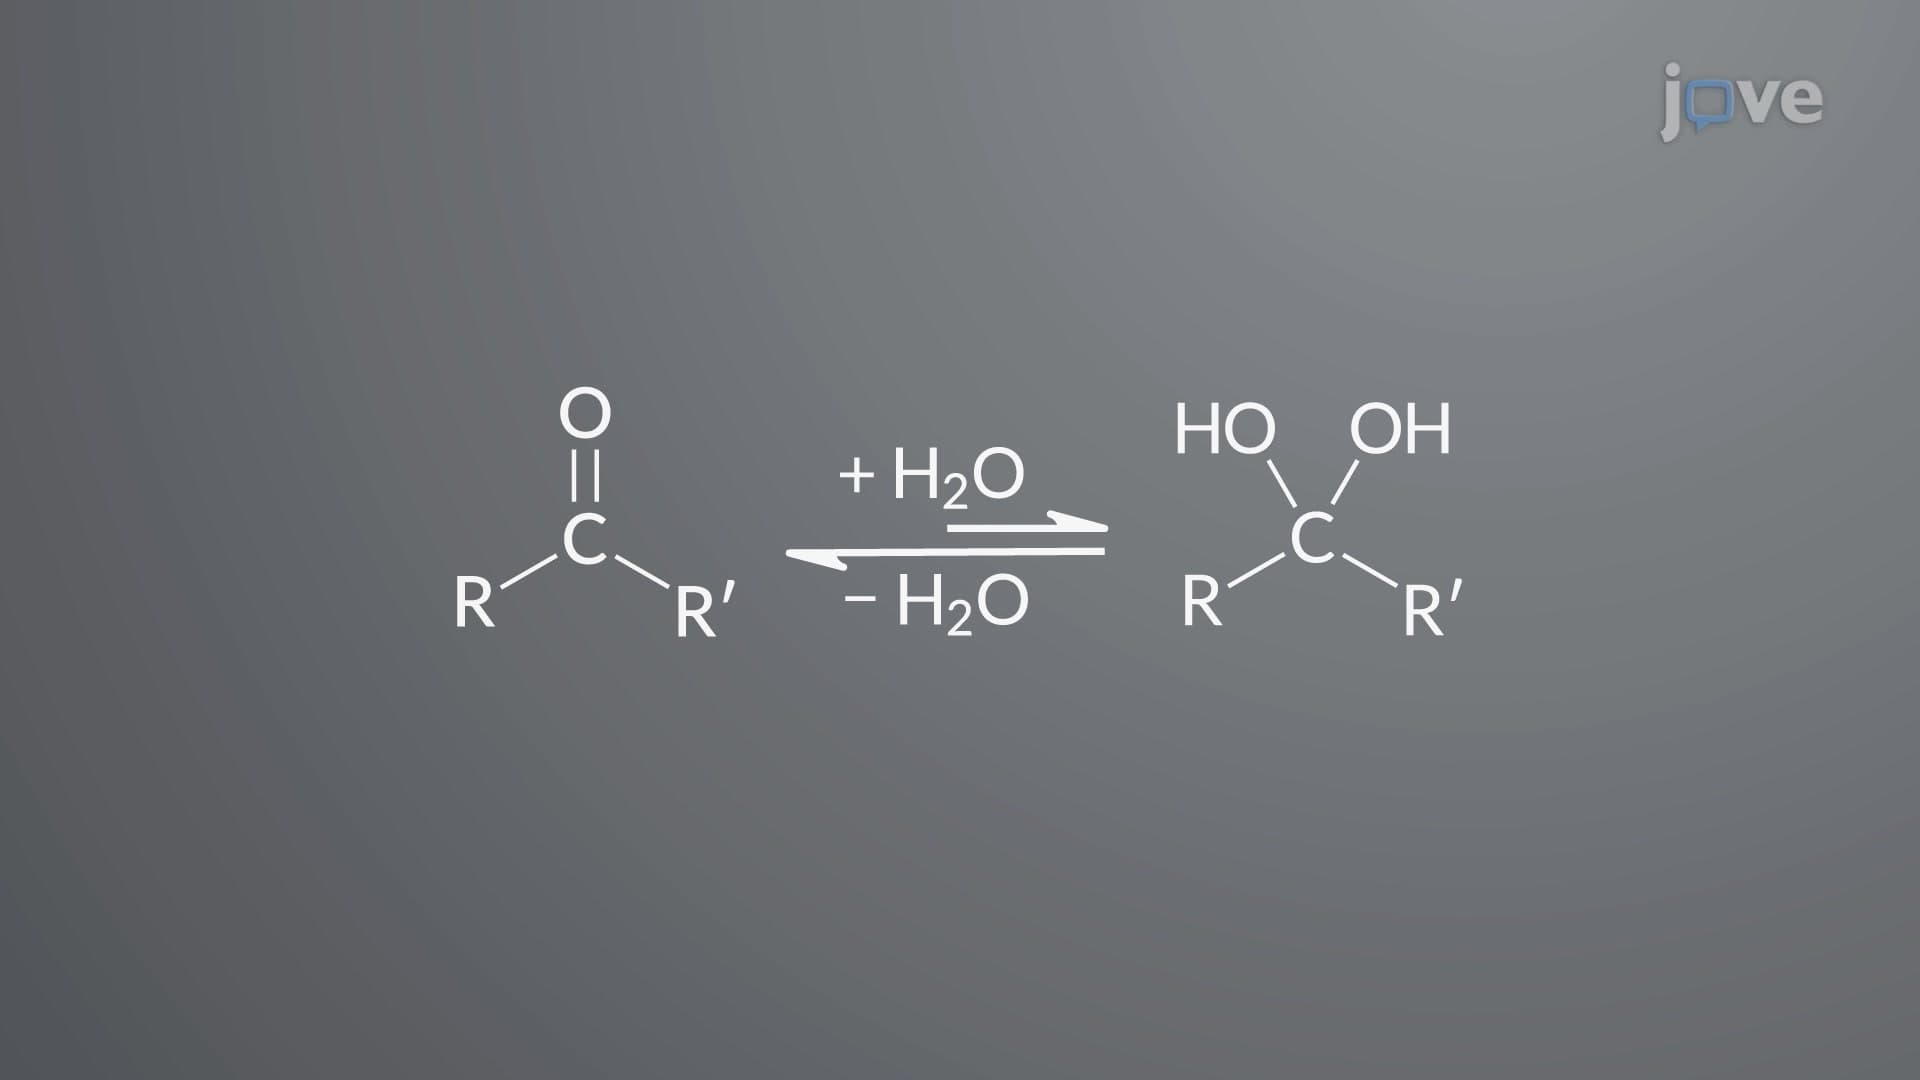
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 02:39
Enthalpy of Solution
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration