Related Experiment Video
Updated: Jun 16, 2025

Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Small basis set density functional theory method for cost-efficient, large-scale condensed matter simulations
Elisabeth Keller1,2, Jack Morgenstein3, Karsten Reuter1
1Fritz Haber Institute of the Max Planck Society, Berlin, Germany.

Abstract:
We present an efficient first-principles based method geared toward reliably predicting the structures of solid materials across the Periodic Table. To this end, we use a density functional theory baseline with a compact, near-minimal min+s basis set, yielding low computational costs and memory demands. Since the use of such a small basis set leads to systematic errors in chemical bond lengths, we develop a linear pairwise correction, available for elements Z = 1-86 (excluding the lanthanide series), parameterized for use with the Perdew-Burke-Ernzerhof exchange-correlation functional. We demonstrate the reliability of this corrected approach for equilibrium volumes across the Periodic Table and the transferability to differently coordinated environments and multi-elemental crystals. We examine relative energies, forces, and stresses in geometry optimizations and molecular dynamics simulations.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 00:56
Density
 01:17
Theory of Metallic Conduction
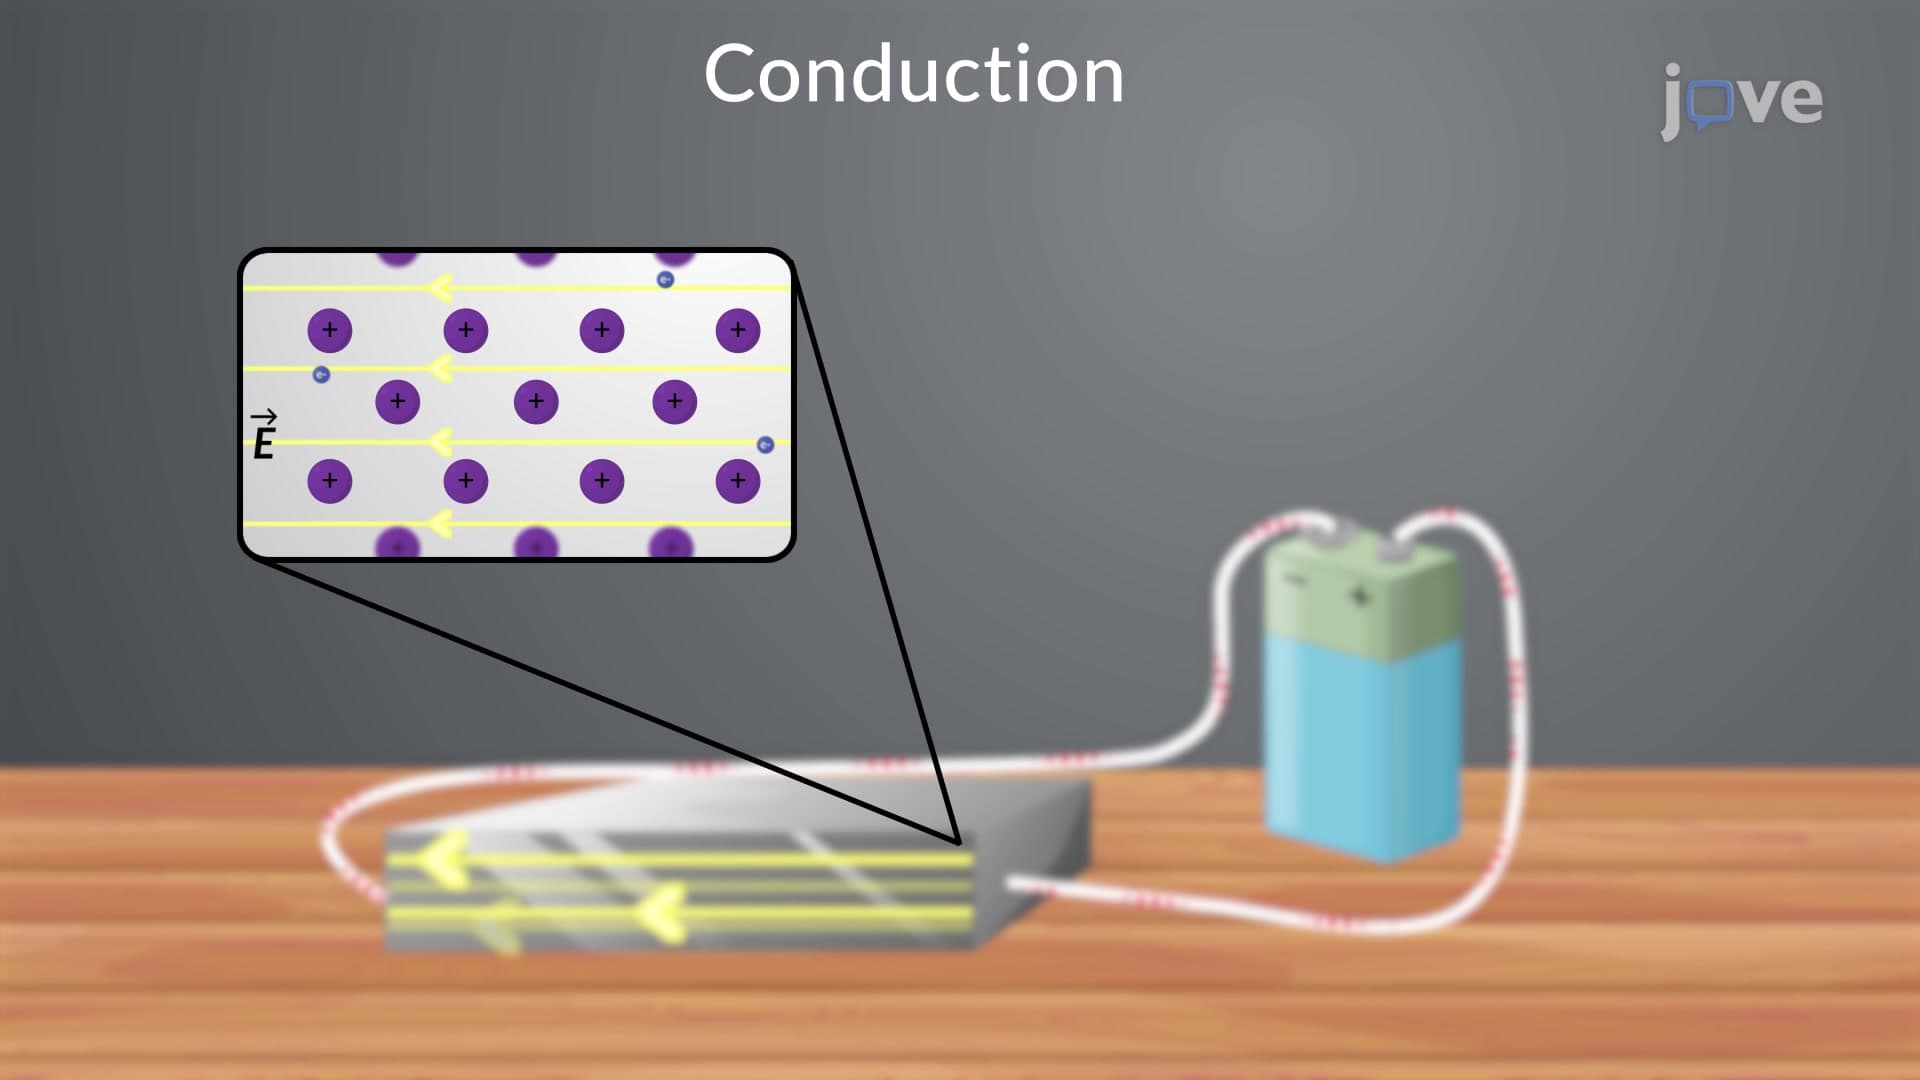
In this theory, Newton's second law of motion is used to determine the acceleration of an electron in the presence of an applied electric field. Then, its velocity is expressed via this acceleration.
An electron moves through the crystal, containing positive ions,...
 01:20
Maxwell-Boltzmann Distribution: Problem Solving
This distribution function f(v) is defined by saying that the expected number N (v1,v2) of particles with speeds between v1 and v2 is given by
 01:10
Van der Waals Equation
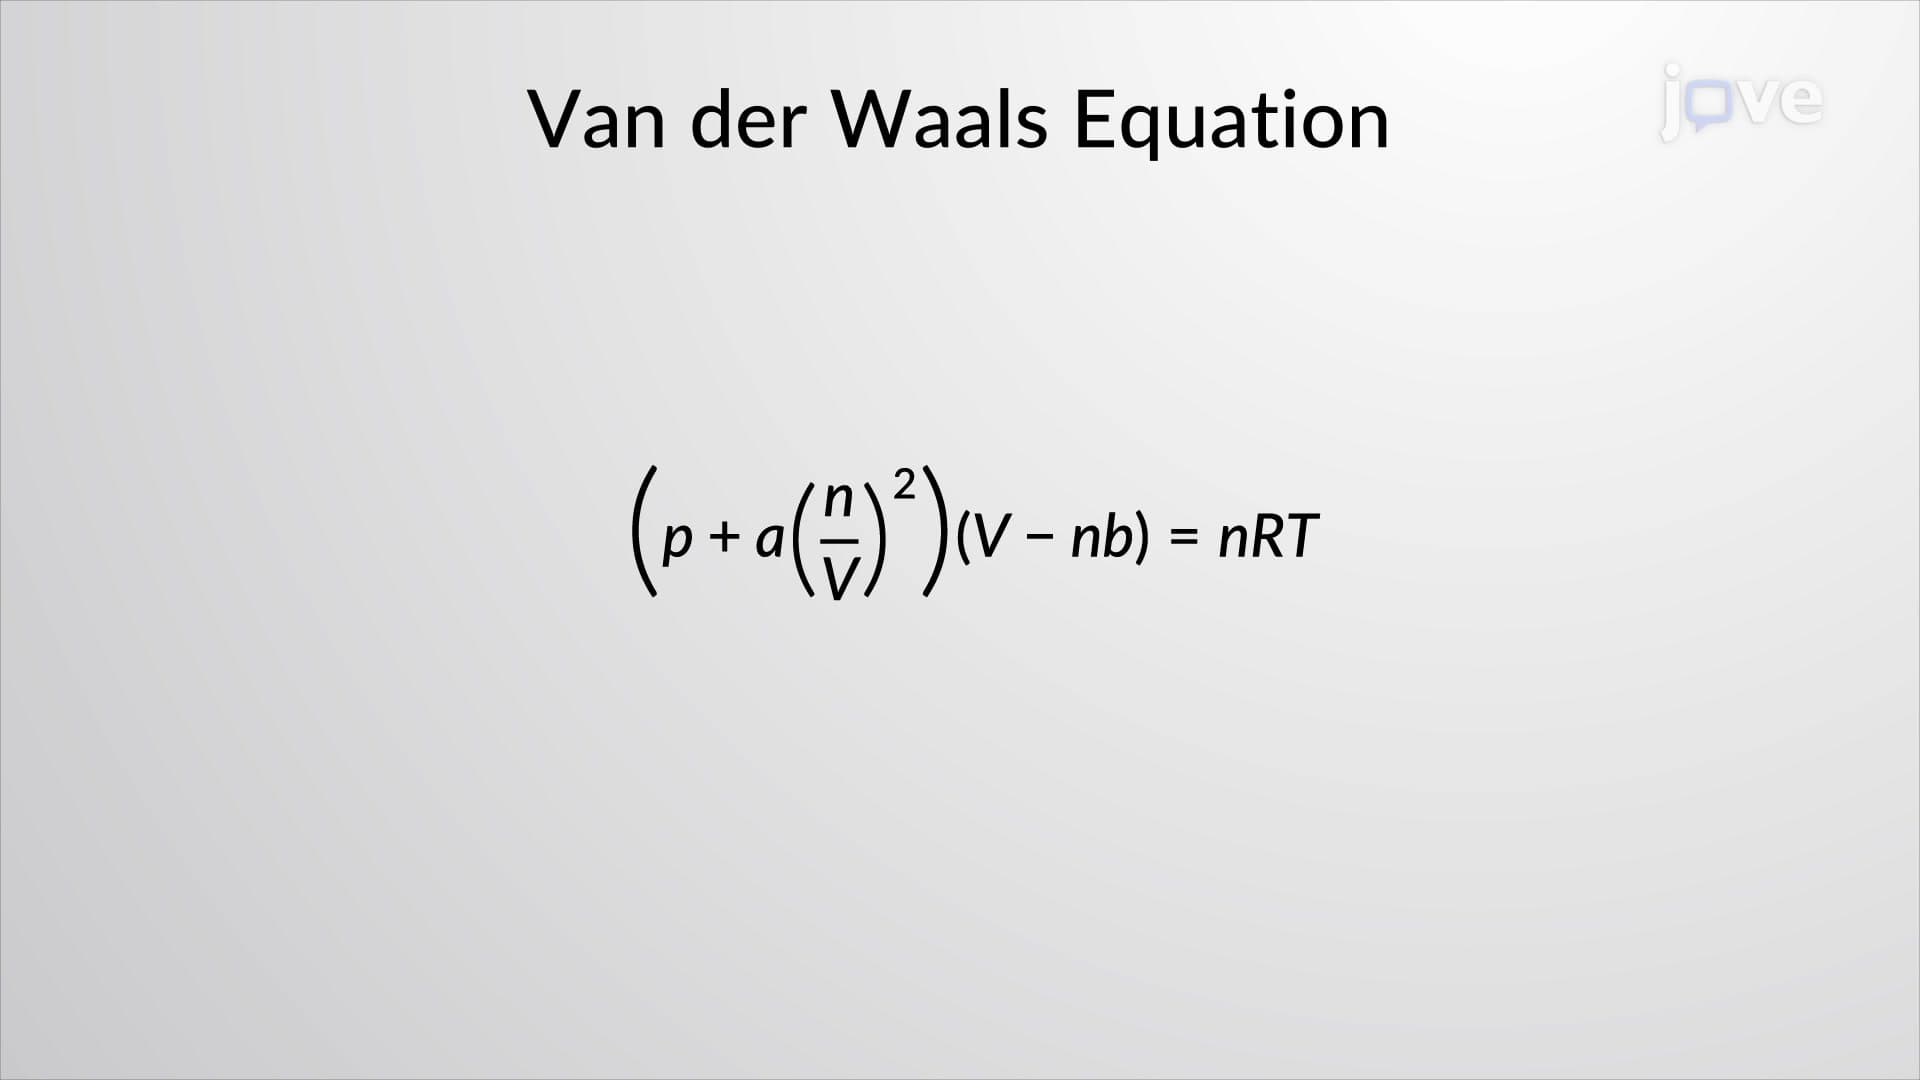
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
Van der Waals Interactions
 01:21
Current Density