Related Experiment Video
Updated: Jun 16, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
DASH properties: Estimating atomic and molecular properties from a dynamic attention-based substructure hierarchy
Marc T Lehner1, Paul Katzberger1, Niels Maeder1
1Department of Chemistry and Applied Biosciences, ETH Zürich, Vladimir-Prelog-Weg 2, 8093 Zürich, Switzerland.

Abstract:
Recently, we presented a method to assign atomic partial charges based on the DASH (dynamic attention-based substructure hierarchy) tree with high efficiency and quantum mechanical (QM)-like accuracy. In addition, the approach can be considered "rule based"-where the rules are derived from the attention values of a graph neural network-and thus, each assignment is fully explainable by visualizing the underlying molecular substructures. In this work, we demonstrate that these hierarchically sorted substructures capture the key features of the local environment of an atom and allow us to predict different atomic properties with high accuracy without building a new DASH tree for each property. The fast prediction of atomic properties in molecules with the DASH tree can, for example, be used as an efficient way to generate feature vectors for machine learning without the need for expensive QM calculations. The final DASH tree with the different atomic properties as well as the complete dataset with wave functions is made freely available.

Related Concept Videos
 02:00
Molecular Models
 02:27
Predicting Molecular Geometry
 03:02
The Aufbau Principle and Hund's Rule
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 01:27
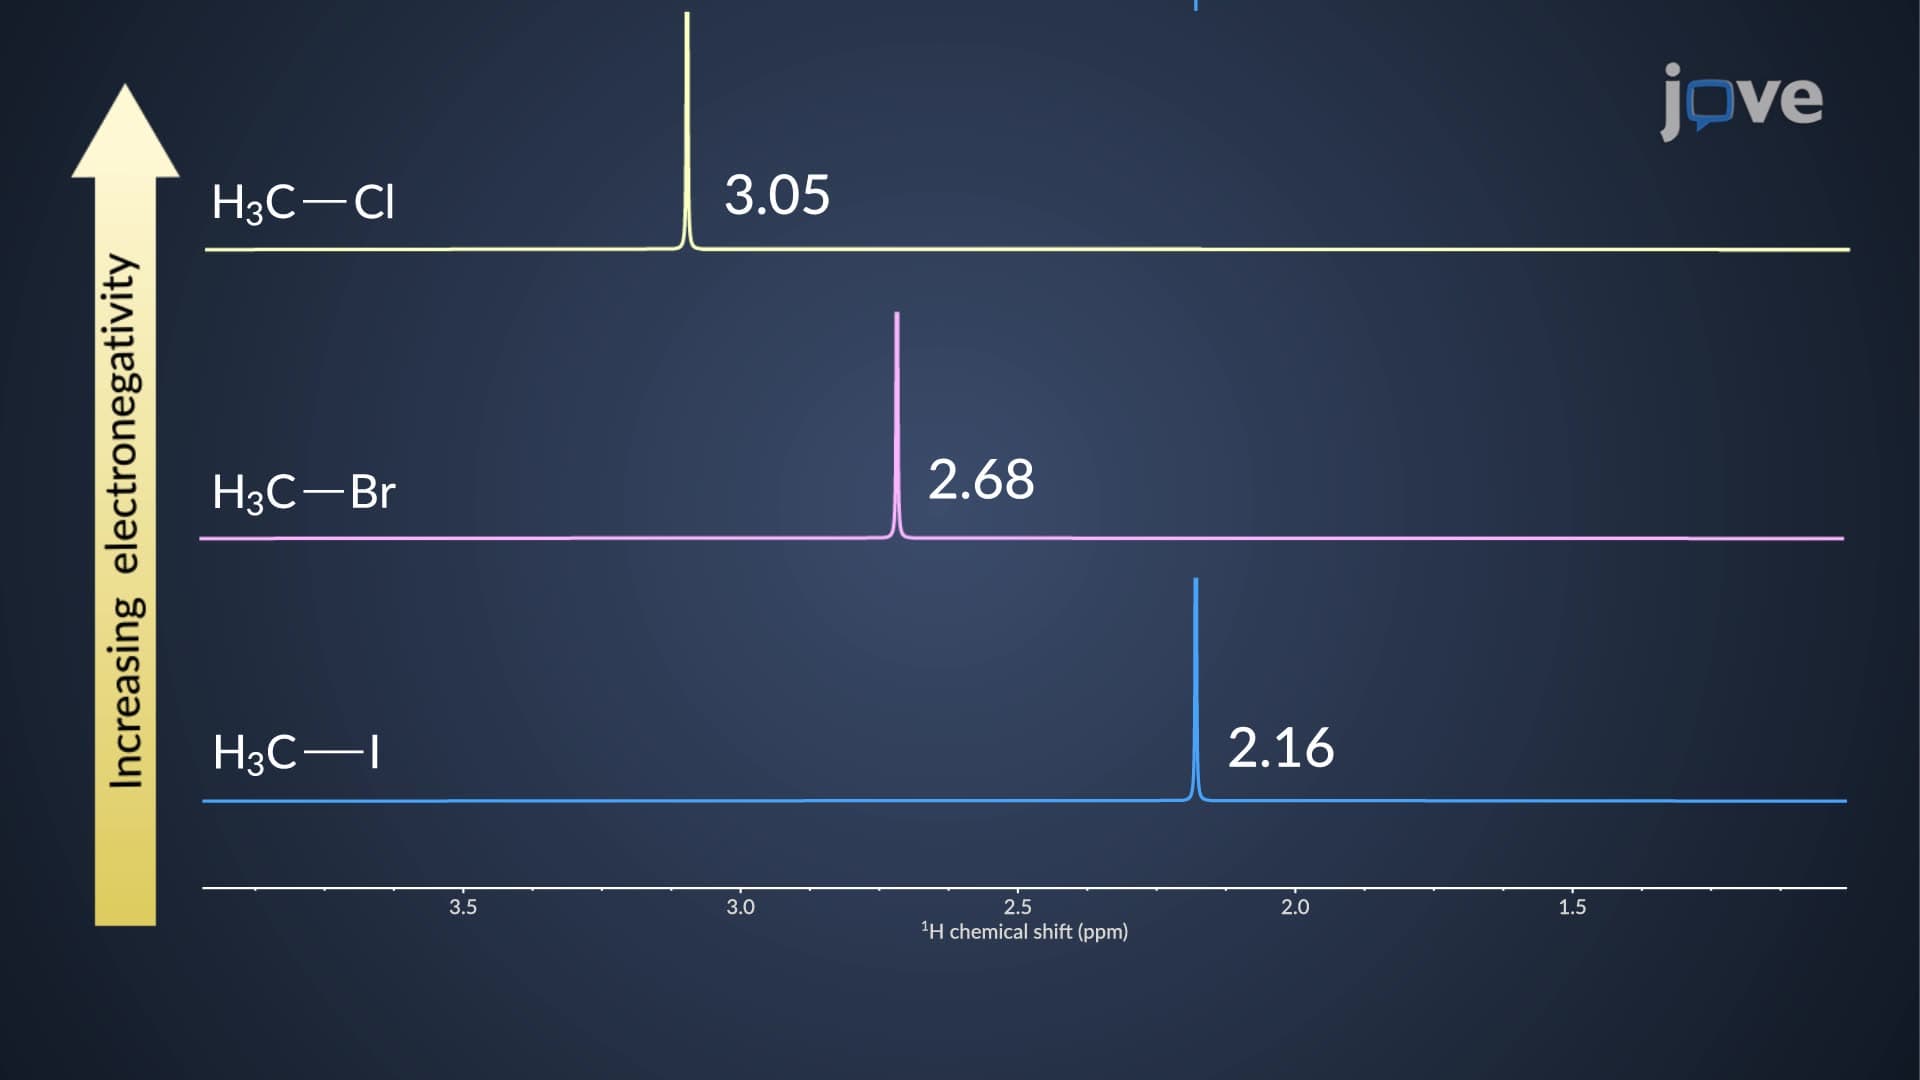
Inductive Effects on Chemical Shift: Overview
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...