Related Experiment Video
Updated: Jun 16, 2025

Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Resolving Conflicting Mechanisms for Photoredox Allylic sp3-CH Arylation Using Deuterium-Labeling and Isotope Effects
Anuradha Dagar1, Tamal Das1, Sharath Chandra Mallojjala1
1Department of Chemistry, Binghamton University, Vestal, New York 13850, United States.

Abstract:
Two conflicting mechanisms have emerged for the direct arylation of allylic C-H bonds enabled by the combined use of thiol and photoredox catalysis. In the original report (Nature, 2015, 519, 74-77), a radical coupling step-between a radical anion of an arene and an allylic radical-is proposed to be the key C-C bond-forming step. A recent mechanistic study (J. Org. Chem. 2022, 87, 223-230) has suggested that the C-C bond formation occurs via radical anion capture by the olefin followed by an H atom transfer (HAT) event to deliver the allylic C-H arylation product. Utilizing cyclohexene-4,4,5,5-d 4 as a mechanistic probe to distinguish between otherwise indistinguishable regioisomeric allylic C-H arylation products in the reaction of cyclohexene and dicyanobenzene, we establish that the radical anion capture-HAT mechanism is not operative. Furthermore, experimental k H/k D studies and DFT calculations lend strong support to the radical coupling mechanism proceeding via irreversible HAT to form the allylic radical of cyclohexene, followed by regioselectivity-determining radical coupling (for unsymmetrical olefins) and facile decyanation.
More Related Videos
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
 06:46
Facile Preparation of 2Z,4E-Dienamides by the Olefination of Electron-deficient Alkenes with Allyl Acetate
Published on: June 21, 2017
Related Concept Videos
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn...
 00:48
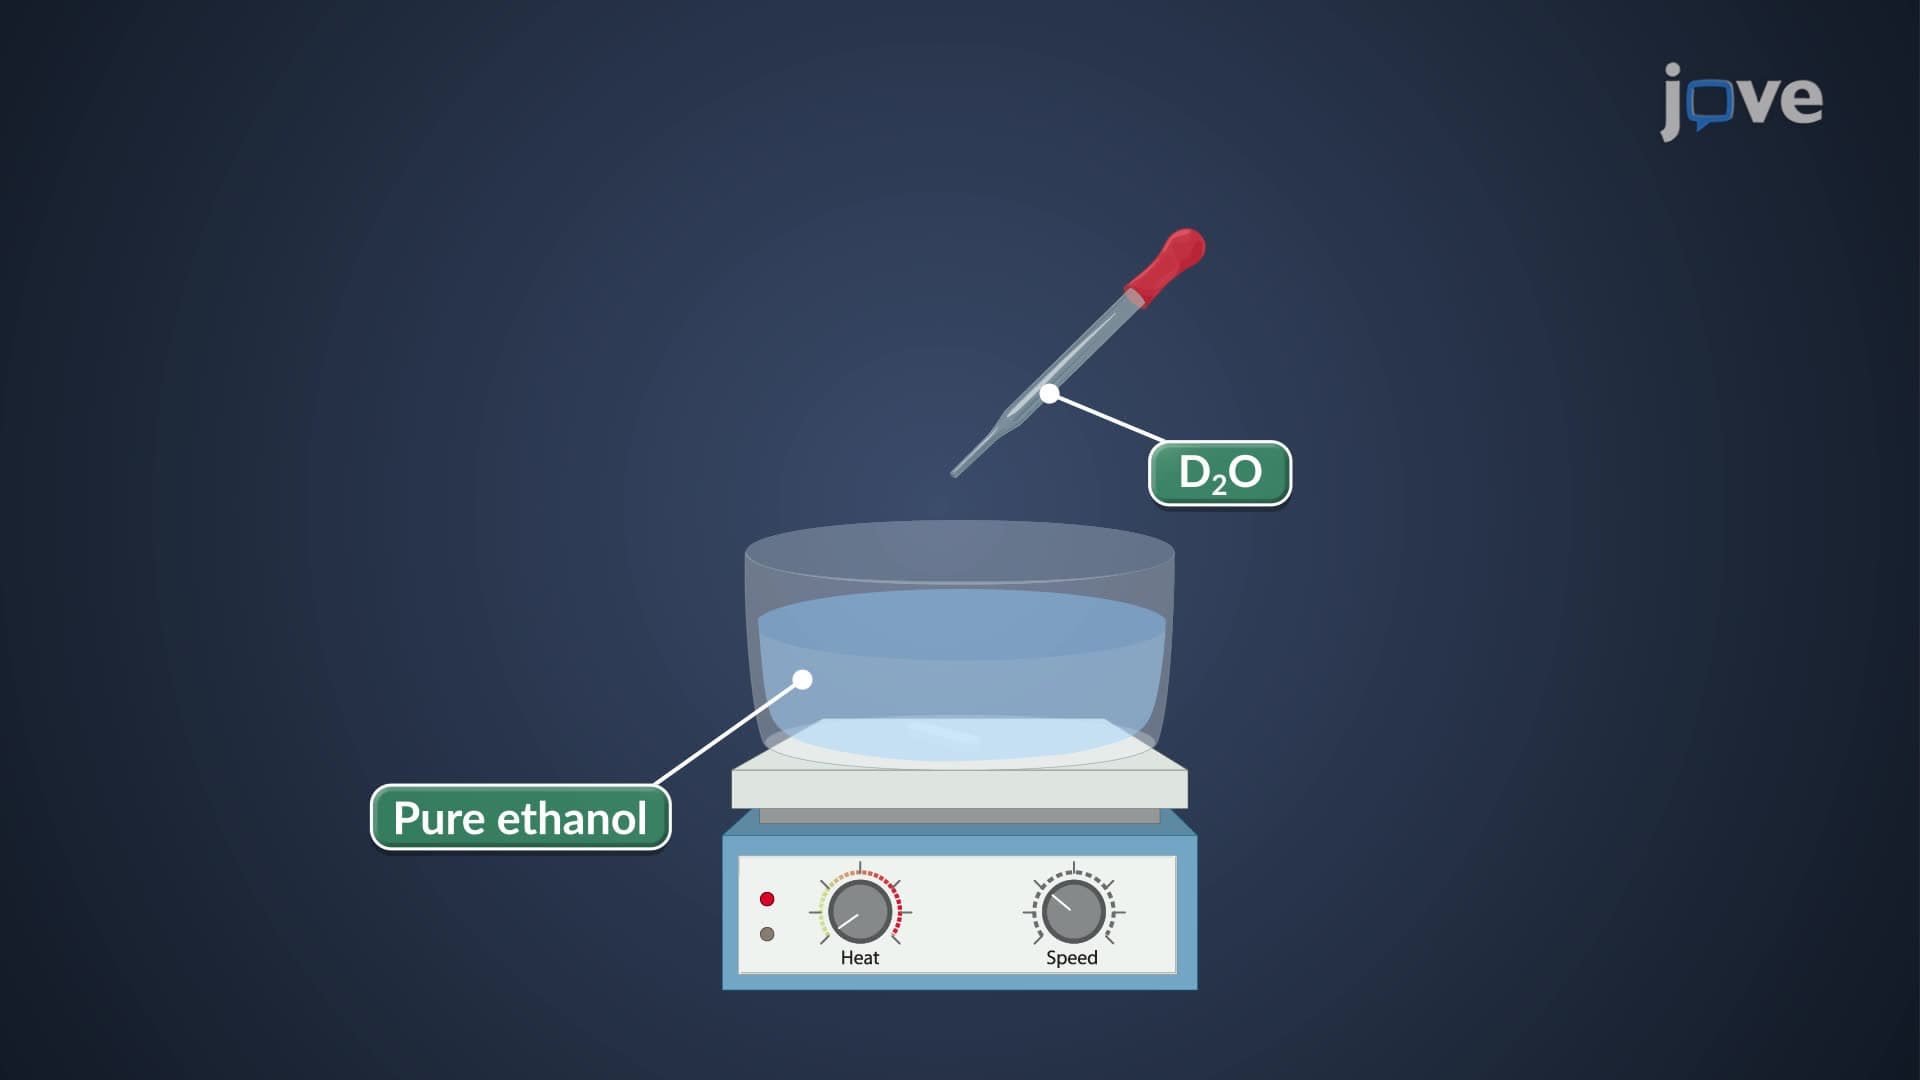
¹H NMR of Labile Protons: Deuterium (²H) Substitution
 02:46
E1 Reaction: Kinetics and Mechanism
 02:45
E2 Reaction: Kinetics and Mechanism
 02:44
Oxidation of Alkenes: Syn Dihydroxylation with Osmium Tetraoxide
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration