Related Experiment Video
Updated: Jun 16, 2025

Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Improved Prediction of Stabilizing Mutations in Proteins by Incorporation of Mutational Effects on Ligand Binding
Srivarshini Ganesan1, Nidhi Mittal1, Akash Bhat2
1Molecular Biophysics Unit (MBU), Indian Institute of Science, Bengaluru, India.

Abstract:
While many computational methods accurately predict destabilizing mutations, identifying stabilizing mutations has remained a challenge, because of their relative rarity. We tested ΔΔG 0 predictions from computational predictors such as Rosetta, ThermoMPNN, RaSP, and DeepDDG, using 82 mutants of the bacterial toxin CcdB as a test case. On this dataset, the best computational predictor is ThermoMPNN, which identifies stabilizing mutations with a precision of 68%. However, the average increase in T m for these predicted mutations was only 1°C for CcdB, and predictions were poorer for a more challenging target, influenza neuraminidase. Using data from multiple previously described yeast surface display libraries and in vitro thermal stability measurements, we trained logistic regression models to identify stabilizing mutations with a precision of 90% and an average increase in T m of 3°C for CcdB. When such libraries contain a population of mutants with significantly enhanced binding relative to the corresponding wild type, there is no benefit in using computational predictors. It is then possible to predict stabilizing mutations without any training, simply by examining the distribution of mutational binding scores. This avoids laborious steps of in vitro expression, purification, and stability characterization. When this is not the case, combining data from computational predictors with high-throughput experimental binding data enhances the prediction of stabilizing mutations. However, this requires training on stability data measured in vitro with known stabilized mutants. It is thus feasible to predict stabilizing mutations rapidly and accurately for any system of interest that can be subjected to a binding selection or screen.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:18
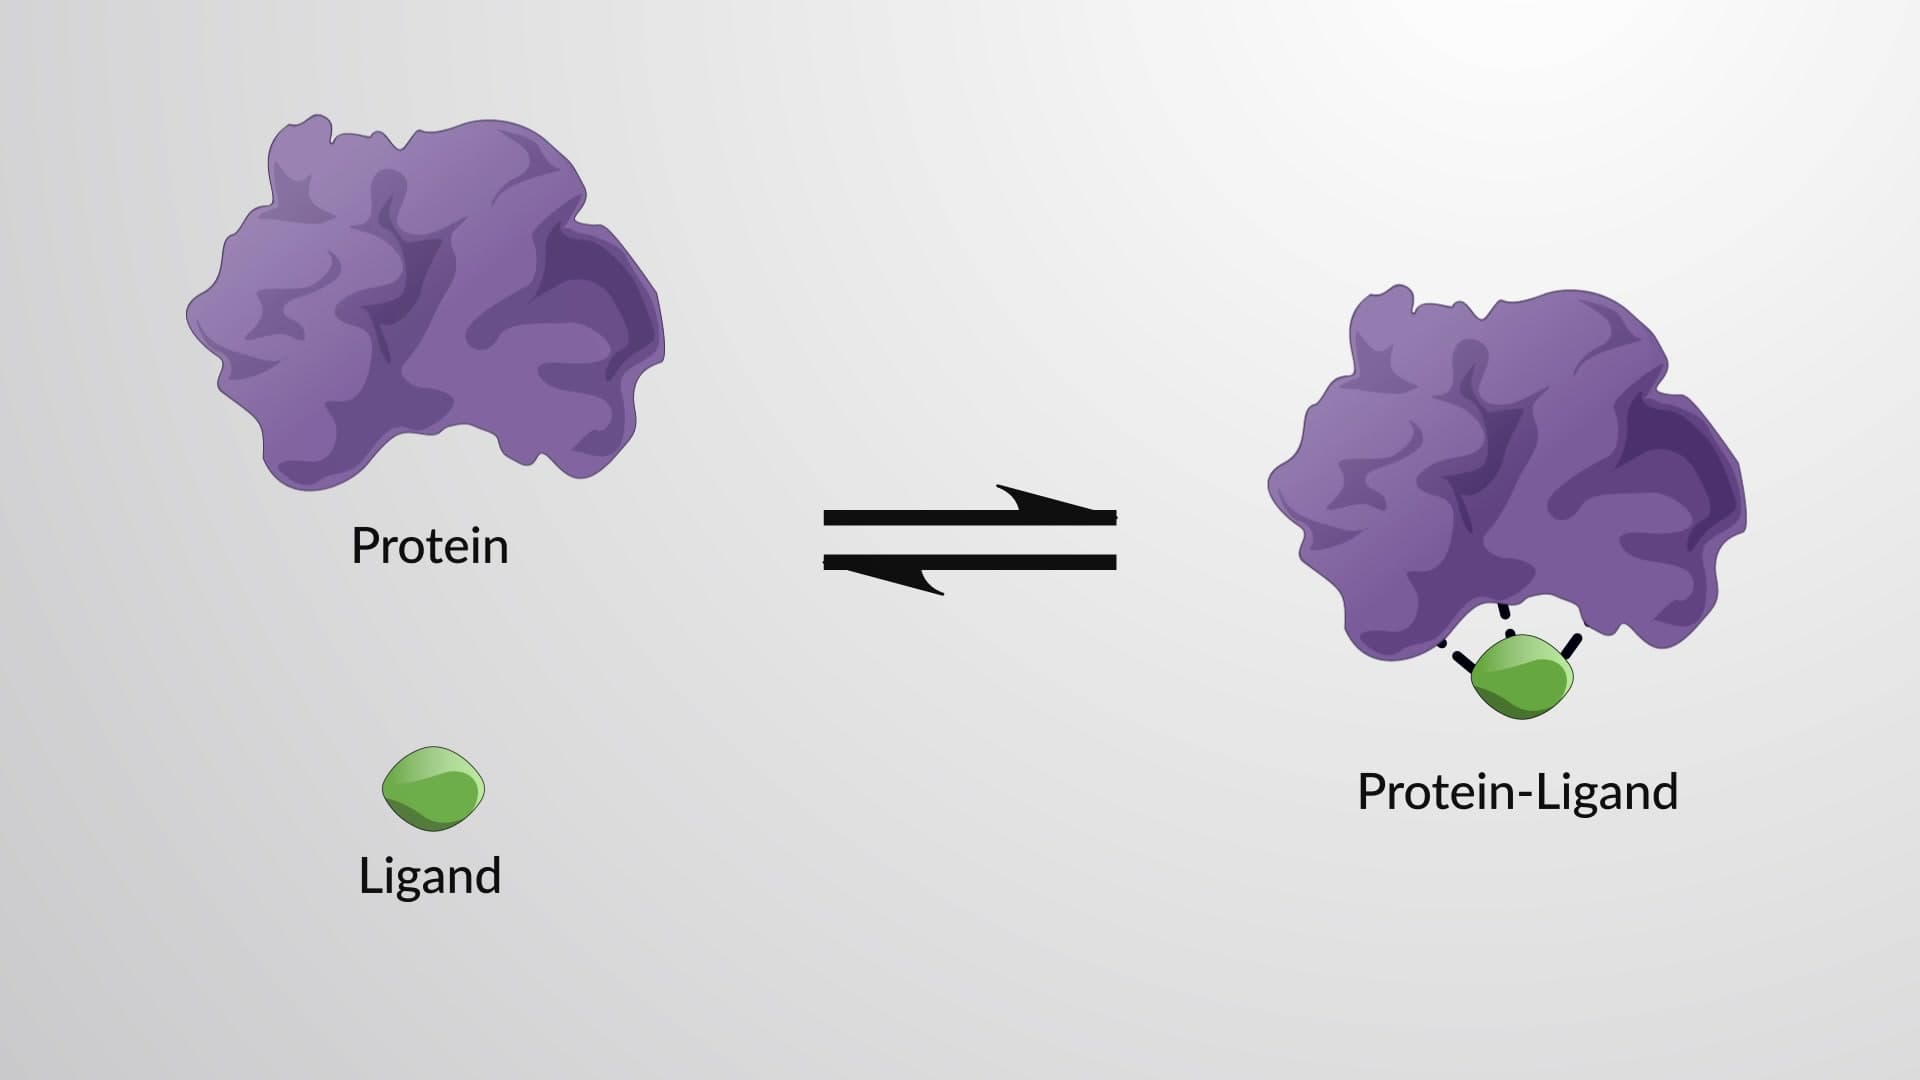
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage
 01:58
Cooperative Allosteric Transitions
 02:04
Covalently Linked Protein Regulators
These groups modify specific amino acids in a protein....