Related Experiment Video
Updated: Jun 15, 2025

From Molecules to Materials: Engineering New Ionic Liquid Crystals Through Halogen Bonding
Published on: March 24, 2018
Unusual quadruple bonds featuring collective interaction-type σ bonds between first octal-row atoms in the
Li-Juan Cui1, Yu-Qian Liu1, Sudip Pan1
1Institute of Atomic and Molecular Physics, Jilin University Changchun 130023 China zcui@jlu.edu.cn sudip@jlu.edu.cn.

Abstract:
Quantum chemical calculations are reported for the complexes of alkaline earth metals AeOLi2 (Ae = Be-Ba) at the BP86-D3(BJ)/def2-QZVPP and CCSD(T)/def2-QZVPPQZVPP levels. The nature of the Ae-OLi2 bond has been analyzed with a variety of methods. The AeOLi2 molecules exhibit an unprecedented σ donor bond Ae→OLi2 where the (n)s2 lone-pair electrons of the Ae atom are donated to vacant O-Li2 antibonding orbitals having the largest coefficient at lithium. This is a covalent bond where the accumulation of the associated electronic charge is located at two positions above and below the Ae-OLi2 axis. The bifurcated component of orbital interactions is structurally related to the recently proposed collective bonding model, but exhibits a completely different type of bonding. The most stable isomer of AeOLi2 has a C 2v geometry and a singlet (1A1) electronic ground state. The bond dissociation energy (BDE) of the Ae-OLi2 bonds exhibits a zig-zag trend from BeOLi2 to BaOLi2, with BeOLi2 having the largest BDE (D e = 73.0 kcal mol-1) and MgOLi2 possessing the lowest BDE (D e = 42.3 kcal mol-1) at the CCSD(T) level. The calculation of the atomic partial charges by the Hirshfeld and Voronoi methods suggests that Be and Mg carry small negative charges in the lighter molecules whereas the heavier atoms Ca-Ba have small positive charges. In contrast, the NBO and QTAIM methods give positive charges for all Ae atoms that are larger for Ca-Ba than that calculated by the Hirshfeld and Voronoi approaches. The molecules AeOLi2 have large dipole moments where the negative end is at the Ae atom with the polarity Ae→OLi2. The largest dipole moments are predicted for the lighter species BeOLi2 and MgOLi2 and the smallest value is calculated for BaOLi2. The calculation of the vibrational spectra shows a significant red-shift toward lower wave numbers for the Ae-OLi2 stretching mode in comparison to diatomic AeO. Besides the Ae→OLi2 σ-donor bonds there are also three dative bonds due to Ae←OLi2 backdonation which consist of one σ bond and two π bonds. The appearance of strong Ae→OLi2 σ donation leads to quadruple bonds AeOLi2 in all systems AeOLi2, even for the lightest species with Ae = Be, Mg. The valence orbitals of Ca, Sr, and Ba, which are involved in the dative interactions, are the (n)s and (n-1)d AOs whereas Be and Mg use their (n)s and (n)p AOs. The EDA-NOCV results are supported by the AdNDP calculations which give four 2c-2e bonding orbitals. Three bonding orbitals have occupation numbers ∼2. One σ orbital has smaller occupation numbers between 1.32 and 1.73 due to the delocalization to the lithium atoms. The analysis of the electronic structure with the ELF method suggests multicenter bonds with mainly trisynaptic and tetrasynaptic basins, which also support the results of the EDA-NOCV calculations.
More Related Videos
 10:42
Combining Solid-state and Solution-based Techniques: Synthesis and Reactivity of ChalcogenidoplumbatesII or IV
Published on: December 29, 2016
 10:27
Simultaneous Multi-surface Anodizations and Stair-like Reverse Biases Detachment of Anodic Aluminum Oxides in Sulfuric and Oxalic Acid Electrolyte
Published on: October 5, 2017
Related Concept Videos
 02:48
Ionic Bonding and Electron Transfer
 02:54
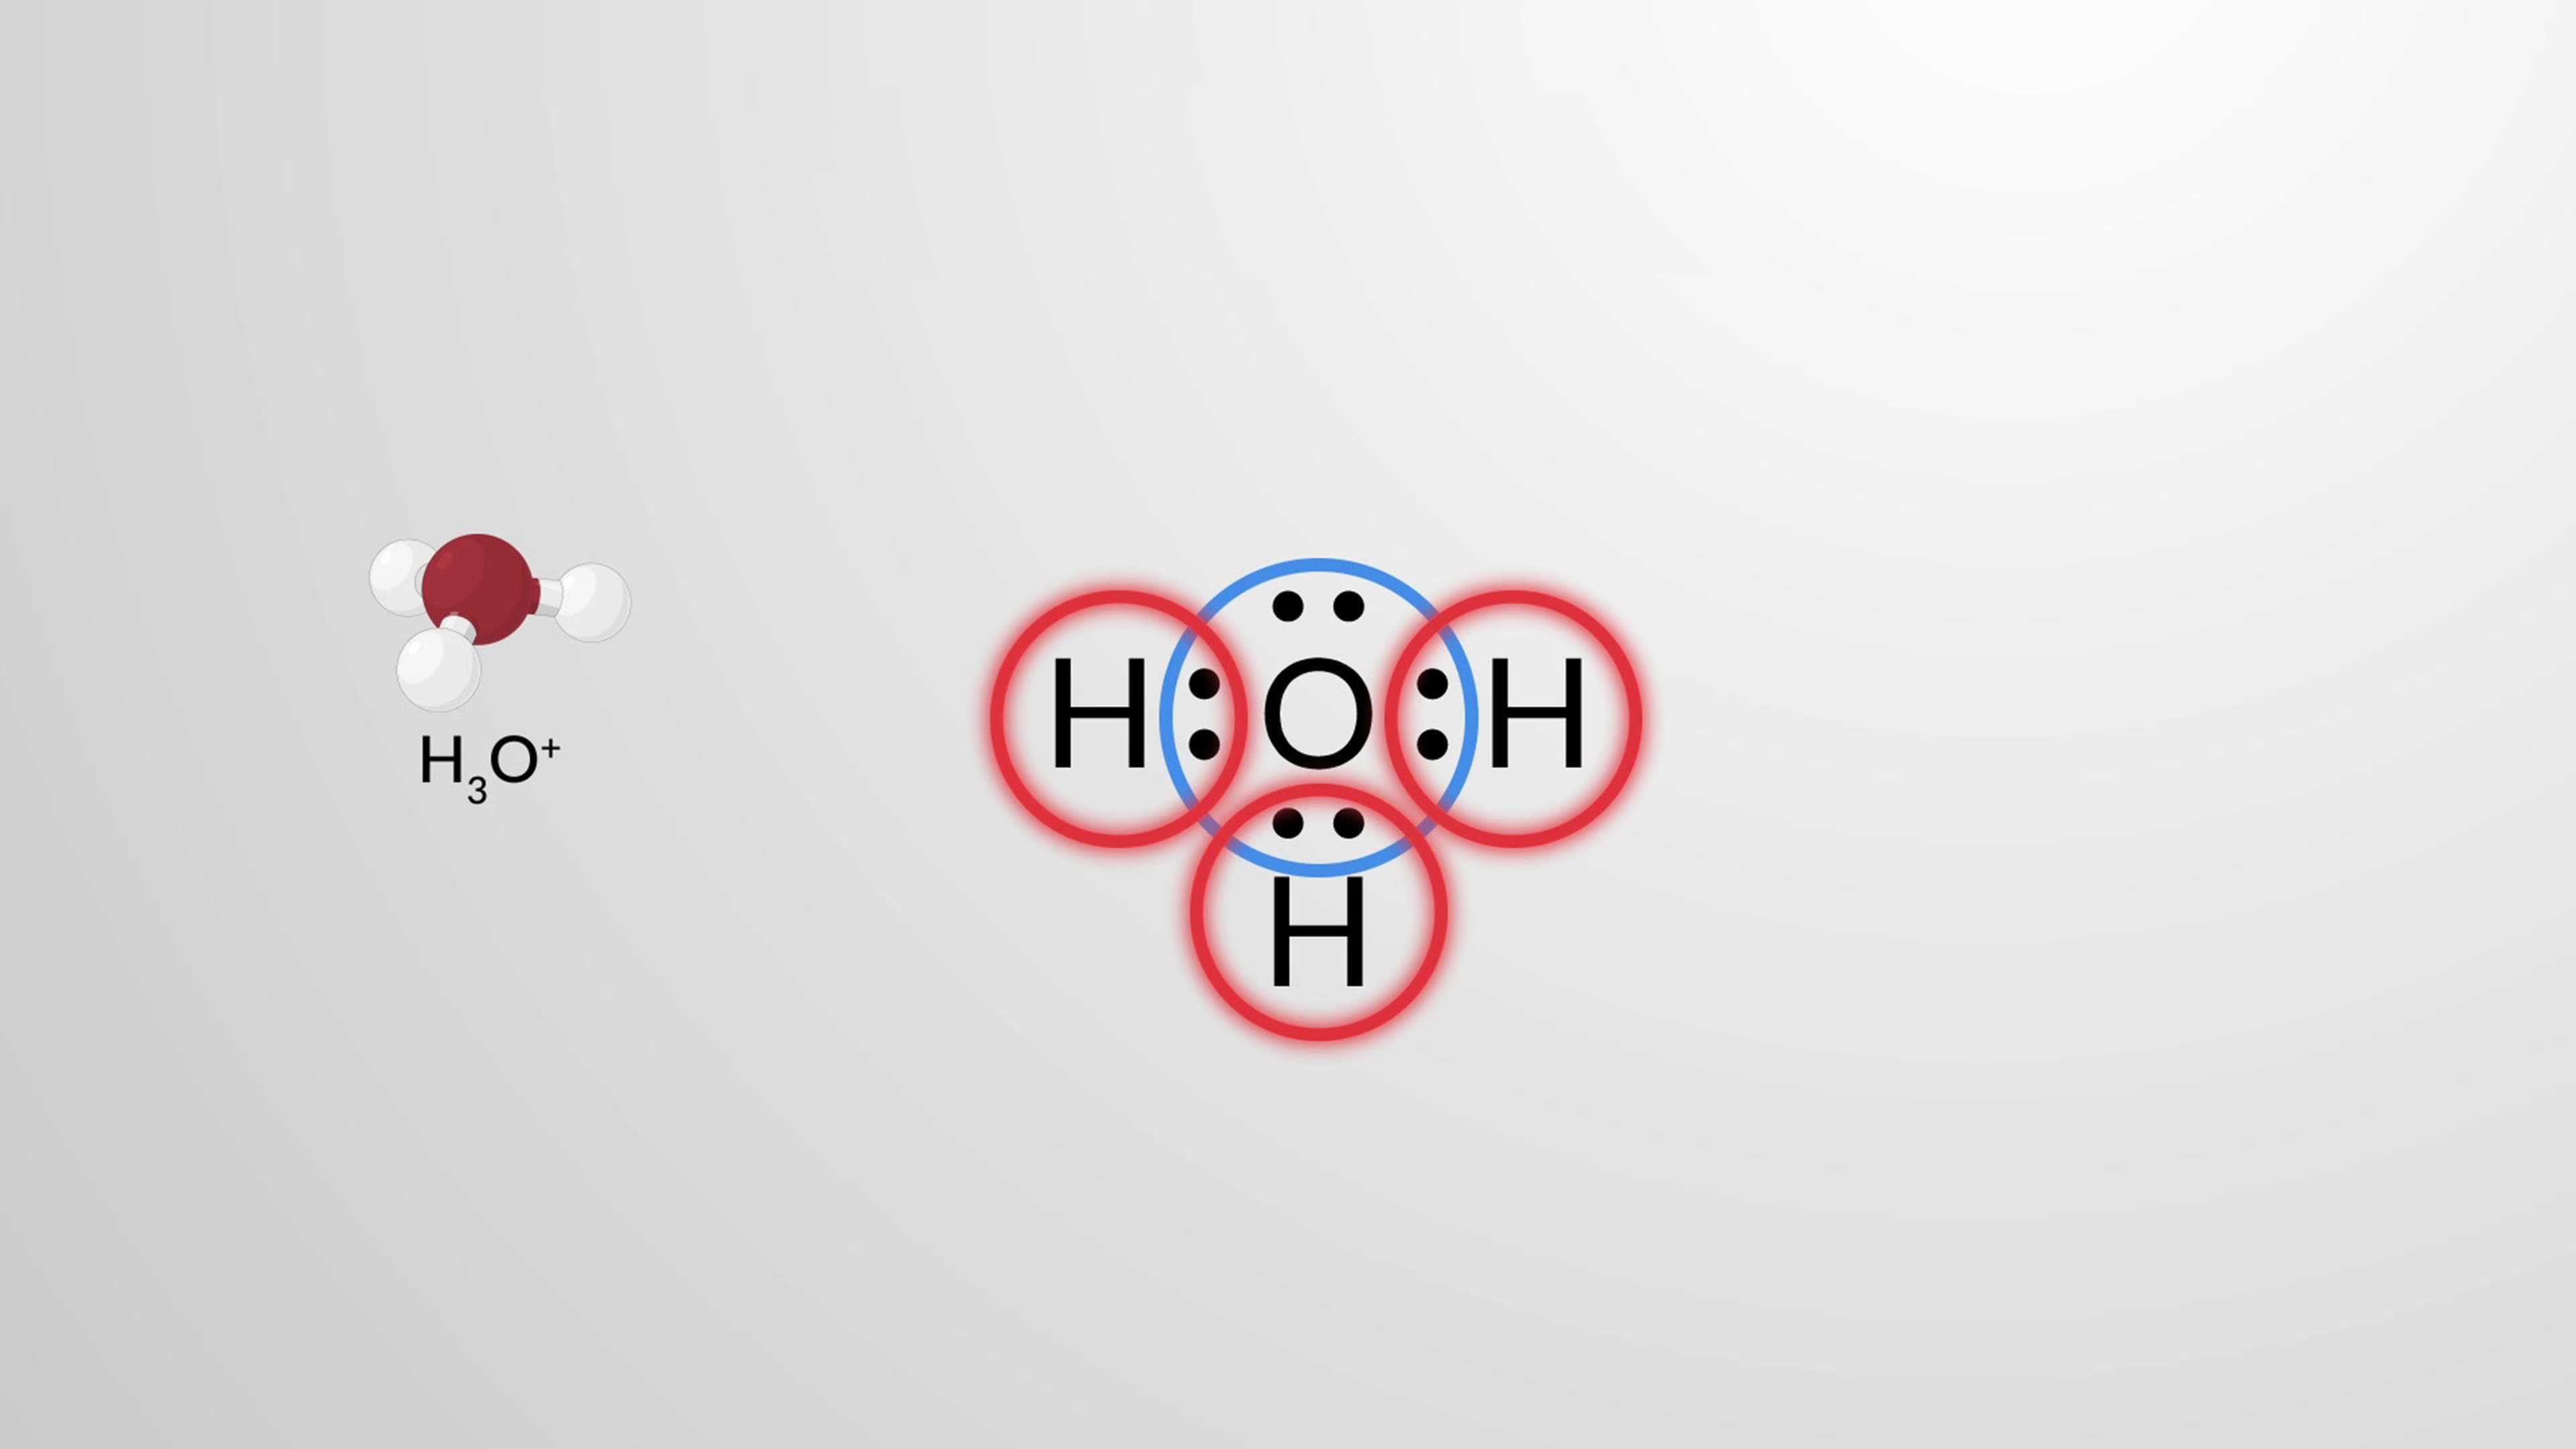
Lewis Structures of Molecular Compounds and Polyatomic Ions
 02:42
Valence Bond Theory
 03:51
Molecular Orbital Theory II
 02:46
Covalent Bonding and Lewis Structures
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...