Defective post-transcriptional modification of tRNA disrupts mitochondrial homeostasis in Leber's hereditary optic neuropathy
Related Experiment Video
Updated: Jun 14, 2025
Optogenetic Phase Transition of TDP-43 in Spinal Motor Neurons of Zebrafish Larvae
Published on: February 25, 2022
Juanjuan Zhang1, Wenxu Li2, Zhen Liu2
1State Key Laboratory of Ophthalmology, Optometry and Vision Science, Eye Hospital, Wenzhou Medical University, Wenzhou, Zhejiang, China; Attardi Institute of Mitochondrial Biomedicine, School of Laboratory Medicine and Life Sciences, Wenzhou Medical University, Wenzhou, Zhejiang, China.

Abstract:
Leber's Hereditary Optic Neuropathy (LHON) is a rare, maternally inherited eye disease, predominantly due to the degeneration of retinal ganglion cells (RGCs). It is associated with a mitochondrial DNA (mtDNA) point mutation. Our previous study identified that the m.15927G > A homoplasmic mutation damaged the highly conserved base pairing (28C-42G) in anticodon stem of tRNAThr, caused deficient t6A modification and significantly decreased efficiency in aminoacylation and steady-state levels of tRNAThr, and led to mitochondrial dysfunction. Meanwhile, mechanisms underlying mtDNA mutations regulate intracellular signaling related to mitochondrial and cellular integrity are less explored. Here, we manifested that defective nucleotide modification induced by the m.15927G > A mutation interfered with the expression of nuclear genes involved in cytoplasmic proteins essential for oxidative phosphorylation system (OXPHOS), thereby impacting the assemble and integrity of OXPHOS complexes. As a result of these mitochondrial dysfunctions, there was an imbalance in mitochondrial dynamics, particularly distinguished by an increased occurrence of mitochondrial fission. Excessive fission compromised the autophagy process, including the initiation phase, formation, and maturation of autophagosomes. Both Parkin-mediated mitophagy and receptor-dependent mitophagy were significantly impaired in cybrids haboring the m.15927G > A mutation. These changes facilitated intrinsic apoptosis, as indicated by increased cytochrome c release and elevated levels of apoptosis-associated proteins (e.g., BAK, BAX, cleaved caspase 9, cleaved caspase 3, and cleaved PARP) in the mutant cybrids. This study demonstrates that the m.15927G > A mutation contributes to LHON by dysregulating OXPHOS biogenesis, aberrant quality control, increased autophagy, inhibited mitophagy, and abnormal apoptosis.
Related Concept Videos
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
 01:48
ATP Synthase: Mechanism
 02:31
Nuclear Export of mRNA
 02:23
RNA Editing
 02:27
Nonsense-mediated mRNA Decay
Usually, Upf3 binds to an Exon Junction Complex (EJC) at mRNA splice sites. If a ribosome fully translates the mRNA,...
 01:19
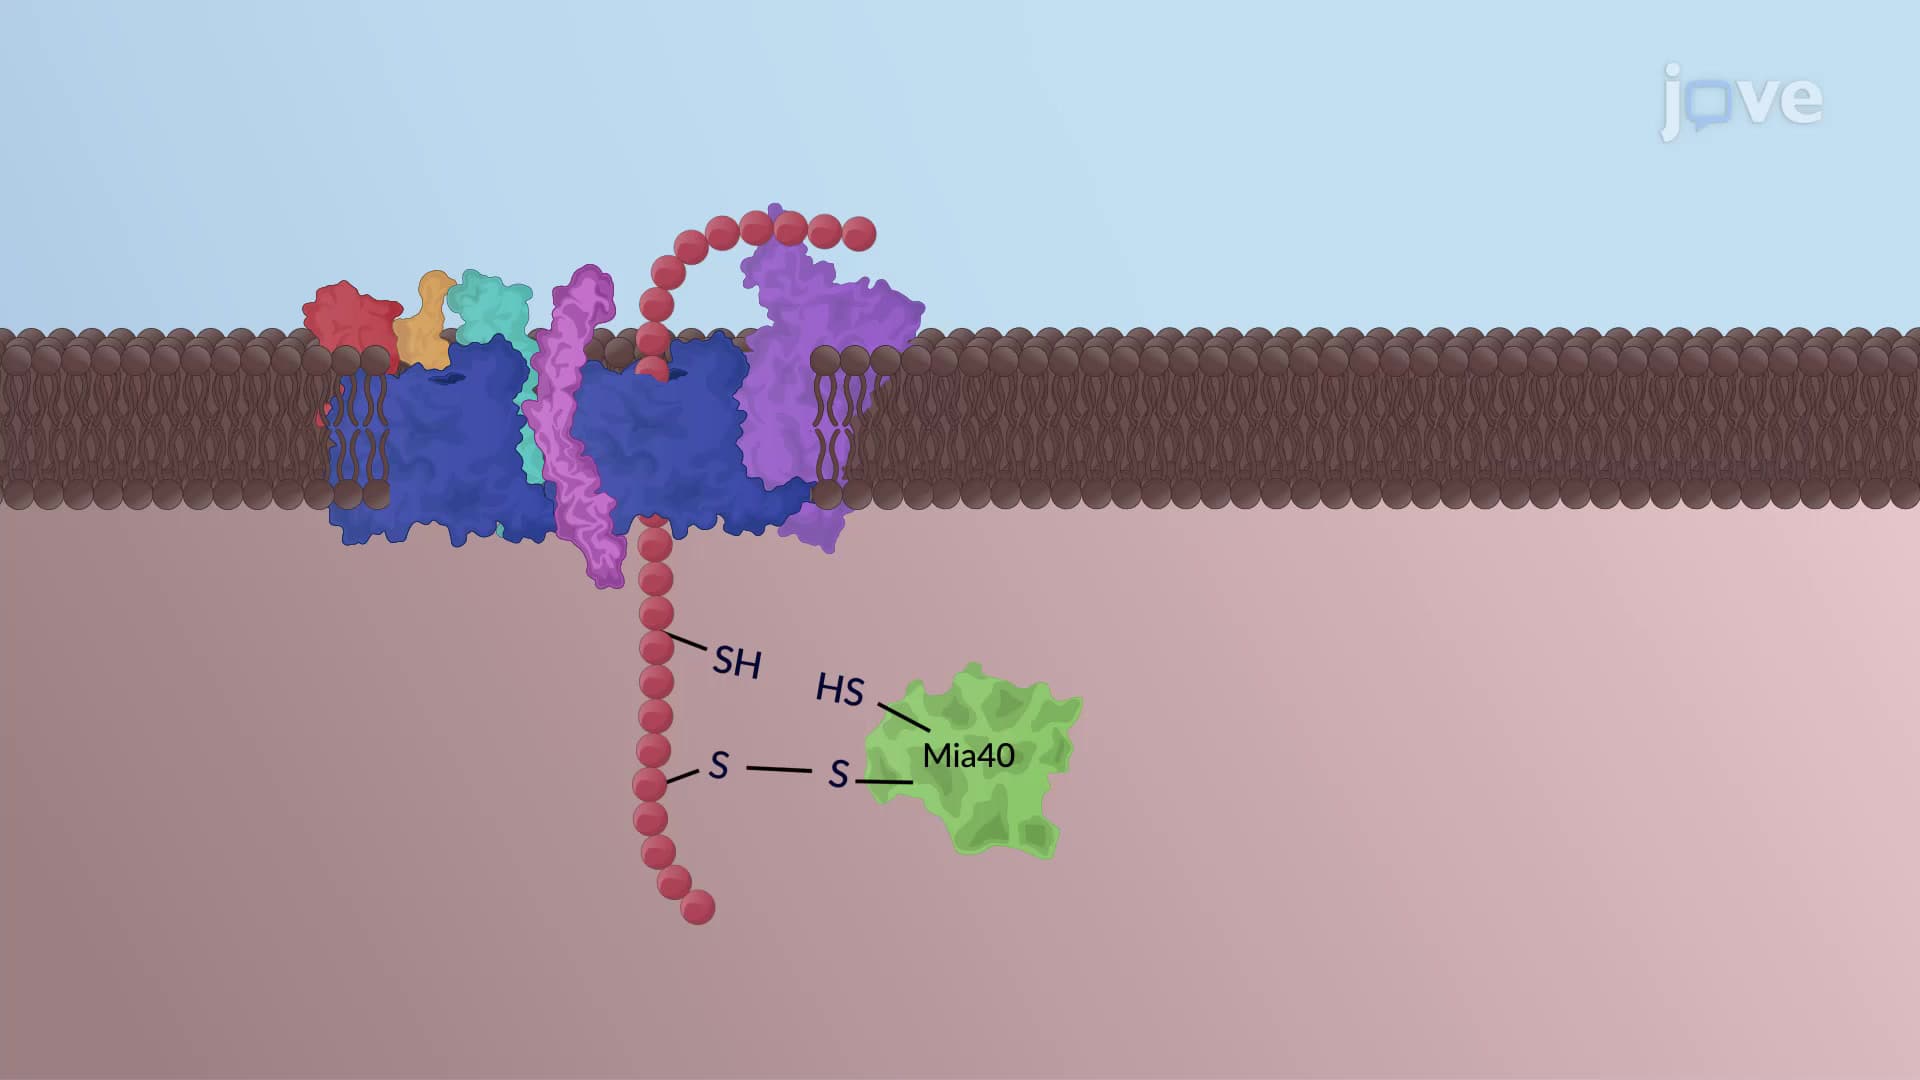
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...