Related Experiment Video
Updated: Jun 12, 2025
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Energy spectra with the Dirac equation of the q-deformed generalized Pöschl-Teller potential via the Feynman approach
Amina Ghobrini1,2, Hocine Boukabcha3, Ismahane Ami1,4
1Department of Physics, Faculty of Sciences, University M'hamed Bougara of Boumerdes, Route de la Gare Ferroviaire, Boumerdes, 35000, Boumerdes, Algeria.

Context:
The diatomic molecules of potassium is widely used in industrial chemicals and alternative energy. Besides that, is very useful for researching molecular interactions and energy states, especially in the context of quantum chemistry and spectroscopy. In the present work, a newly proposed diatomic potential model within relativistic and non-relativistic quantum mechanics has been considered, to obtain corresponding energy eigenvalues and related normalized eigenfunctions.
Methods:
The Dirac equation has been solved for an arbitrary spin-orbit quantum number using the path integral technique with the -deformed generalized Pöschl-Teller potential . By including a Pekeris-type approximation to handle the centrifugal factor, it was possible to obtain the spin and pseudospin-symmetric solution of the relativistic energy eigenvalues and wave equation. To assess the correctness of this work, Maple software was used to present some numerical findings for various values of and . With the constraint , it was shown that in the situation of pseudospin symmetry, only bound states exist with negative energy. In the non-relativistic limits, the non-relativistic ro-vibrational energy expression of the diatomic molecule is derived from the relativistic energy equation under spin symmetry. Under Varshni conditions, both vibrational and ro-vibrational energies of the molecule were computed and compared with the data. The average absolute percentage deviations from the data obtained for the potassium molecule are . This demonstrates that the model is a very consistent model to study and characterize diatomic molecules.

Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 01:20
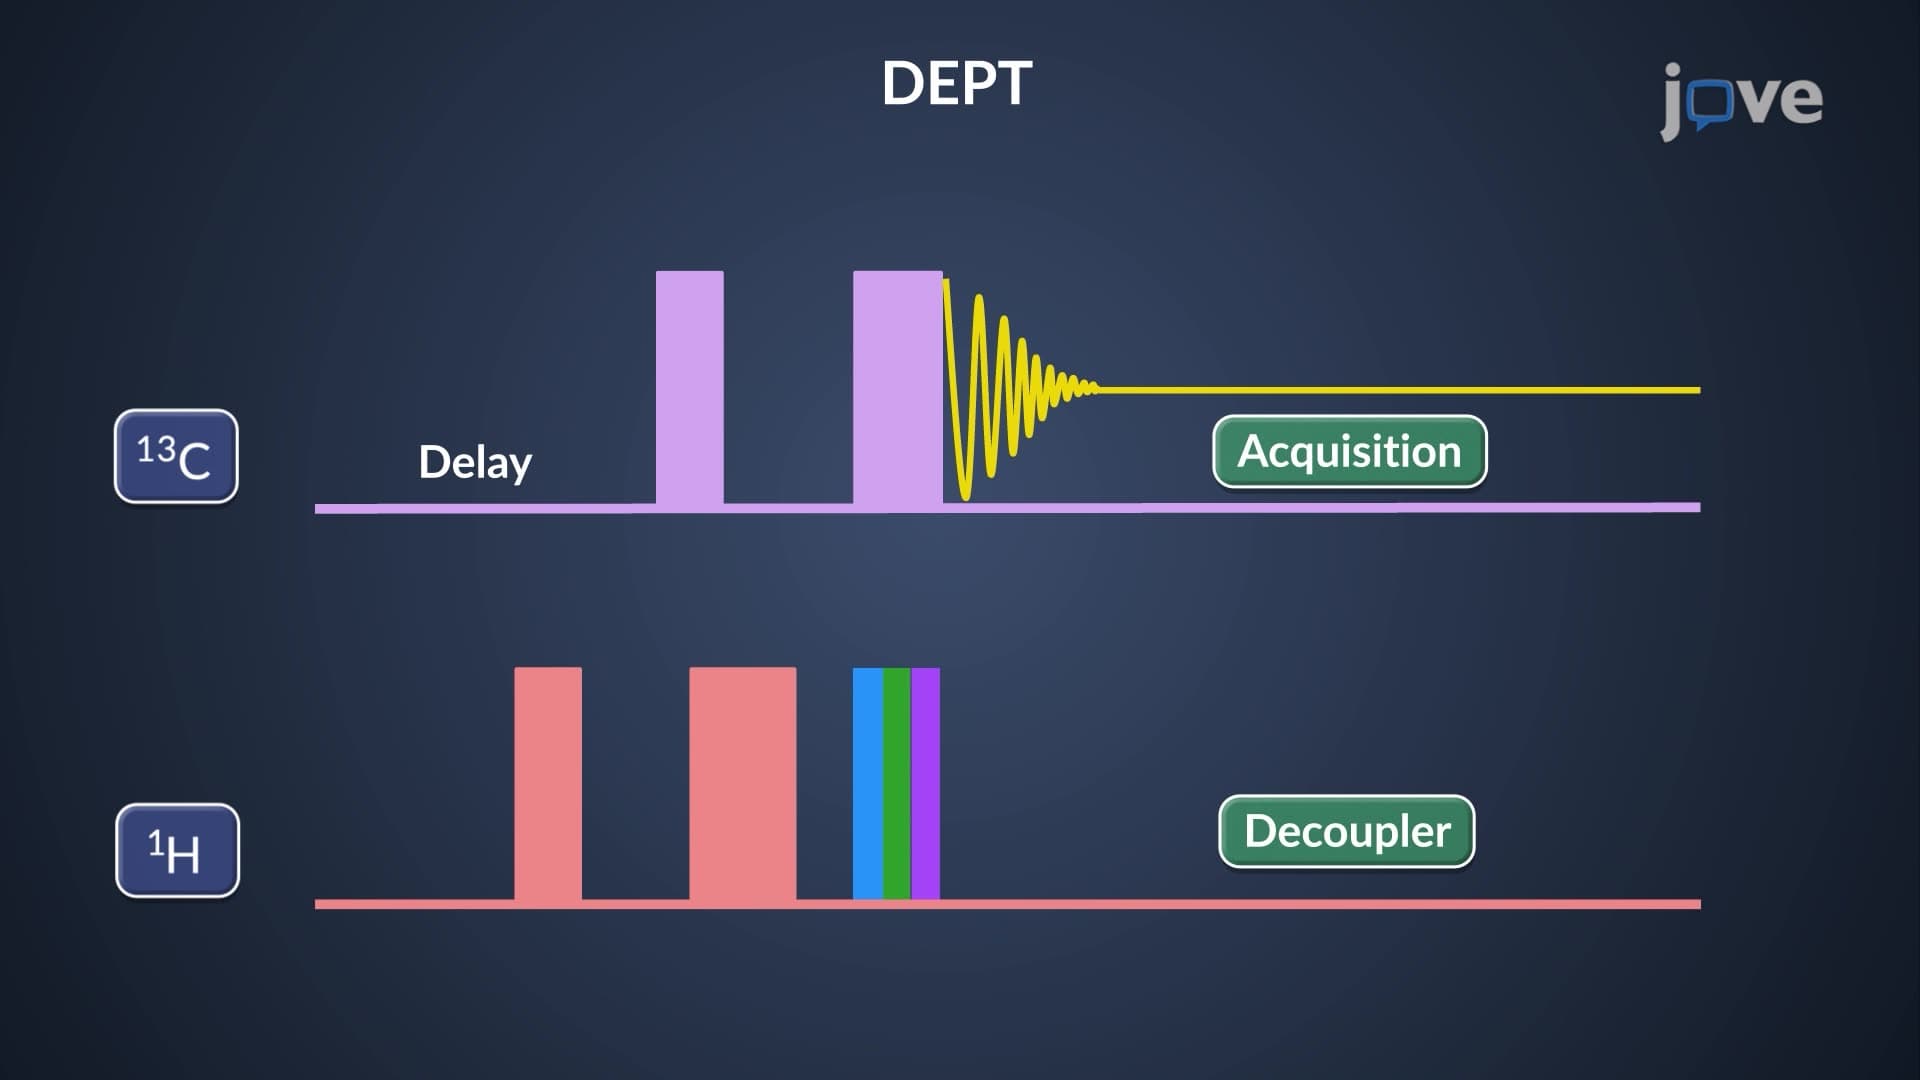
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:32
The de Broglie Wavelength
 03:06
The Pauli Exclusion Principle
 01:29
Potential Due to a Polarized Object
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...