Related Experiment Video
Updated: Jun 12, 2025

Novel 3D/VR Interactive Environment for MD Simulations, Visualization and Analysis
Published on: December 18, 2014
Constructing potential energy surface for carbon-chain containing systems using the radial angular network with
1University Rennes, CNRS, IPR (Institut de Physique de Rennes) - UMR 6251, F-35000 Rennes, France.

Abstract:
Investigating molecular excitation induced by collisions requires the prior determination of accurate analytical potential energy surfaces for the colliding partners. For carbon-chain molecules, such as cyanopolyynes, this has been a longstanding challenge, resulting in the absence of rate coefficients for HC5N, HC7N, HC9N, and others, induced by collisions with He. To overcome this bottleneck, we introduce a new approach: the Radial Angular Network with Gradual Expansion (RANGE). This method jointly connects the construction of ab initio interaction potentials with the determination of their analytical forms. We use the HC3N-He molecular complex as a reference to assess the reliability of our method, given that its analytical potential has been derived using various methods. Additionally, we apply the RANGE approach to construct the analytical representation of the interaction potential for HC5N-He and HC7N-He. The analysis of the analytical potentials reveals three systematic trends: (i) the anisotropy increases with the length of the carbon chain, (ii) the number of local minima correlates with the number of carbon atoms, and (iii) the shallowest local minimum is consistently located at ∼30 cm-1 below the dissociation limit of the complex. Using the time-independent quantum mechanical close-coupling formalism, we briefly estimate the propensity rules governing the excitation of HC3N, HC5N, and HC7N induced by collisions with He. Consequently, the three collisional systems exhibit the same propensity rule, favoring Δj = 2 transitions.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 07:16
Author Spotlight: Development of a Novel Finite Element Analysis Model for Improved Orthognathic Surgical Techniques
Published on: October 20, 2023
Related Concept Videos
 01:17
Radical Chain-Growth Polymerization: Chain Branching
 01:10
Radical Chain-Growth Polymerization: Overview
 01:09
Radical Chain-Growth Polymerization: Mechanism
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...
 02:29
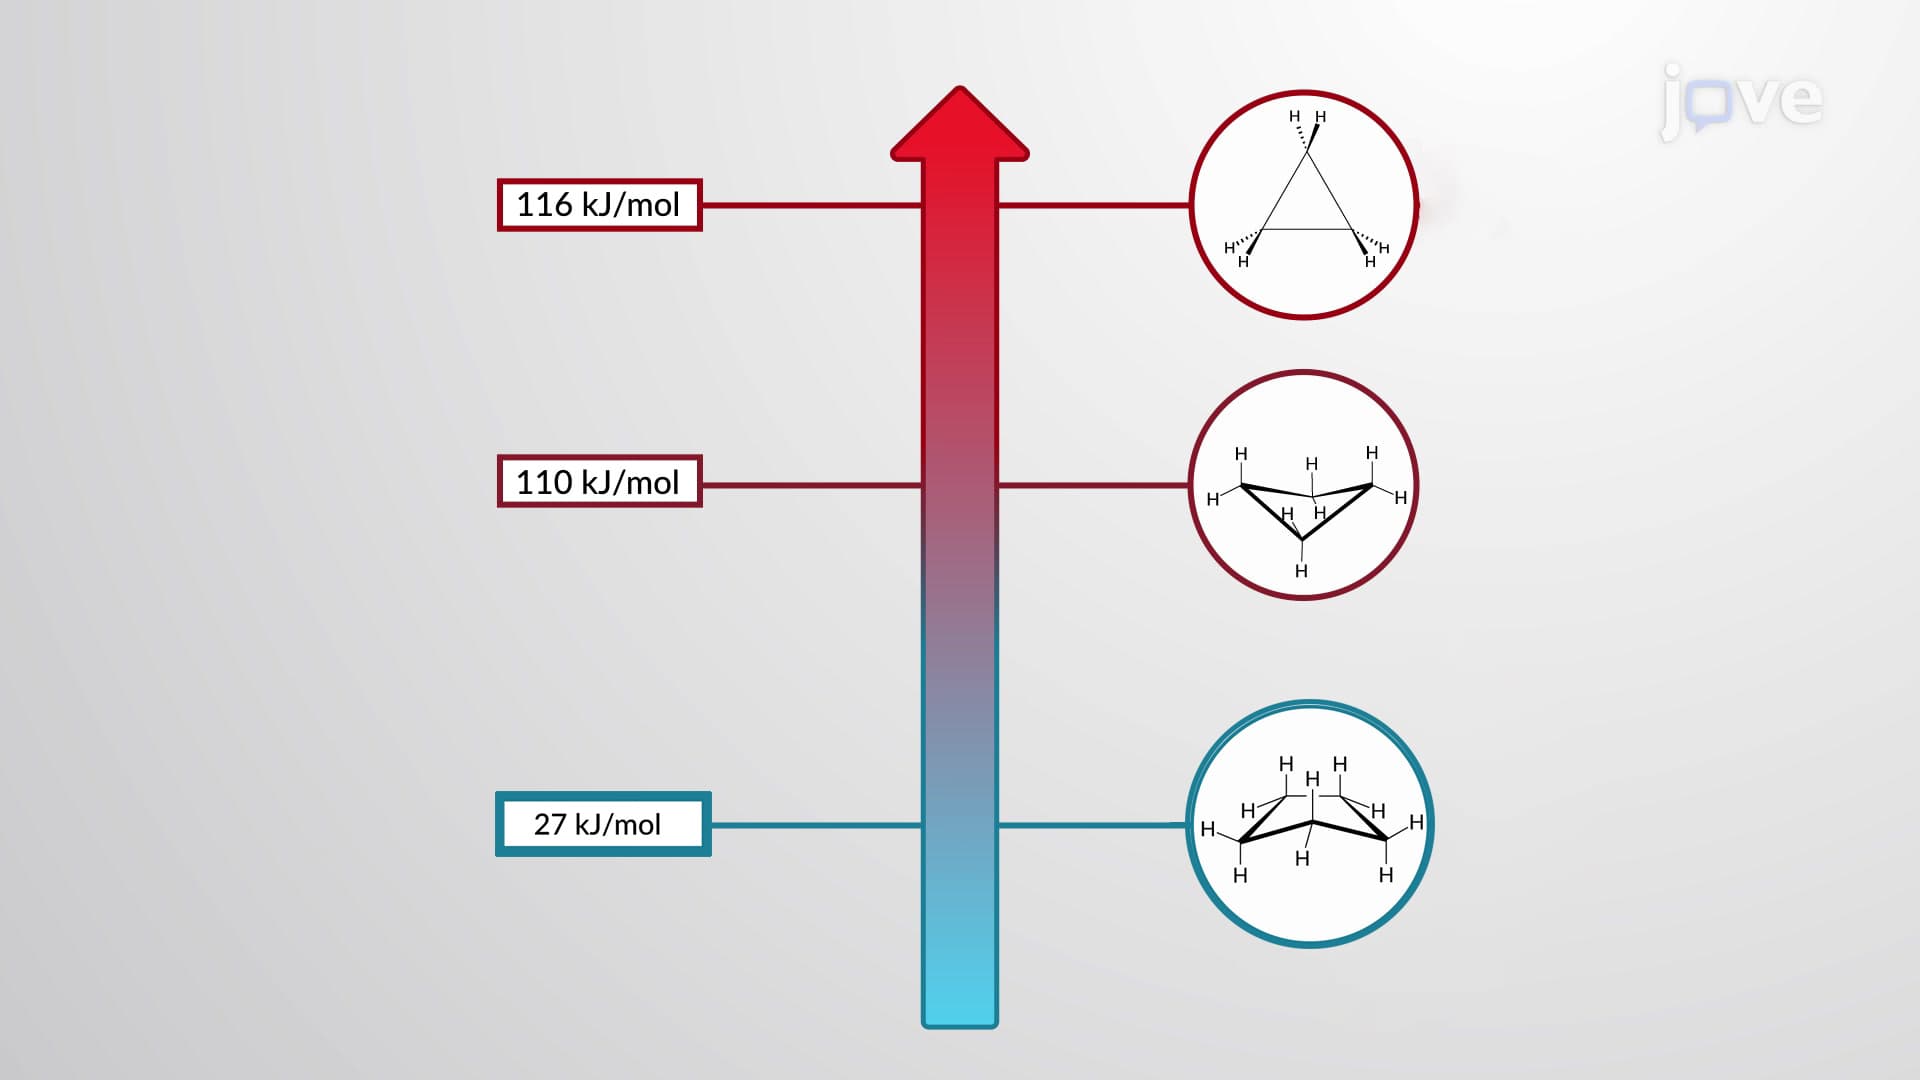
Conformations of Cycloalkanes
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...