Related Experiment Video
Updated: Jun 12, 2025

Author Spotlight: Decoding Mitochondrial Aging
Published on: June 30, 2023
Primary mitochondrial diseases
Chiara Pizzamiglio1, Michael G Hanna1, Robert D S Pitceathly1
1Department of Neuromuscular Diseases, UCL Queen Square Institute of Neurology, London, United Kingdom; NHS Highly Specialised Service for Rare Mitochondrial Disorders, Queen Square Centre for Neuromuscular Diseases, National Hospital for Neurology and Neurosurgery, London, United Kingdom.

Abstract:
Primary mitochondrial diseases (PMDs) are a heterogeneous group of hereditary disorders characterized by an impairment of the mitochondrial respiratory chain. They are the most common group of genetic metabolic disorders, with a prevalence of 1 in 4,300 people. The presence of leukoencephalopathy is recognized as an important feature in many PMDs and can be a manifestation of mutations in both mitochondrial DNA (classic syndromes such as mitochondrial encephalomyopathy, lactic acidosis, and stroke-like episodes; myoclonic epilepsy with ragged-red fibers [RRFs]; Leigh syndrome; and Kearns-Sayre syndrome) and nuclear DNA (mutations in maintenance genes such as POLG, MPV17, and TYMP; Leigh syndrome; and mitochondrial aminoacyl-tRNA synthetase disorders). In this chapter, PMDs associated with white matter involvement are outlined, including details of clinical presentations, brain MRI features, and elements of differential diagnoses. The current approach to the diagnosis of PMDs and management strategies are also discussed. A PMD diagnosis in a subject with leukoencephalopathy should be considered in the presence of specific brain MRI features (for example, cyst-like lesions, bilateral basal ganglia lesions, and involvement of both cerebral hemispheres and cerebellum), in addition to a complex neurologic or multisystem disorder. Establishing a genetic diagnosis is crucial to ensure appropriate genetic counseling, multidisciplinary team input, and eligibility for clinical trials.
Insights
Primary mitochondrial diseases (PMDs) are genetic metabolic disorders affecting the respiratory chain. Leukoencephalopathy, indicated by specific MRI findings, suggests PMD and requires genetic diagnosis for proper management.
Area of Science:
- Neurology
- Genetics
- Metabolic Disorders
Background:
- Primary mitochondrial diseases (PMDs) are common genetic metabolic disorders impacting the mitochondrial respiratory chain.
- Leukoencephalopathy is a significant feature in many PMDs, stemming from mitochondrial or nuclear DNA mutations.
- PMDs affect approximately 1 in 4,300 individuals.
Purpose of the Study:
- To outline PMDs associated with white matter involvement.
- To detail clinical presentations, MRI findings, and differential diagnoses for these conditions.
- To discuss diagnostic approaches and management strategies for PMDs.
Main Methods:
- Review of PMDs with leukoencephalopathy, including genetic causes (mtDNA and nDNA).
- Analysis of clinical and neuroimaging (brain MRI) features.
- Discussion of diagnostic criteria and genetic testing approaches.
Main Results:
- Specific brain MRI features (e.g., cyst-like lesions, basal ganglia involvement) aid in suspecting PMD in patients with leukoencephalopathy.
- A complex neurological or multisystem disorder combined with characteristic MRI findings supports a PMD diagnosis.
- Genetic diagnosis is essential for personalized care and clinical trial eligibility.
Conclusions:
- PMDs with leukoencephalopathy require consideration based on clinical and MRI evidence.
- Establishing a genetic diagnosis is critical for patient management, counseling, and research participation.
- Multidisciplinary input and genetic confirmation are key for addressing these complex disorders.
More Related Videos
 06:07
Author Spotlight: Establishing a New Fluorescence-Based Protocol for In Vivo Mitochondrial Morphology Analysis in Parkinson's Disease
Published on: June 23, 2023
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase COX/SDH Double-labeling Histochemistry
Published on: November 23, 2011
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:48
ATP Synthase: Mechanism
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:37
Mitochondria
 01:19
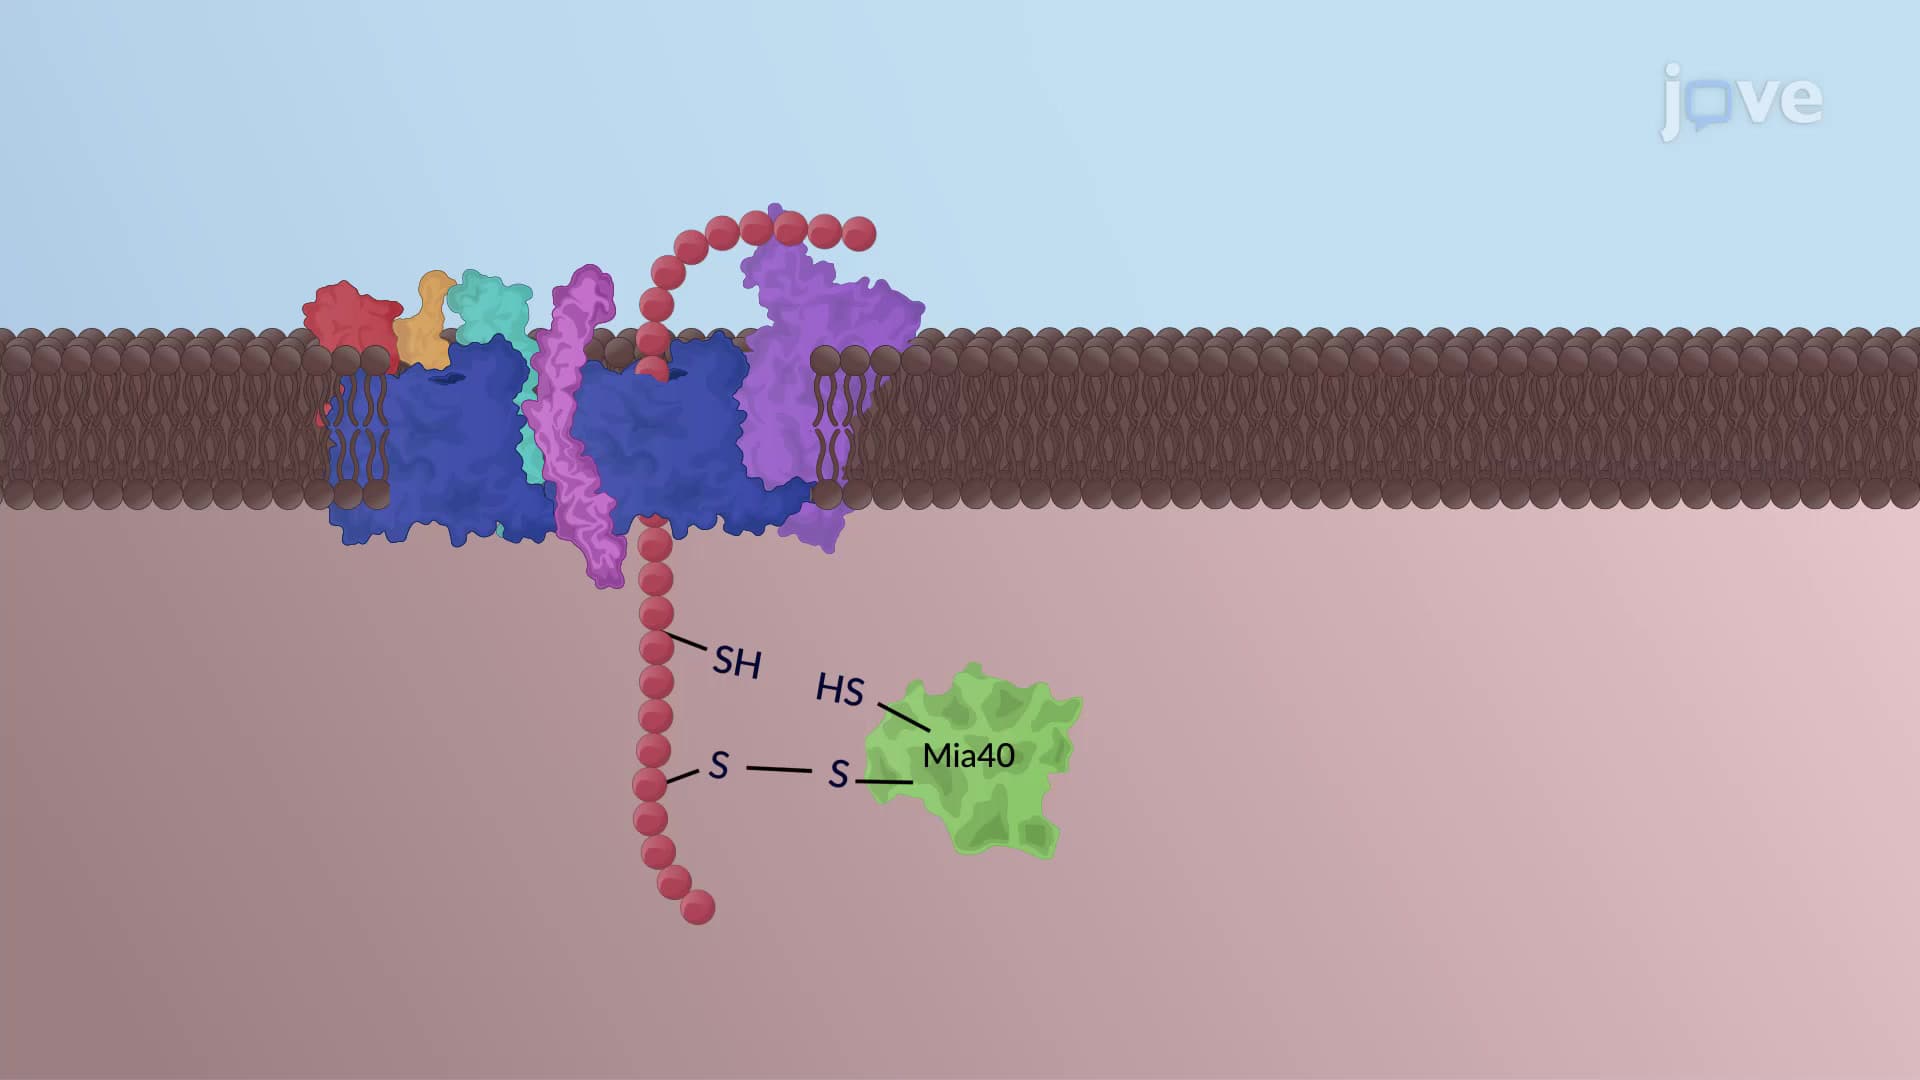
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
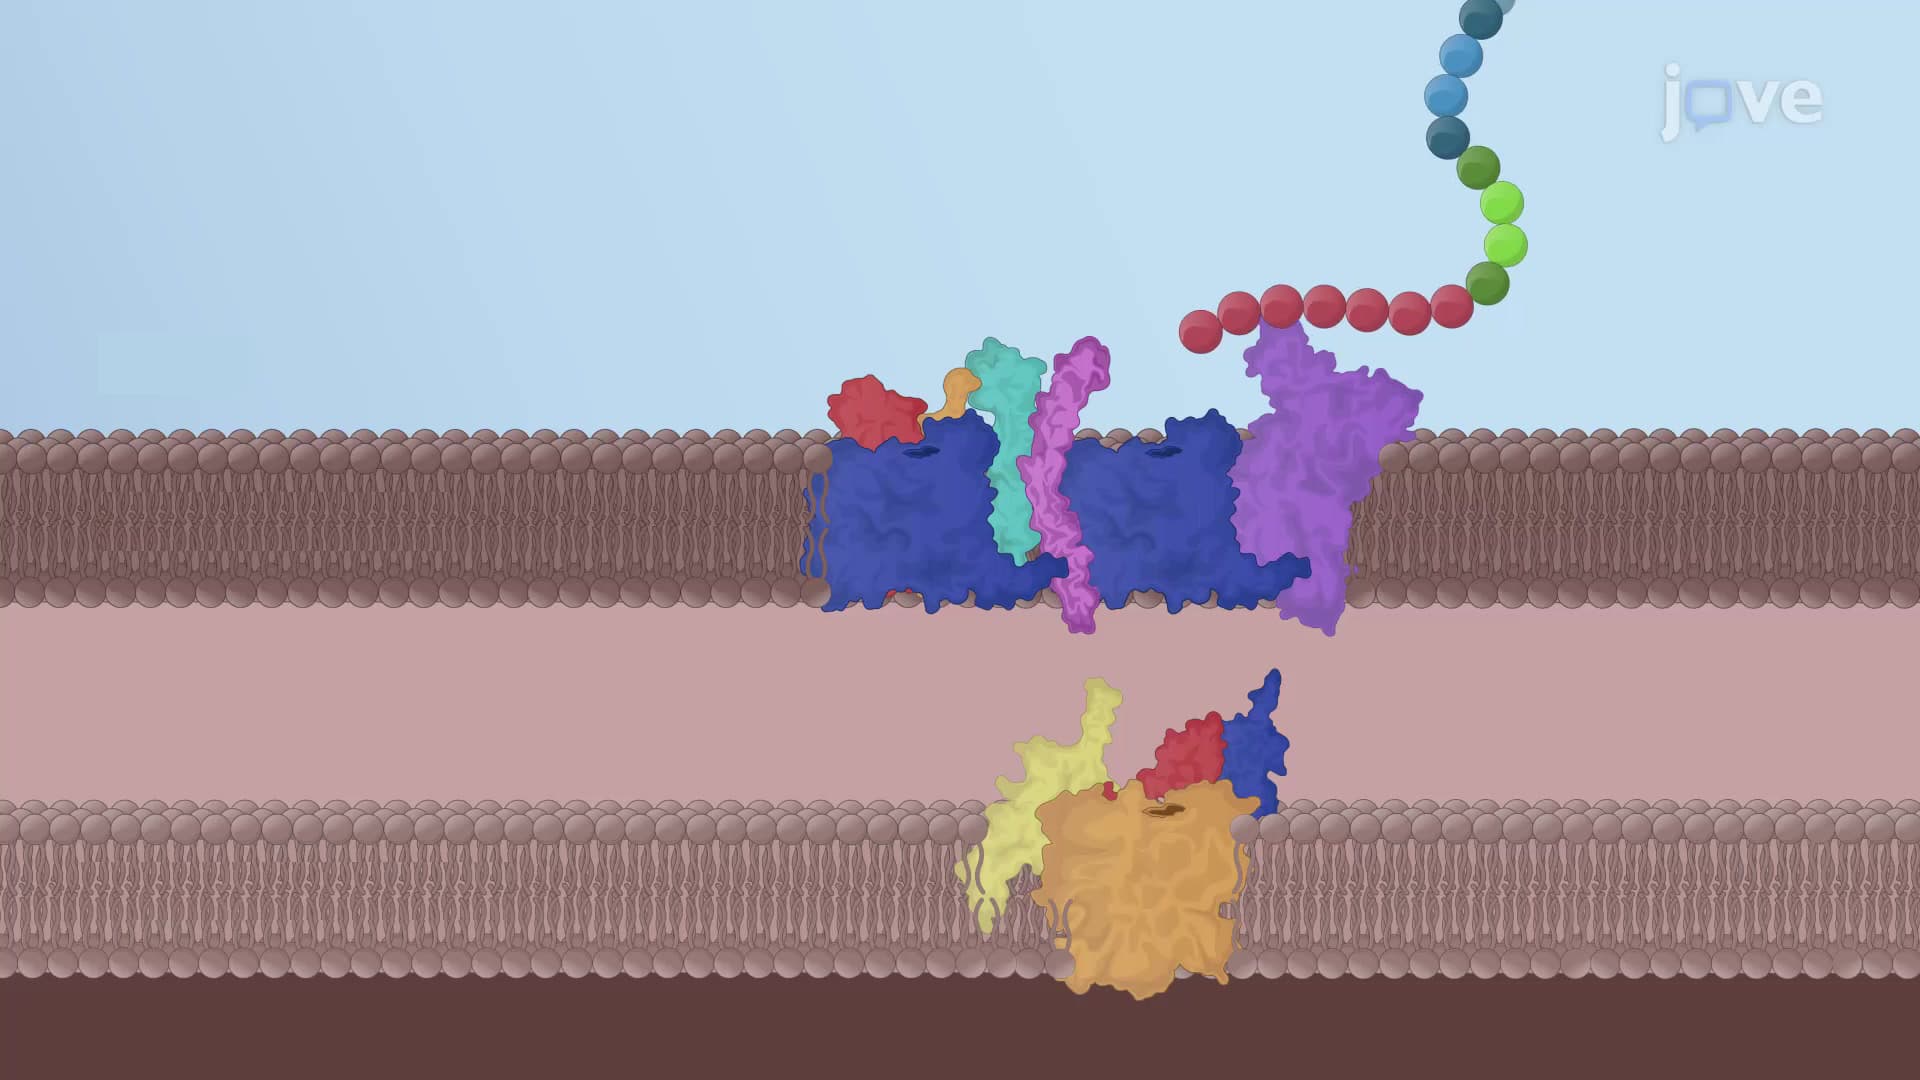
Mitochondrial Precursor Proteins
Most of the mitochondrial...