Related Experiment Video
Updated: Jun 11, 2025
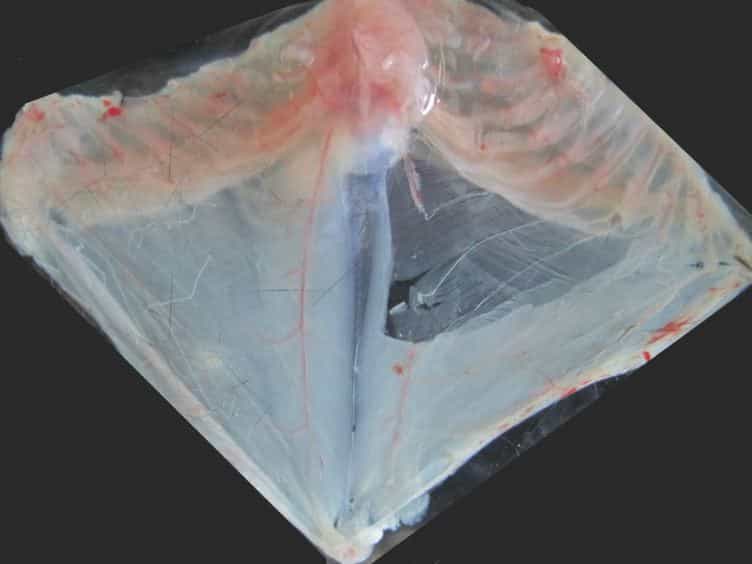

Dissection of the Transversus Abdominis Muscle for Whole-mount Neuromuscular Junction Analysis
Published on: January 11, 2014
[Clinical characteristics and genetics functional analysis of two children with Spinal muscular atrophy]
Wenchen Huang1, Jinli Bai, Hong Wang
1Department of Medical Genetics, Capital Institute of Pediatrics, Beijing 100020, China. hurry_qu@sina.com.

Objetive:
To explore the characteristics of SMN1 gene variants and carry out functional verification for two children with Spinal muscular atrophy (SMA).
Methods:
Two male children with complicated SMA diagnosed at the Children's Hospital Affiliated to Capital Institute of Pediatrics respectively in July 2021 and April 2022 due to delayed or retrograde motor development were selected as the study subjects. Clinical data of the children were collected. Primary culture of skin fibroblasts was carried out, and peripheral blood samples were collected from both children and their parents. Multiplex ligation-dependent probe amplification, combined long-range PCR and nested PCR, and Sanger sequencing were carried out to detect the copy number and variants of the SMN1 gene. Absolute quantitative real-time PCR, Western blotting and immunofluorescence were used to determine the transcriptional level of the SMN gene, expression of the SMN protein, and the number of functional SMN protein complexes (gems body), respectively. This study was approved by the Children's Hospital Affiliated to Capital Institute of Pediatrics (Ethics No. SHERLLM2021009).
Results:
Child 1, a 1-year-old boy, was clinically diagnosed with type 1 SMA. Child 2, a 2-and-a-half-year-old boy, was clinically diagnosed with type 3 SMA. Both children were found to harbor a paternally derived SMN1 deletion and a maternally derived SMN1 gene variant, namely c.824G>T (p.Gly275Val) and c.884A>T (p.*295Leu). Compared with the normal controls and carriers, the levels of full-length SMN1 transcripts in their peripheral blood and skin fibroblast cell lines were significantly decreased (P < 0.05), and the levels of SMN protein normalized to that of β-actin, and the numbers of gems bodies in the primary fibroblast cells were also significantly lower (P < 0.05). Based on the guidelines from the American College of Medical Genetics and Genomics, both variants were classified as likely pathogenic (PS3+PM3+PM5+PP3; PS3+PM3+PM4+PP3). Following the diagnosis, both children had received nusinersen treatment. Although their motor function was improved, child 1 still died at the age of 2 due to severe pulmonary infection. The walking ability of child 2 was significantly improved, and his prognosis appeared to be good.
Conclusion:
Two cases of clinically complicated SMA have been confirmed by genetic testing and experimental studies, which has provided a reference for their accurate treatment.
More Related Videos
 08:59
Real-Time Fluorescent Measurement of Synaptic Functions in Models of Amyotrophic Lateral Sclerosis
Published on: July 16, 2021
 06:35
In Vivo Electrophysiological Measurement of Compound Muscle Action Potential from the Forelimbs in Mouse Models of Motor Neuron Degeneration
Published on: June 15, 2018
Related Concept Videos
 01:21
Satellite Stem Cells and Muscular Dystrophy
 01:28
Disorders of the Skeletal Muscle
Musculoskeletal disorders
Musculoskeletal disorders involve injuries and conditions affecting the skeletal muscles and associated connective tissues. These disorders can arise from acute biomechanical stresses or chronic overuse and can occur across different age groups. Common injuries include sprains, fractures, and muscular strains, often resulting from...
 01:43
Sex-linked Disorders
 01:22
Lysosomal Hydrolases