Related Experiment Video
Updated: Jun 11, 2025
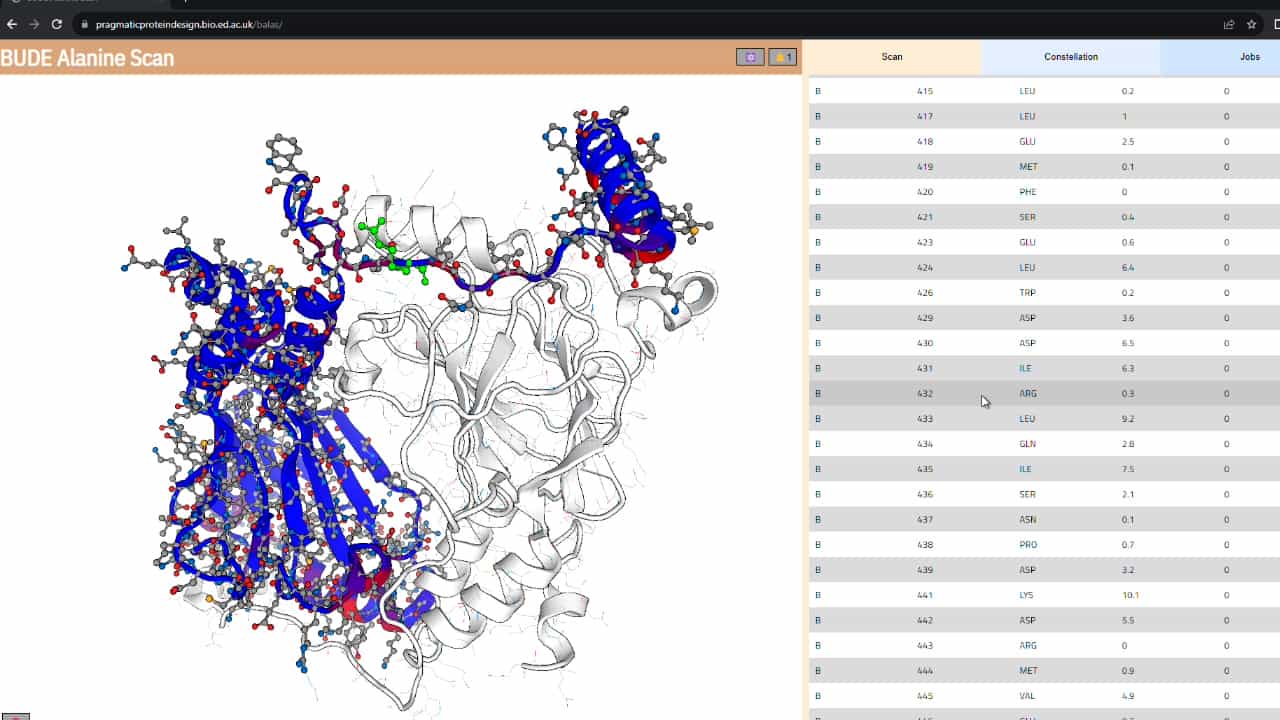
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Structure-based prediction of protein-nucleic acid binding using graph neural networks
Jared M Sagendorf1,2, Raktim Mitra1, Jiawei Huang1
1Department of Quantitative and Computational Biology, University of Southern California, Los Angeles, CA 90089 USA.

Abstract:
Protein-nucleic acid (PNA) binding plays critical roles in the transcription, translation, regulation, and three-dimensional organization of the genome. Structural models of proteins bound to nucleic acids (NA) provide insights into the chemical, electrostatic, and geometric properties of the protein structure that give rise to NA binding but are scarce relative to models of unbound proteins. We developed a deep learning approach for predicting PNA binding given the unbound structure of a protein that we call PNAbind. Our method utilizes graph neural networks to encode the spatial distribution of physicochemical and geometric properties of protein structures that are predictive of NA binding. Using global physicochemical encodings, our models predict the overall binding function of a protein, and using local encodings, they predict the location of individual NA binding residues. Our models can discriminate between specificity for DNA or RNA binding, and we show that predictions made on computationally derived protein structures can be used to gain mechanistic understanding of chemical and structural features that determine NA recognition. Binding site predictions were validated against benchmark datasets, achieving AUROC scores in the range of 0.92-0.95. We applied our models to the HIV-1 restriction factor APOBEC3G and showed that our model predictions are consistent with and help explain experimental RNA binding data.
Supplementary Information:
The online version contains supplementary material available at 10.1007/s12551-024-01201-w.
Related Concept Videos
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:04Protein-Protein Interfaces
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...