Related Experiment Video
Updated: Jun 10, 2025

Diagonal Method to Measure Synergy Among Any Number of Drugs
Published on: June 21, 2018
Quantitative Prediction of Drug-Drug Interactions Caused by CYP3A Induction Using Endogenous Biomarker
Hiroaki Takubo1, Toshio Taniguchi2, Yukihiro Nomura2
1DMPK Research Laboratories, Central Pharmaceutical Research Institute, Japan Tobacco Inc., Osaka, Japan hiroaki.takubo@jt.com.

Abstract:
Evaluation of the CYP3A induction risk is important in early drug development stages. This study focused on 4β-hydroxycholesterol (4β-HC) as an endogenous biomarker of drug-drug interactions (DDIs) caused by CYP3A induction. We investigated a new approach using 4β-HC for quantitative prediction of DDIs caused by CYP3A induction based on the mechanistic static pharmacokinetic (MSPK) model. The induction ratio, i.e., the ratio of plasma 4β-HC or 4β-HC/cholesterol (4β-HC/C) with and without a coadministered CYP3A inducer, and the ratio of the area under the plasma concentration-time curve (AUCR), i.e., the ratio of the AUC of plasma CYP3A substrate drugs with and without a coadministered CYP3A inducer, were collected. The scaling factor (d) in the MSPK model was calculated from the induction ratio of 4β-HC or 4β-HC/C based on the systemic term in the MSPK model. The AUCR of 18 CYP3A substrates with and without coadministration of seven CYP3A inducers were then predicted by substituting the calculated d value into the MSPK model. This approach showed that approximately 84% of the predicted AUCR values were within a twofold range of the observed values, showing that this approach can be a good tool to quantitatively predict DDIs caused by CYP3A induction. SIGNIFICANCE STATEMENT: A concise approach to predict drug interactions with adequate accuracy is preferable in the early drug development stage. In this study, a new approach using 4β-hydroxycholesterol for quantitative prediction of drug-drug interactions caused by CYP3A induction was investigated. The predictability was verified using seven CYP3A inducers and 18 substrates.
More Related Videos
 10:44
Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
 15:04
Potentiation of Anticancer Antibody Efficacy by Antineoplastic Drugs: Detection of Antibody-drug Synergism Using the Combination Index Equation
Published on: January 19, 2019
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:23
Drug Concentrations: Measurements
Plasma...
 01:14
Time Course of Drug Effect
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...
 01:09
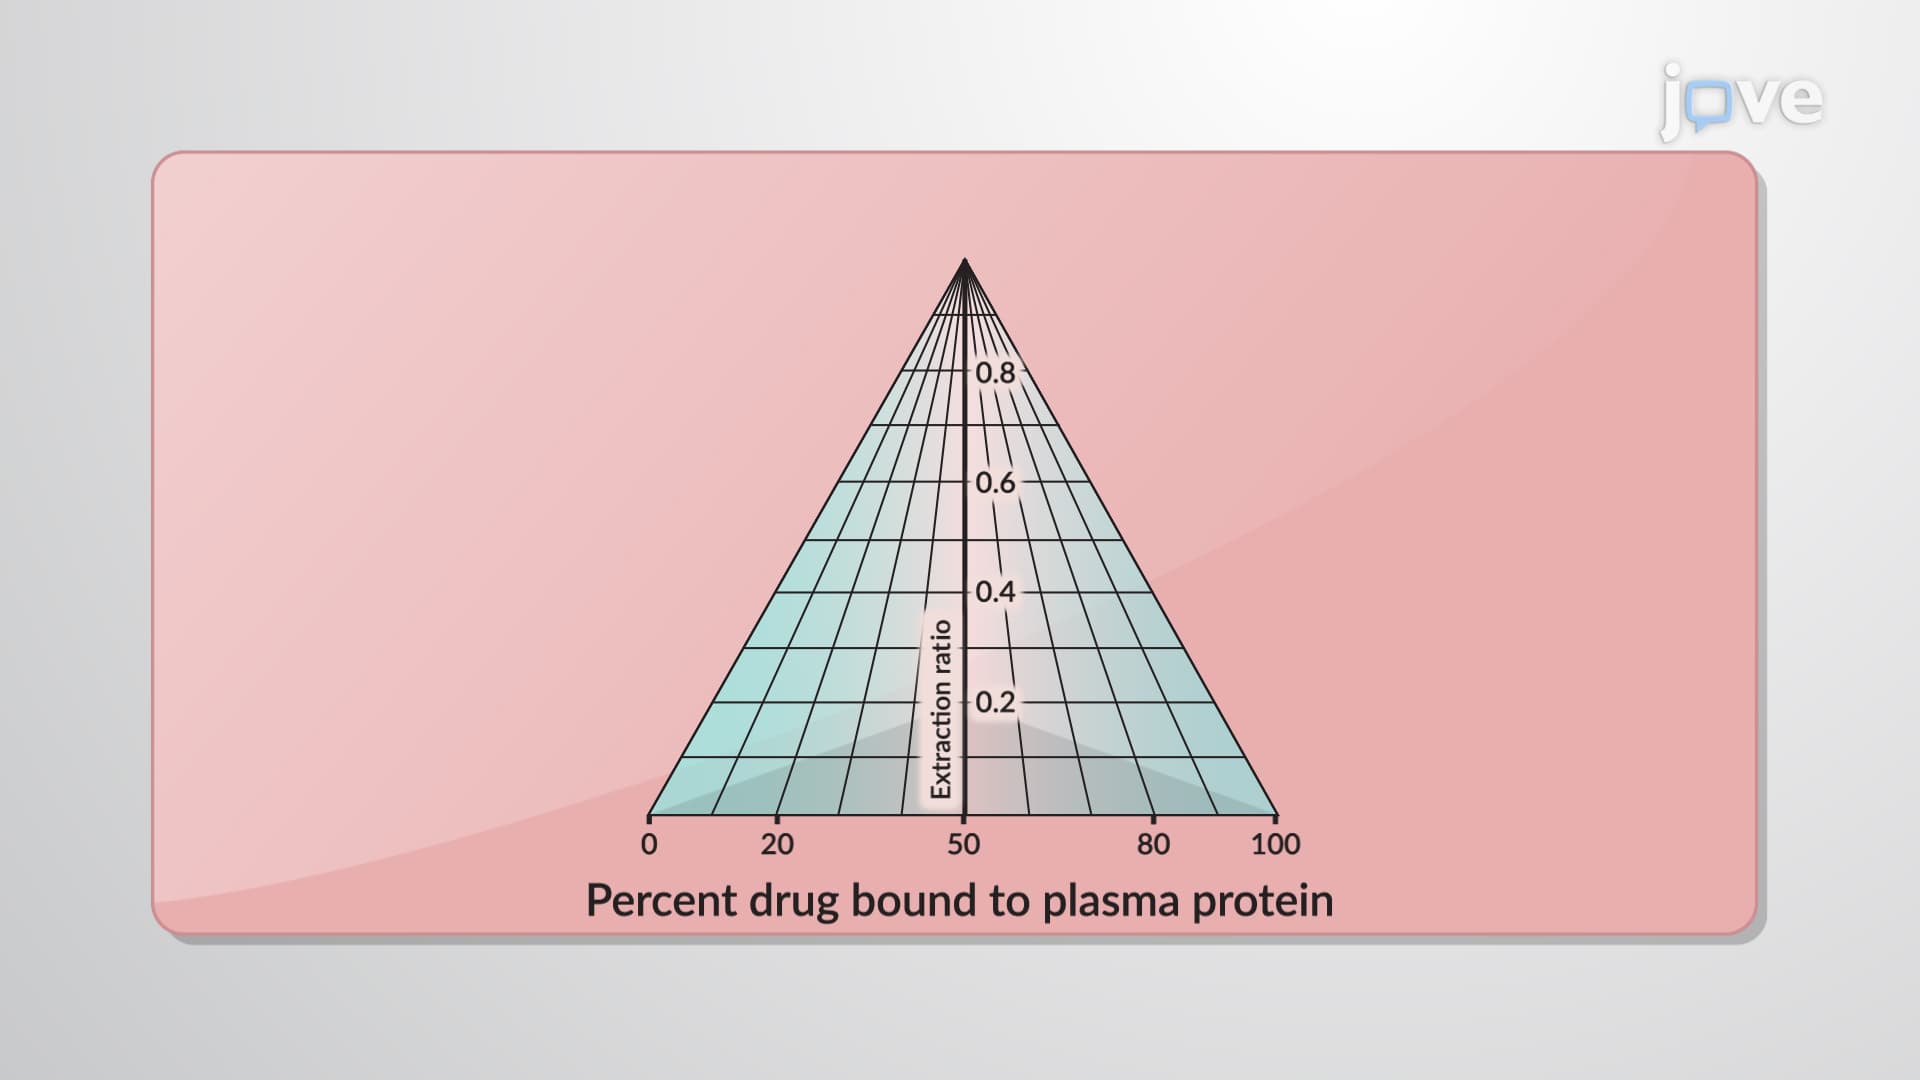
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...