Related Experiment Video
Updated: Jun 10, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Evaluating the Chemical Reactivity of DFT-Simulated Liquid Water with Hydrated Electrons via the Dual Descriptor
William R Borrelli1, Xiaoyan Liu1, Benjamin J Schwartz1
1Department of Chemistry and Biochemistry, University of California, Los Angeles, Los Angeles, California 90095-1569, United States.

Abstract:
Modeling the various properties of liquid water, particularly its reactivity, has been a longstanding challenge for simulation methods. Recently, ab initio simulations based on density functional theory (DFT) have come to the fore as tenable methods for calculating the properties and reactivity of water, with varying degrees of success for different exchange-correlation functionals. In particular, hybrid-GGA and meta-GGA functionals have been shown to reproduce many of the structural, dynamical, and energetic properties of water to a high degree of accuracy relative to their computational cost. Here, we show that the dual descriptor (DD) measure of nucleophilicity and electrophilicity, which is sometimes used to elucidate organic chemistry reaction mechanisms, can also be used to characterize the reactivity of DFT-simulated liquid water. The DD is especially apt for understanding the reactivity of excess electrons with water as its calculation explicitly involves adding and removing an excess electron from a reference system. We use the DD to explore the reactivity of water simulated using three different DFT functionals: the LDA functional (LDA), a hybrid-GGA functional (PBE0), and a hybrid meta-GGA functional (SCAN0). Using the DD, we show that the SCAN0 functional with the standard 25% Hartree-Fock exchange produces simulated liquid water with many regions that are far more reactive than either PBE0 or LDA. To understand the implications of these highly reactive regions, we then add a strong nucleophile in the form of an excess electron and find that although PBE0 and LDA predict stable hydrated electrons, the excess electron reacts nearly instantaneously with SCAN0 water via proton abstraction to form a hydrogen atom and hydroxide ion. We show that the DD provides the ability to not only predict whether or not liquid water will react with a hydrated electron but also which particular waters will be involved solely from analyzing pure water configurations generated with each functional. We rationalize this result in terms of the known trap-seeking behavior of injected hydrated electrons, which are able to find the most electronegative region in bulk water. These results highlight the utility of the dual descriptor as a fast and interpretable method for investigating condensed-phase reactivity with excess electrons.
More Related Videos
 10:28
Probing the Structure and Dynamics of Interfacial Water with Scanning Tunneling Microscopy and Spectroscopy
Published on: May 27, 2018
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:42
Aqueous Solutions and Heats of Hydration
When ionic compounds dissolve in water, the ions in the solid separate and disperse uniformly throughout the solution because water molecules surround and solvate the ions, reducing the strong electrostatic forces between them. This process...
 03:13
Intermolecular Forces
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 02:45
Acid-Catalyzed Hydration of Alkenes
 01:20
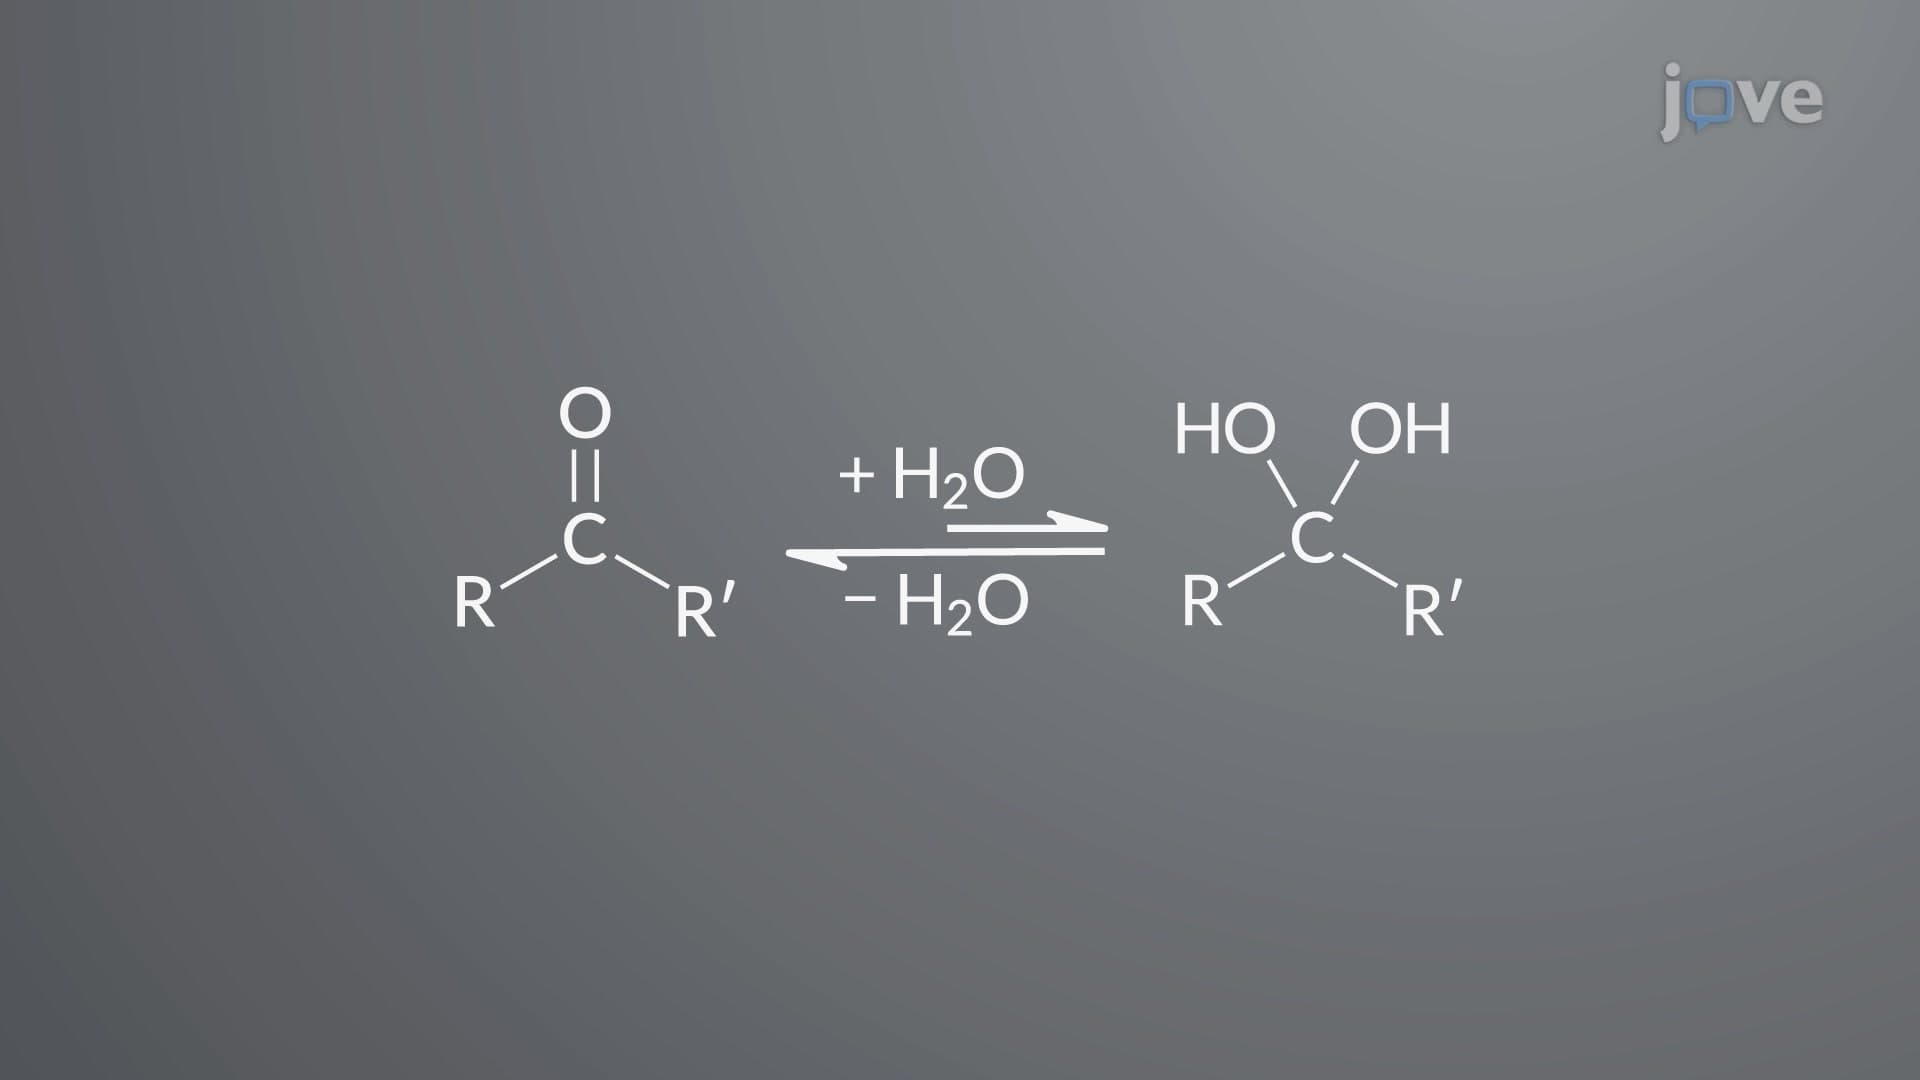
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 02:30
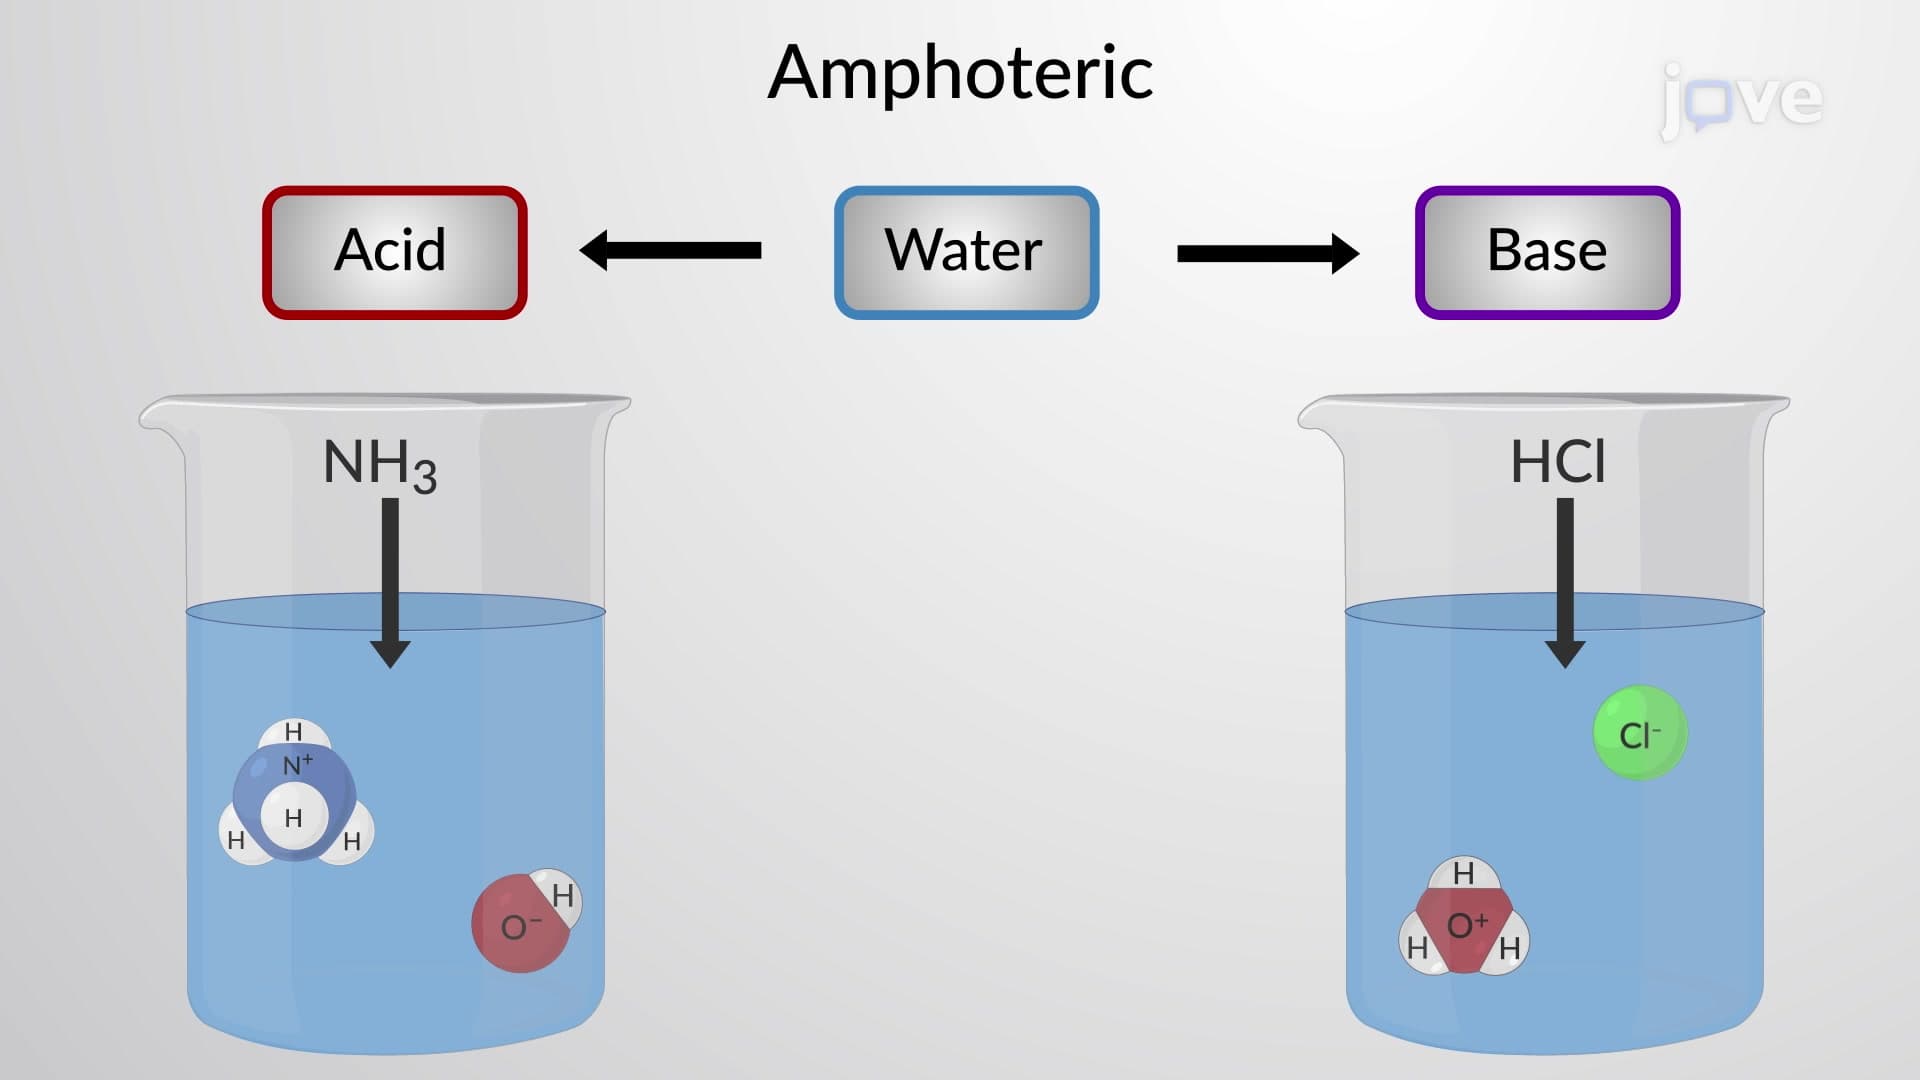
Water: A Bronsted-Lowry Acid and Base