Related Experiment Video
Updated: Jun 9, 2025

Synthesis and Characterization of an Aspirin-fumarate Prodrug that Inhibits NFκB Activity and Breast Cancer Stem Cells
Published on: January 18, 2017
Clinical Pharmacology of Asciminib: A Review
Matthias Hoch1, Felix Huth2, Paul William Manley2
1Novartis Biomedical Research, Fabrikstrasse 2, 4056, Basel, Switzerland. Matthias.hoch@novartis.com.

Asciminib is a first-in-class allosteric inhibitor of the kinase activity of BCR::ABL1, specifically targeting the ABL myristoyl pocket (STAMP). This review focuses on the pharmacokinetic (PK) and pharmacodynamic data of asciminib, which is approved at a total daily dose of 80 mg for the treatment of adult patients with chronic myeloid leukemia in chronic phase who are either resistant or intolerant to ≥ 2 tyrosine kinase inhibitors or those harboring the T315I mutation (at a dose of 200 mg twice daily). Asciminib is predicted to be almost completely absorbed from the gut, with an absolute bioavailability (F) of approximately 73%. It should be administered in a fasted state, as food (particularly high-fat meals) reduces exposure. Asciminib displays a slightly greater than dose-proportional increase in exposure, with no time-dependent changes in PK observed following repeated dosing. This drug shows low clearance (6.31 L/h), with a moderate volume of distribution (111 L) and high human plasma protein binding (97.3%). The apparent terminal elimination half-life (t1/2) across studies was estimated to be between 7 and 15 h. The PK of asciminib is not substantially affected by body weight, age, gender, race, or renal or hepatic impairment. Asciminib is primarily metabolized via CYP3A4-mediated oxidation (36.0%) and UGT2B7- and UGT2B17-mediated glucuronidation (13.3% and 7.8%, respectively); biliary secretion via breast cancer resistance protein contributes to about 31.1% to total systemic clearance, which is mainly through hepatic metabolism and biliary secretion through the fecal pathway, with renal excretion playing a minor role. The potential for PK drug interaction for asciminib both as a victim and a perpetrator has been summarized here based on clinical and predicted drug-drug interaction studies. Robust exposure-response models characterized asciminib exposure-efficacy and exposure-safety relationships. In patients without the T315I mutation, the exposure-efficacy analysis of the time course of BCR::ABL1IS percentages highlighted the existence of a slightly positive, albeit not clinically significant, relationship. Higher exposure was required for efficacy in patients harboring the T315I mutation compared with those who did not. The exposure-safety relationship analysis showed no apparent association between exposure and adverse events of interest over the broad range of exposure or dose levels investigated. Asciminib has also been shown to have no clinically relevant effect on cardiac repolarization. Here, we review the clinical pharmacology data available to date for asciminib that supported its clinical development program and regulatory applications.
Asciminib is a first-in-class allosteric inhibitor of the kinase activity of BCR::ABL1, specifically targeting the ABL myristoyl pocket (STAMP). This review focuses on the pharmacokinetic (PK) and pharmacodynamic data of asciminib, which is approved at a total daily dose of 80 mg for the treatment of adult patients with chronic myeloid leukemia in chronic phase who are either resistant or intolerant to ≥ 2 tyrosine kinase inhibitors or those harboring the T315I mutation (at a dose of 200 mg twice daily). Asciminib is predicted to be almost completely absorbed from the gut, with an absolute bioavailability (F) of approximately 73%. It should be administered in a fasted state, as food (particularly high-fat meals) reduces exposure. Asciminib displays a slightly greater than dose-proportional increase in exposure, with no time-dependent changes in PK observed following repeated dosing. This drug shows low clearance (6.31 L/h), with a moderate volume of distribution (111 L) and high human plasma protein binding (97.3%). The apparent terminal elimination half-life (t1/2) across studies was estimated to be between 7 and 15 h. The PK of asciminib is not substantially affected by body weight, age, gender, race, or renal or hepatic impairment. Asciminib is primarily metabolized via CYP3A4-mediated oxidation (36.0%) and UGT2B7- and UGT2B17-mediated glucuronidation (13.3% and 7.8%, respectively); biliary secretion via breast cancer resistance protein contributes to about 31.1% to total systemic clearance, which is mainly through hepatic metabolism and biliary secretion through the fecal pathway, with renal excretion playing a minor role. The potential for PK drug interaction for asciminib both as a victim and a perpetrator has been summarized here based on clinical and predicted drug-drug interaction studies. Robust exposure-response models characterized asciminib exposure-efficacy and exposure-safety relationships. In patients without the T315I mutation, the exposure-efficacy analysis of the time course of BCR::ABL1IS percentages highlighted the existence of a slightly positive, albeit not clinically significant, relationship. Higher exposure was required for efficacy in patients harboring the T315I mutation compared with those who did not. The exposure-safety relationship analysis showed no apparent association between exposure and adverse events of interest over the broad range of exposure or dose levels investigated. Asciminib has also been shown to have no clinically relevant effect on cardiac repolarization. Here, we review the clinical pharmacology data available to date for asciminib that supported its clinical development program and regulatory applications.
More Related Videos
 11:44
Cellular Membrane Affinity Chromatography Columns to Identify Specialized Plant Metabolites Interacting with Immobilized Tropomyosin Kinase Receptor B
Published on: January 19, 2022
 09:39
Drug-induced Sensitization of Adenylyl Cyclase: Assay Streamlining and Miniaturization for Small Molecule and siRNA Screening Applications
Published on: January 27, 2014
Related Concept Videos
 01:24
Cholinergic Antagonists: Pharmacokinetics
 01:31
Direct-Acting Cholinergic Agonists: Pharmacokinetics
 01:24
Antiarrhythmic Drugs: Class II Agents as β-Adrenergic Blockers
 01:17
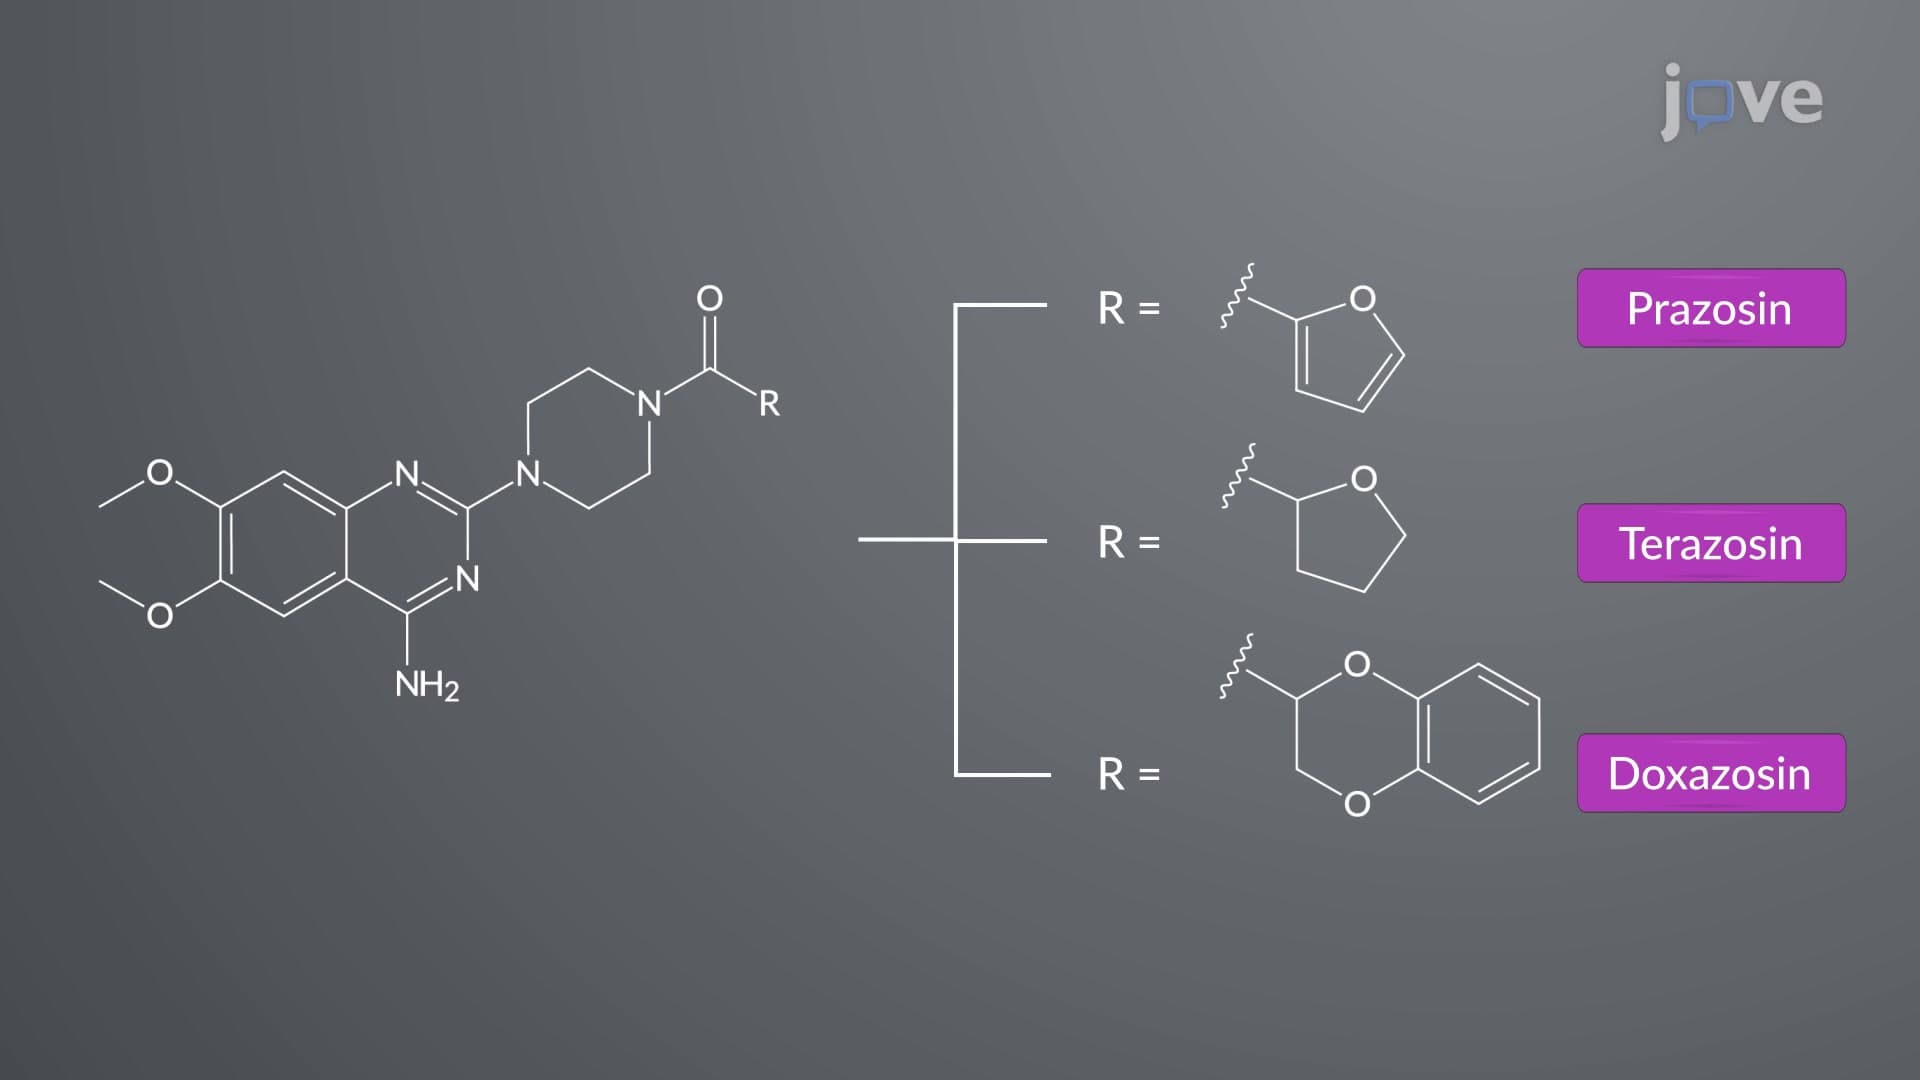
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
 01:26
Heart Failure Drugs: Inhibitors of Renin-Angiotensin System
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...