

A Multipole-Based Reactive Force Field for Hydrocarbons
Junben Weng1,2, Hongqiang Cui1,2, Da Zheng1,2
1Laboratory of Molecular Modeling and Design, State Key Laboratory of Molecular Reaction Dynamics, Dalian Institute of Chemical Physics, Chinese Academy of Sciences, Dalian 116023, China.

Abstract:
The computational complexity of quantum chemistry methods has prompted the development of reactive force fields, facilitating practical applications of molecular dynamics simulations for large-scale reactive systems. Current reactive force fields typically employ intricate corrections based on prior chemical knowledge, which severely impedes their further advancement. This study presents a new atomic multipole-based reactive model with bond free (OPERATOR). The force field is constructed on a simple, physically motivated model within the AMOEBA framework that closely resembles the physical representation of the chemical reaction processes. In the force field, the atomic multipoles are generated dynamically according to the atomic environments, aiming to effectively capture significant changes in the electrostatic environments during chemical reactions. Subsequently, atomic multipole-based charge penetration, polarization, and charge transfer effects are incorporated into the force field to describe the complex electrostatic interactions in the system. The force field also includes van der Waals interactions and three-body potentials. In addition, to extend these nonreactive interactions to chemical reactions, the atom distribution multipole moments are used to characterize different chemical environments. The force field has been optimized using the dataset of potential energy surfaces (PESs) of hydrocarbons derived from DFT results of millions of conformations with six degrees of freedom (DOFs). The results demonstrate that the new force field effectively replicates both the monopoles and the energies. In comparison to ReaxFF, the new force field exhibits comparable or superior performance. Furthermore, molecular dynamics simulations of n-heptane decomposition effectively reproduce the primary products and reactions observed in the experiments. Given the simplicity and physically motivated nature of the model, it is expected that the new force field will be utilized in future studies to investigate chemical reaction mechanisms involving more elements.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 05:57
Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
Related Concept Videos
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:14
Combustion Energy: A Measure of Stability in Alkanes and Cycloalkanes
Alkanes undergo combustion in the presence of excess oxygen and high-temperature conditions to give carbon dioxide and water. A combustion reaction is the energy source in natural gas, liquified...
 02:33
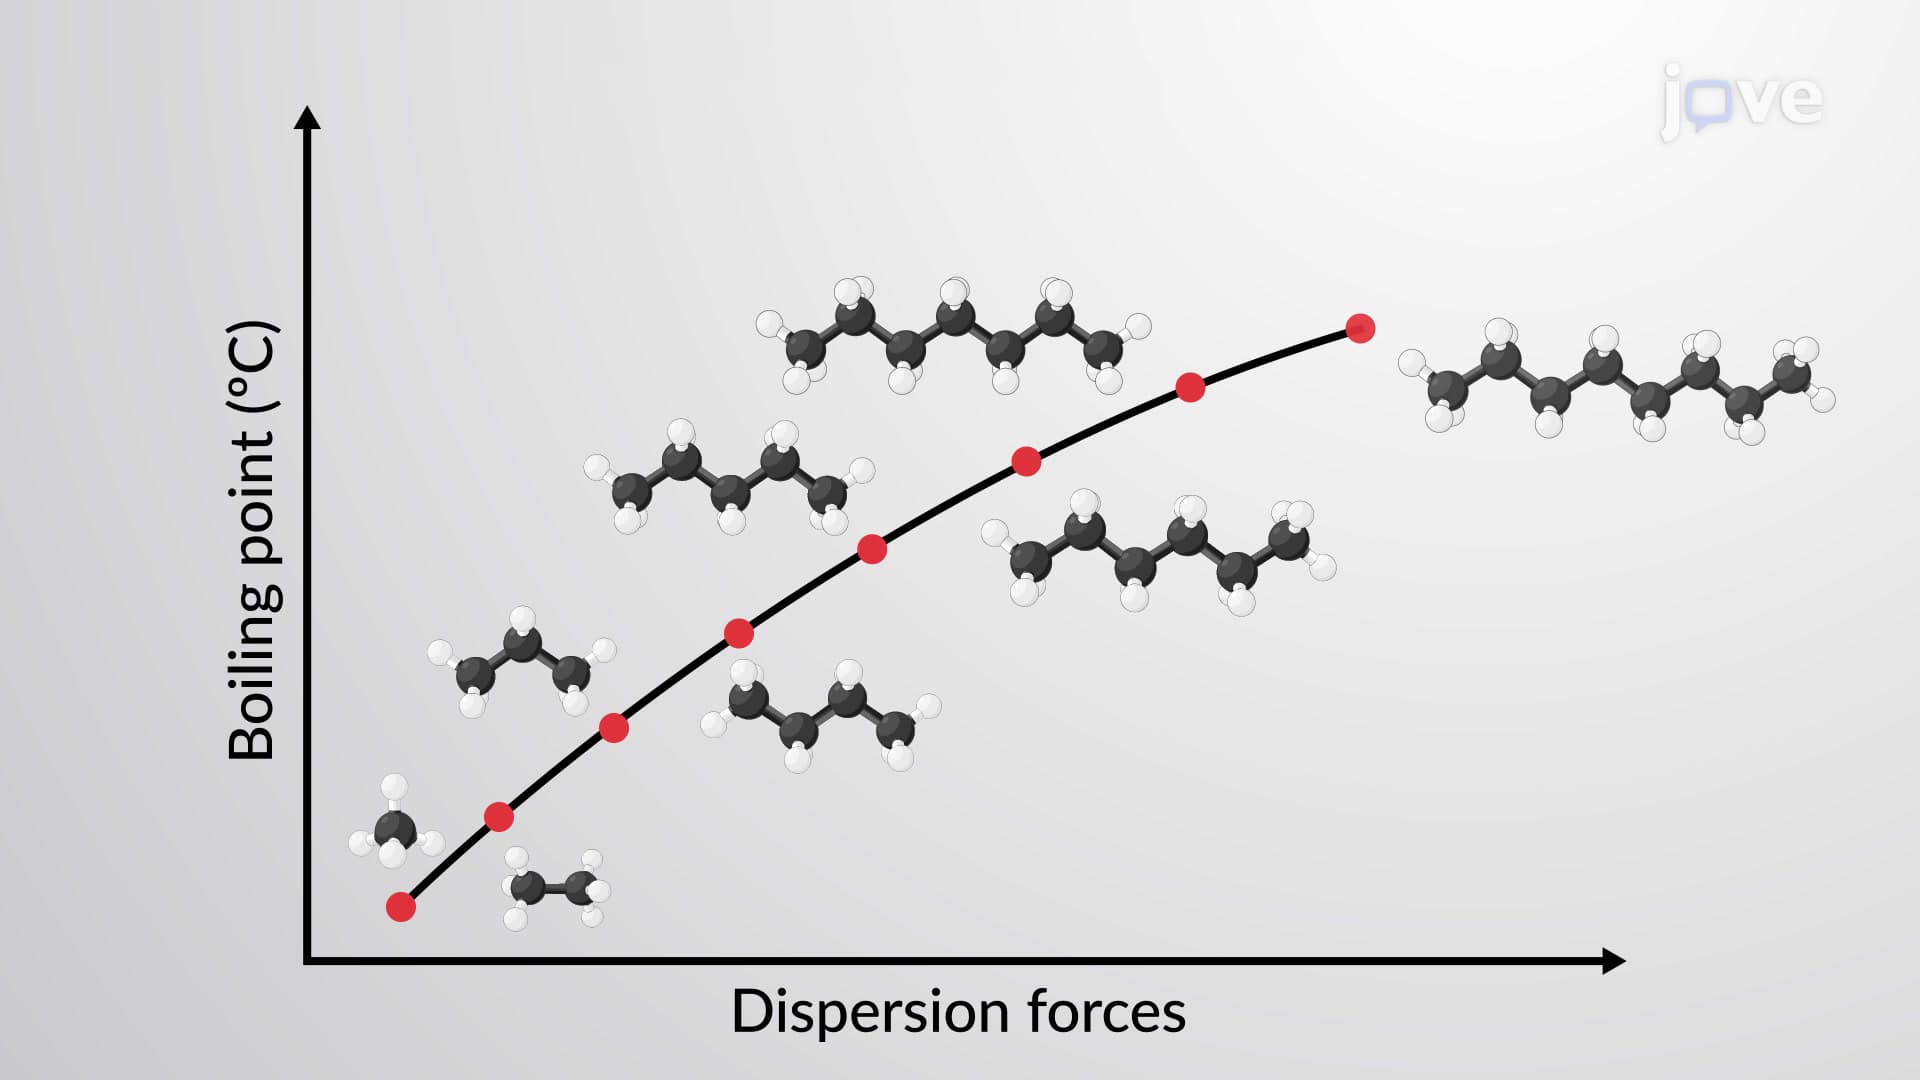
Physical Properties of Alkanes
 01:13
Force and Potential Energy in One Dimension
 01:27
π Electron Effects on Chemical Shift: Overview
 01:20
Two-Dimensional Force System