Related Experiment Video
Updated: Jun 7, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Non-adiabatic Couplings in Surface Hopping with Tight Binding Density Functional Theory: The Case of Molecular Motors
Gonzalo Díaz Mirón1, Carlos R Lien-Medrano2, Debarshi Banerjee1,3
1Condensed Matter and Statistical Physics, The Abdus Salam International Centre for Theoretical Physics, 34151 Trieste, Italy.

Abstract:
Nonadiabatic molecular dynamics (NAMD) has become an essential computational technique for studying the photophysical relaxation of molecular systems after light absorption. These phenomena require approximations that go beyond the Born-Oppenheimer approximation, and the accuracy of the results heavily depends on the electronic structure theory employed. Sophisticated electronic methods, however, make these techniques computationally expensive, even for medium size systems. Consequently, simulations are often performed on simplified models to interpret the experimental results. In this context, a variety of techniques have been developed to perform NAMD using approximate methods, particularly density functional tight binding (DFTB). Despite the use of these techniques on large systems, where ab initio methods are computationally prohibitive, a comprehensive validation has been lacking. In this work, we present a new implementation of trajectory surface hopping combined with DFTB, utilizing nonadiabatic coupling vectors. We selected the methaniminium cation and furan systems for validation, providing an exhaustive comparison with the higher-level electronic structure methods. As a case study, we simulated a system from the class of molecular motors, which has been extensively studied experimentally but remains challenging to simulate with ab initio methods due to its inherent complexity. Our approach effectively captures the key photophysical mechanism of dihedral rotation after the absorption of light. Additionally, we successfully reproduced the transition from the bright to dark states observed in the time-dependent fluorescence experiments, providing valuable insights into this critical part of the photophysical behavior in molecular motors.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 01:22
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...
 01:10
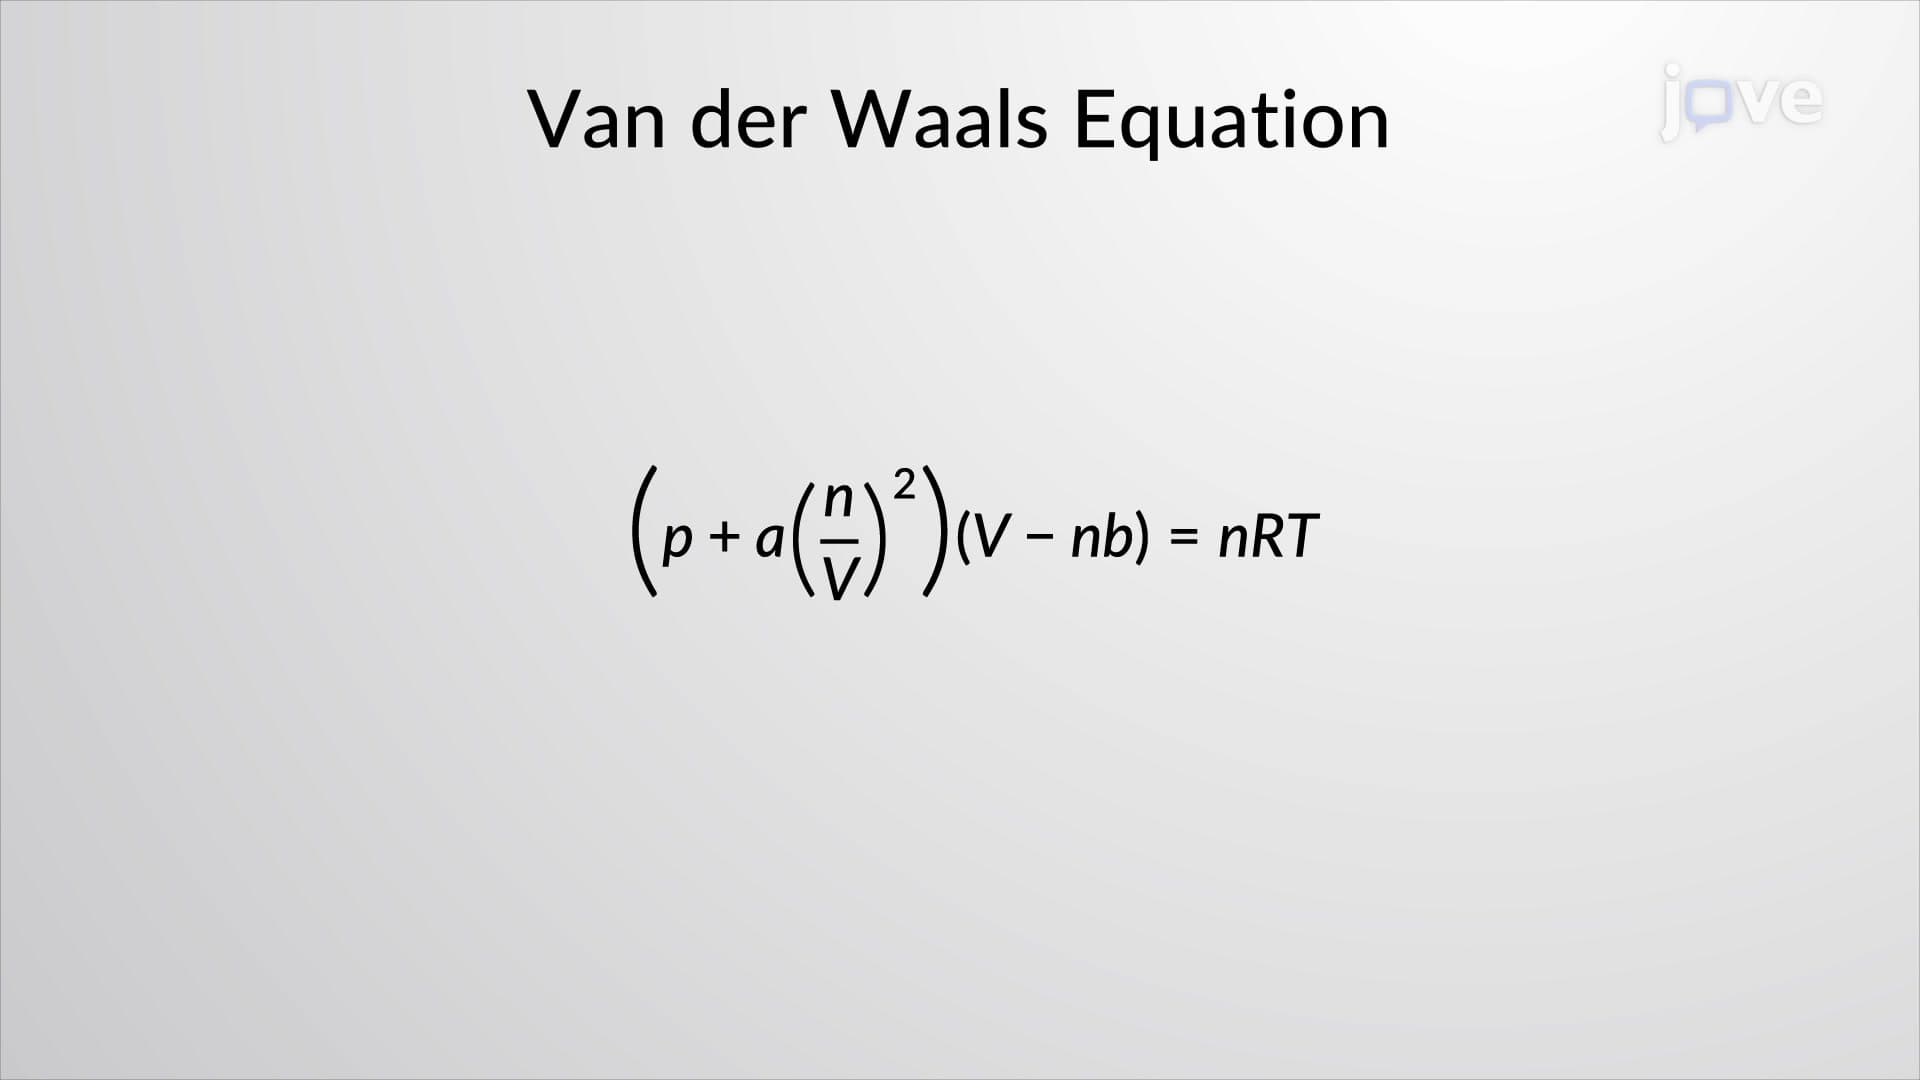
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:35
Hybridization of Atomic Orbitals II