

Anisotropic Dielectric Screened Range-Separated Hybrid Density Functional Theory Calculations of Charge Transfer
Chandrima Chakravarty1, Maximilian A C Saller1, Hüseyin Aksu2
1Department of Chemistry and Biochemistry, Kent State University, Kent, Ohio 44242, United States.

Abstract:
A density functional theory framework is developed to study electronic excited states affected by an anisotropic dielectric environment. In particular, an anisotropic dielectric screened range-separated hybrid (SRSH[r]) functional is defined and combined with an anisotropic polarizable continuum model (PCM) implemented through a generalized Poisson equation solver. We develop the SRSH-PCM(r) approach and use it to quantify the effect of anisotropy on an excited charge transfer (CT) state energy. In particular, the dielectric interface effect on the CT state within a donor-acceptor molecular complex of antrancene and tetracyanoquinodimethane is studied. The donor-acceptor complex and the dielectric interface are used to represent the interface between thin films consisting of these materials. We report the effect of such a dielectric interface on the energy of a CT and follow its dependence on the donor-acceptor distance. We also benchmark the anisotropy-affected energy by comparing to homogeneous dielectric calculated energies. Due to the planar interface, the anisotropic energies are expected to to match with those obtained based on isotropic calculations of the larger dielectric constant at large enough distances. The approach is applicable, in general, to more complicated dielectric constant distributions as expected to be found in actual interfaces of such thin films or in other systems, for example, for CT processes within photosystems.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:18
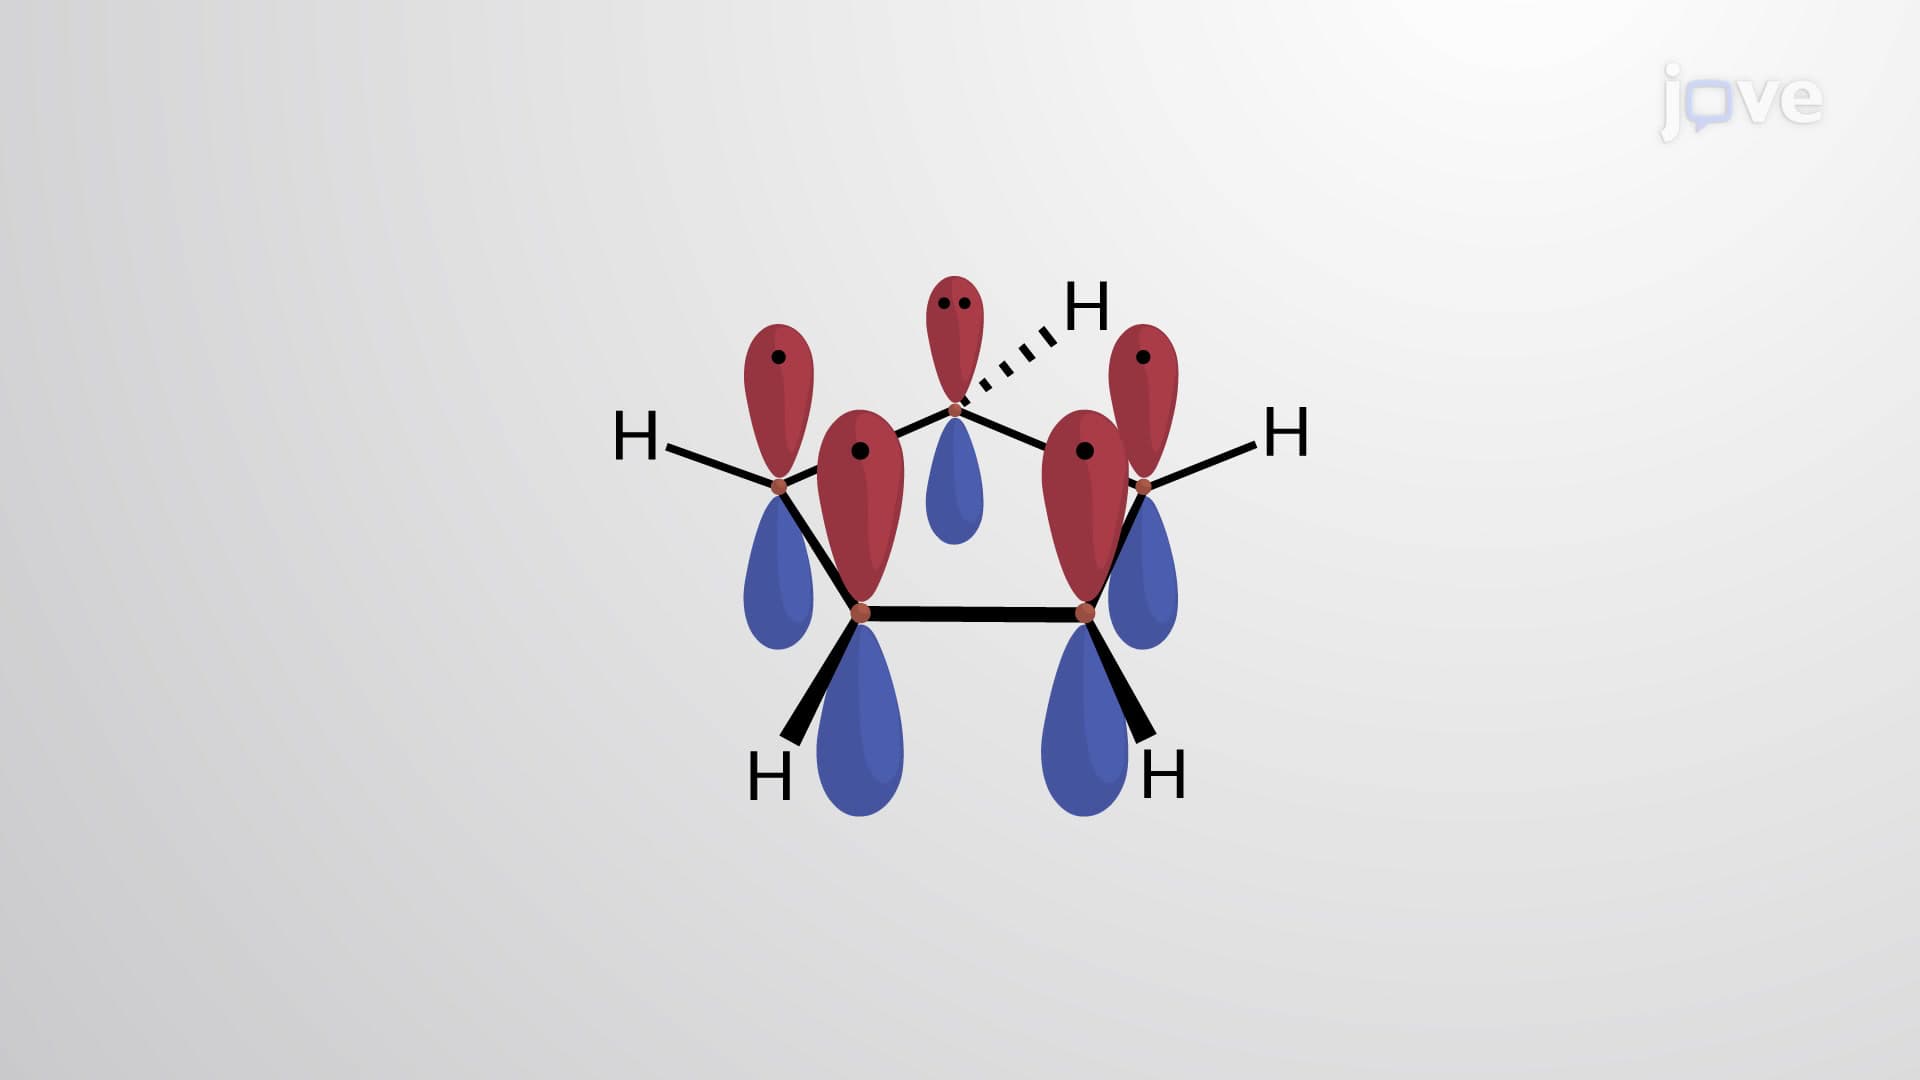
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...