Related Experiment Video
Updated: Jun 5, 2025

Targeting Cysteine Thiols for in Vitro Site-specific Glycosylation of Recombinant Proteins
Published on: October 4, 2017
A novel mutation, Ile344Asn, in microsomal triglyceride transfer protein abolishes binding to protein disulfide
Swati Valmiki1, Cindy Bredefeld2, M Mahmood Hussain1
1Department of Foundations of Medicine, NYU Grossman Long Island School of Medicine, Mineola, NY, USA.

Abstract:
Microsomal triglyceride transfer protein (MTP) plays crucial roles in the assembly and secretion of apolipoprotein B-containing lipoproteins and loss of function MTP variants are associated with abetalipoproteinemia, a disease characterized by the absence of these lipoproteins. MTP is a heterodimeric protein of two subunits, MTP and protein disulfide isomerase (PDI). In this study, we report a proband with abetalipoproteinemia who was monitored annually for 10 years in her third decade and had very low plasma lipids and undetectable apoB-containing lipoproteins. Genetic testing revealed biallelic variants in the MTTP gene. She has a well-documented nonsense mutation Gly865∗ that does not interact with the PDI subunit. She also has a novel missense MTP mutation, Ile344Asn. We show that this mutation abrogates lipid transfer activity in MTP and does not support apolipoprotein B secretion. This residue is present in the central α-helical domain of MTP and the substitution of Ile with Asn at this position disrupts interactions between MTP and PDI subunits. Ile344 is away from the known MTP:PDI interacting sites identified in the crystal structure of MTP suggesting that MTP:PDI interactions are more dynamic than previously envisioned. Identification of more missense mutations will enhance our understanding of the structure-function of MTP and the role of critical residues in these interactions between the two subunits. This knowledge may guide us in developing novel treatment modalities to reduce plasma lipids and atherosclerosis.
Insights
Microsomal triglyceride transfer protein (MTP) mutations cause abetalipoproteinemia. A novel MTP mutation, Ile344Asn, disrupts MTP:PDI interactions, impairing lipid transfer and apolipoprotein B secretion.
Area of Science:
- Lipid metabolism and lipoprotein assembly
- Genetic disorders of lipid transport
- Protein structure-function relationships
Background:
- Microsomal triglyceride transfer protein (MTP) is essential for apolipoprotein B-lipoprotein assembly.
- MTP variants causing loss of function lead to abetalipoproteinemia, a condition with absent apoB-lipoproteins.
- MTP functions as a heterodimer with protein disulfide isomerase (PDI).
Purpose of the Study:
- To investigate the molecular basis of abetalipoproteinemia in a proband with biallelic MTTP variants.
- To characterize a novel MTP missense mutation (Ile344Asn) and its impact on MTP function and MTP:PDI interactions.
- To explore the structure-function relationship of MTP and its interactions with PDI.
Main Methods:
- Clinical monitoring and genetic testing of an abetalipoproteinemia proband.
- In vitro assessment of lipid transfer activity and apolipoprotein B secretion for the novel MTP mutant.
- Analysis of MTP:PDI subunit interactions in relation to the Ile344Asn mutation.
Main Results:
- The proband presented with very low plasma lipids and undetectable apoB-lipoproteins.
- Genetic analysis revealed a known nonsense mutation (Gly865∗) and a novel missense mutation (Ile344Asn) in the MTTP gene.
- The Ile344Asn mutation abrogated MTP lipid transfer activity and apolipoprotein B secretion, indicating a loss of function.
- This mutation disrupts MTP:PDI subunit interactions, suggesting MTP:PDI interactions are more complex than previously understood.
Conclusions:
- The novel MTP Ile344Asn missense mutation is pathogenic, causing abetalipoproteinemia by impairing MTP function and MTP:PDI interactions.
- This finding expands our understanding of MTP structure-function and the dynamics of MTP:PDI interactions.
- Further characterization of MTP mutations may inform the development of novel therapeutic strategies for hyperlipidemia and atherosclerosis.
Related Concept Videos
 01:26
Protein Modifications in the RER
Broadly, these modifications can be categorized into four main categories — glycosylation, formation of disulfide bonds, assembly of protein subunits, and specific proteolytic cleavages like removal of signal...
 01:45
Tail-anchoring of Proteins in the ER Membrane
 01:32
Export of Misfolded Proteins out of the ER
 01:19
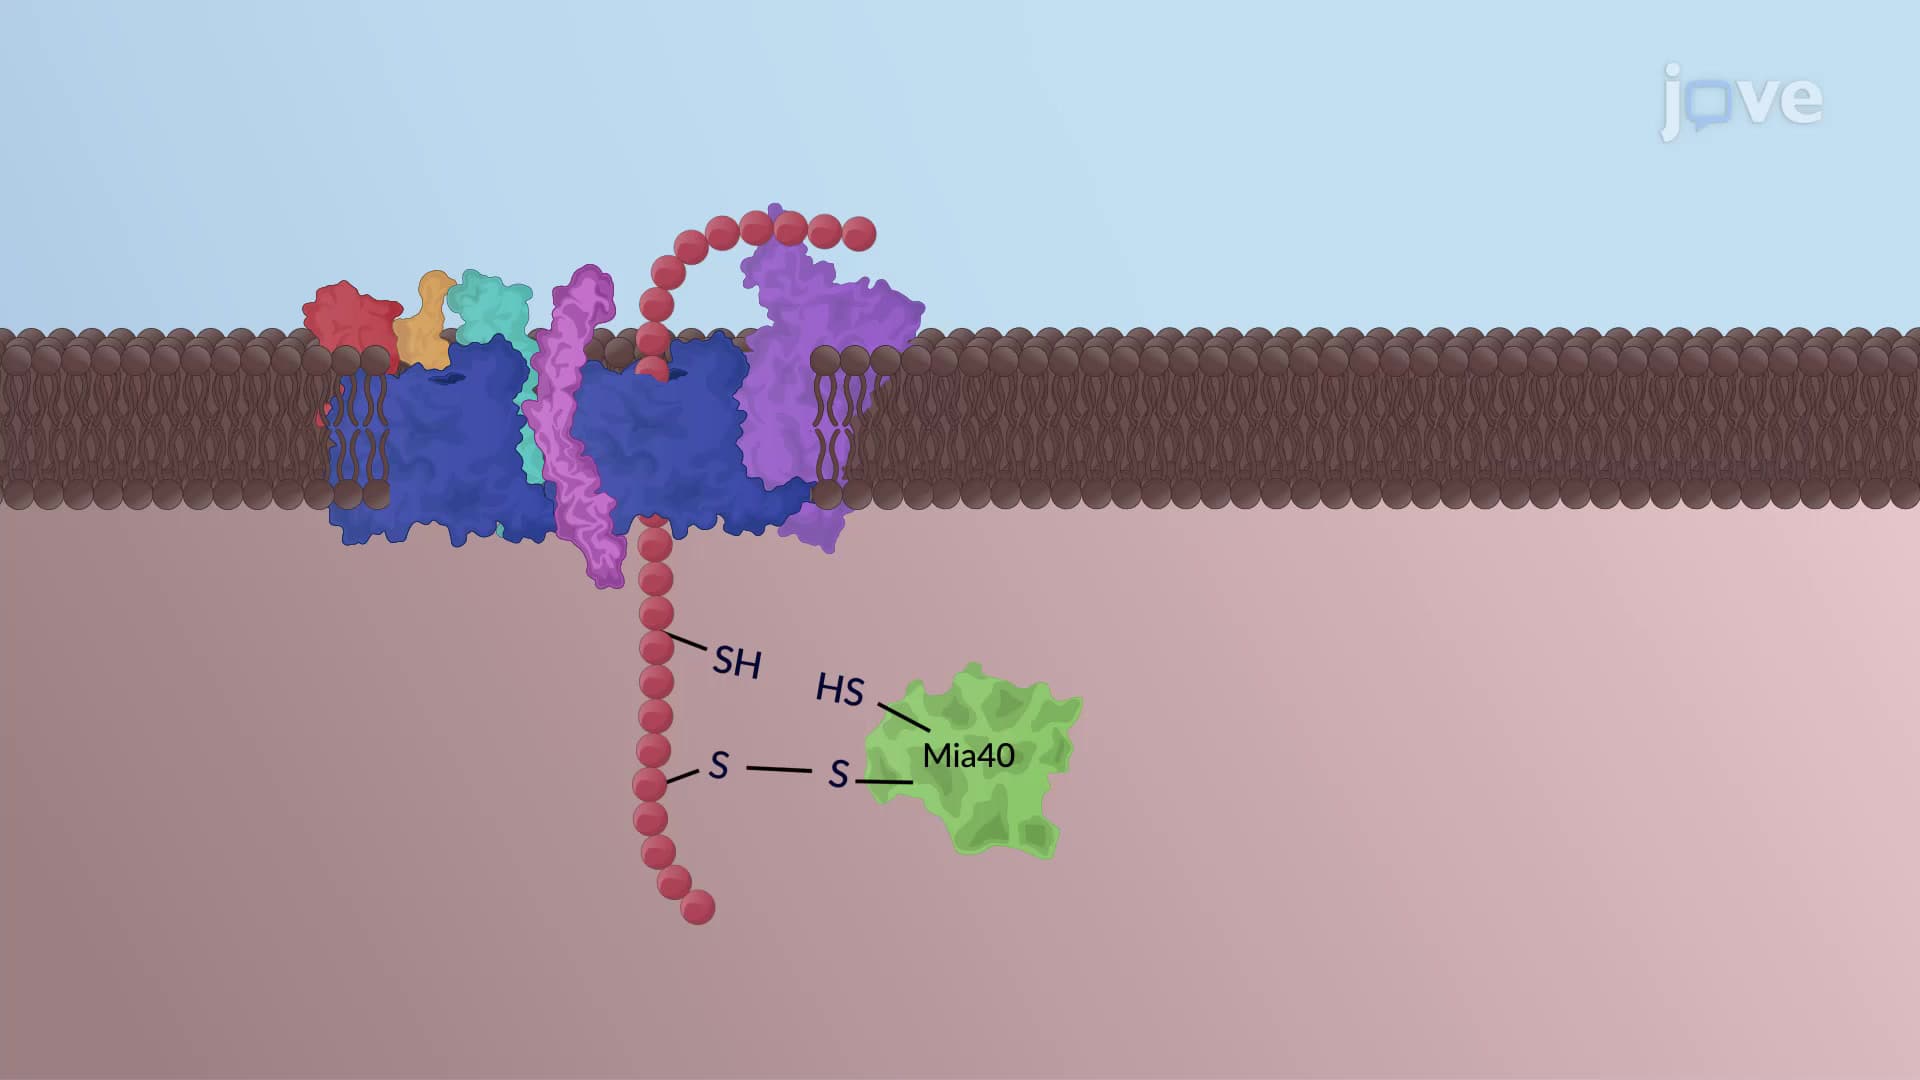
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
 01:19
Membrane Asymmetry Regulating Transporters
Flippase
Eukaryotic flippases are type-IV P-type ATPases or P4-ATPases belonging to P-type ATPase family proteins that are membrane-bound pumps involved in the ATP-mediated transport of ions and molecules across the membrane. Flippases flip specific phospholipids from the outer to the inner leaflet of a membrane. All P4-ATPases have one...