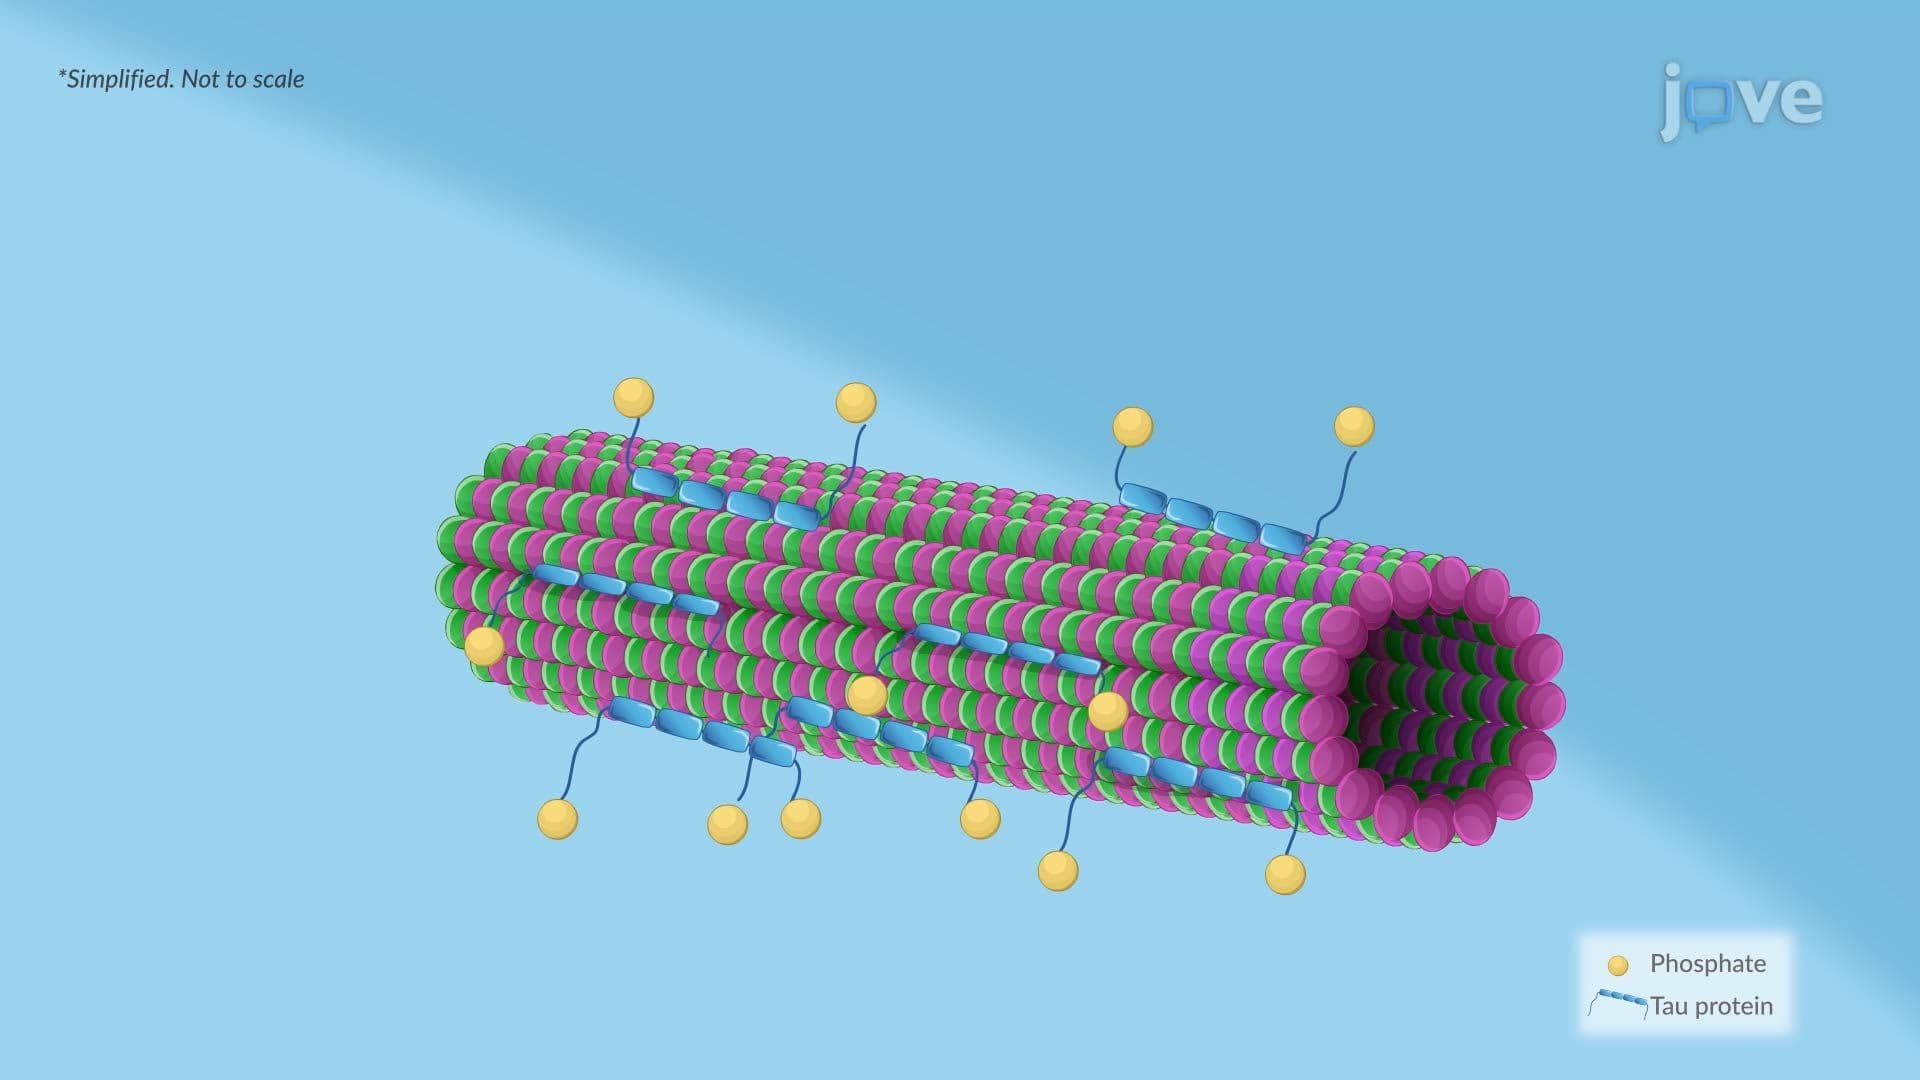
Related Experiment Video
Updated: Jun 19, 2026

Experimental Demyelination and Remyelination of Murine Spinal Cord by Focal Injection of Lysolecithin
Published on: March 26, 2015
Demyelination-derived lysophosphatidylserine promotes microglial dysfunction and neuropathology in a mouse model of
Yubo Zhou1,2, Zonghui Huang2,3, Bolong Lin2
1Department of Geriatrics, Gerontology Institute of Anhui Province, The First Affiliated Hospital of University of Science and Technology of China, Division of Life Sciences and Medicine, University of Science and Technology of China, Hefei, 230001, China.

Abstract:
Microglia dysfunction-associated neuroinflammation is an important driver of Alzheimer's disease (AD), but the mechanism is poorly understood. Here, we show that demyelination promotes neuroinflammation and cognitive impairment via the lysophosphatidylserine (LysoPS)-GPR34 axis in AD. Demyelination is observed at the early stage and is accompanied by an increase in LysoPS in myelin debris in a 5xFAD mouse model of AD. Reducing the content of LysoPS in myelin or inhibiting its receptor GPR34 via genetic or pharmacological approaches can reduce microglial dysfunction and neuroinflammation and improve microglial Aβ phagocytosis, subsequently resulting in less Aβ deposition and memory restoration in 5xFAD mice. Furthermore, increased LysoPS production and microglial GPR34 expression were also observed in the brains of AD patients. These results reveal the pathogenic role of demyelination-derived LysoPS in microglial dysfunction and AD pathology and suggest that blocking GPR34 as a therapeutic strategy beyond targeting Aβ.
Insights
Demyelination in Alzheimer's disease (AD) drives neuroinflammation and cognitive decline through the lysophosphatidylserine (LysoPS)-GPR34 pathway. Blocking GPR34 may offer a novel therapeutic approach for AD.
Area of Science:
- Neuroscience
- Immunology
- Pathology
Background:
- Microglia dysfunction and neuroinflammation are key drivers of Alzheimer's disease (AD), but underlying mechanisms remain unclear.
- Early-stage demyelination is observed in AD models and patients, suggesting a potential link to disease progression.
Purpose of the Study:
- To investigate the role of demyelination-associated lysophosphatidylserine (LysoPS) and its receptor GPR34 in AD pathogenesis.
- To explore the therapeutic potential of targeting the LysoPS-GPR34 axis in AD.
Main Methods:
- Utilized a 5xFAD mouse model of AD to study demyelination, LysoPS levels, and microglial function.
- Employed genetic and pharmacological inhibition of the GPR34 receptor.
- Analyzed brain tissues from AD patients for LysoPS production and GPR34 expression.
Main Results:
- Demyelination in 5xFAD mice correlated with increased LysoPS in myelin debris, exacerbating neuroinflammation and cognitive impairment.
- Reducing LysoPS or inhibiting GPR34 improved microglial function, enhanced Aβ phagocytosis, reduced Aβ deposition, and restored memory.
- Elevated LysoPS and GPR34 expression were found in human AD brains.
Conclusions:
- Demyelination-derived LysoPS plays a pathogenic role in microglial dysfunction and AD progression.
- Targeting GPR34 presents a promising therapeutic strategy for Alzheimer's disease, independent of direct Aβ targeting.
More Related Videos
 09:33
Quantitative 3D In Silico Modeling q3DISM of Cerebral Amyloid-beta Phagocytosis in Rodent Models of Alzheimer's Disease
Published on: December 26, 2016
 04:55
A Stably Established Two-Point Injection of Lysophosphatidylcholine-Induced Focal Demyelination Model in Mice
Published on: May 11, 2022