Related Experiment Video
Updated: Jun 3, 2025
Inkjet Printing All Inorganic Halide Perovskite Inks for Photovoltaic Applications
Published on: January 22, 2019
Full analysis for the optical properties of the Cs2XInBr6 (X = Cu or Ag) double perovskites using both GW and
Fatemeh Negahdari1, Ali Mokhtari2,3, Vishtasb Soleimanian1,4
1Department of Physics, Faculty of Sciences, Shahrekord University, P.O. Box 115, Shahrekord, Iran.

Context:
Exploration for renewable and environmentally friendly energy sources has become a major challenge to overcome the depletion of fossil fuels and their environmental hazards. Therefore, solar cell technology, as an alternative solution, has attracted the interest of many researchers. In the present work, the Cs2XInBr6 (X = Cu or Ag) compounds as lead-free halide perovskites have been studied due to their direct energy gap in the range of solar energy, thermodynamic stability, low effective mass of electrons, and high absorption coefficient. The calculated optical gap and static refractive index about 1.35 (1.51) eV and 1.47 (1.41), respectively using BS (GW) approach for the Cs2AgInBr6 compound were in good agreement with experimental data. The threshold absorption was estimated about 1.03 (1.22) using BS (GW) approach (which correspond to the optical gap) for the Cs2CuInBr6 compound. Both compounds have small (< 0.35) reflection coefficient in the infrared, visible and UV regions and high absorption coefficient (105 cm-1). In the infrared and visible regions, the absorption coefficient of the Cu-based compound is much larger than the other, therefore this material can be more useful as an absorbent layer in solar cells (SC). Due to the fact that the spectrum of sunlight that reaches the earth includes 47% infrared, 46% visible and 7% ultraviolet, the Cu-based compound is more efficient for the SCs and the Cs2AgInBr6 compound is more suitable in the design of detectors.
Methods:
The electronic, structural and optical properties of Cs2XInBr6 (X = Cu or Ag) compounds have been calculated and analyzed using the Abinit computational package based on density functional theory (DFT). The ultrasoft pseudopotentials within the framework of the generalized gradient approximation (GGA) are adopted for the electron exchange-correlation potential and are employed for the calculations of the electronic, and optical properties as well. The wave functions have expanded on a plane-wave basis set to cutoff energy of 950 eV and 64 k-points with a k-mesh of 4 × 4 × 4 are considered for the integrations over the Brillouin zone. The behavior of the real and imaginary parts of the dielectric function and other optical quantities have been simulated using both RPA-GW (random phase approximation with GW energies) and Bethe-Salpeter formalisms.
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:18
X-ray Crystallography
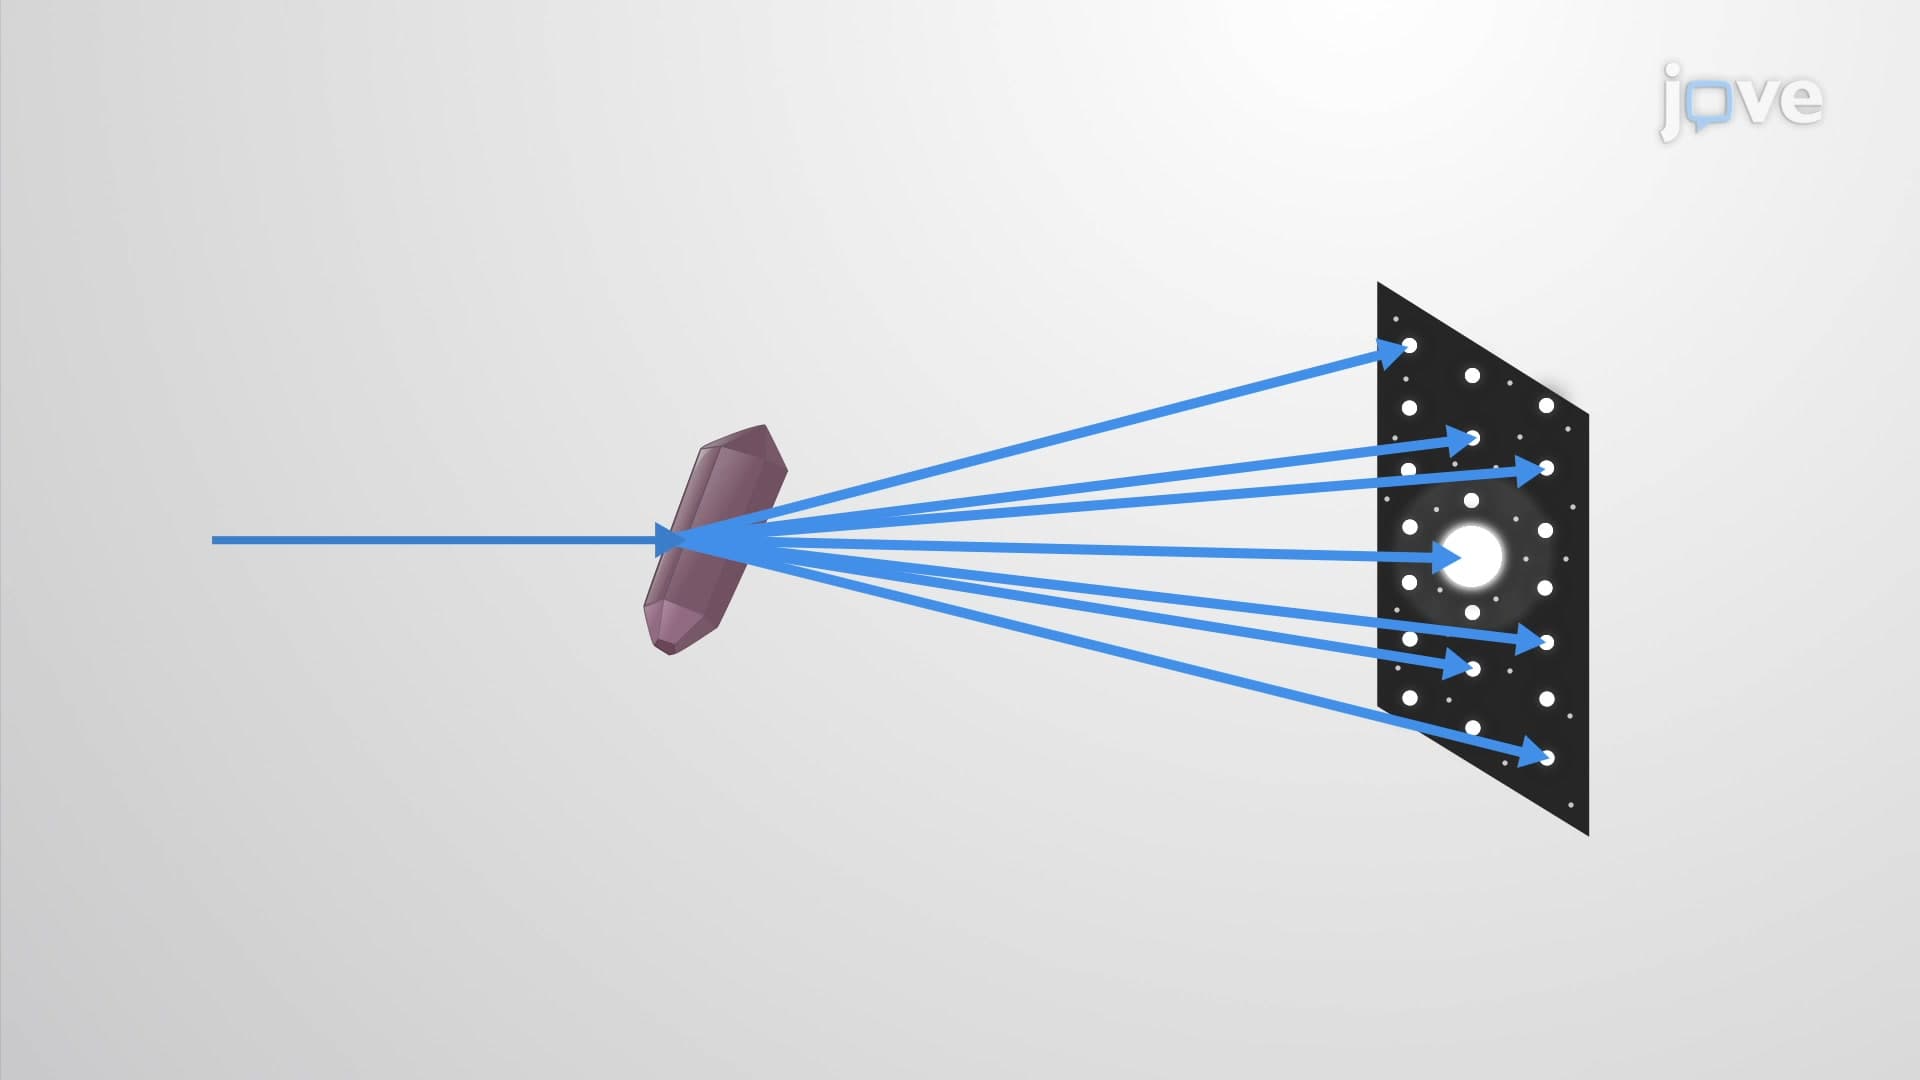
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...