Related Experiment Video
Updated: Jun 3, 2025
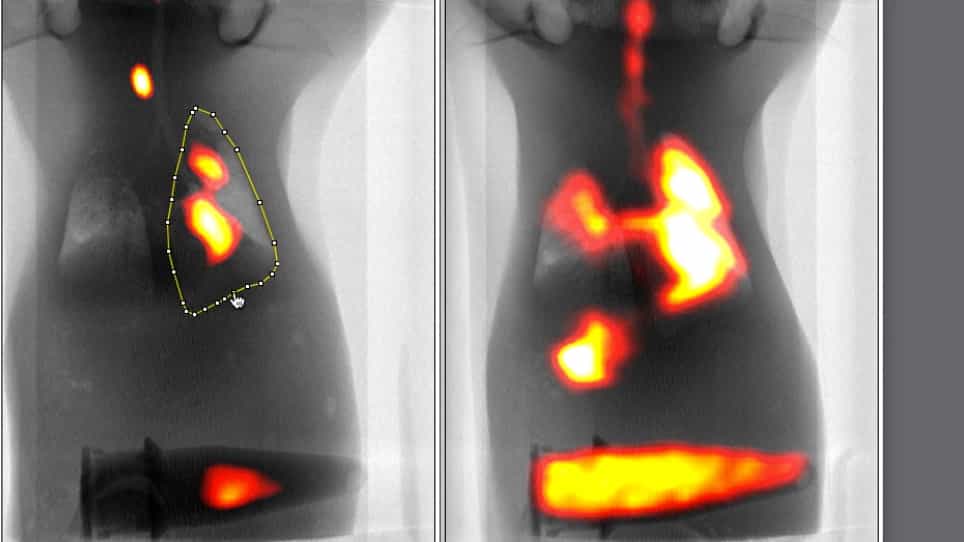

In vivo Evaluation of Mucociliary Clearance in Mice
Published on: December 18, 2020
Does using the Lung Clearance Index (LCI) inform clinical decisions in children with cystic fibrosis?
Lucy Perrem1, Stephanie Jeanneret-Manning2, Stephanie D Davis3
1Children's Health Ireland, Dublin, Ireland; University College Dublin, Dublin 4, Ireland.

Introduction:
The Lung Clearance Index (LCI) is an established research test, but its role in clinical decision-making is not well defined. This study estimated the proportion of treatment decisions that are changed or supported by the added information provided by LCI.
Methods:
A mixed methods prospective observational study was conducted in North America. Providers were invited to participate in a clinical vignette survey consisting of 10 hypothetical scenarios involving pediatric cystic fibrosis (CF) management. First, they made a clinical decision based on information captured in routine clinical visits. Then, the LCI value was made available, and providers were asked whether the LCI changed or supported their decision. A prospective study was also conducted at three CF centres to determine how often physicians make pulmonary treatment decisions at CF clinic visits and how often they perceive additional lung function data would be helpful for these decisions.
Results:
We received 522 vignette responses from 62 participants. LCI changed the decision in 18.4 % of cases, supported the decision in 57.1 % and did not impact the decision in 24.5 %. Data from patient encounters in the prospective study demonstrated that changes to pulmonary treatments were considered in 98/322 (30.4 %) visits; additional lung function information could potentially have helped in 64.3 % of the treatment decisions.
Conclusion:
LCI changes or supports a significant proportion of treatment decisions. Providers perceive that additional information about lung function could be helpful at the majority of encounters where changes in treatment are considered.
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:47
Lung Capacity
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History
 01:24
Respiratory Capacities
One key metric is the Inspiratory Capacity (IC), which represents the maximum amount of air that can be inhaled with full effort. IC is calculated by summing the tidal volume and inspiratory reserve volume, typically ranging from 2.4 to 3.6 liters.
The Functional Residual Capacity (FRC) represents the air in the...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:20
Chronic Obstructive Pulmonary Disease-I: Introduction