Related Experiment Video
Updated: Jun 2, 2025

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Ab initio calculations of electric field gradients in H-bond rich molecular crystals with nearly experimental
1Jožef Stefan Institute, Jamova 39, 1000 Ljubljana, Slovenia.

Abstract:
Ab initio calculations of electric field gradients (EFGs) in molecular crystals have advanced significantly due to the gauge including projector augmented wave (GIPAW) formalism, which accounts for the infinite periodicity in crystals. However, theoretical accuracies still lag behind experimental ones, making it challenging to distinguish experimentally distinguishable similar structures, a deficiency largely attributed to the limitation of GIPAW codes to generalized gradient approximation (GGA) density functional theory (DFT) functionals. In this study, we investigate whether hybrid DFT functionals can enhance the EFG calculation accuracy and the associated geometry optimization. Using the many-body expansion method, we focus on nitrogen EFGs in amino acids with complex H-bonding, which are often poorly described with GGA functionals. Our results show that both functionals provide highly accurate calculations that surpass current studies and approach experimental precision. The accuracies are also almost three times higher than available GIPAW/GGA calculations in the literature. However, we show that this difference is not due to the GGA functional but rather due to the improper selection of the nitrogen quadrupole moment.
More Related Videos


Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:27
π Electron Effects on Chemical Shift: Overview
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:04
Hydrogen Bonds
 02:18
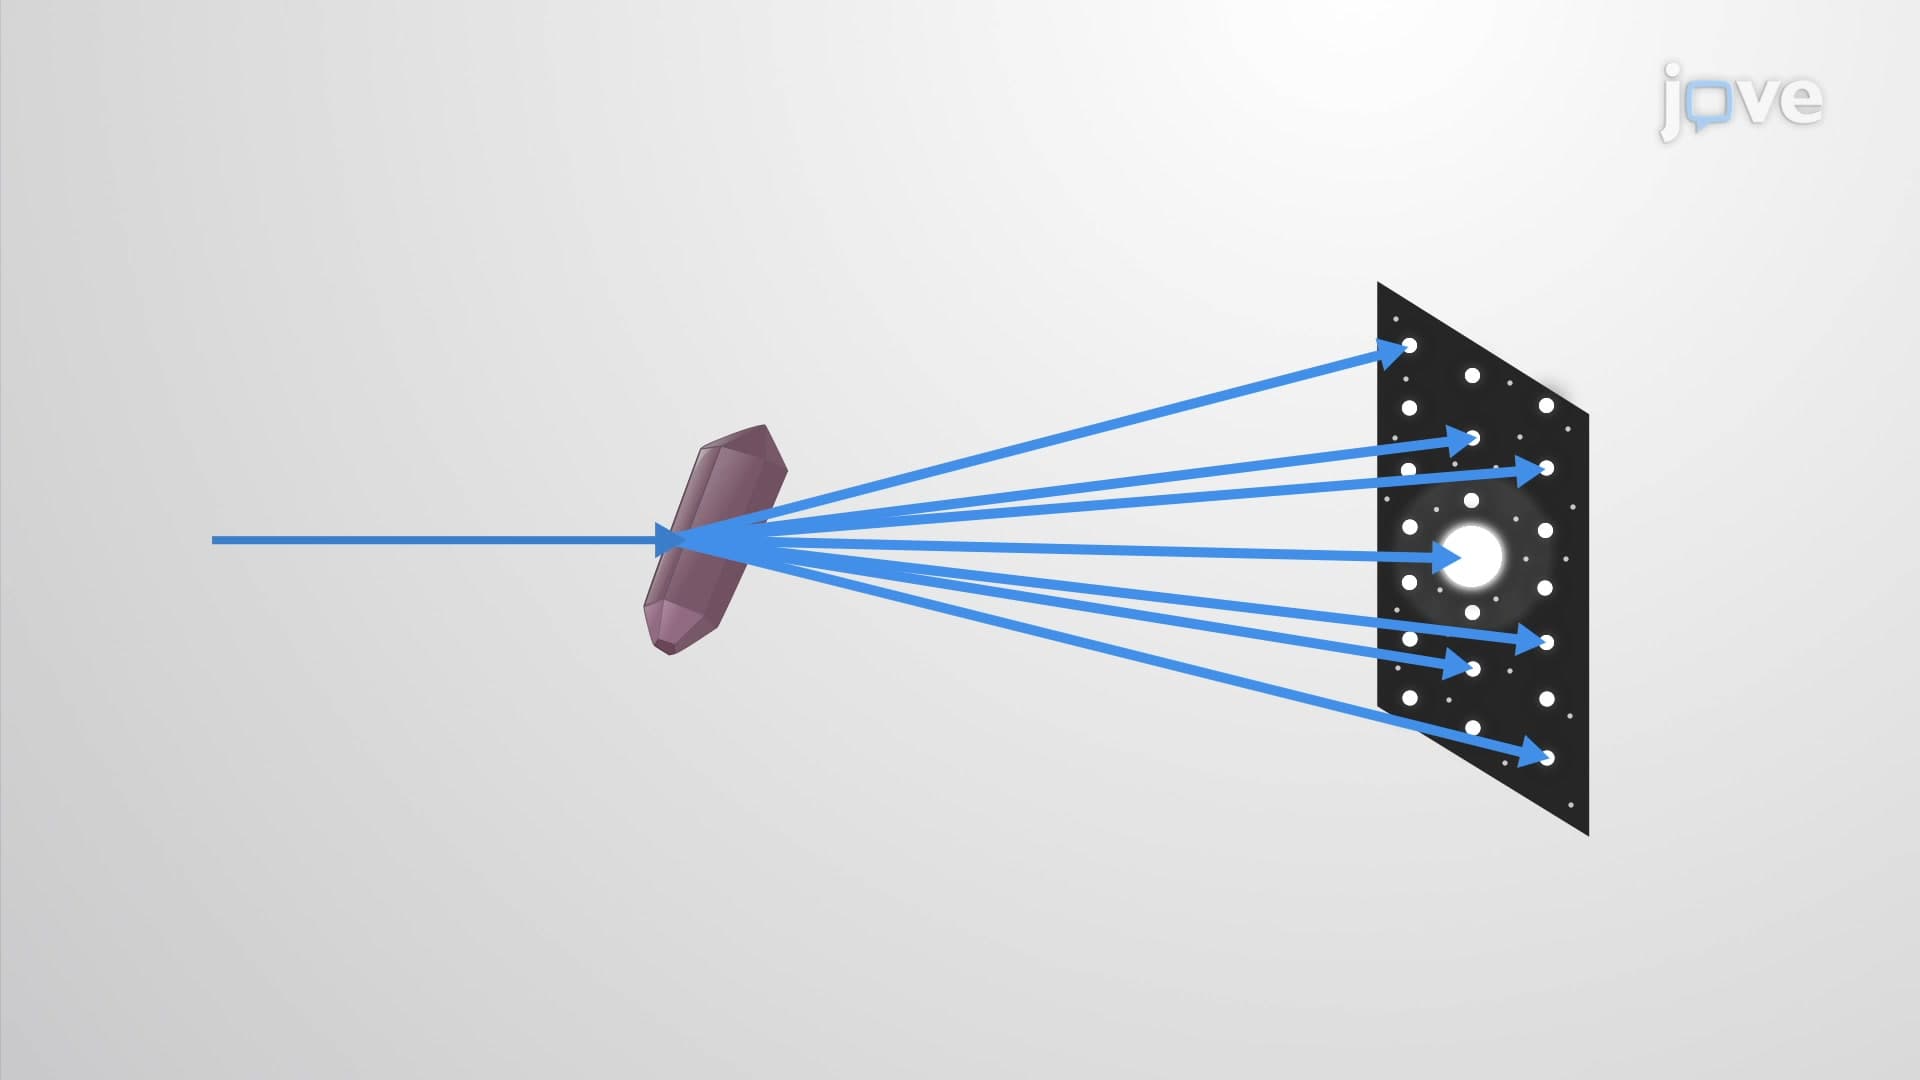
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:36
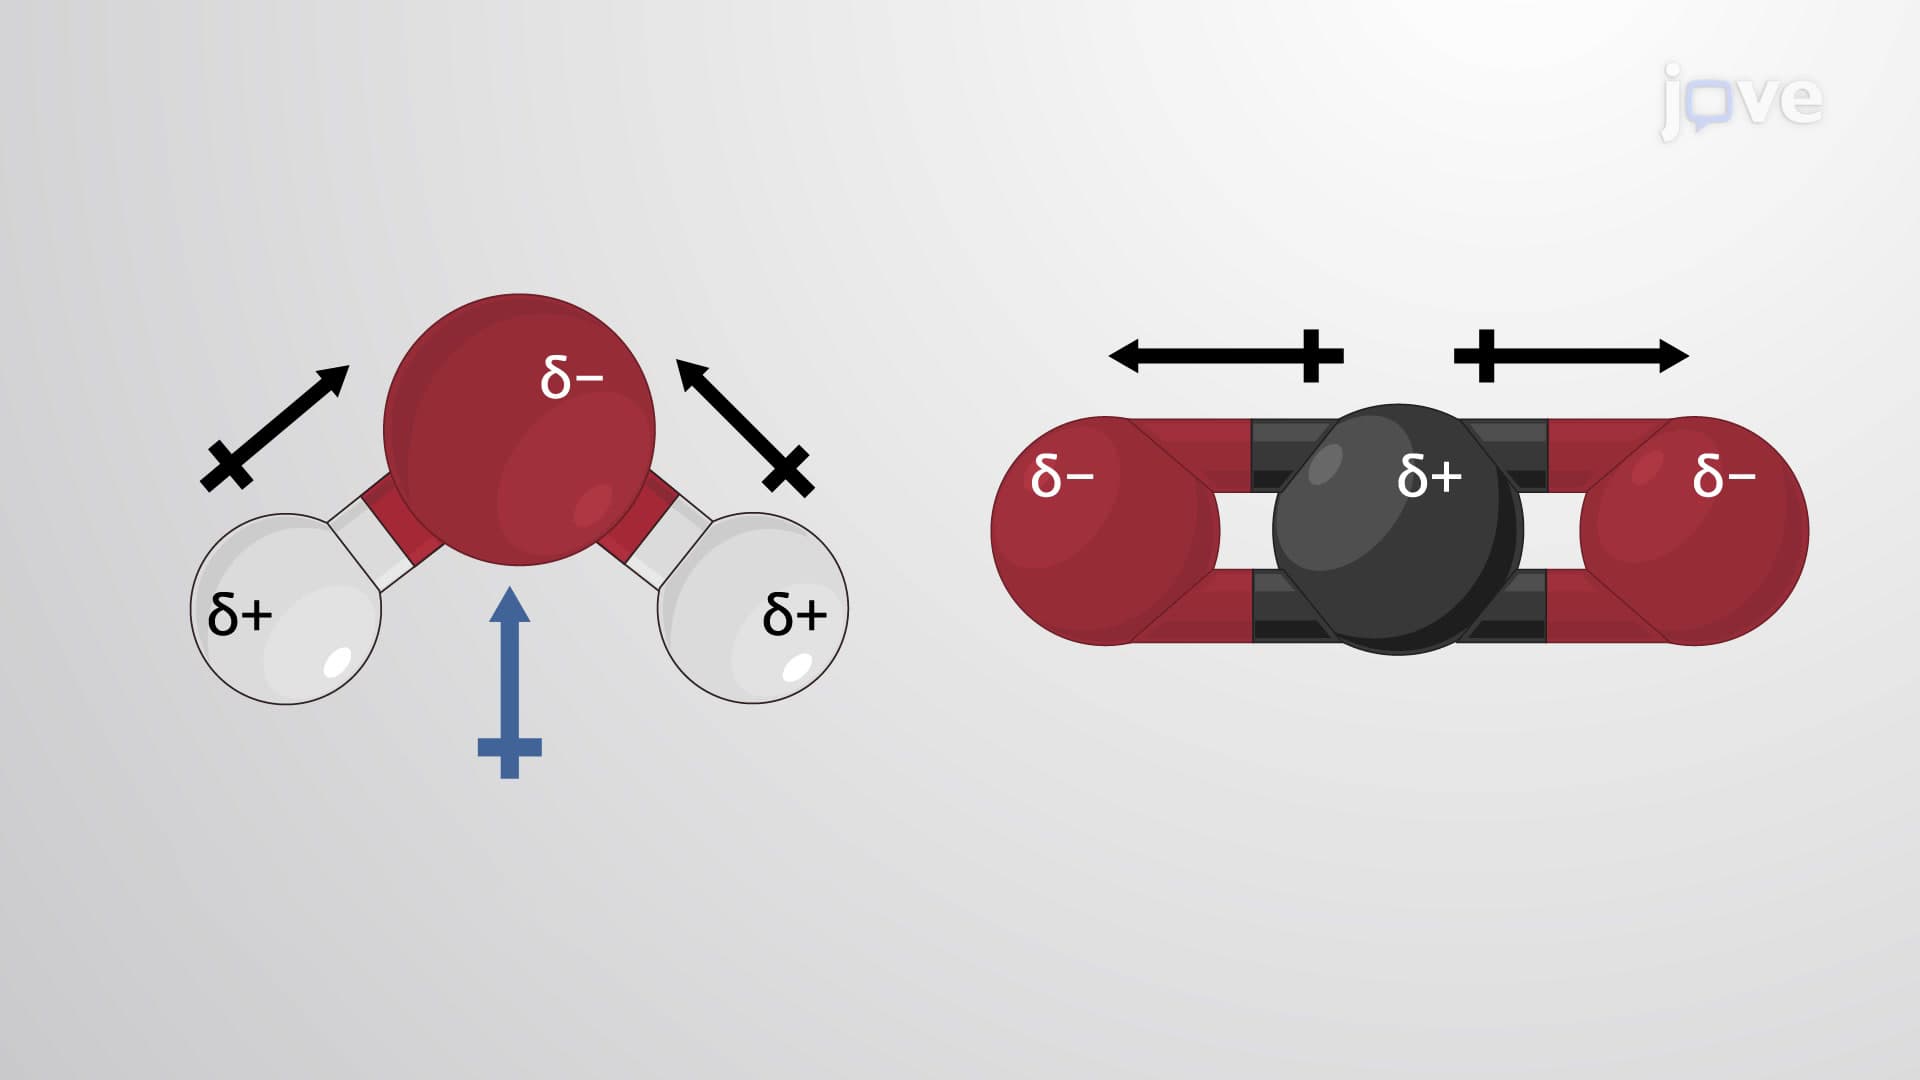
Molecular Geometry and Dipole Moments