Related Experiment Video
Updated: Jun 1, 2025
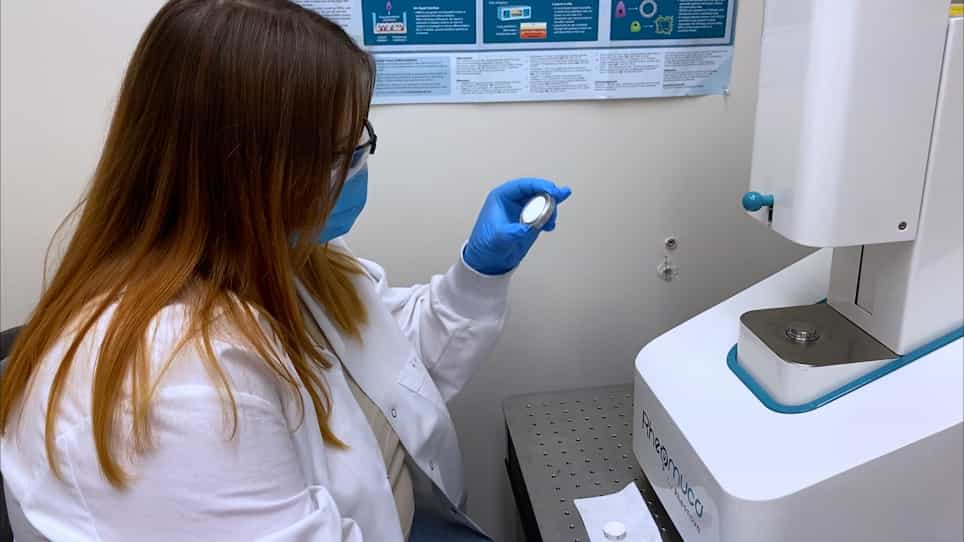

Rapid Viscoelastic Characterization of Airway Mucus Using a Benchtop Rheometer
Published on: April 21, 2022
Nanoscale Viscometry Reveals an Inherent Mucus Defect in Cystic Fibrosis
Olga Ponomarchuk1, Francis Boudreault1, Ignacy Gryczynski2
1Centre de recherche du Centre hospitalier de l'Université de Montréal (CRCHUM), Montréal, Québec H2X 0A9, Canada.

Abstract:
The abnormally viscous and thick mucus is a hallmark of cystic fibrosis (CF). How the mutated CF gene causes abnormal mucus remains an unanswered question of paramount interest. Mucus is produced by the hydration of gel-forming mucin macromolecules that are stored in intracellular granules prior to release. Current understanding of mucin/mucus structure before and after secretion remains limited, and contradictory models exist. Here, we used a molecular viscometer and fluorescence lifetime imaging of human bronchoepithelial cells (Normal and CF) to measure nanometer-scale viscosity. We found significantly elevated intraluminal nanoviscosity in a population of CF mucin granules, indicating an intrinsic, presecretory mucin defect. Nanoviscosity influences protein conformational dynamics and function. Its elevation along the protein secretory pathway could arise from molecular overcrowding, impacting mucin's post-translational processing, hydration, and mucus rheology after release. The nanoviscosity of secreted CF mucus was elevated compared to that of non-CF. Interestingly, it was higher after release than in granules. Validation experiments indicate that reduced mobility of water hydrating mucin macromolecules may contribute to the high nanoviscosity in mucus and mucin granules. This suggests that mucins have a weakly ordered state in granules but adopt a highly ordered, nematic crystalline structure when secreted. This challenges the traditional view of mucus as a porous agarose-like gel and suggests an alternative model for mucin organization before and after secretion. Our study also indicates that endoplasmic reticulum stress due to molecular overcrowding could contribute to mucus pathogenesis in CF cells. It encourages the development of therapeutics that target presecretory mechanisms in CF and other muco-obstructive lung diseases.
More Related Videos

Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...