Related Experiment Video
Updated: Jun 16, 2026

Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
Discovery of Severe Acute Respiratory Syndrome Coronavirus 2 Main Protease Inhibitors through Rational Design of
Huiying Jiang1, Heping Xia1, Zhonghua Wang2
1Department of Chemistry, University of Shanghai for Science and Technology, Shanghai, P. R. China.

Abstract:
As severe acute respiratory syndrome coronavirus 2 (SARS‑CoV‑2) variants continue to emerge, there is an urgent need to develop more effective antiviral drugs capable of combating the COVID-19 pandemic. The main protease (Mpro) of SARS-CoV-2 is an evolutionarily conserved drug discovery target. The present study mainly focused on chemoinformatics computational methods to investigate the efficacy of our newly designed trifluoromethyl-1,3,4-oxadiazole amide derivatives as SARS-CoV-2 Mpro inhibitors. Drug-likeness absorption, distribution, metabolism, excretion, and toxicity analysis, molecular docking simulation, density functional theory (DFT), and molecular dynamics simulation methods were included. A comprehensive drug-likeness analysis was performed on the 14 newly designed compounds (1a-1n), and this series of small molecule inhibitors showed potential anti-SARS-CoV-2 activity. In order to reveal the mechanism of drug interaction, these novel compounds were classified by structure, and molecular docking simulations were performed. The results showed good interactions and identified the key amino acid residue GLY-143. Further DFT analysis using B3LYP-D3BJ functional and 6-311 + + G (d, p) basis set was performed to optimize the optimal configuration of the Mpro inhibitors, and the infrared spectrum of the vibration frequency was analyzed to clearly understand the structure and stability of the drug. The electrostatic potential map was analyzed to predict the reactivity of functional groups and protein-substrate interactions. The frontier molecular orbital analysis and density of states map showed the reactivity level and stability of the drug itself, among which 1i had the smallest energy gap difference (ΔEgap = 3.64 ev), showing good reactivity. The analysis of global reactivity descriptors such as electrophilic index (ω) and chemical potential (μ) also showed that our newly designed Mpro inhibitors had stronger interactions. Molecular dynamics simulation further revealed the stable binding of the Mpro inhibitors in a solvent environment. The binding free energy results calculated by Molecular Mechanics / Poisson Boltzmann Surface Area (MM/PBSA) all exceeded the Food and Drug Administration-approved standard reference drug (Nirmatrelvir), and the free energy landscape and principal component analysis also further described the energy sites formed during the binding process between the drug molecule and the ligand-protein and the changes in conformation. These new series of small molecule inhibitors studied in this work will provide the necessary theoretical basis for the synthesis and activity evaluation of novel SARS-CoV-2 Mpro inhibitors.
More Related Videos

Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:29
Coronavirus
 01:29
Respiratory Syncytial Virus Disease
 01:30
Inhibitors of Viral Protein Synthesis
 01:25
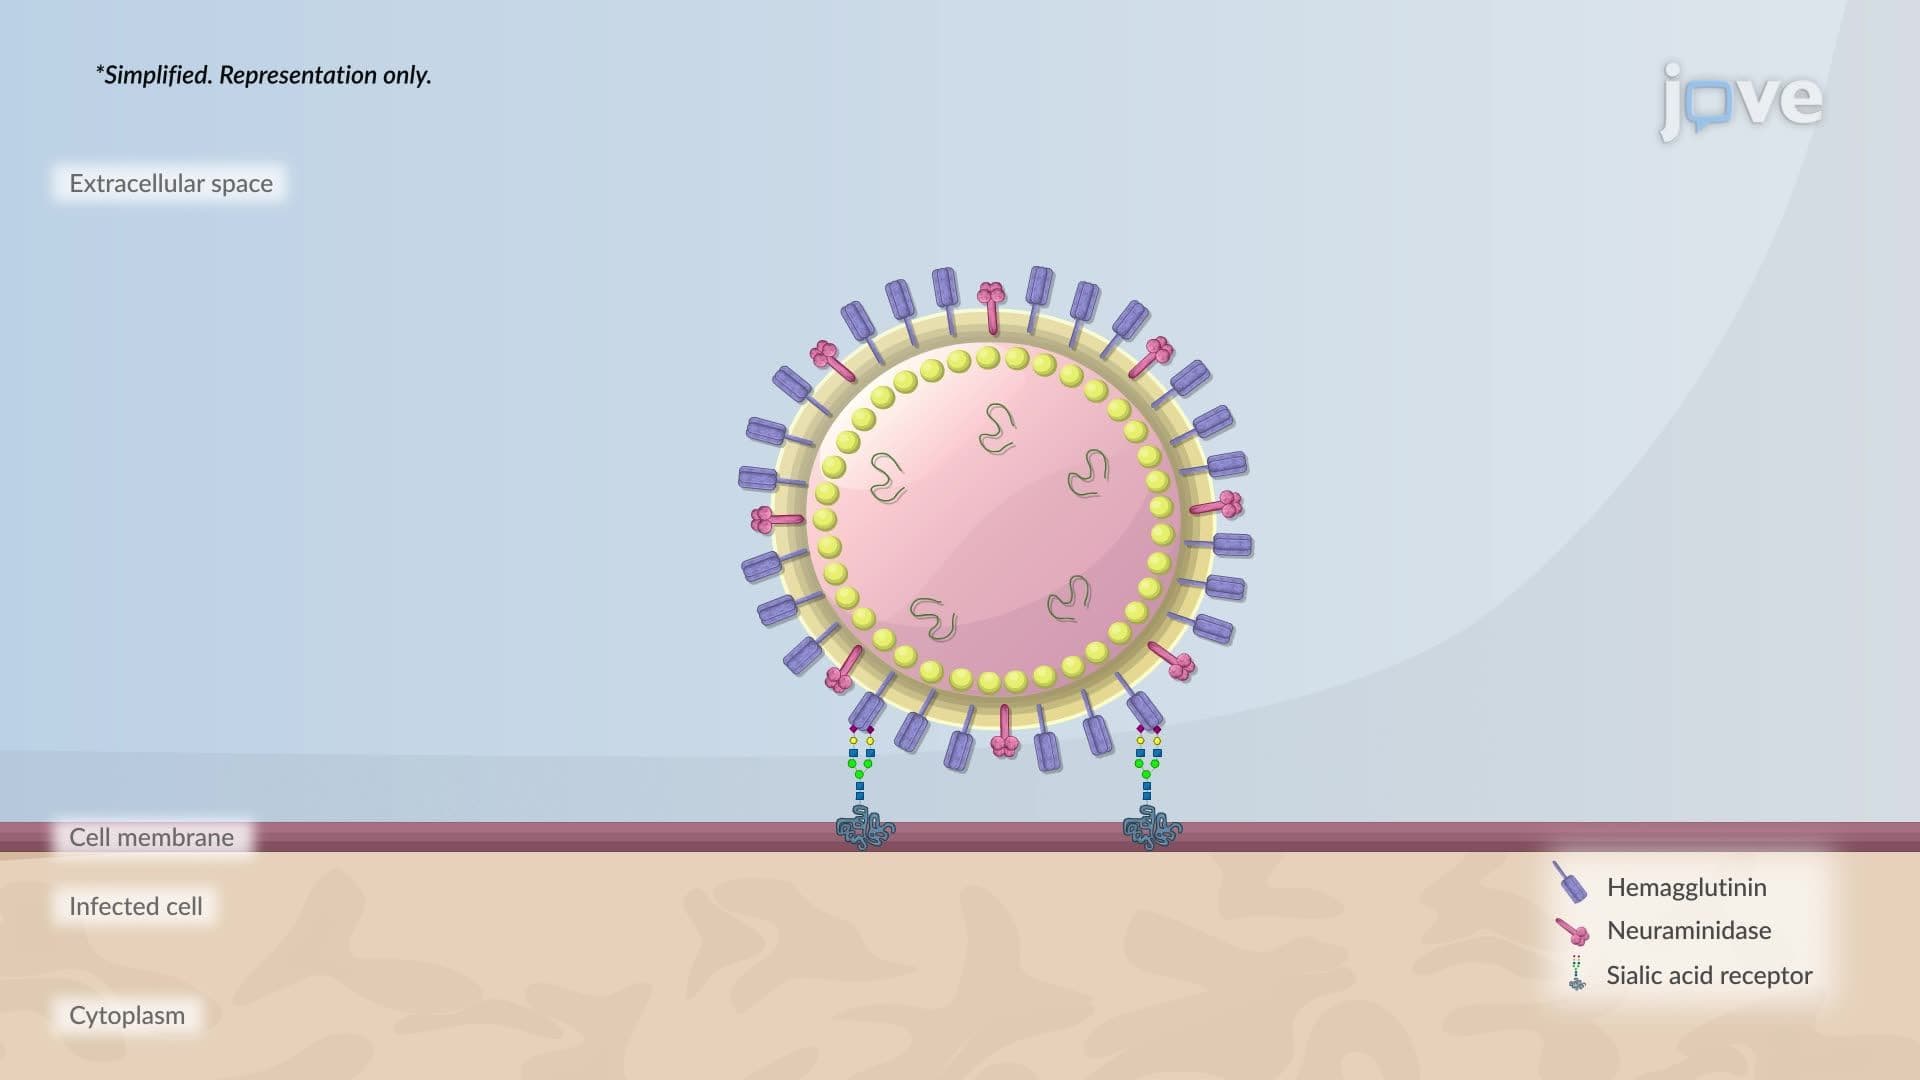
Inhibitors Of Virion Release
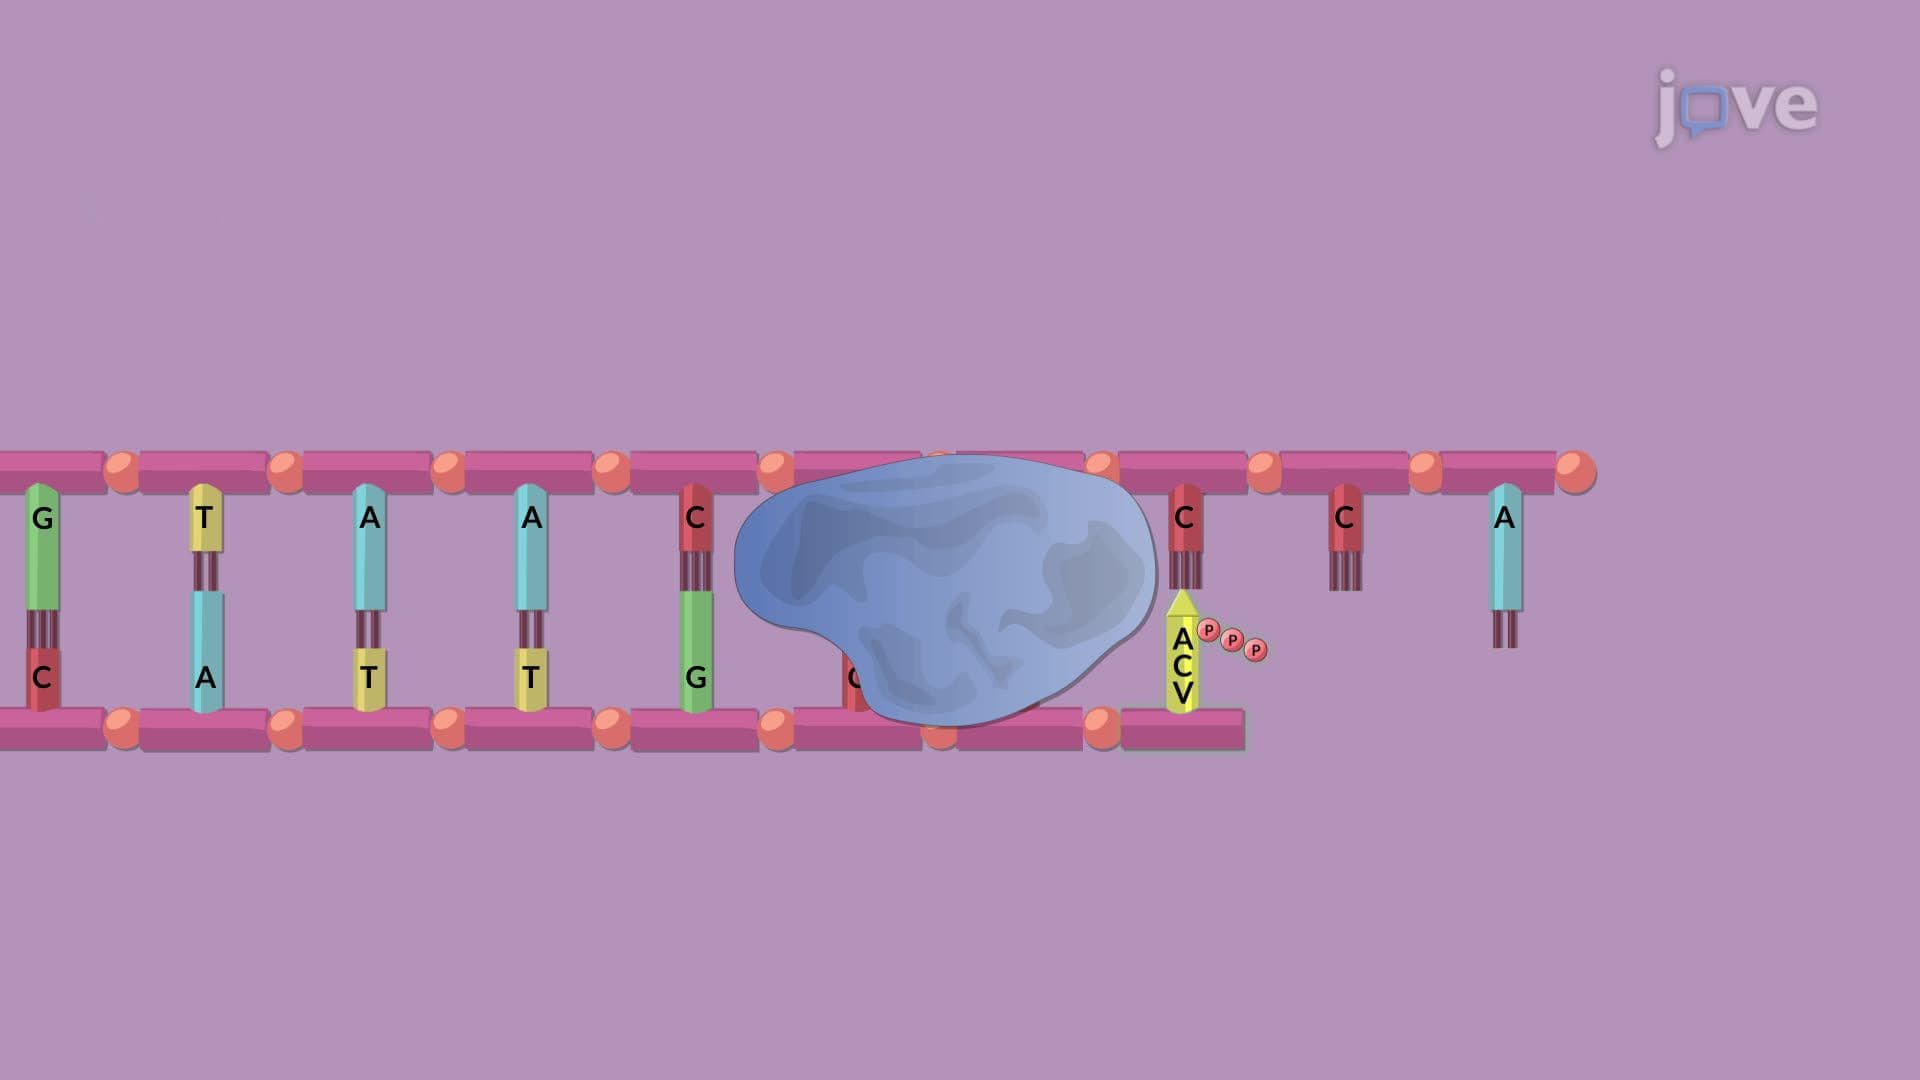 01:22
Antiviral Nucleoside Inhibitors