Related Experiment Video
Updated: May 31, 2025

Quantification of the Immunosuppressant Tacrolimus on Dried Blood Spots Using LC-MS/MS
Published on: November 8, 2015
Effect of Target-Mediated Disposition on Iptacopan Clinical Pharmacokinetics in Participants with Normal or Impaired
Elise Burmeister Getz1, Richard R Stein2,3, Martin Fink3
1Biomedical Research, Novartis, Emeryville, California, USA.

Iptacopan, a first-in-class complement factor B inhibitor acting proximally in the alternative complement pathway, has been shown to be safe and effective for patients with complement-mediated diseases. Iptacopan selectively binds with high affinity to factor B, a soluble, plasma-based, hepatically produced protein. Factor B is abundant in the circulation but can be saturated at the iptacopan clinical dose of 200 mg twice daily. Iptacopan pharmacokinetics (PK) are influenced by target binding. This target-mediated drug disposition (TMDD) behavior makes PK data useful for understanding target occupancy and motivates modeling of drug-target binding to connect exposure with pharmacological effect. A phase I hepatic impairment (HI) PK study measuring both total and unbound iptacopan PK profiles provided an opportunity to characterize the effect of variation in target concentration (due to varying hepatic function) on iptacopan PK. HI caused no change in total iptacopan exposure but increased unbound iptacopan exposure 1.38- to 3.72-fold in participants with mild, moderate, or severe HI relative to demographically matched participants with normal hepatic function, with the largest increases in severe HI. A two-site competitive binding model was developed to elucidate the relationship between iptacopan PK and factor B occupancy to characterize exposure thresholds for maximal target engagement. The model was used to assess alternative dose regimens to provide insight into how to approach dose recommendations for patients with severe HI. This study provides an example of small-molecule TMDD, a behavior typically associated with targeted biologics; its importance is too often underappreciated in small-molecule drug development.
Iptacopan, a first-in-class complement factor B inhibitor acting proximally in the alternative complement pathway, has been shown to be safe and effective for patients with complement-mediated diseases. Iptacopan selectively binds with high affinity to factor B, a soluble, plasma-based, hepatically produced protein. Factor B is abundant in the circulation but can be saturated at the iptacopan clinical dose of 200 mg twice daily. Iptacopan pharmacokinetics (PK) are influenced by target binding. This target-mediated drug disposition (TMDD) behavior makes PK data useful for understanding target occupancy and motivates modeling of drug-target binding to connect exposure with pharmacological effect. A phase I hepatic impairment (HI) PK study measuring both total and unbound iptacopan PK profiles provided an opportunity to characterize the effect of variation in target concentration (due to varying hepatic function) on iptacopan PK. HI caused no change in total iptacopan exposure but increased unbound iptacopan exposure 1.38- to 3.72-fold in participants with mild, moderate, or severe HI relative to demographically matched participants with normal hepatic function, with the largest increases in severe HI. A two-site competitive binding model was developed to elucidate the relationship between iptacopan PK and factor B occupancy to characterize exposure thresholds for maximal target engagement. The model was used to assess alternative dose regimens to provide insight into how to approach dose recommendations for patients with severe HI. This study provides an example of small-molecule TMDD, a behavior typically associated with targeted biologics; its importance is too often underappreciated in small-molecule drug development.
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:14
Hepatic Drug Clearance: Role of Transporters
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:09
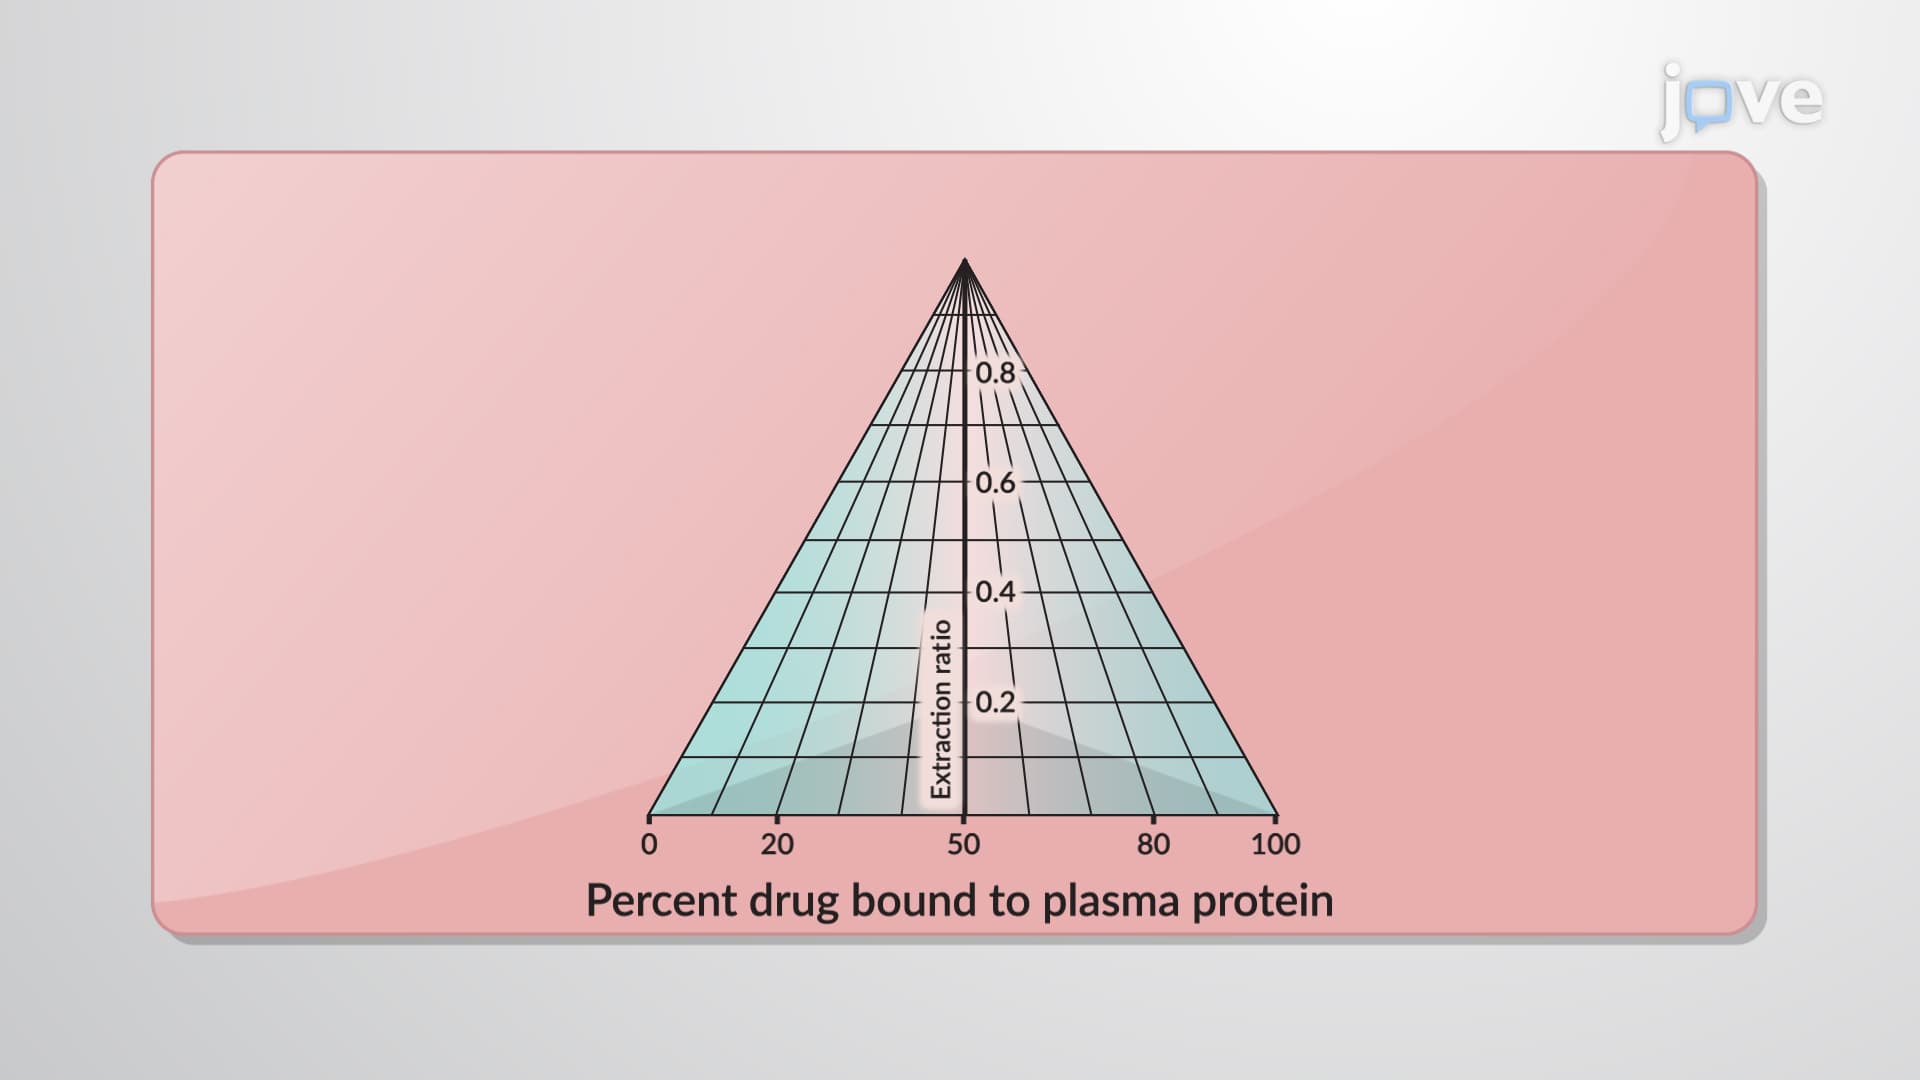
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:21
Factors Affecting Drug Response: Overview
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the...