Related Experiment Video
Updated: May 29, 2025

Preparation of 6-aminocyclohepta-2,4-dien-1-one Derivatives via Tricarbonyltroponeiron
Published on: August 12, 2019
Comparison of a Nonheme Iron Cyclopropanase with a Homologous Hydroxylase Reveals Mechanistic Features Associated
Yu-Cong Zheng1, Xiaojun Li1, Lide Cha1
1Department of Chemistry, North Carolina State University, Raleigh, North Carolina 27695, United States.

Abstract:
Despite the diversity of reactions catalyzed by mononuclear iron and 2-oxoglutarate-dependent enzymes, the factors that lead to diverse reaction outcomes beyond canonical hydroxylation remain elusive. Cyclopropanation reactions are of particular interest not only due to the prevalence of cyclopropane moieties in pharmaceuticals but also due to the chemistry that allows cyclopropanation to outcompete oxygen rebound. HrmJ is one such cyclopropanase from the biosynthetic pathway of hormaomycin; however, a homologue is herein discovered that instead catalyzes C-hydroxylation of the same nitro enolate substrate. These enzymes were reconstituted with Mn(II) and V(IV)═O as mimics of the resting (Fe(II)) and reactive (Fe(IV)═O) intermediate states, respectively. Corresponding crystal structures of the cyclopropanase bound with a substrate imply H atom transfer via an offline π-pathway. In contrast, analogous structural analysis of the hydroxylase implies H atom abstraction likely proceeds through a σ-pathway. Preparation of isotopically labeled substrates and stopped-flow kinetic analyses indicate that while the pro-S hydrogen of C4 is abstracted in both enzymes, the Fe(IV)═O intermediate reacts ca. 17-fold faster in the active site of the hydroxylase, consistent with the mechanistic assignments. These results also support a correlation between the mechanism of H atom transfer and the subsequent fate of the substrate radical once generated. A subtle difference in substrate positioning not only affects the H atom abstraction pathway but also allows the nitro enolate moiety to intercept the resulting substrate radical in the active site of the cyclopropase, thereby facilitating intramolecular C-C bond formation in a stereoselective manner.
More Related Videos
 10:01
Protein Film Infrared Electrochemistry Demonstrated for Study of H2 Oxidation by a [NiFe] Hydrogenase
Published on: December 4, 2017
 08:31
Anaerobic Protein Purification and Kinetic Analysis via Oxygen Electrode for Studying DesB Dioxygenase Activity and Inhibition
Published on: October 3, 2018
Related Concept Videos
 01:27
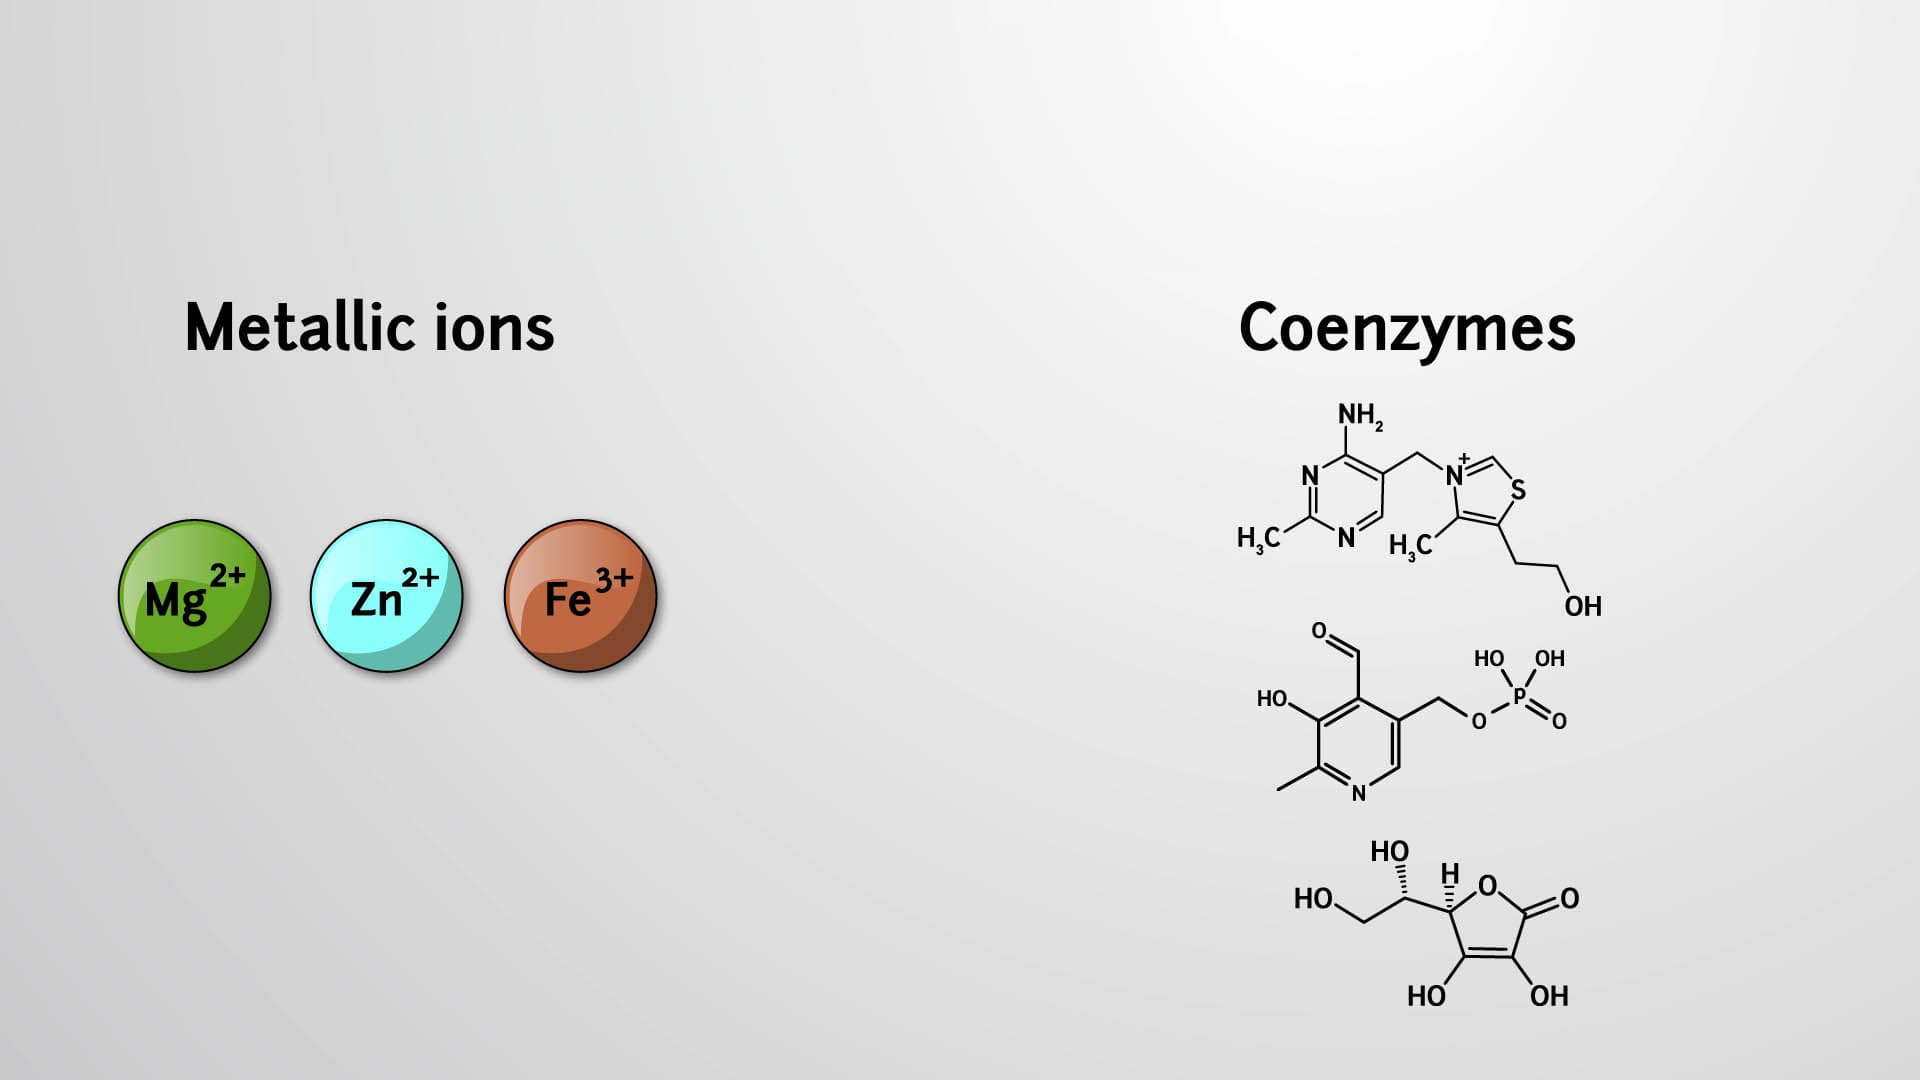
Cofactors and Coenzymes
 01:17
Drug Metabolism: Phase I Reactions
 02:44
Oxidation of Alkenes: Syn Dihydroxylation with Osmium Tetraoxide
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 01:17
Oxidation of Phenols to Quinones
o-hydroxy phenols are oxidized to o-quinones and p-hydroxy phenols to p-quinones. Such redox reactions involve the transfer of two electrons and two protons. The reversible redox...
 01:13
Introduction to Mechanisms of Enzyme Catalysis