Related Experiment Video
Updated: May 29, 2025

Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Spectroscopic (FT-IR and FT-Raman) and quantum chemical study on monomer and dimer of benznidazole from DFT and
Tirth Raj Paneru1,2, Manoj Kumar Chaudhary3, Poonam Tandon4
1Central Department of General Science, Far Western University, Mahendranagar, 10400, Nepal.

Abstract:
This work presents the quantum chemical calculations of the monomer and dimer of benznidazole using density functional theory (DFT) at the B3LYP/6-311++G(d,2p) level of theory. A one-dimensional potential energy surface scan was carried out across flexible bonds to find the minimum energy structure. The structure with minimum energy was taken as a monomer and dimer is constructed based on intermolecular hydrogen bonding N-H…O. The vibrational analysis was conducted by comparing the calculated FT-IR and FT-Raman spectra of the monomer and dimer with the experimental ones. The red shift in the spectra of amide and carbonyl functional groups indicates their involvement in intermolecular hydrogen bonding in crystal packing, while the other peaks showed good agreement with the experimental result. The intra- and intermolecular interactions in the monomer and dimer were analyzed using various tools. The steric effects and van der Waals forces in the dimer were found to be more effective than the monomer. The dimer in the gaseous medium was found to have a lower Frontier molecular orbital energy (ΔEL-H) value than the monomer, suggesting that it is more reactive in a gaseous medium. The ELF value for hydrogen in monomer and dimer around the ring was found to be more which confirms that the electrons in these regions are more localized. The negative value of the overlap population density of states (OPDOS) both in monomer and dimer indicate that there are anti-bonding orbitals between the acetamide and the benzyl groups of the compound. The drug potential of benznidazole was evaluated by molecular docking with carbonic anhydrase XII, which shows the highest binding affinity of (-8.3 kcal/mol) with 6YH8, indicating that benznidazole is its potent inhibitor.
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:34
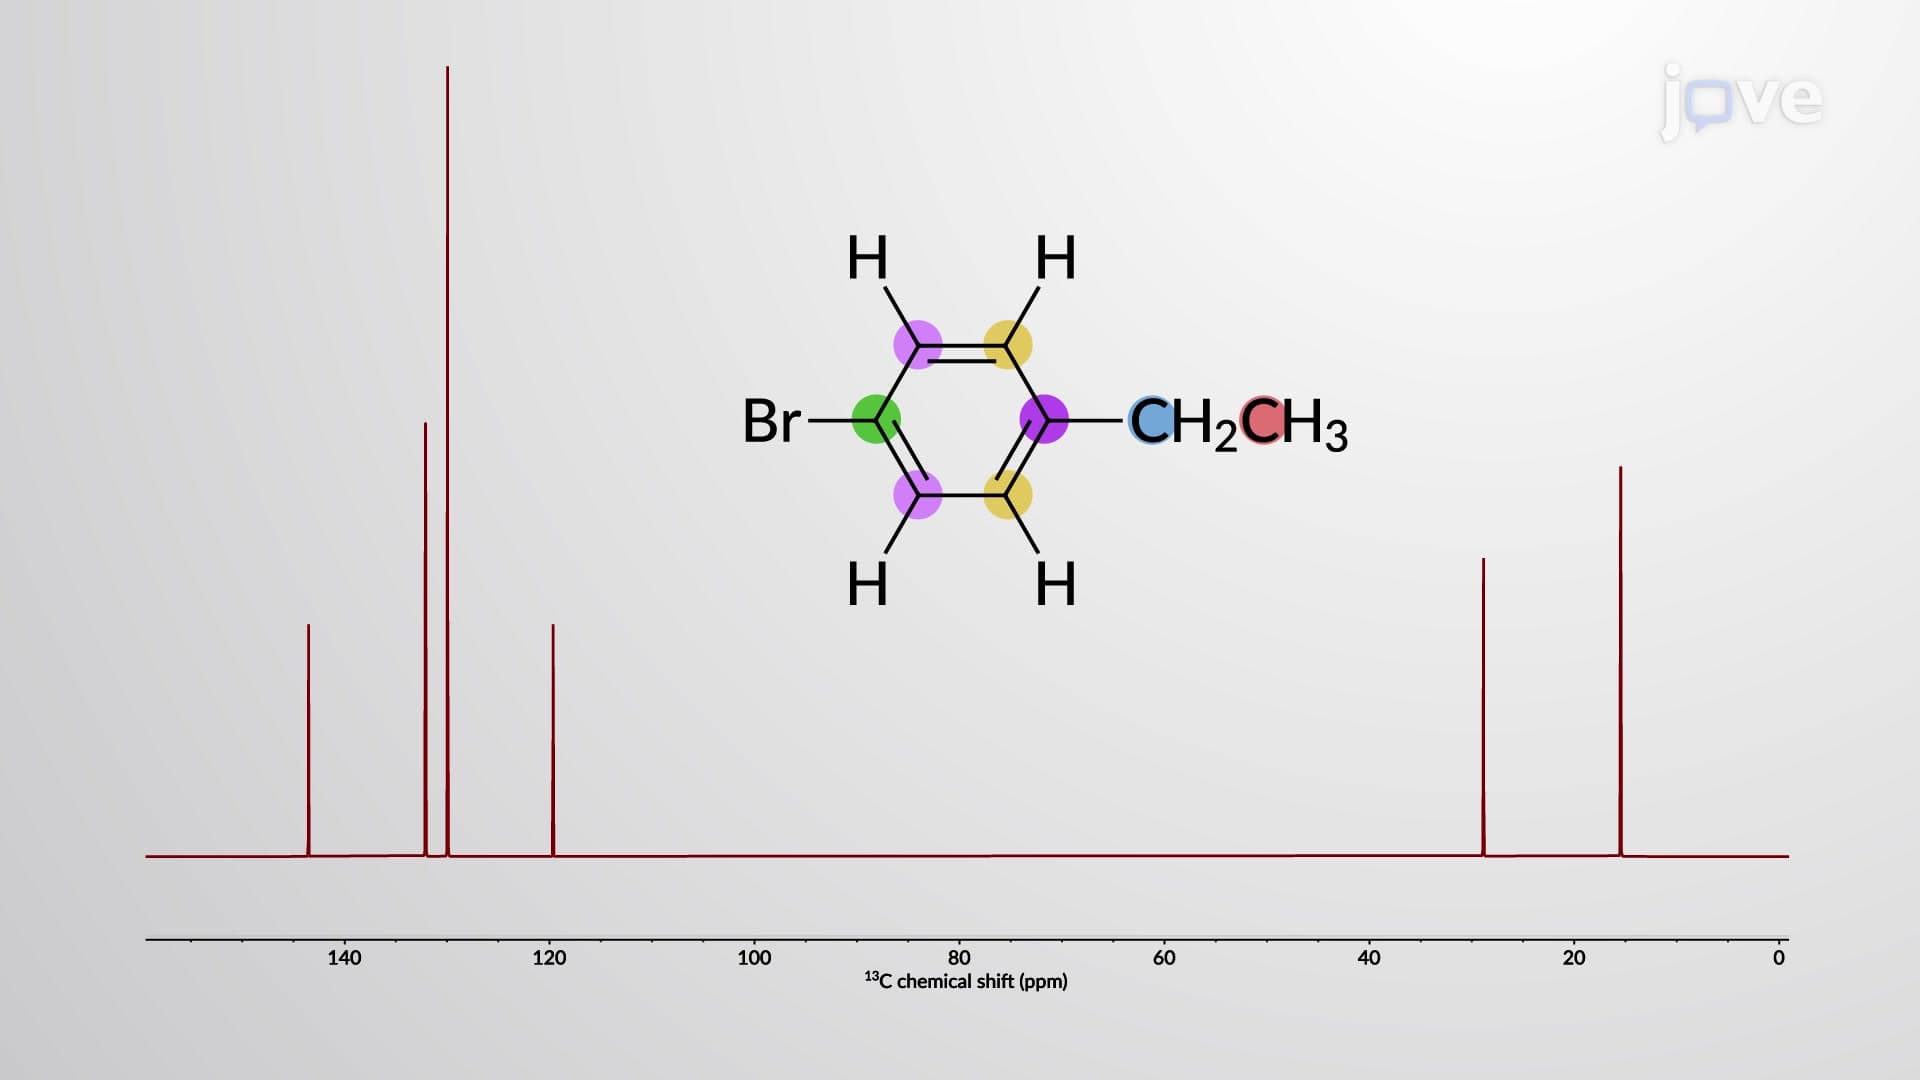
NMR Spectroscopy of Benzene Derivatives
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
Directing and Steric Effects in Disubstituted Benzene Derivatives
 03:51
Molecular Orbital Theory II
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules