Ligand impacts on band edge energies and excited state splittings of silicane
Related Experiment Video
Updated: May 27, 2025
Dependence of Laser-induced Breakdown Spectroscopy Results on Pulse Energies and Timing Parameters Using Soil Simulants
Published on: September 23, 2013
Guoying Yao1,2, Ekadashi Pradhan2, Zhenyu Yang1
1MOE Laboratory of Bioinorganic and Synthetic Chemistry, Lehn Institute of Functional Materials, School of Chemistry, IGCME, Sun Yat-Sen University, Guangzhou, Guangdong, 510275, China. yangzhy63@mail.sysu.edu.cn.

Abstract:
Two-dimensional (2D) silicanes are promising semiconductors for applications in optoelectronics and photochemistry. Covalent functionalization presents a facile strategy to customize silicanes to attain desired properties. A comprehensive understanding of the impacts of ligands on silicanes is thus pivotal. In this study, we perform density functional theory (DFT) and time-dependent DFT (TDDFT) calculations to investigate the effects of three typical classes of ligands: (1) σ-withdrawing and π-donating, (2) σ-withdrawing and π-withdrawing, and (3) σ-donating and π-withdrawing, on the geometric structure, electronic structure, and band edge excited states of silicanes. Covalent functionalization of silicanes enables a wide range of band edge energies. The band gaps can be tuned between indirect and direct, and the underlying mechanism is explained for the first time. Additionally, TDDFT calculations confirm that the band edge optical absorptions can be adjusted by the ligands broadly from the near-infrared to the visible light region. These desirable properties enhance the functionalities of silicanes. Methodologically, we discuss the applicability of using the conventional one-particle orbital model to describe the excited states of silicanes, and nail down the ∼0.1 eV inaccuracy of the model in describing excited state splittings. We present empirical functions for quick estimations of band edge energies of covalently functionalized silicanes. Through careful comparisons, we justify the use of the carbon-adapted σ/π-donating/accepting indices of typical ligands in silicon chemistry and present a set of silicon-based indices for future prudent applications in inorganic computational studies of silicon-based materials.
More Related Videos
Related Concept Videos
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:35
Band Theory
The energy difference between these bands is known as the band gap.
Conductor, Semiconductor,...
 01:20
Types of Semiconductors
 01:12
Fermi Level Dynamics
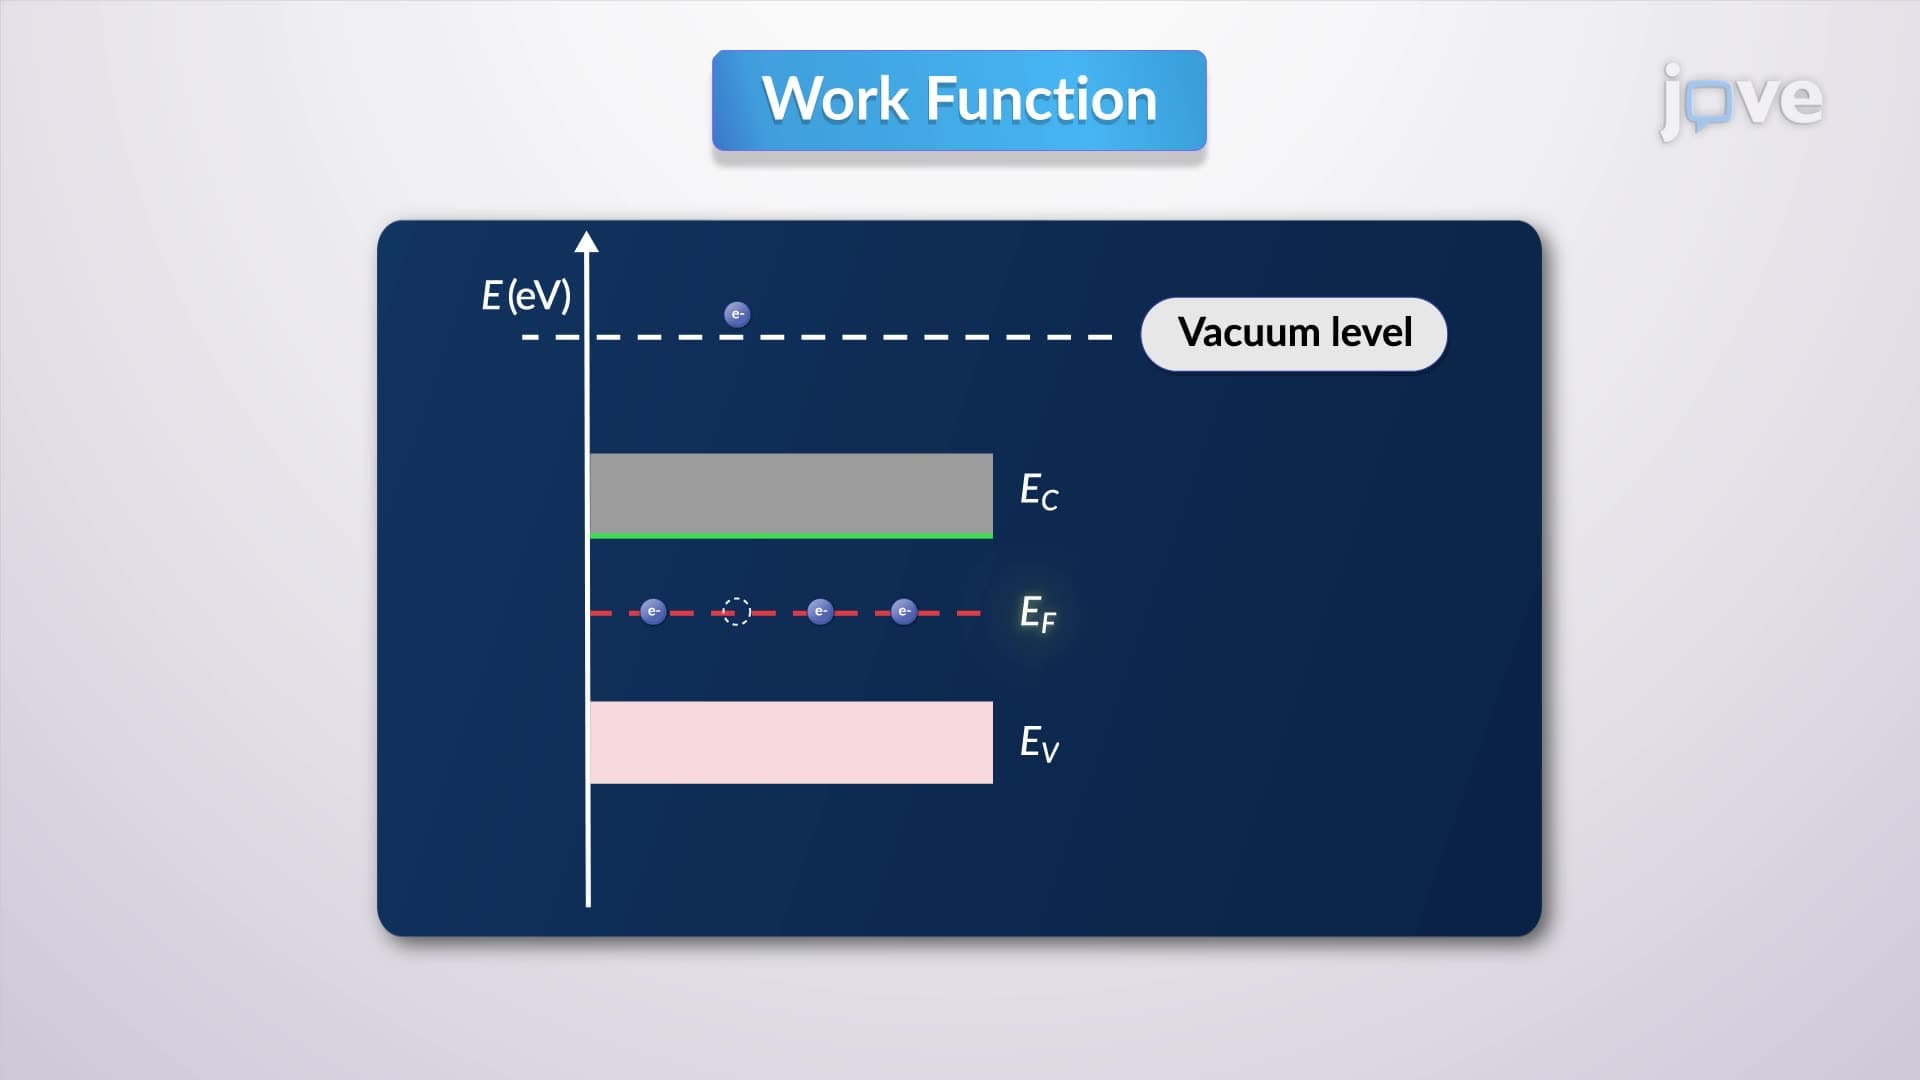
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...