Related Experiment Video
Updated: May 27, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accurate prediction of electron correlation energies of topological atoms by delta learning from the Müller
Prasanta Bandyopadhyay1, Bienfait K Isamura1, Paul L A Popelier1
1Department of Chemistry, The University of Manchester, Manchester M13 9PL, United Kingdom.

Abstract:
FFLUX is a polarizable machine-learning force field that deploys pre-trained kernel-based models of quantum topological properties in molecular dynamics simulations. Despite a track record of successful applications, this unconventional force field still uses Lennard-Jones parameters to account for dispersion effects when performing in-bulk simulations. However, optimal Lennard-Jones parameters are system-dependent and not easy to calibrate. Fortunately, physics-informed dispersion energies can be obtained from the two-particle density matrix (2PDM) of any system using correlated wavefunctions. The only challenge is that the 2PDM is a humongous object whose calculation is very time-consuming and memory-greedy. In this proof-of-concept study, we utilize the Δ-learning method to address both problems using a small set of water trimers. More specifically, we obtain pure two-electron correlation energies with the aug-cc-pVDZ basis set at the cost of Müller-approximated 2PDM calculated at a very small basis set, 6-31+G(d). We also benchmark different Δ-learning tasks designed by changing the baseline and target method and/or the basis set. Our experiments suggest that two-electron correlation energies of weakly relaxed water trimers can be accurately predicted via Δ-learning with a maximum absolute error of 1.30 ± 0.32 kJ/mol traded against a colossal computational speed-up of roughly 40 times.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:45
The Quantum-Mechanical Model of an Atom
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 01:18
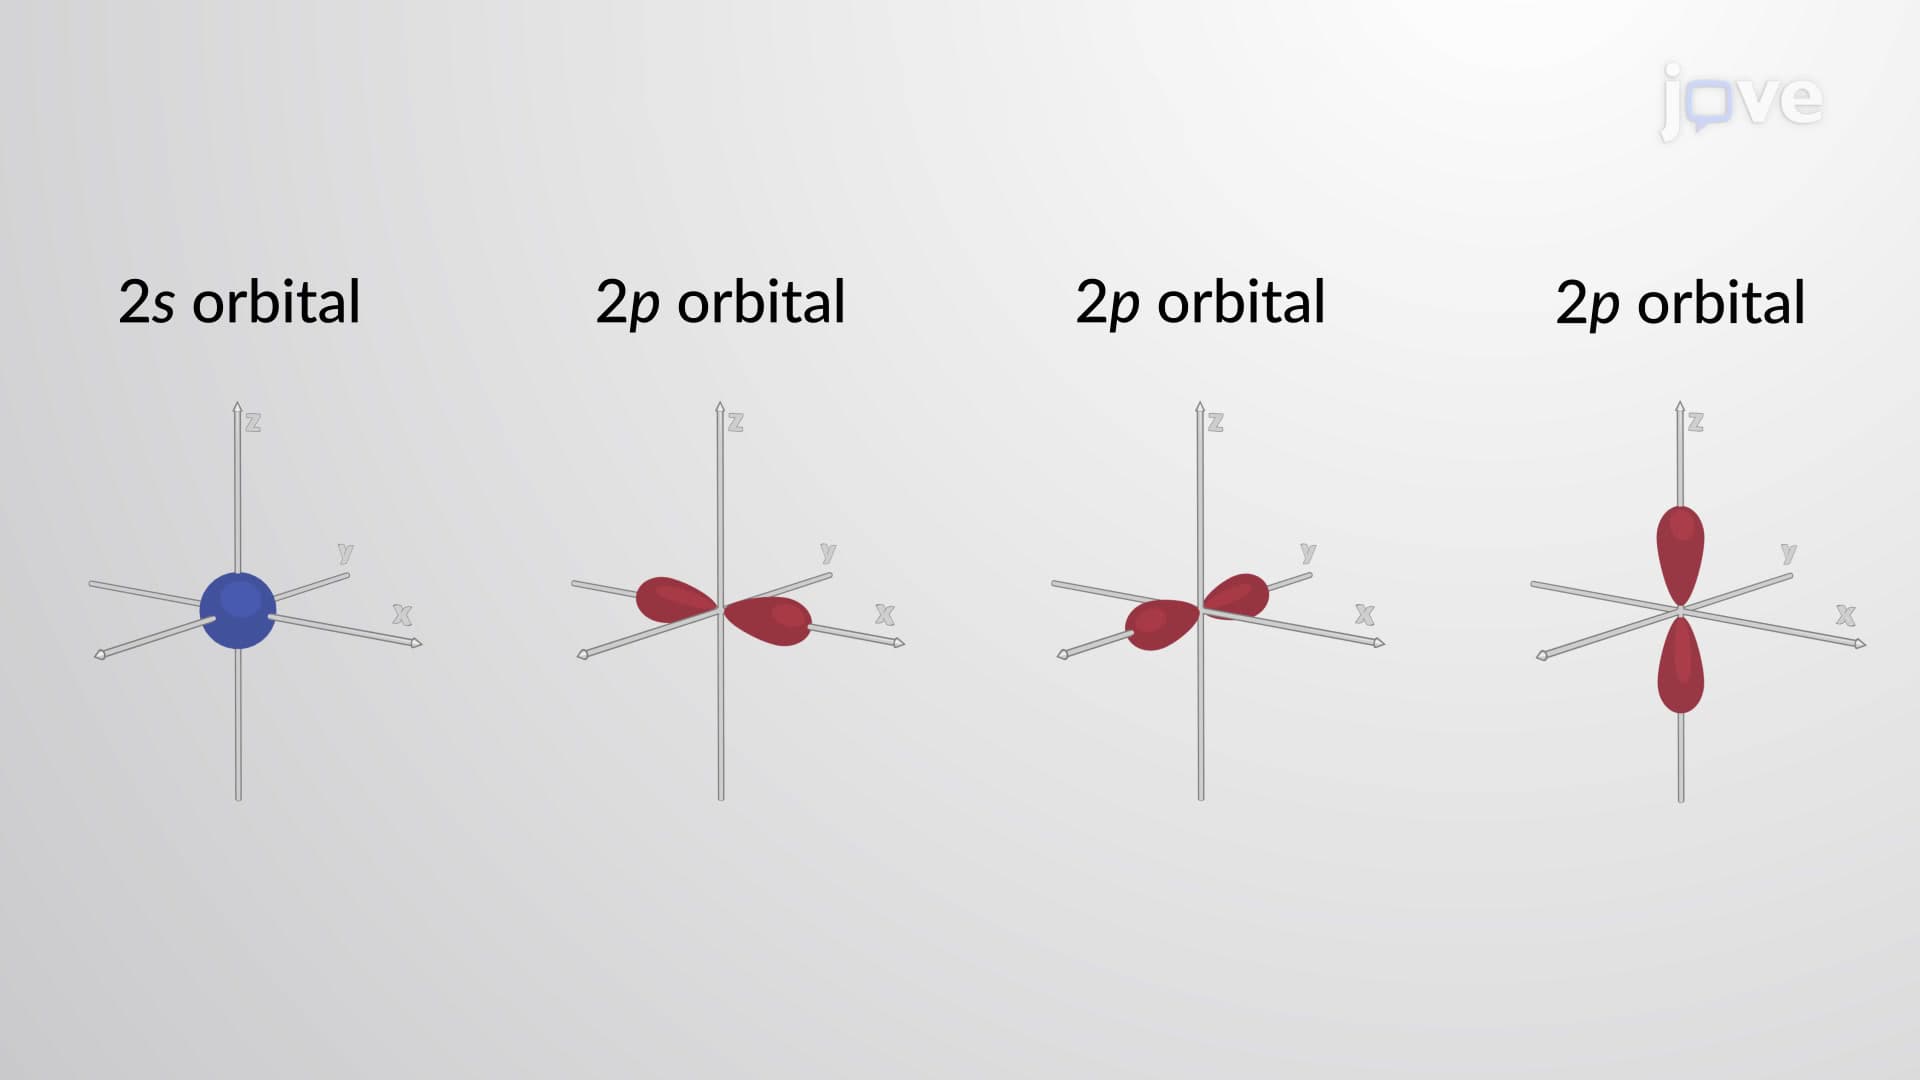
Electron Orbital Model
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
 02:18
The Bohr Model