Related Experiment Video
Updated: May 27, 2025
Quantifying the Binding Interactions Between CuII and Peptide Residues in the Presence and Absence of Chromophores
Published on: April 5, 2022
Predicting Reduction Potentials of Blue Copper Proteins Using Quantum Mechanical Calculations
Maryam Haji Dehabadi1, Mehdi Irani1, Ulf Ryde2
1Department of Chemistry, University of Kurdistan, Sanandaj 66177-15175, Iran.

Abstract:
We have calculated redox potentials of 12 blue copper protein sites comparing 64 computational methods, systematically varying the quantum mechanics (QM) system size, dielectric constants, density functional, and basis sets. All methods were based on structures optimized with combined QM and molecular mechanics (QM/MM) approaches. The redox potentials were evaluated using 10 quality metrics. The best results for relative potentials were achieved using a QM system of intermediate size (∼70 atoms), the TPSS density functional, and a SV(P) basis set, using QM-cluster calculations in a continuum solvent with a dielectric constant of 20, yielding a mean absolute deviation of 0.09 V and a maximum deviation of 0.26 V. For absolute redox potentials, methods using larger QM systems (∼340 atoms), the B3LYP density functional, and larger basis sets perform better, achieving mean signed errors down to -0.27 V. Compared to previous studies on iron-sulfur clusters, redox potentials for blue copper proteins show improved accuracy due to their narrower potential range and simpler coordination environments, but systematic errors are system-dependent. This study underscores the challenges of modeling redox-active sites in proteins and highlights the effectiveness of QM-cluster calculations in a continuum solvent in balancing computational cost with predictive power.
More Related Videos
 14:44
Structure and Coordination Determination of Peptide-metal Complexes Using 1D and 2D 1H NMR
Published on: December 16, 2013
 06:53
Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Related Concept Videos
 03:02
Standard Electrode Potentials
 01:30
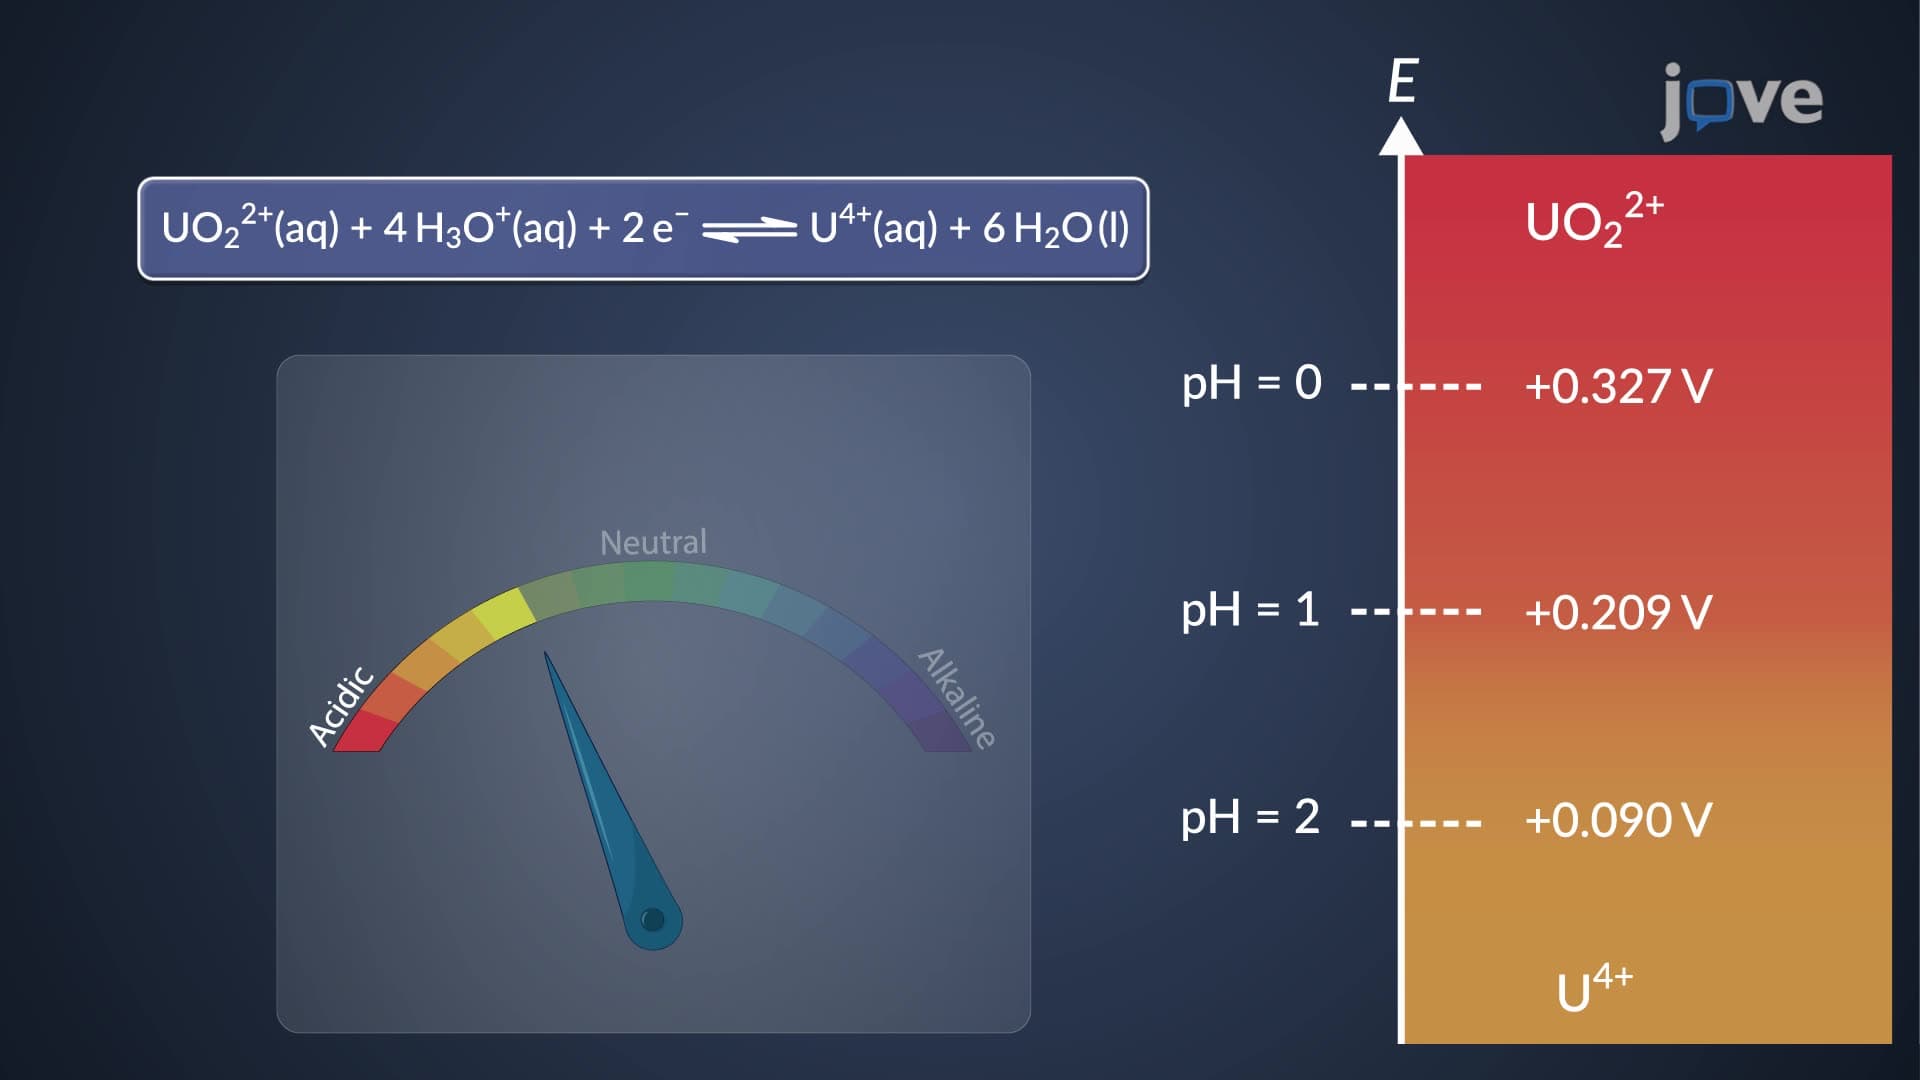
Ladder Diagrams: Redox Equilibria
Consider the Fe3+/Fe2+ half-reaction, which has a standard-state potential of +0.771 V. At potentials more positive than +0.771 V, Fe3+ predominates, whereas Fe2+...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 00:56
Extraction: Advanced Methods
 01:08
Electrodeposition
Electrodeposition can...